Local alternatives to Becke weights for molecular quadrature
Related Experiment Video
Updated: Nov 3, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Peter M W Gill1, Kosta D Tsoukalas1
1School of Chemistry, University of Sydney, Sydney, NSW 2006, Australia.

Abstract:
Most density functional theory (DFT) calculations employ Becke's methodology to partition the space in and around an n-atom molecule into n overlapping quasi-atomic regions. In this way, a single molecular integral for the energy (or other property) is reduced to a sum of n atomic integrals, each of which can then be approximated using a local grid in spherical polar coordinates. Unfortunately, the formal cost of Becke's partition is O(n2N), where N is the total number of grid points, and this can become a computational bottleneck when applied to large molecules. We propose two modifications of Becke's approach, each of which is formally exact and depends only on knowledge of the Delaunay neighbors of each nucleus. The two modifications can be used separately to produce schemes whose costs are O(nN) or together to produce a scheme whose cost is only O(N). Numerical tests for a variety of molecules and atomic grids suggest that the O(N) scheme should be implemented in all DFT software packages.
Related Concept Videos
 01:08
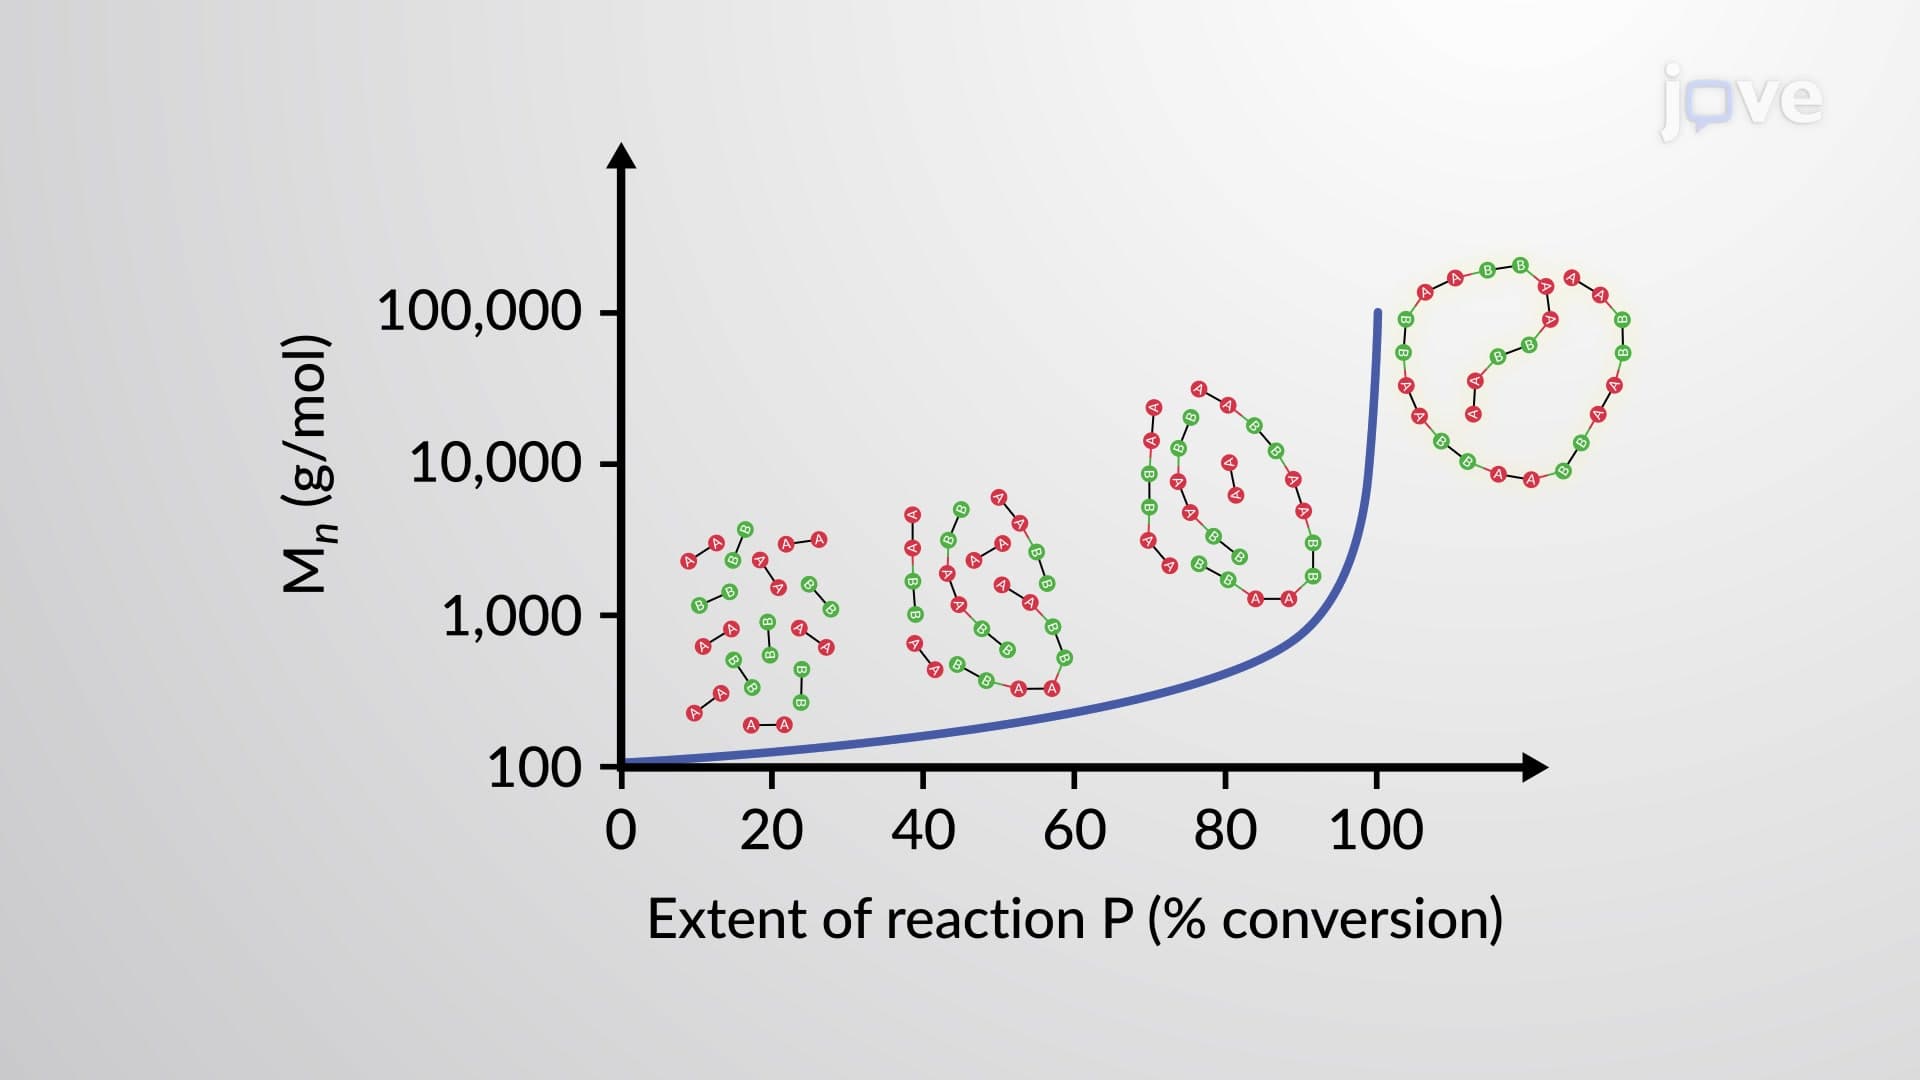
Molecular Weight of Step-Growth Polymers
As the step-growth polymerization involves step-wise condensation of monomers, the molecular weight also builds up eventually. Consequently, high molecular weight polymers are obtained at the late stages of the polymerization, where 99% of monomers have been consumed.
The extent of the...
 01:27
Determination of Molar Masses of Polymers II
 01:15
High-Resolution Mass Spectrometry (HRMS)
 01:19
Mass Analyzers: Common Types
 01:01
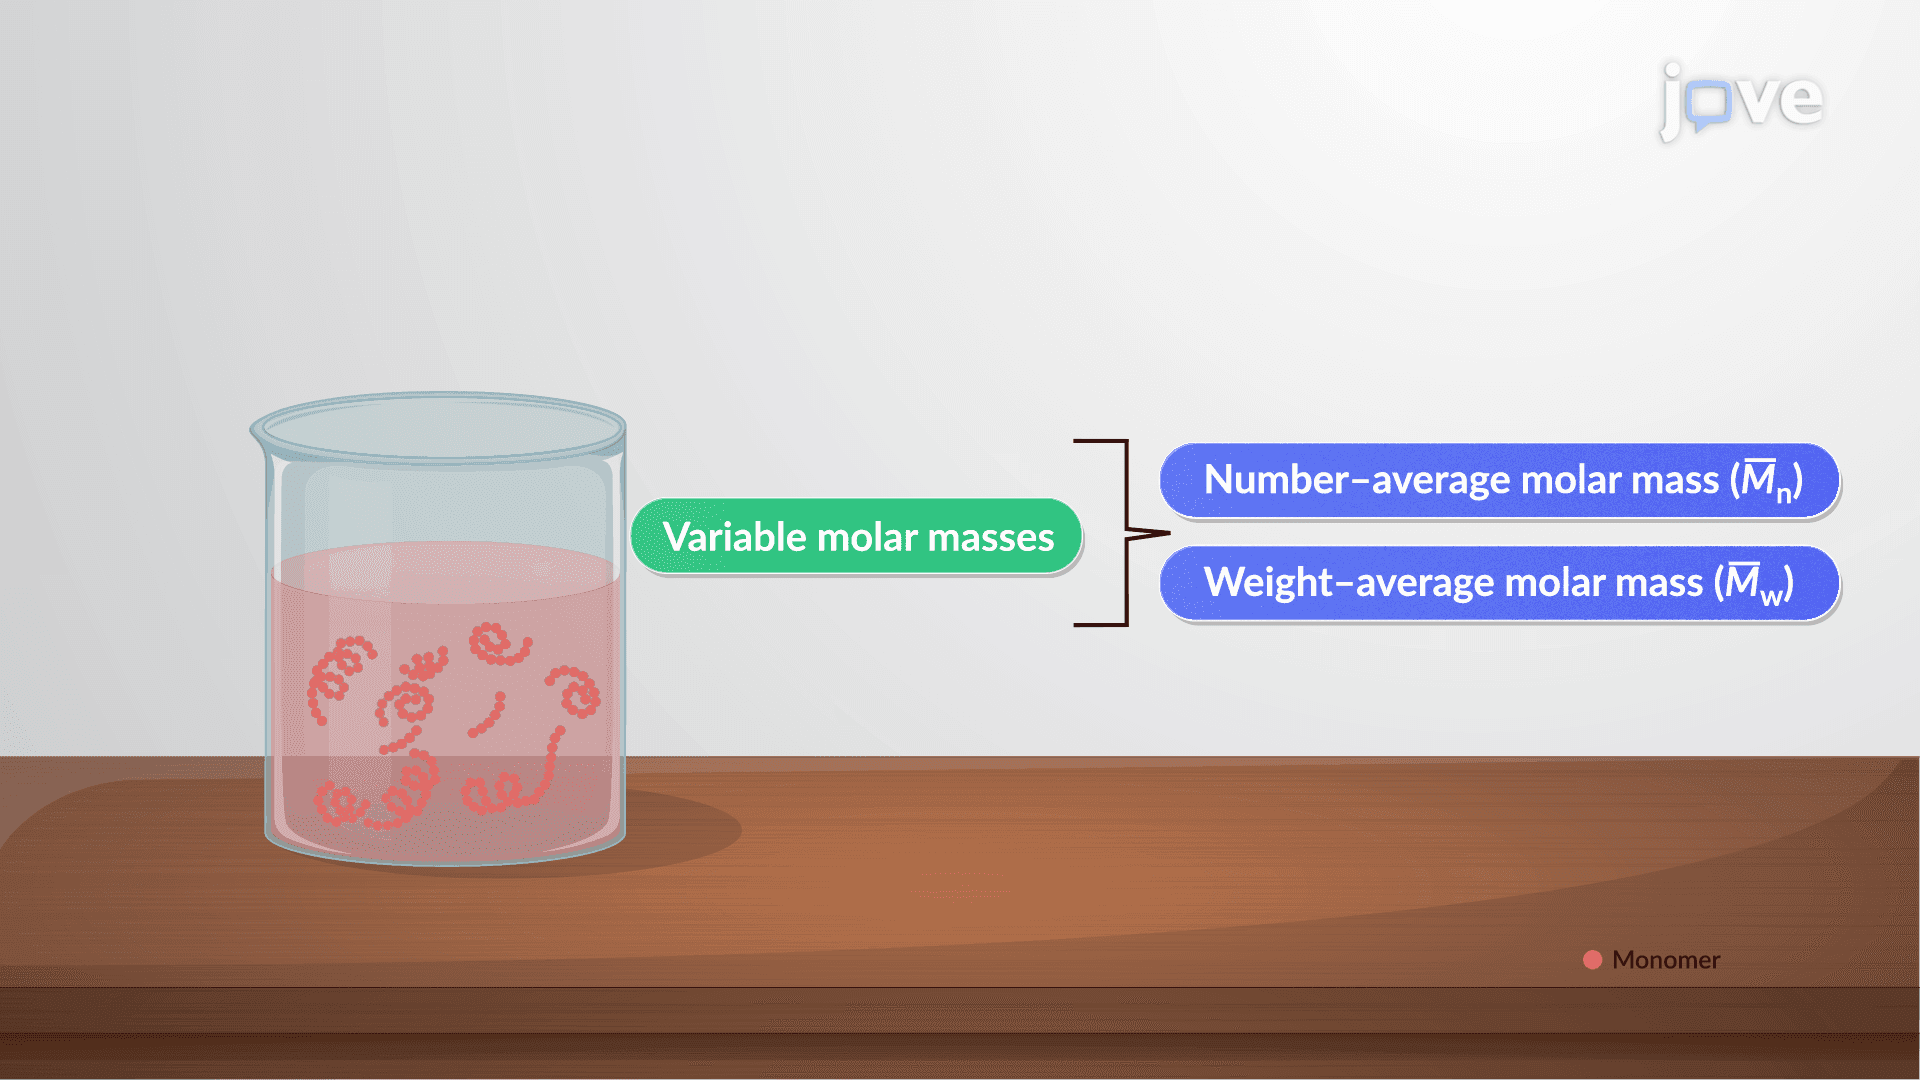
Polymers: Defining Molecular Weight
The number average molecular weight (Mn) is the summation of the number...
 01:24
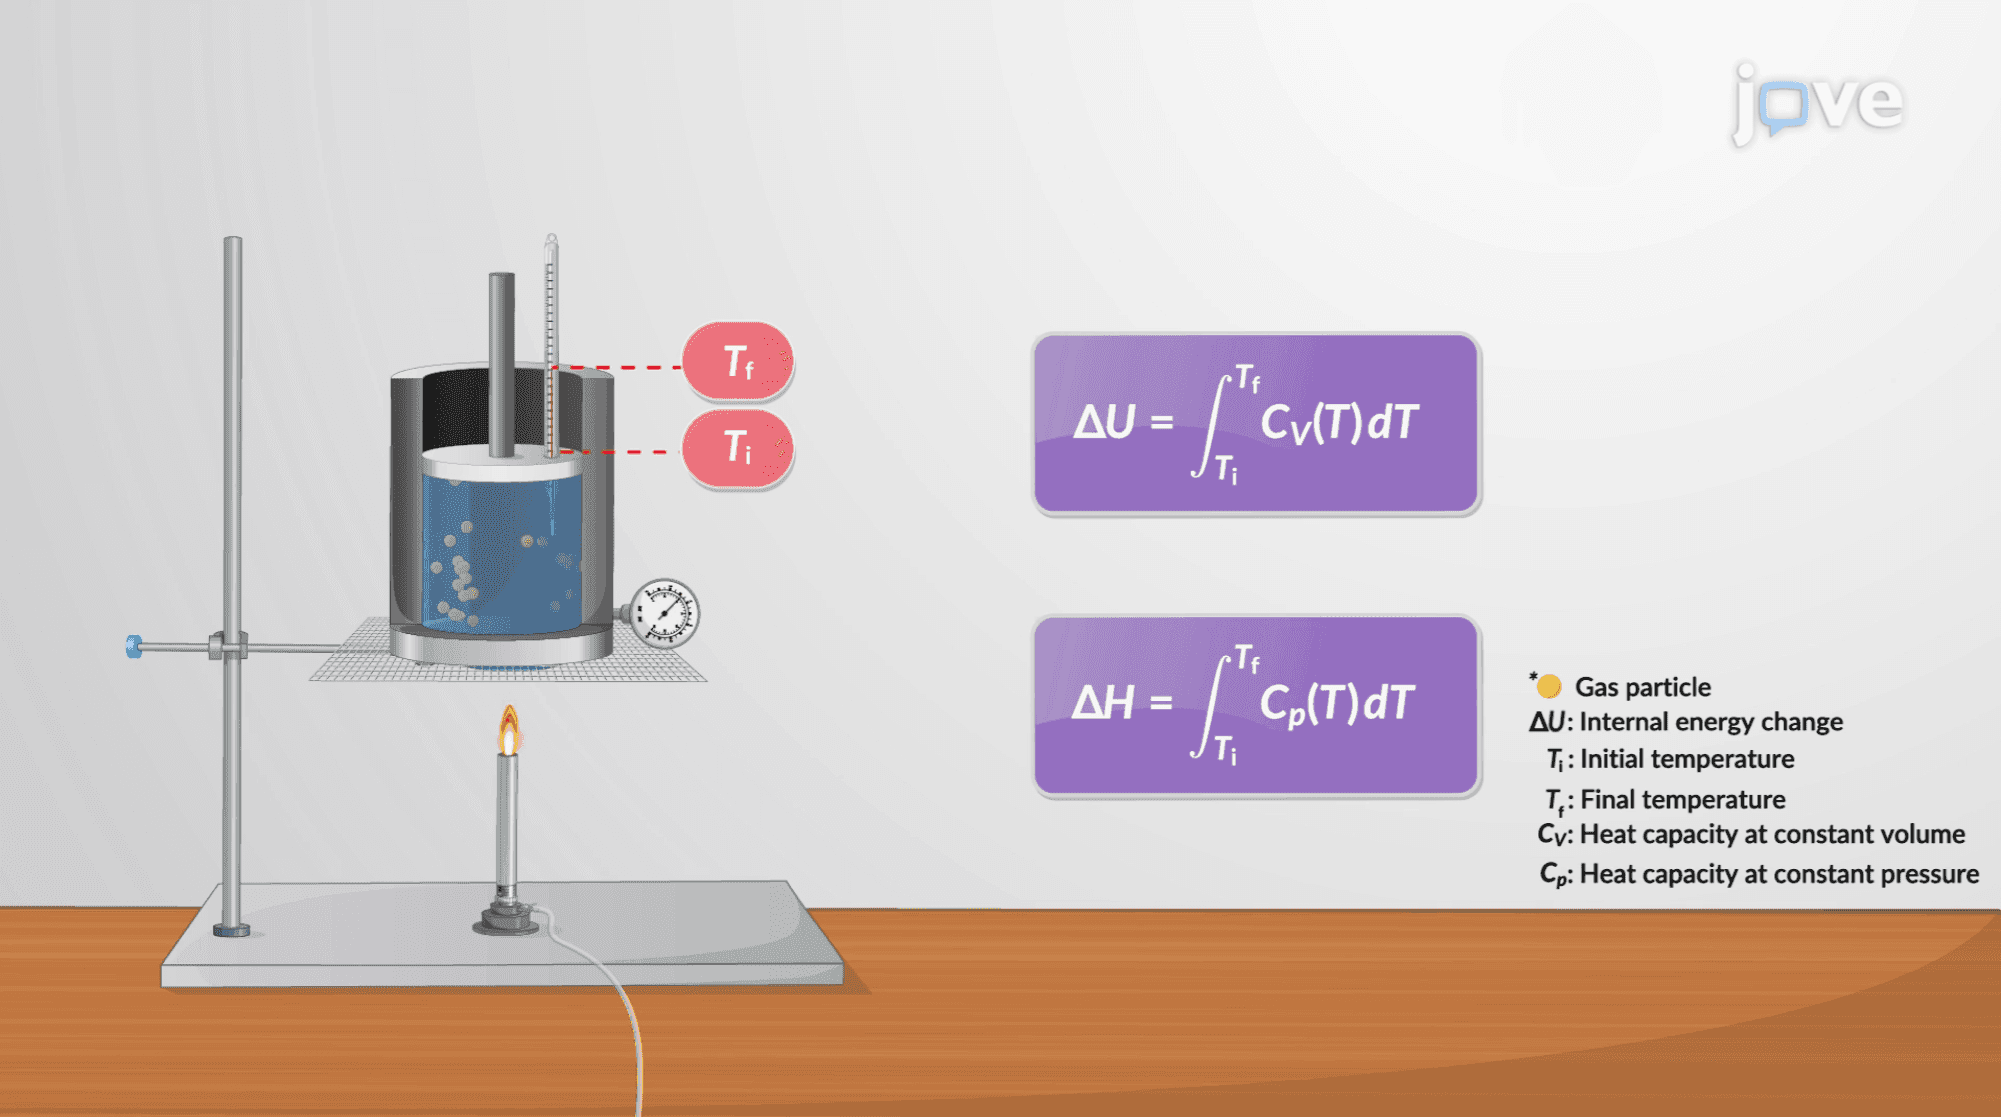
Calculation of First-Law Quantities II