Accurate ab initio structure and vibration-rotation dynamics of the beryllium dimer
1Department of Chemistry, Adam Mickiewicz University, 61-614 Poznań, Poland.

Abstract:
The accurate potential energy function of the beryllium dimer in its ground electronic state has been obtained. The valence-electron correlation energy was calculated using the coupled-cluster approach, up to the CCSDTQ level of approximation (equivalent in this case to the full-configuration-interaction level of theory) with the augmented correlation-consistent basis sets as large as aug-cc-pV10Z. The core-electron correlation effects were estimated up to the CCSDTQ level using the core-valence basis sets as large as aug-cc-pCV7Z. The scalar relativistic, adiabatic, and non-adiabatic effects on spectroscopic constants of Be2 were also discussed. In particular, the adiabatic binding energy De of Be2 was predicted without invoking any extrapolation procedures to be 935 cm-1, with an uncertainty of ±2 cm-1. The vibrational term values and the effective rotational constants of Be2 were predicted to be within the corresponding experimental error bars.
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 02:52
VSEPR Theory and the Basic Shapes
 01:18
Structure of Benzene: Molecular Orbital Model
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:27
Predicting Molecular Geometry
 01:07
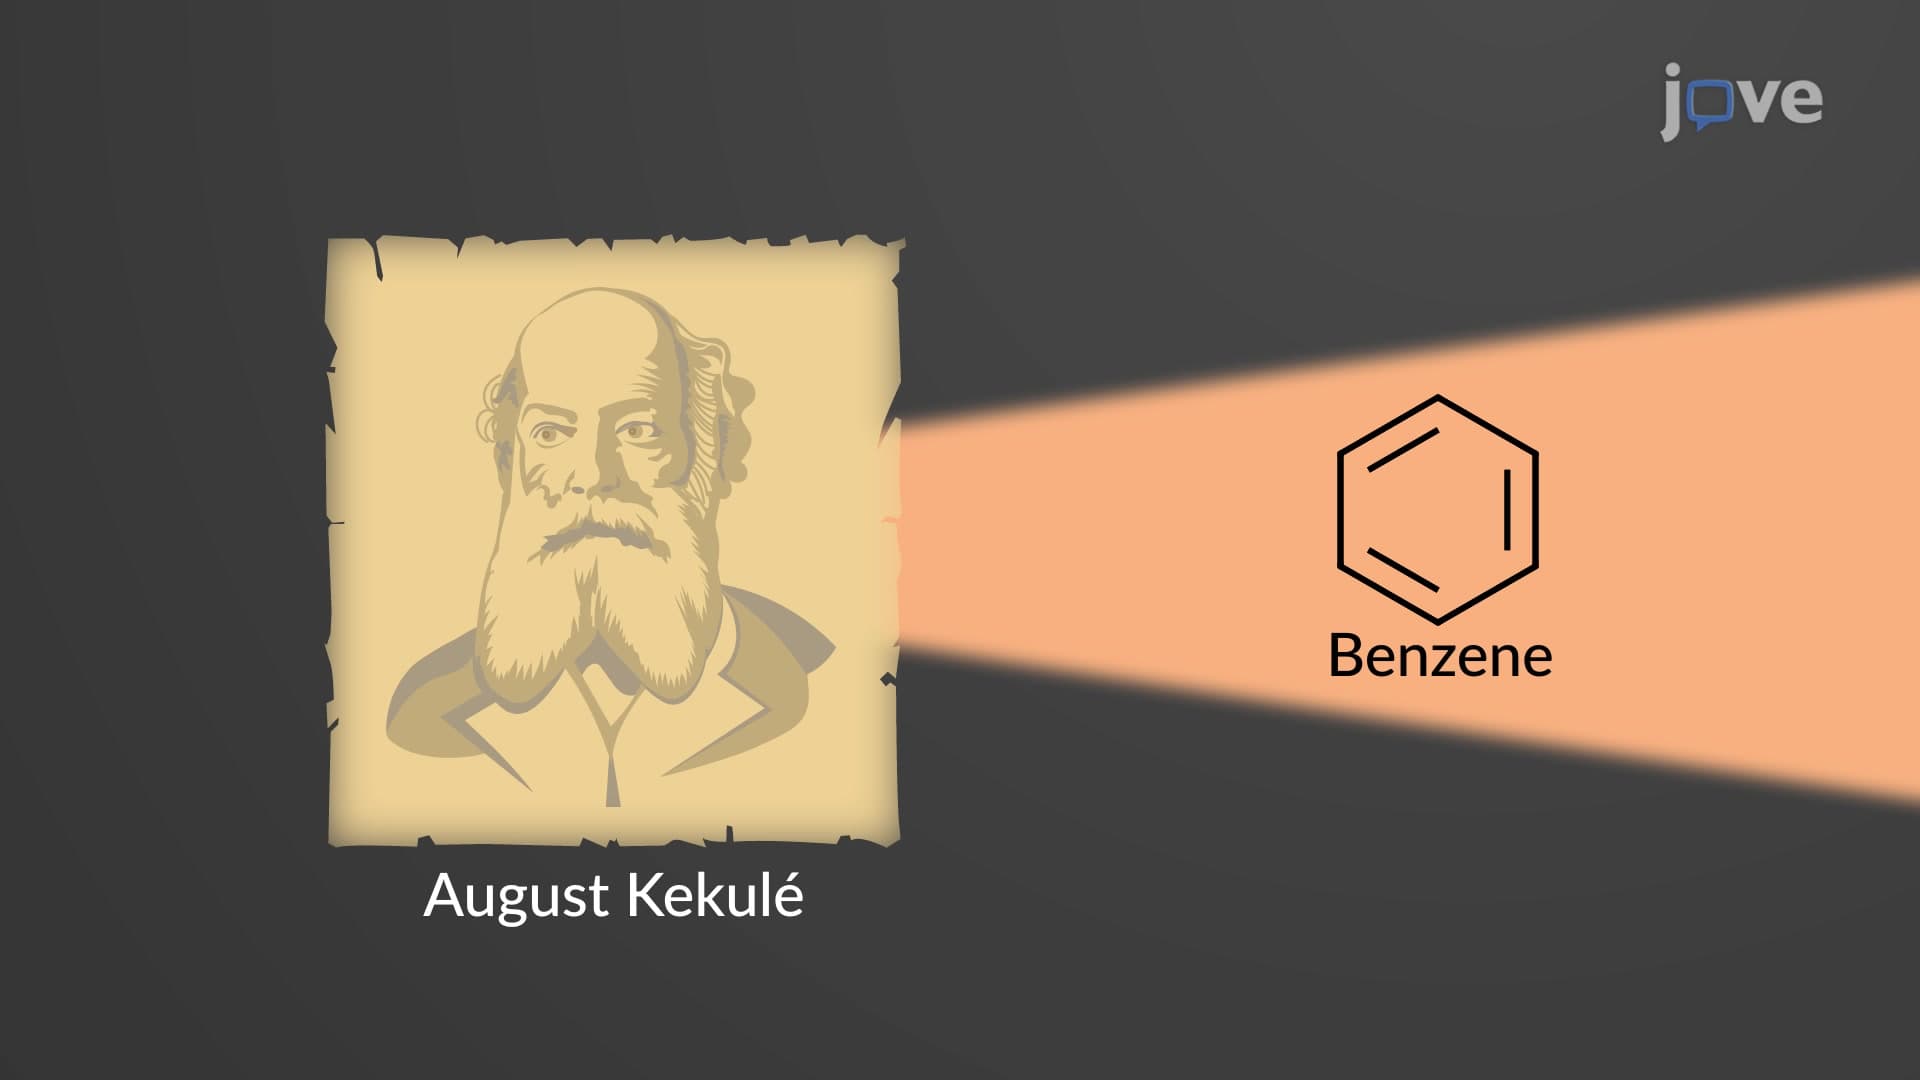
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.