DFT study of actinide dimers (An2, An = Ac-Lr): the role of dispersion interactions
Related Experiment Video
Updated: Nov 3, 2026
Controlling the Size, Shape and Stability of Supramolecular Polymers in Water
Published on: August 2, 2012
1Nikolaev Institute of Inorganic Chemistry, Siberian Branch of Russian Academy of Science, Novosibirsk, Russia.

Abstract:
A systematic quantum chemical study of the complete series of homonuclear actinide dimers (An2, where An = Ac-Lr) was carried out using noncollinear unrestricted spin-orbit relativistic density functional theory within the zeroth-order regular approximation. This work focuses on the evolution of chemical bonding across the 5f series and, in particular, on the potentially critical role of dispersion interactions in determining the structural parameters of heavy actinide dimers. For the early actinides (Ac2-Pu2), the bonding is predominantly covalent, and dispersion corrections have negligible impact on equilibrium geometries. In contrast, the middle actinides (Am2, Bk2-No2) exhibit a transition to weakly bound van der Waals systems, where the inclusion of Grimme's D3(BJ) dispersion correction appears to be a major factor, shortening bond lengths. Notably, Cm2 and Lr2 demonstrate a reversion to non-dispersion-dominated, driven by the involvement of 6d and 7p valence electrons, respectively. The structural trends are additionally analyzed by a topological analysis of the electron density within the quantum theory of atoms in molecules framework. The obtained results reveal a complex, non-monotonic alternation of bondings across the actinide series and highlight the importance of accounting for dispersion forces when modeling heavy element systems with localized 5f electrons.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 07:24
Hyperspectral Imaging as a Tool to Study Optical Anisotropy in Lanthanide-Based Molecular Single Crystals
Published on: April 14, 2020
Related Concept Videos
 01:24
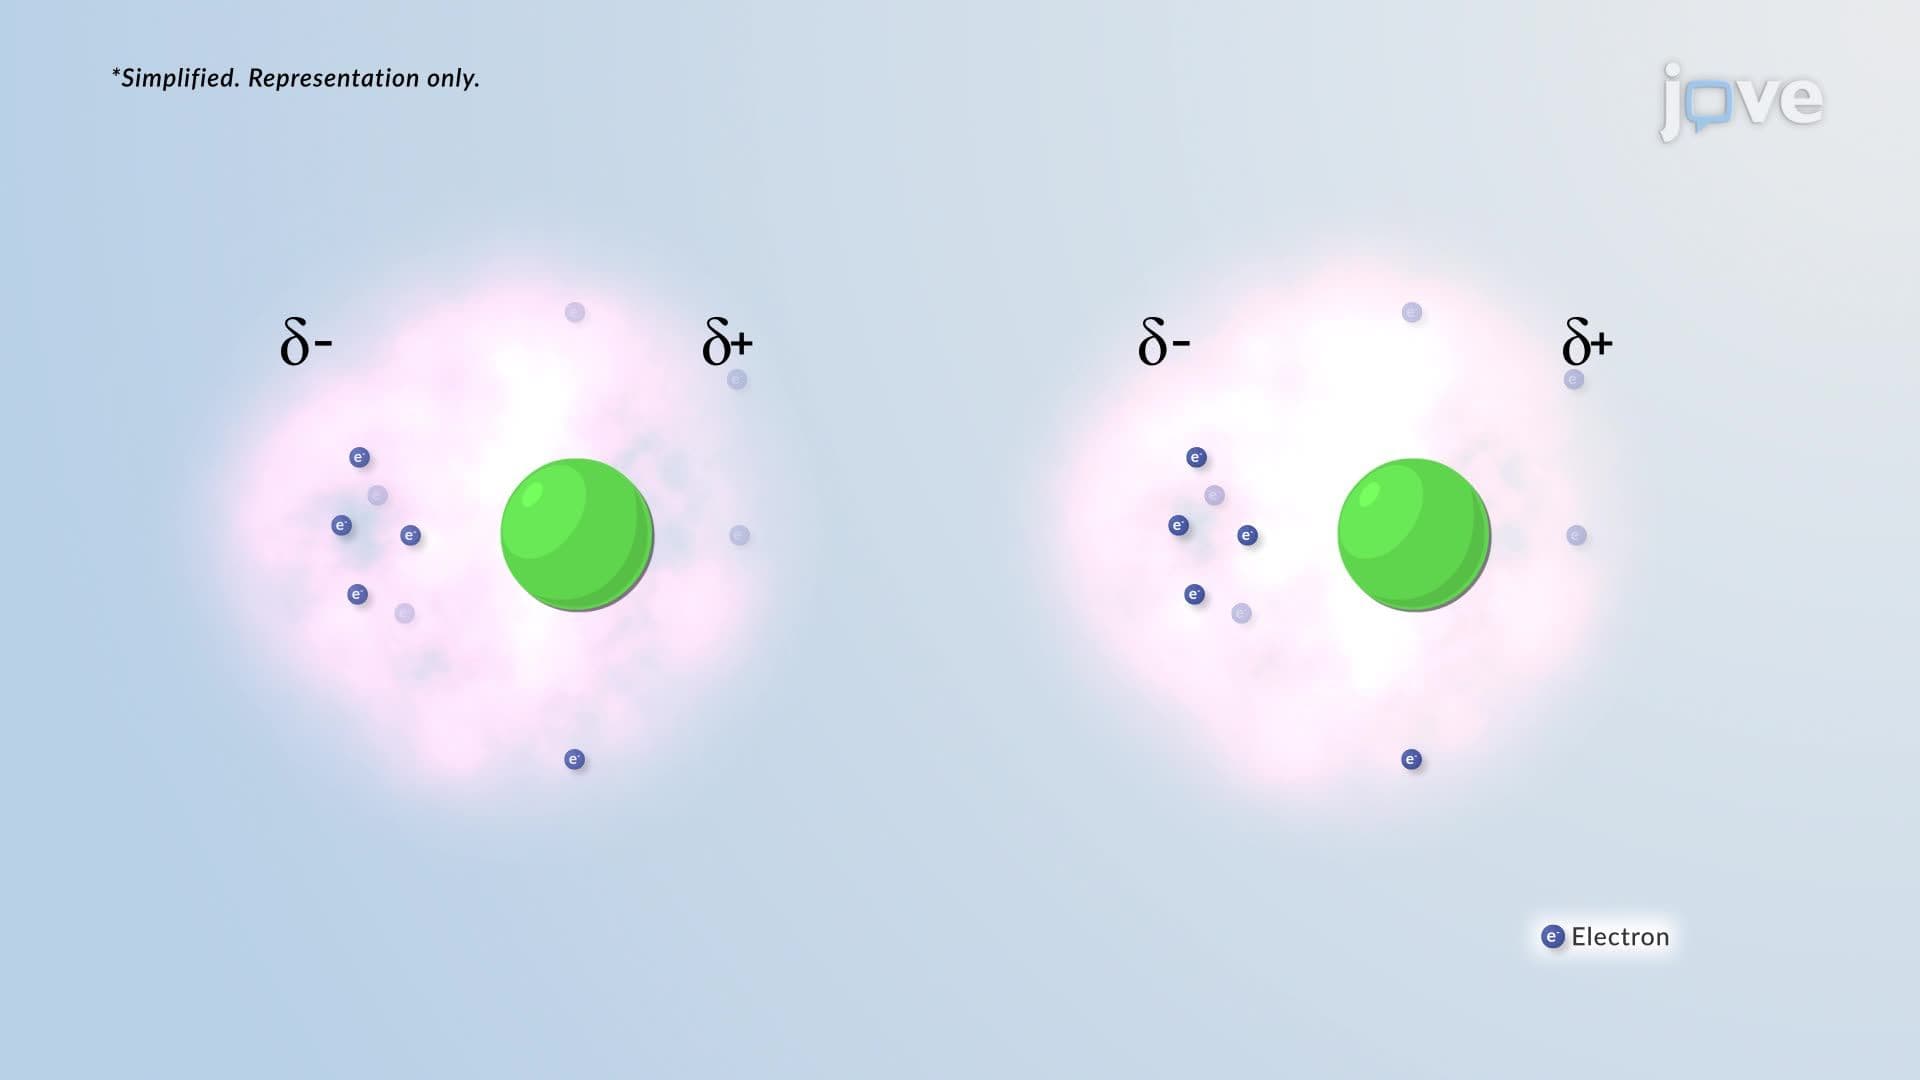
Van der Waals Interactions
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 03:13
Intermolecular Forces
 03:51
Molecular Orbital Theory II