Related Experiment Video
Updated: Oct 13, 2026
Expression and Purification of the Cystic Fibrosis Transmembrane Conductance Regulator Protein in Saccharomyces cerevisiae
Published on: March 10, 2012
Trypsin binding activity of alpha 2-macroglobulin in cystic fibrosis and other lung diseases

Abstract:
Abnormalities of the interaction between alpha 2-macroglobulin and proteases have been suspected in cystic fibrosis. We measured the binding activity for trypsin of alpha 2-macroglobulin in 65 patients with cystic fibrosis, 41 obligate heterozygotes for cystic fibrosis, 18 children with asthma, 21 adult patients with chronic obstructive pulmonary disease, and 21 healthy control subjects. The assay was based on the observation that, once it is bound to alpha 2-macroglobulin, trypsin is no longer inhibited by soybean trypsin inhibitor. The bound trypsin retains activity against synthetic substrates such as N-alpha-benzoyl-arginine-p-nitroanilide. We found a moderately increased molar binding ratio of trypsin to alpha 2-macroglobulin in all patient groups and heterozygotes compared to healthy control subjects. The absolute concentration of alpha 1-macroglobulin was related to age rather than to the disease of the patient. Our data argue against a defect in protease-binding activity of alpha 2-macroglobulin in cystic fibrosis. The moderately increased binding activity observed could have been related to a structural difference in alpha 2-macroglobulin or to binding of ligands. This finding however, was not unique to patients with cystic fibrosis; the binding activity was increased in heterozygotes and in patients with other pulmonary diseases.
Insights
This study found no defect in alpha 2-macroglobulin protease-binding in cystic fibrosis patients. Increased binding activity was observed in cystic fibrosis, heterozygotes, and other lung diseases, suggesting a non-specific effect.
Area of Science:
- Biochemistry
- Pulmonary Medicine
- Genetics
Background:
- Abnormal interactions between alpha 2-macroglobulin and proteases are implicated in cystic fibrosis (CF).
- Alpha 2-macroglobulin is a key protease inhibitor involved in regulating proteolytic activity.
- Understanding these interactions is crucial for CF pathogenesis research.
Purpose of the Study:
- To investigate the protease-binding activity of alpha 2-macroglobulin in patients with cystic fibrosis.
- To compare binding activity in CF patients, heterozygotes, and individuals with other pulmonary conditions.
- To determine if a defect in alpha 2-macroglobulin function contributes to cystic fibrosis.
Main Methods:
- Measured the binding activity of trypsin to alpha 2-macroglobulin using a functional assay.
- Assayed samples from 65 cystic fibrosis patients, 41 heterozygotes, 18 children with asthma, 21 COPD patients, and 21 healthy controls.
- Utilized the substrate N-alpha-benzoyl-arginine-p-nitroanilide to quantify bound trypsin activity.
Main Results:
- A moderately increased molar binding ratio of trypsin to alpha 2-macroglobulin was observed in all patient groups and heterozygotes compared to controls.
- Absolute alpha 2-macroglobulin concentration correlated with age, not disease status.
- The increased binding activity was not specific to cystic fibrosis, also appearing in heterozygotes and other pulmonary diseases.
Conclusions:
- The study provides evidence against a defect in alpha 2-macroglobulin protease-binding activity in cystic fibrosis.
- The observed increase in binding activity may be due to structural variations or ligand binding, not a CF-specific defect.
- Findings suggest the altered binding is a common response in various pulmonary conditions and genetic carriers.
Related Concept Videos
 01:13
Enzyme-linked Receptors
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:23
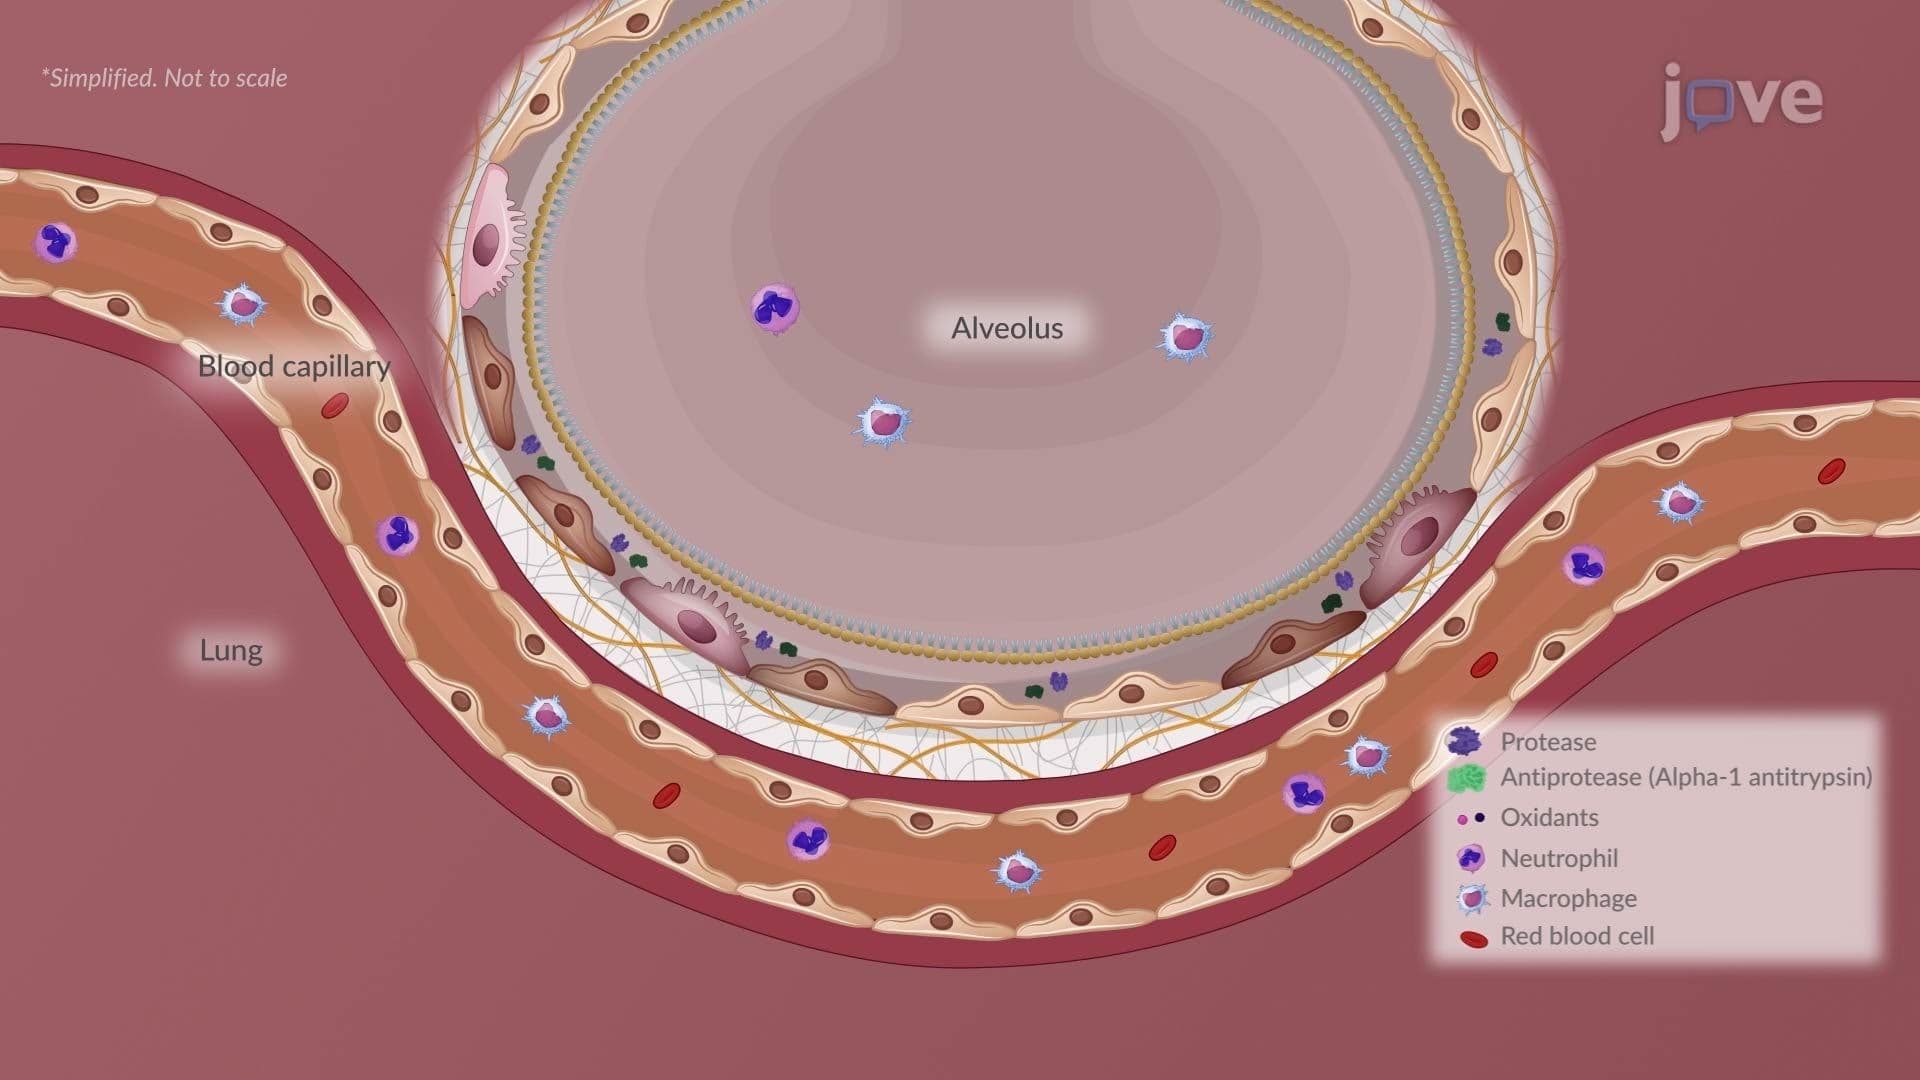
Chronic Obstructive Pulmonary Disease II: Emphysema