Protein binding displacement interactions and their clinical importance

Abstract:
The binding of drugs to proteins is an important pharmacokinetic parameter. Many methods are available for the study of drug protein binding phenomena and there are also many ways to interpret the binding data. Although much emphasis has been placed on the binding of drugs in the plasma, binding also takes place in the tissues. Displacement interactions involving plasma or tissue binding sites have been implicated as the causative mechanisms in many drug interactions. However, the importance of plasma binding displacement as a mechanism of drug interactions. However, the importance of plasma binding displacement as a mechanism of drug interaction has been overestimated and overstated, being based largely on in vitro data. Because displaced drug can normally distribute out of the plasma compartment, increases of free drug concentrations are usually transient and therefore will not give rise to changed pharmacological effects in the patient. Those clinically important drug interactions formerly considered to be caused via displacement from plasma binding sites usually have another interaction mechanism involved; commonly decreased metabolism or renal elimination also takes place. Plasma binding displacement interactions, however, do become important clinically in certain specific situations, namely, when the displacing drug is administered quickly to the patient by the intravenous route, during therapeutic drug monitoring, and in certain drug disposition studies which involve the use of a heparin lock for blood sampling. Tissue binding displacement interactions have a greater potential to cause adverse effects in the patient as in this case drug will be forced from extravascular sites back into the plasma. The resulting increased drug plasma levels will lead to enhanced pharmacological effects and, possibly, frank toxicity. Displacement of drugs from binding sites simultaneously in both the plasma and in the tissues will combine the effects seen after displacement from the separate areas. Due to decreased binding in both areas, the free drug concentration in the plasma will increase leading to overactivity of the displaced drug.
Related Concept Videos
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:18
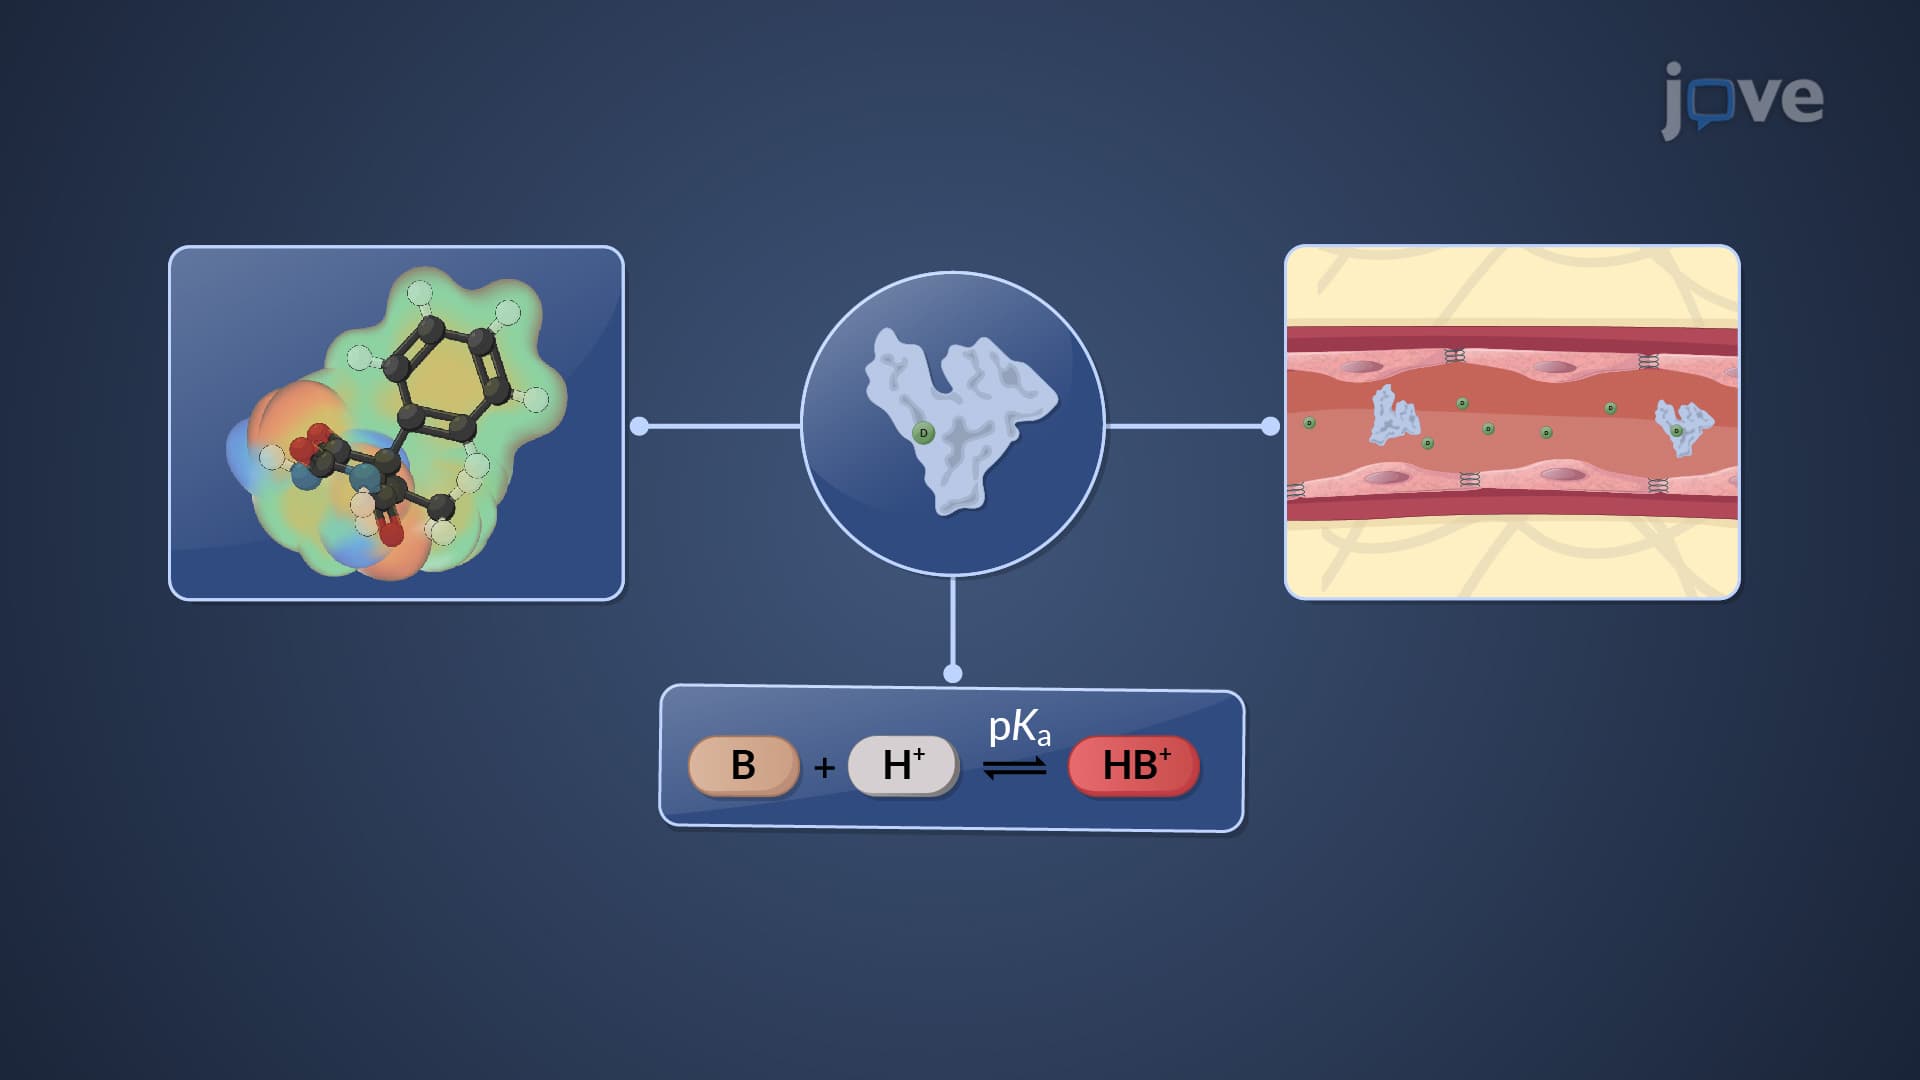
Factors Affecting Protein-Drug Binding: Drug-Related Factors
One crucial factor in drug-protein binding is the drug's lipophilicity or its affinity for fat. More lipophilic drugs tend to have higher binding extents. For example, highly lipophilic drugs like cloxacillin exhibit substantial protein binding, with as much as 95% of the drug binding to proteins. In contrast,...
 01:20
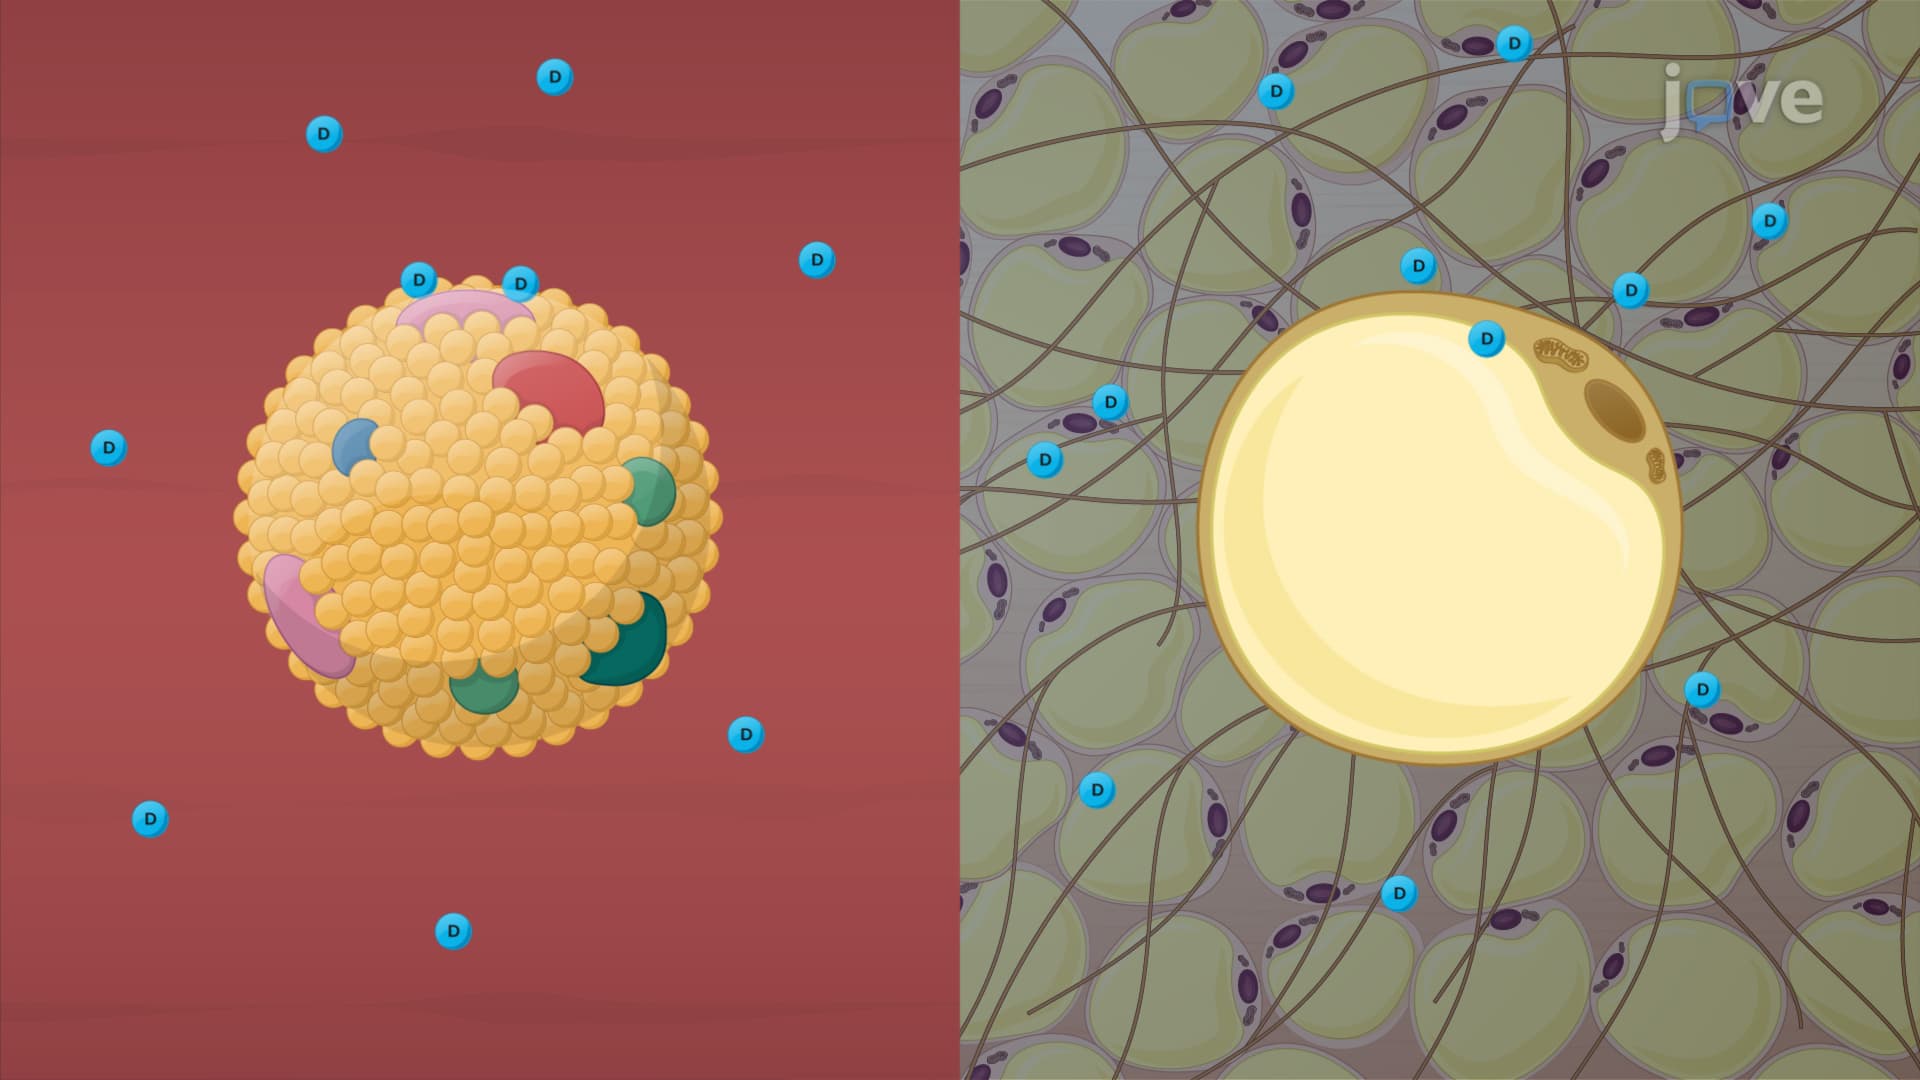
Factors Affecting Protein-Drug Binding: Protein-Related Factors
The physicochemical properties of a drug play a significant role in its ability to bind to proteins. Lipophilic drugs, which dissolve in fats, oils, and lipids, can be bound by...
 01:23
Factors Affecting Protein-Drug Binding: Drug Interactions
Displacement interactions can have varying outcomes, ranging from toxicity to virtually...
 01:29
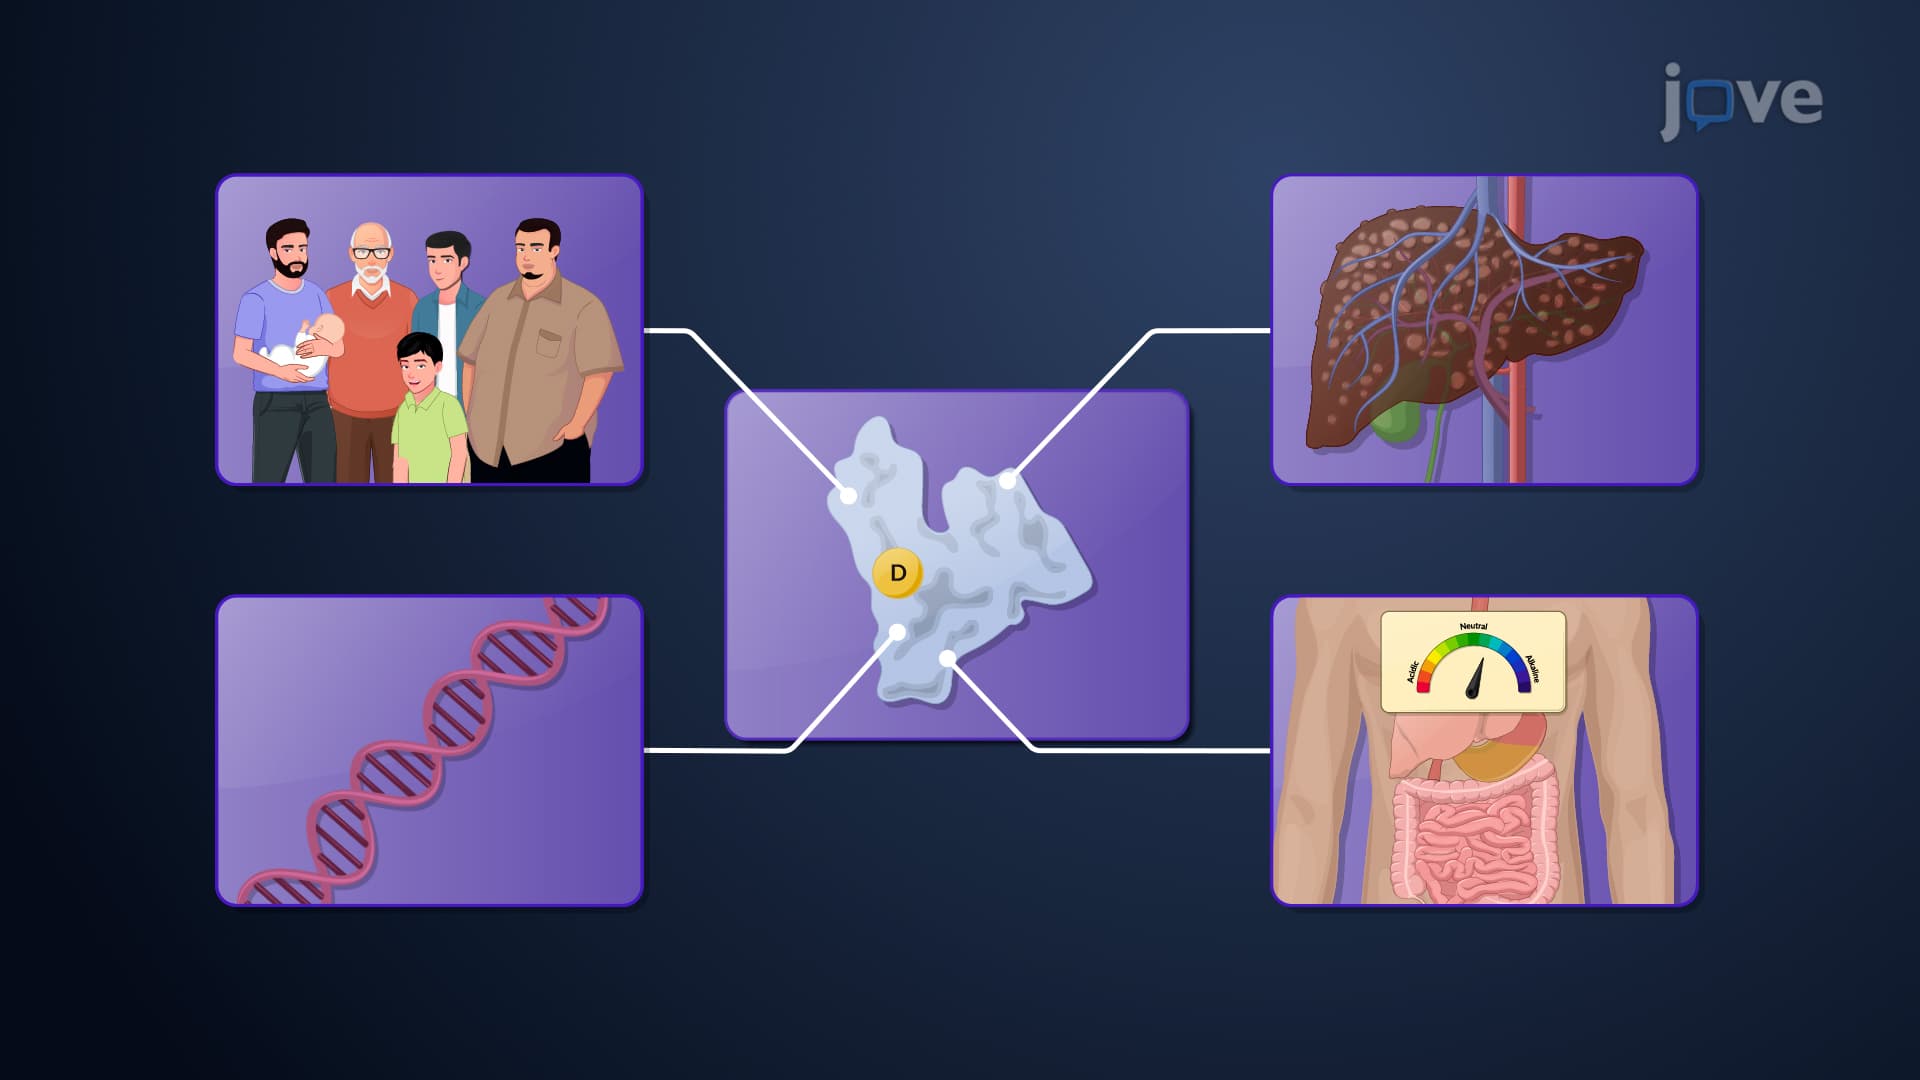
Factors Affecting Protein-Drug Binding: Patient-Related Factors
Age stands as a key determinant in protein-drug binding. Neonates, characterized by low albumin content, experience heightened concentrations of unbound drugs such as phenytoin and...