Related Experiment Video
Updated: May 30, 2026

Construction and Systematical Symmetric Studies of a Series of Supramolecular Clusters with Binary or Ternary Ammonium Triphenylacetates
Published on: February 15, 2016
Ab initio molecular orbital calculations of electron distribution in tetramethylammonium ion

Abstract:
Ab initio molecular orbital calculations of electron distribution in tetramethylammonium ion and its uncharged isoelectronic analogue, neopentane, have been carried out. Comparison of the two compounds permits a detailed description of the delocalization of the positive charge of tetramethylammonium ion. The van der Waals surface of this ion is found to be characterized by "patches" of positive charge associated with the methyl groups, interspersed with essentially neutral regions. The consequences of this nonspherical charge distribution for interaction with anions have been explored by calculations of the interaction energy of fluoride ion with tetramethylammonium ion and neopentane in several mutual orientations. The lowest-energy orientation is found to be one in which the anion approaches a "face" of the tetrahedral cation (opposite to a C--N bond direction). The origins of this preference and the electron redistribution produced by the interaction with fluoride are discussed. The tetramethylammonium ion is clearly not a featureless positively charged sphere but will have appreciable geometrical specificity in its interaction with a presumed anionic group on the acetylcholine receptor.
More Related Videos
 08:12
Low Pressure Vapor-assisted Solution Process for Tunable Band Gap Pinhole-free Methylammonium Lead Halide Perovskite Films
Published on: September 8, 2017
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 04:01
VSEPR Theory and the Effect of Lone Pairs
 03:24
Hybridization of Atomic Orbitals I
 03:51
Molecular Orbital Theory II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:18
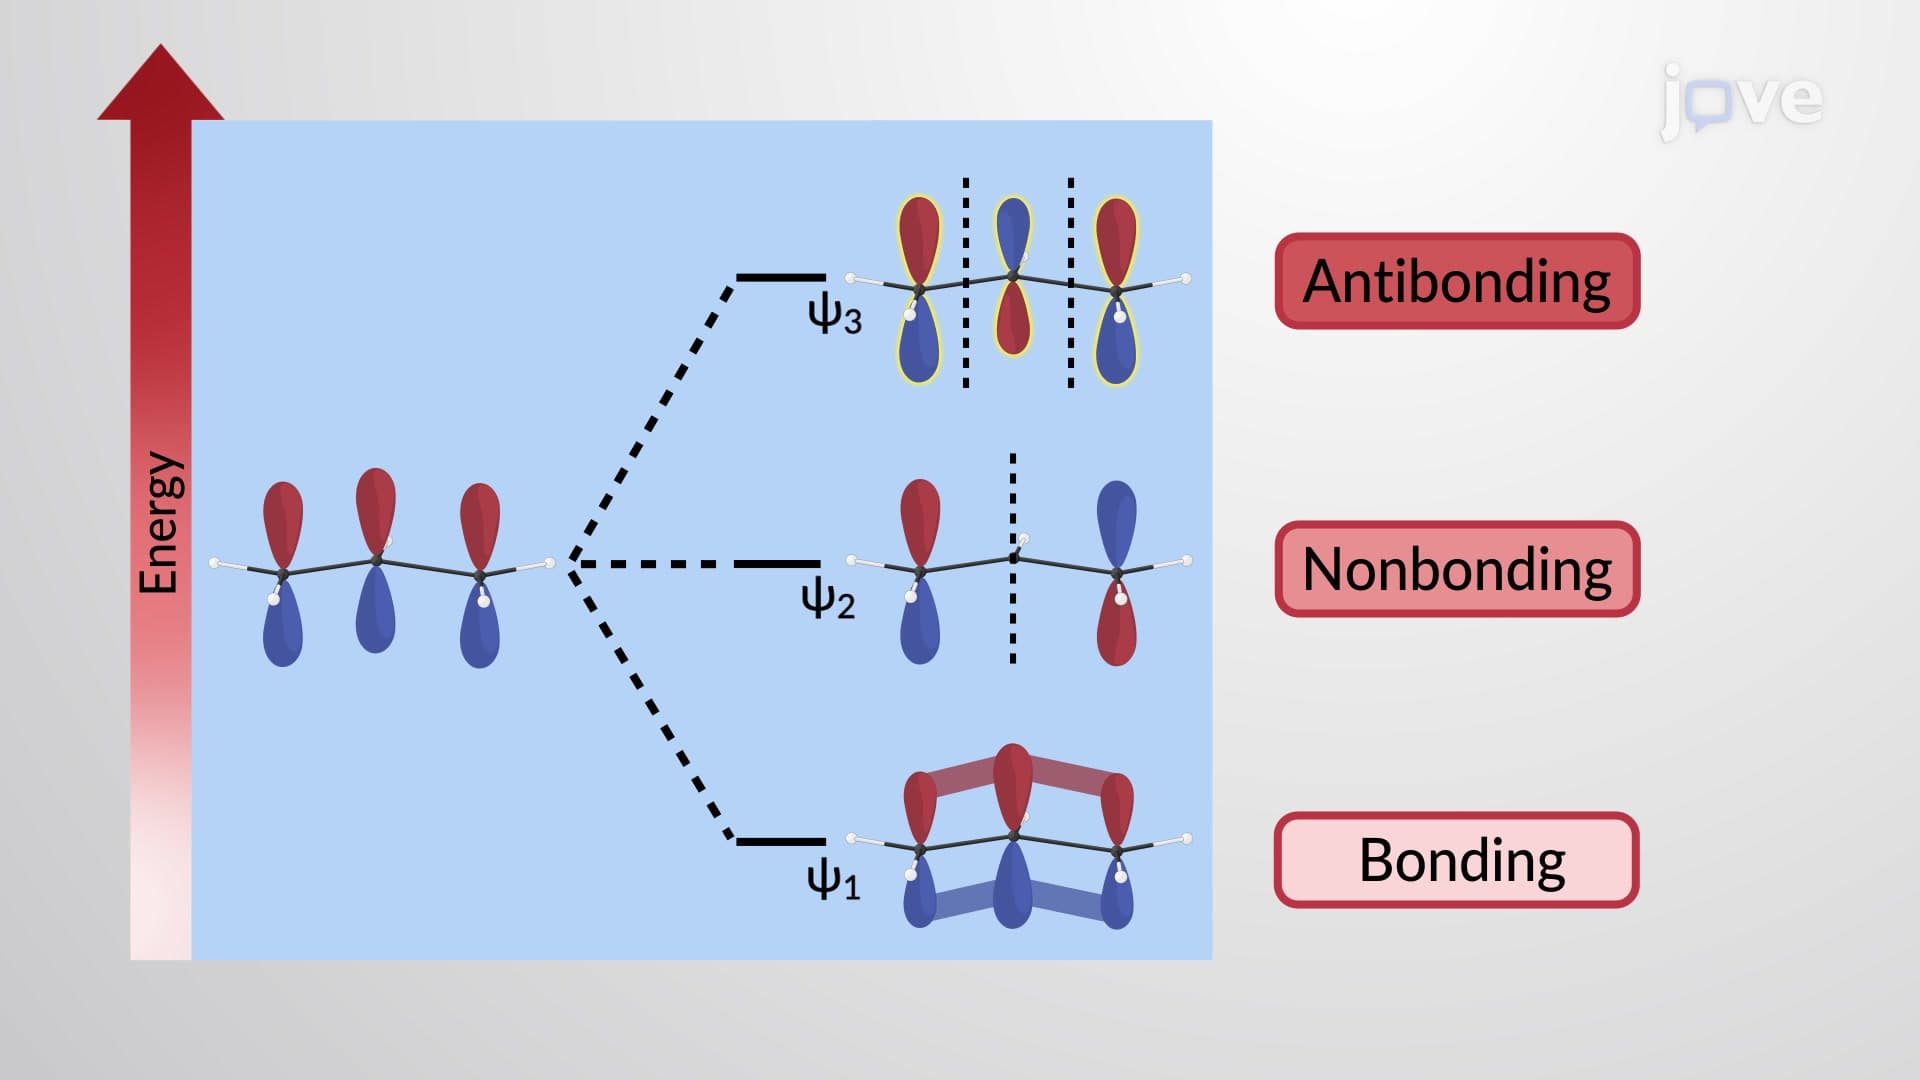
π Molecular Orbitals of the Allyl Cation and Anion
 01:19
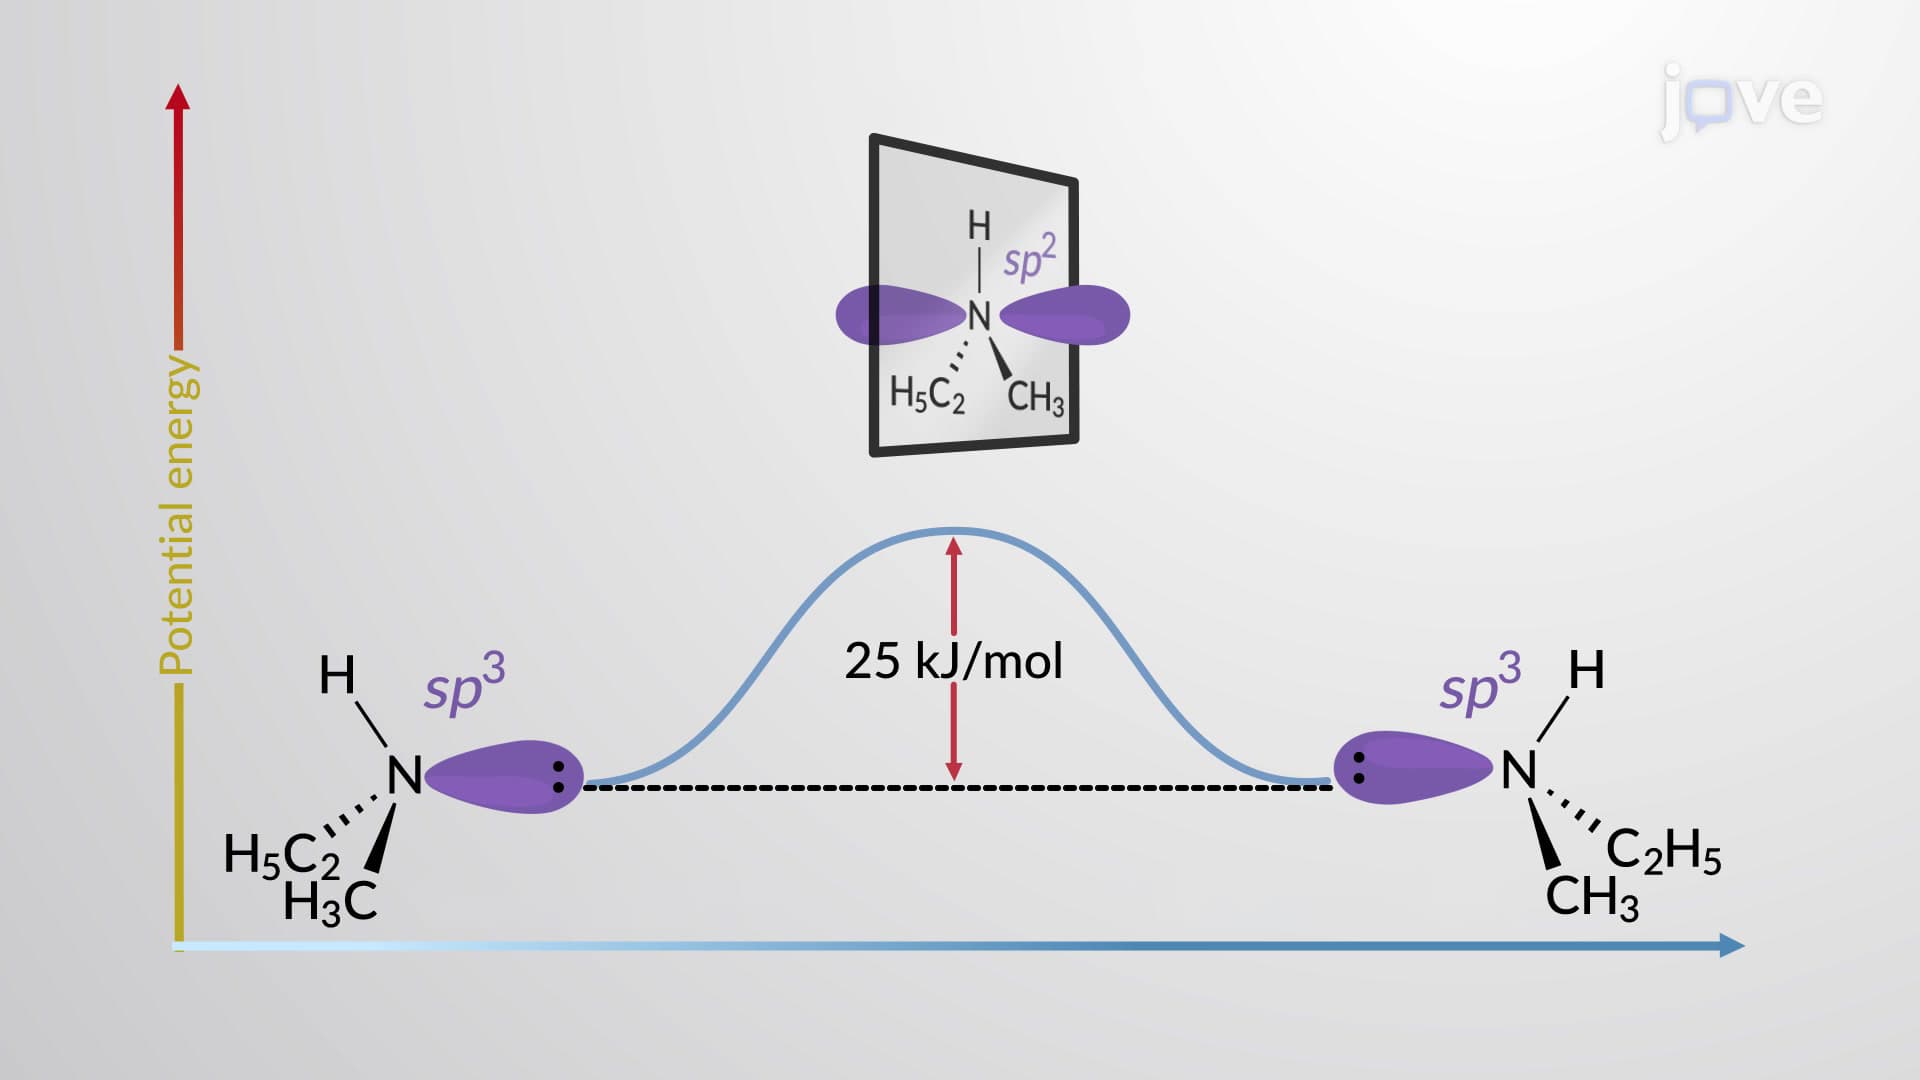
Structure of Amines