Related Experiment Video
Updated: Jul 28, 2026

Quantification of Atherosclerosis in Mice
Published on: June 12, 2019
[Familial apolipoprotein B-100 defect, a newly discovered lipid metabolism disorder]
1III. interní klinika 1. LF UK a VFN, Praha.

Abstract:
Familial defective apolipoprotein B-100 is a newly revealed genetic disorder which leads to a rise of atherogenic LDL lipoproteins. It is probably due to the replacement of a single amino acid in the huge apoliprotein B-100 molecule, i.e. substitution of glutamine by arginine in position 3,500. Thus altered LDL-lipoproteins are unable to bind with the LDL-receptor. As a result of the mentioned metabolic disorder a slightly or markedly elevated plasma cholesterol develops which very probably leads to premature manifestation of atherosclerosis. The disease is transmitted by autosomal dominant inheritance and its incidence in the population is estimated to amount to 1:500, i.e. a similar rate as familial hypercholesterolaemia. In the submitted paper some historical facts are presented which led to the detection of the disease, methods which are used for its detection, and the author presents also results of the first, so still limited clinical investigations.
Related Concept Videos
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:24
Overview of Lipid Metabolism
Lipolysis: The Breakdown of Lipids:
Lipolysis is the process of breaking down lipids, particularly triglycerides, into glycerol and fatty acids. This process typically occurs in the adipose tissue and is triggered by various hormones, including glucagon and...
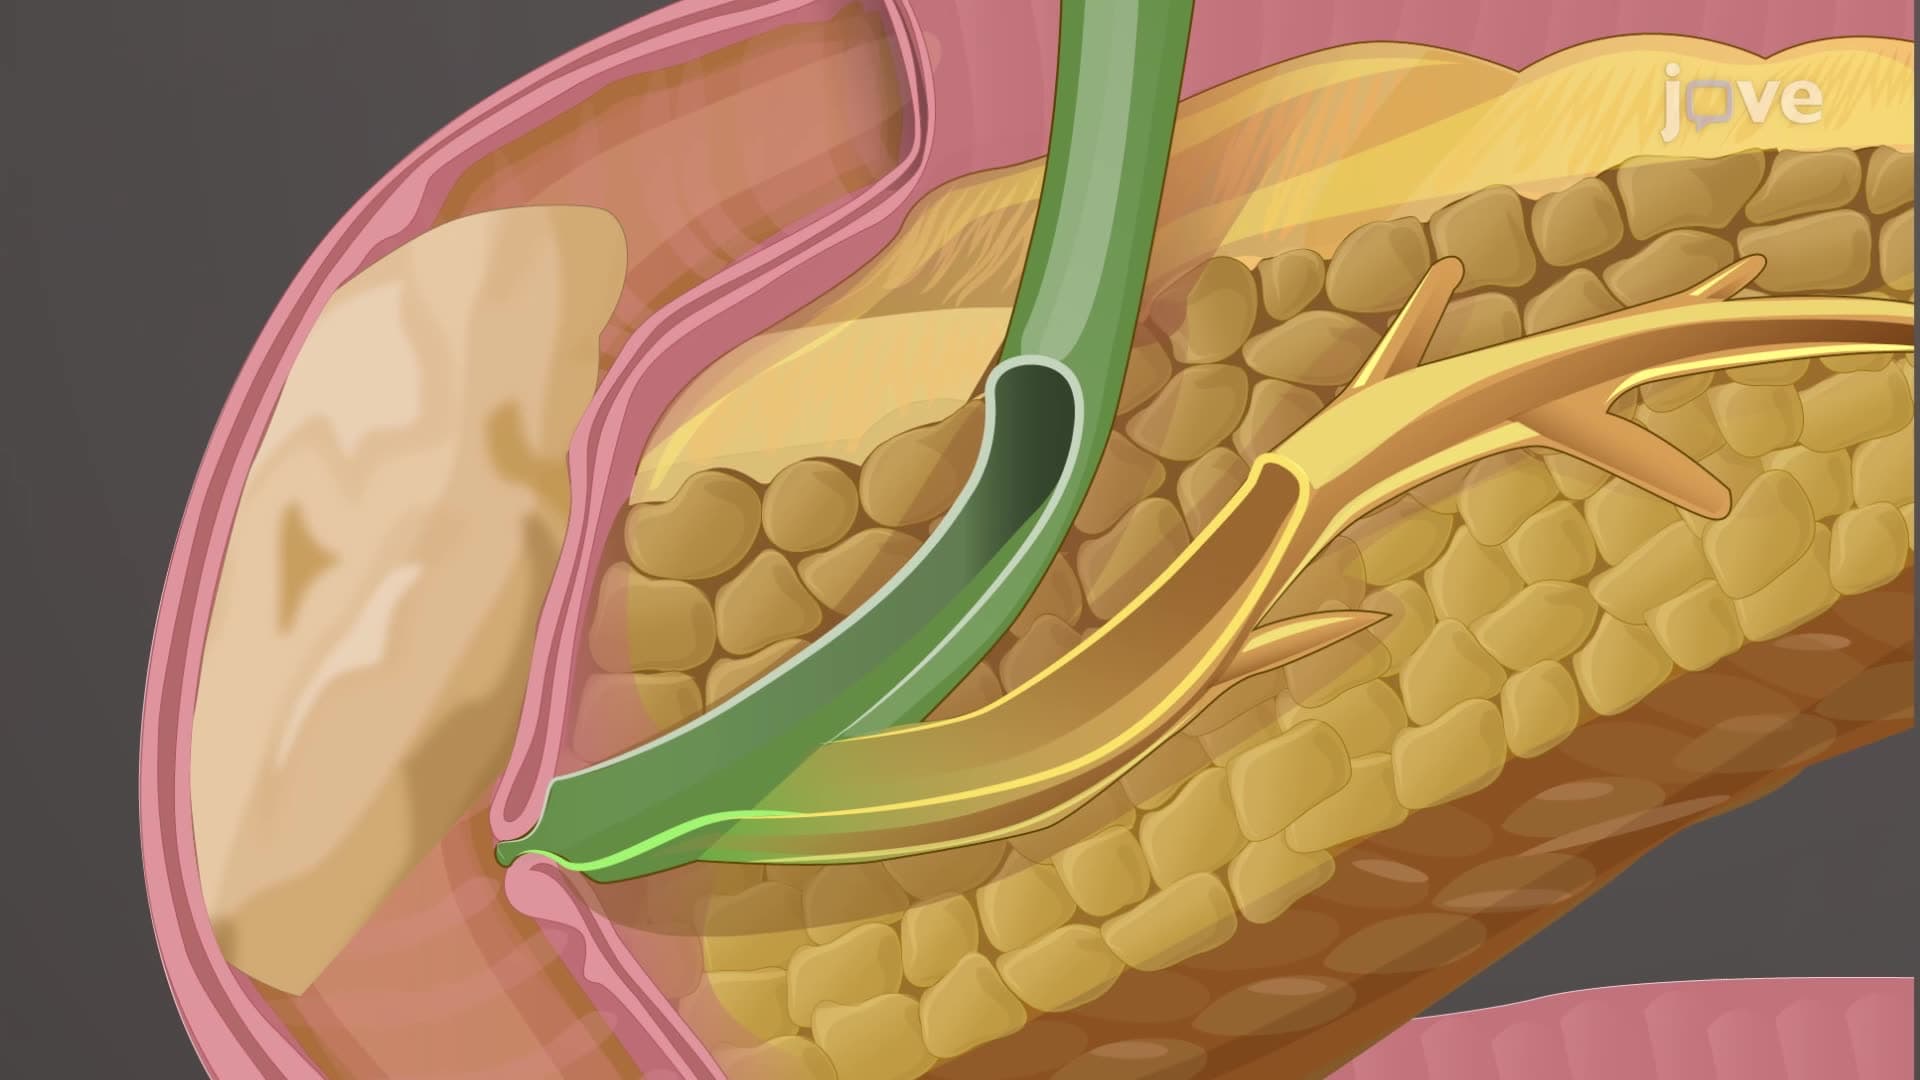 01:24
Lipid Absorption
These breakdown products bind with bile salts and lecithin to form micelles, which quickly pass between microvilli to come in close contact with the apical...
 01:20
Inborn Errors of Metabolism
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:25
Lipid Catabolism