Related Experiment Video
Updated: Jul 31, 2026
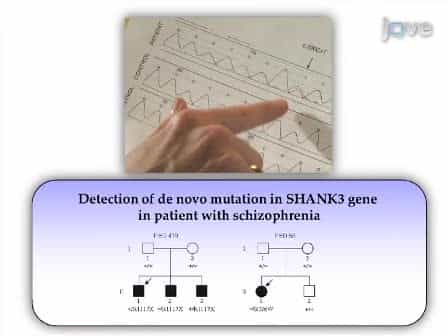

A Strategy to Identify de Novo Mutations in Common Disorders such as Autism and Schizophrenia
Published on: June 15, 2011
Dejerine-Sottas neuropathy in mother and son with same point mutation of PMP22 gene
V V Ionasescu1, C C Searby, R Ionasescu
1Department of Pediatrics, University of Iowa Hospitals and Clinics, Iowa City 52242, USA.

We studied a 25-year-old black woman with healthy parents and her 2-year, 11-month-old son. Her motor development was delayed and she started to walk with support when she was 6 years old. She never walked independently and had always used a wheelchair. Neurological evaluation showed severe weakness and atrophy of her feet, legs, and hands, bilateral pes cavus and hammertoes, corrected scoliosis, hypesthesia for proprioception and vibration sense in both feet and ankles, and areflexia. She had normal intelligence. Her son also had delayed motor milestones and was still unable to stand and walk independently at almost 3 years. Neurological evaluation revealed diffuse muscle hypotonia and weakness with generalized areflexia and normal intelligence. No muscle atrophies or feet deformities were noticed. Nerve conduction velocities showed significant slowing (less than 5 m/s) with prolonged distal latencies (above 30 ms). Compound motor action potential amplitudes were markedly reduced. Electromyography revealed polyphasic motor unit potentials. Molecular genetic studies indicated a Trembler type missense point mutation of exon 4 of the peripheral myelin protein 22 gene that led to the substitution of a spartic acid for glycine in both the mother and her son. Her parents showed normal DNA studies.
We studied a 25-year-old black woman with healthy parents and her 2-year, 11-month-old son. Her motor development was delayed and she started to walk with support when she was 6 years old. She never walked independently and had always used a wheelchair. Neurological evaluation showed severe weakness and atrophy of her feet, legs, and hands, bilateral pes cavus and hammertoes, corrected scoliosis, hypesthesia for proprioception and vibration sense in both feet and ankles, and areflexia. She had normal intelligence. Her son also had delayed motor milestones and was still unable to stand and walk independently at almost 3 years. Neurological evaluation revealed diffuse muscle hypotonia and weakness with generalized areflexia and normal intelligence. No muscle atrophies or feet deformities were noticed. Nerve conduction velocities showed significant slowing (less than 5 m/s) with prolonged distal latencies (above 30 ms). Compound motor action potential amplitudes were markedly reduced. Electromyography revealed polyphasic motor unit potentials. Molecular genetic studies indicated a Trembler type missense point mutation of exon 4 of the peripheral myelin protein 22 gene that led to the substitution of a spartic acid for glycine in both the mother and her son. Her parents showed normal DNA studies.
More Related Videos
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
 06:35
In Vivo Electrophysiological Measurement of Compound Muscle Action Potential from the Forelimbs in Mouse Models of Motor Neuron Degeneration
Published on: June 15, 2018
Related Concept Videos
 01:35
Pedigree Analysis
 02:59
Animal Mitochondrial Genetics
 01:26
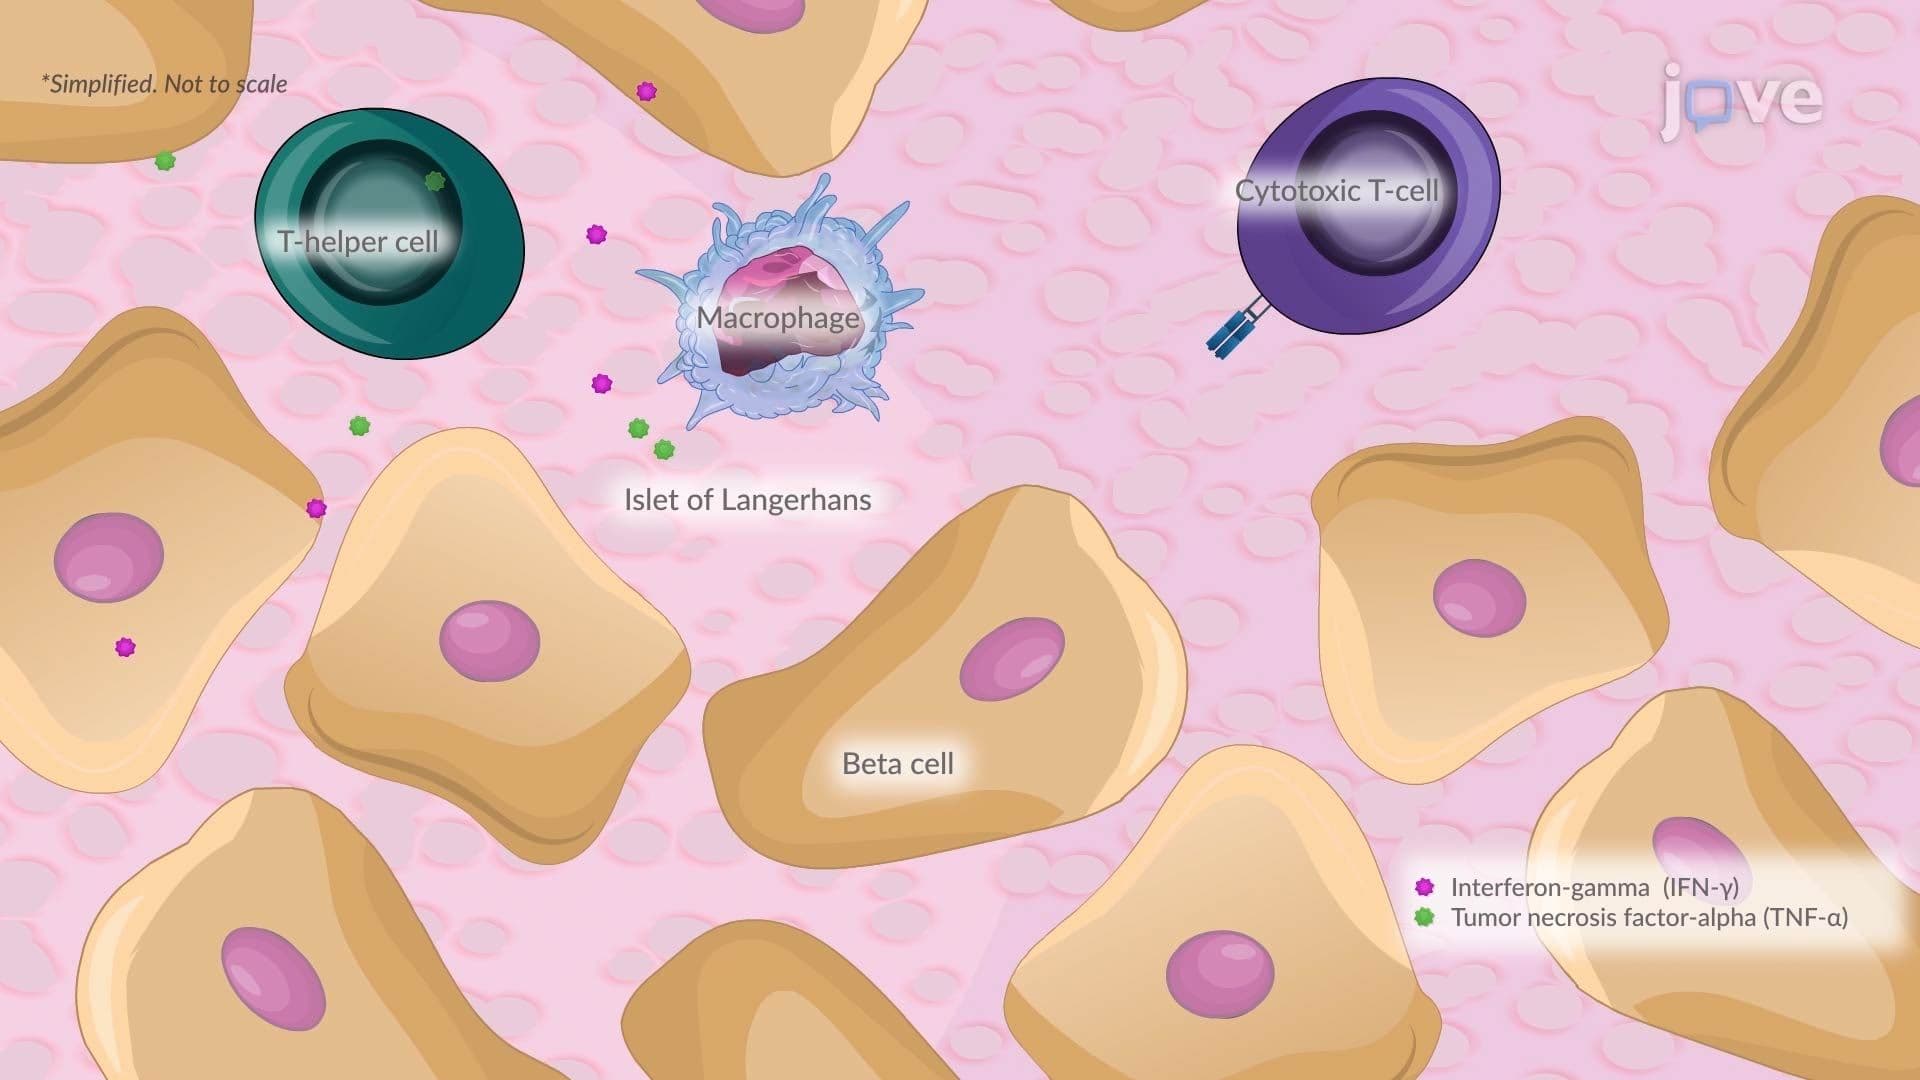
Type I Diabetes II: Pathophysiology
 01:31
Diabetic Foot Ulcer
 01:28
Diabetic Nephropathy
 01:22
Diabetic Neuropathy