Related Experiment Video
Updated: Jul 27, 2026
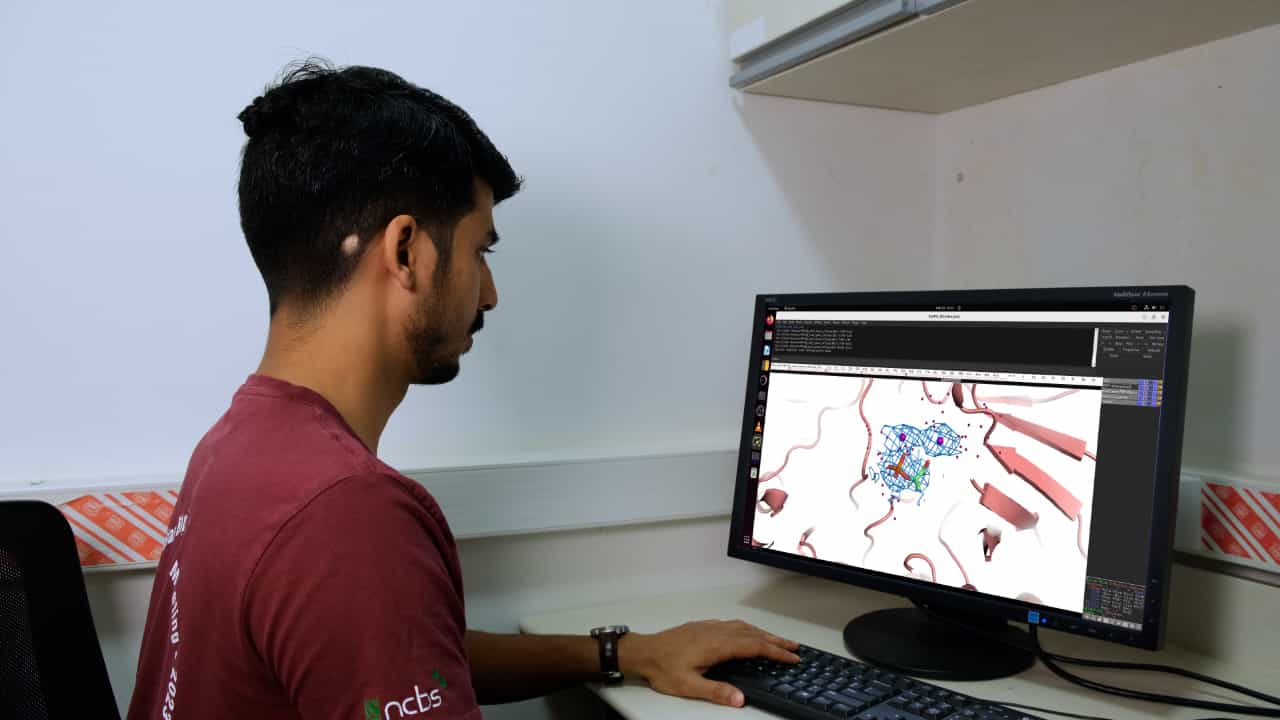
Modeling Ligands into Maps Derived from Electron Cryomicroscopy
Published on: July 19, 2024
Mode matches and their locations in the hydrophobic free energy sequences of peptide ligands and their receptor
A J Mandell1, K A Selz, M F Shlesinger
1The Cielo Institute, 486 Sunset Drive, Asheville, NC 28804, USA.

Abstract:
Patterns in sequences of amino acid hydrophobic free energies predict secondary structures in proteins. In protein folding, matches in hydrophobic free energy statistical wavelengths appear to contribute to selective aggregation of secondary structures in "hydrophobic zippers." In a similar setting, the use of Fourier analysis to characterize the dominant statistical wavelengths of peptide ligands' and receptor proteins' hydrophobic modes to predict such matches has been limited by the aliasing and end effects of short peptide lengths, as well as the broad-band, mode multiplicity of many of their frequency (power) spectra. In addition, the sequence locations of the matching modes are lost in this transformation. We make new use of three techniques to address these difficulties: (i) eigenfunction construction from the linear decomposition of the lagged covariance matrices of the ligands and receptors as hydrophobic free energy sequences; (ii) maximum entropy, complex poles power spectra, which select the dominant modes of the hydrophobic free energy sequences or their eigenfunctions; and (iii) discrete, best bases, trigonometric wavelet transformations, which confirm the dominant spectral frequencies of the eigenfunctions and locate them as (absolute valued) moduli in the peptide or receptor sequence. The leading eigenfunction of the covariance matrix of a transmembrane receptor sequence locates the same transmembrane segments seen in n-block-averaged hydropathy plots while leaving the remaining hydrophobic modes unsmoothed and available for further analyses as secondary eigenfunctions. In these receptor eigenfunctions, we find a set of statistical wavelength matches between peptide ligands and their G-protein and tyrosine kinase coupled receptors, ranging across examples from 13.10 amino acids in acid fibroblast growth factor to 2.18 residues in corticotropin releasing factor. We find that the wavelet-located receptor modes in the extracellular loops are compatible with studies of receptor chimeric exchanges and point mutations. A nonbinding corticotropin-releasing factor receptor mutant is shown to have lost the signatory mode common to the normal receptor and its ligand. Hydrophobic free energy eigenfunctions and their transformations offer new quantitative physical homologies in database searches for peptide-receptor matches.
More Related Videos
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage
00:49Ligand Binding and Linkage
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...