Hydrogen bonding in helical polypeptides from molecular dynamics simulations and amide hydrogen exchange analysis: alamethicin and melittin in methanol
Related Experiment Video
Updated: Oct 30, 2026
Analyzing Protein Dynamics Using Hydrogen Exchange Mass Spectrometry
Published on: November 29, 2013
R B Sessions1, N Gibbs, C E Dempsey
1Biochemistry Department and Centre for Molecular Recognition, Bristol University, School of Medical Sciences, United Kingdom.

Abstract:
Molecular dynamics simulations of ion channel peptides alamethicin and melittin, solvated in methanol at 27 degrees C, were run with either regular alpha-helical starting structures (alamethicin, 1 ns; melittin 500 ps either with or without chloride counterions), or with the x-ray crystal coordinates of alamethicin as a starting structure (1 ns). The hydrogen bond patterns and stabilities were characterized by analysis of the dynamics trajectories with specified hydrogen bond angle and distance criteria, and were compared with hydrogen bond patterns and stabilities previously determined from high-resolution NMR structural analysis and amide hydrogen exchange measurements in methanol. The two alamethicin simulations rapidly converged to a persistent hydrogen bond pattern with a high level of 3(10) hydrogen bonding involving the amide NH's of residues 3, 4, 9, 15, and 18. The 3(10) hydrogen bonds stabilizing amide NH's of residues C-terminal to P2 and P14 were previously proposed to explain their high amide exchange stabilities. The absence, or low levels of 3(10) hydrogen bonds at the N-terminus or for A15 NH, respectively, in the melittin simulations, is also consistent with interpretations from amide exchange analysis. Perturbation of helical hydrogen bonding in the residues before P14 (Aib10-P14, alamethicin; T11-P14, melittin) was characterized in both peptides by variable hydrogen bond patterns that included pi and gamma hydrogen bonds. The general agreement in hydrogen bond patterns determined in the simulations and from spectroscopic analysis indicates that with suitable conditions (including solvent composition and counterions where required), local hydrogen-bonded secondary structure in helical peptides may be predicted from dynamics simulations from alpha-helical starting structures. Each peptide, particularly alamethicin, underwent some large amplitude structural fluctuations in which several hydrogen bonds were cooperatively broken. The recovery of the persistent hydrogen bonding patterns after these fluctuations demonstrates the stability of intramolecular hydrogen-bonded secondary structure in methanol (consistent with spectroscopic observations), and is promising for simulations on extended timescales to characterize the nature of the backbone fluctuations that underlie amide exchange from isolated helical polypeptides.
More Related Videos
 11:32
A Hydrogen-Deuterium Exchange Mass Spectrometry (HDX-MS) Platform for Investigating Peptide Biosynthetic Enzymes
Published on: May 4, 2020
 05:57
Synthesizing Amino Acids Modified with Reactive Carbonyls in Silico to Assess Structural Effects Using Molecular Dynamics Simulations
Published on: April 26, 2024
Related Concept Videos
 01:04
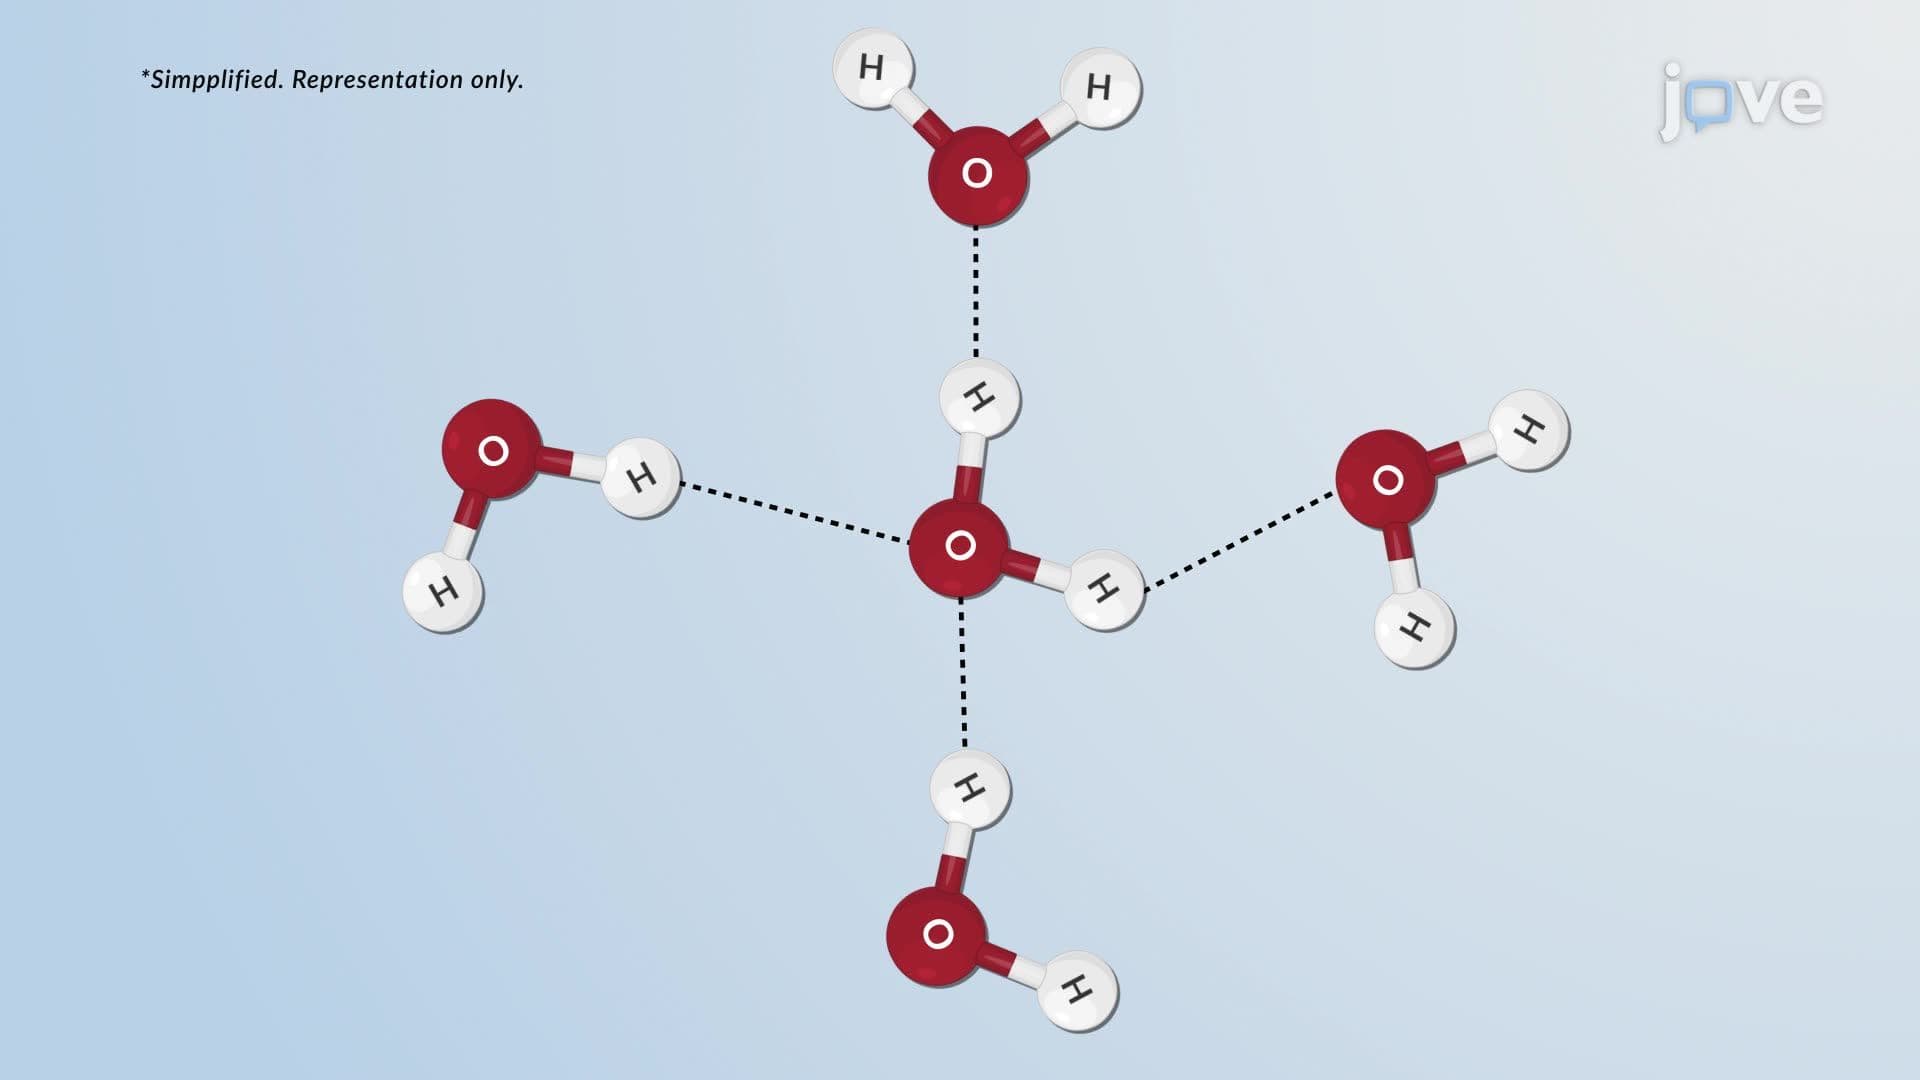
Hydrogen Bonds
 00:26
Hydrogen Bonds
 01:21
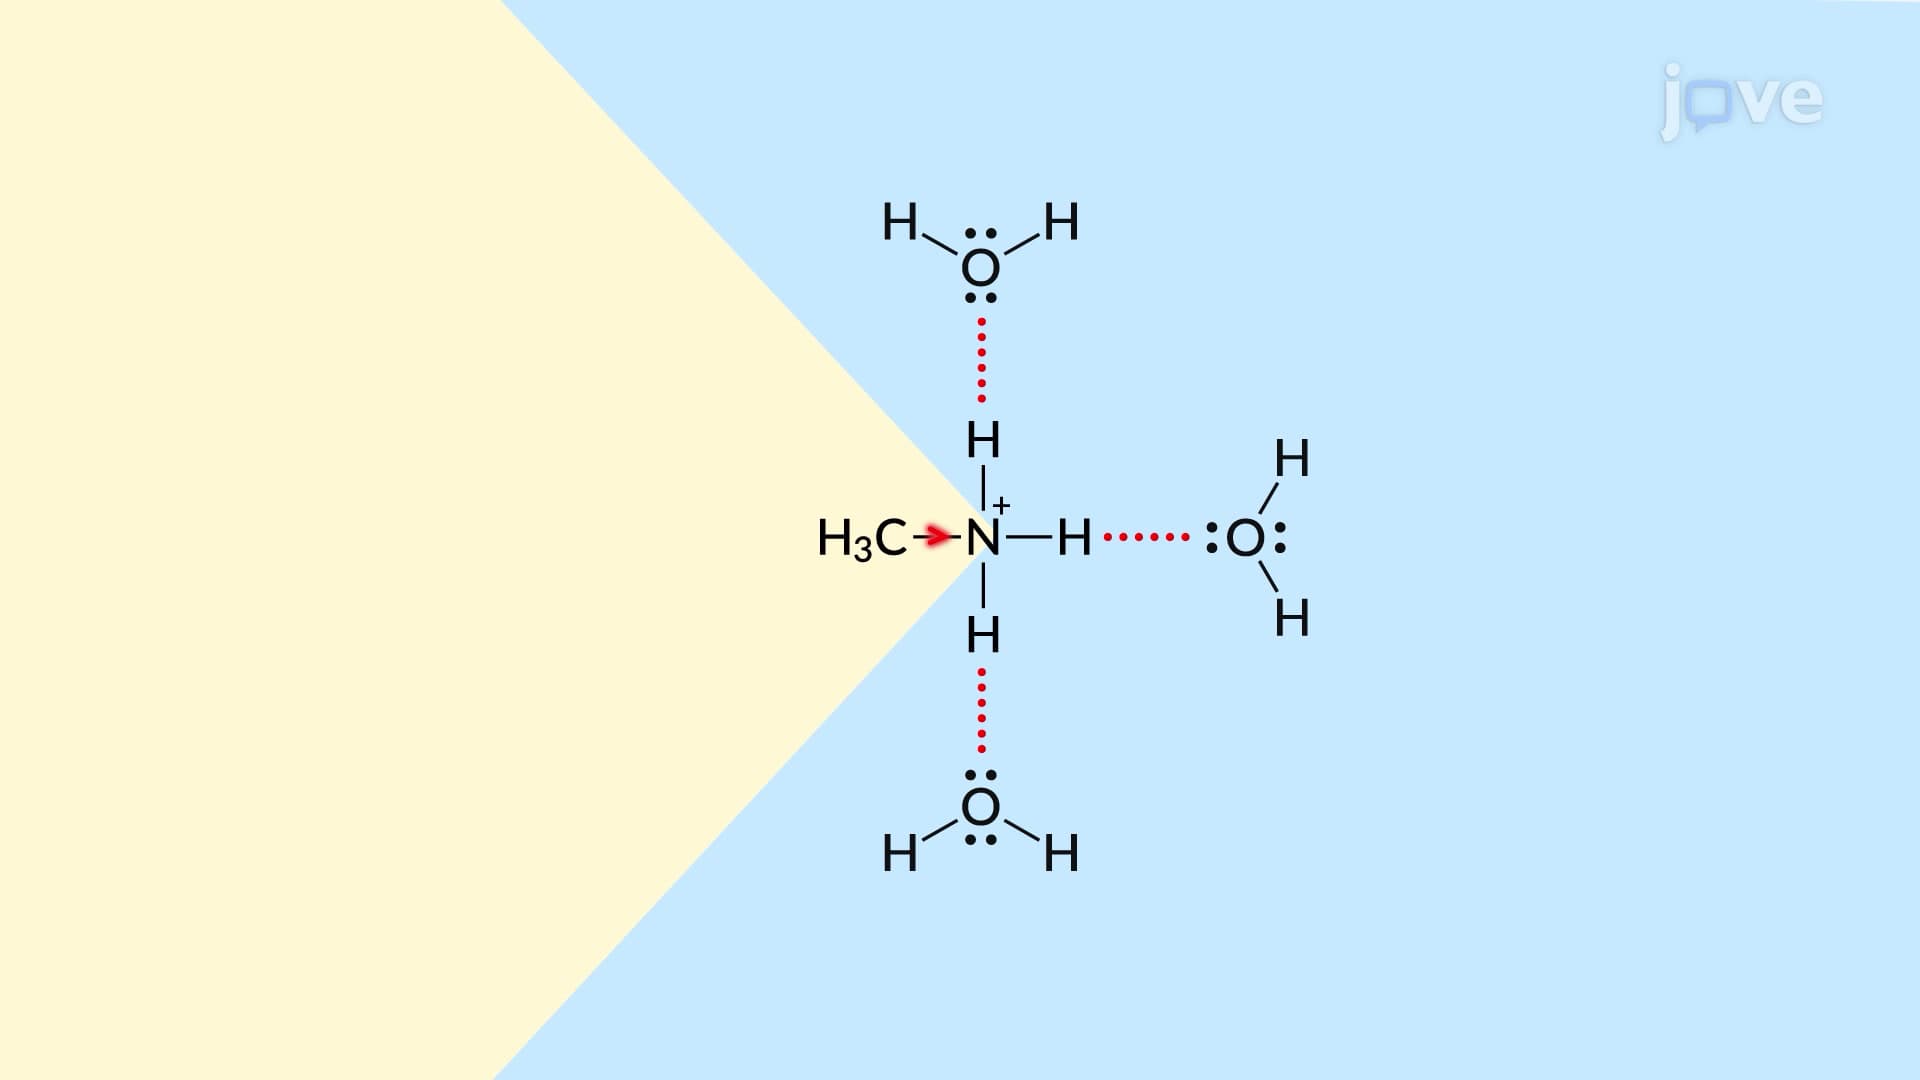
Basicity of Aliphatic Amines
To measure the basicity of amines, two conventions are generally used. The first defines Kb as the basicity constant for the deprotonation reaction of water by the amine, as presented in Figure 1. Conventionally, lower Kb indicates higher...
 01:26
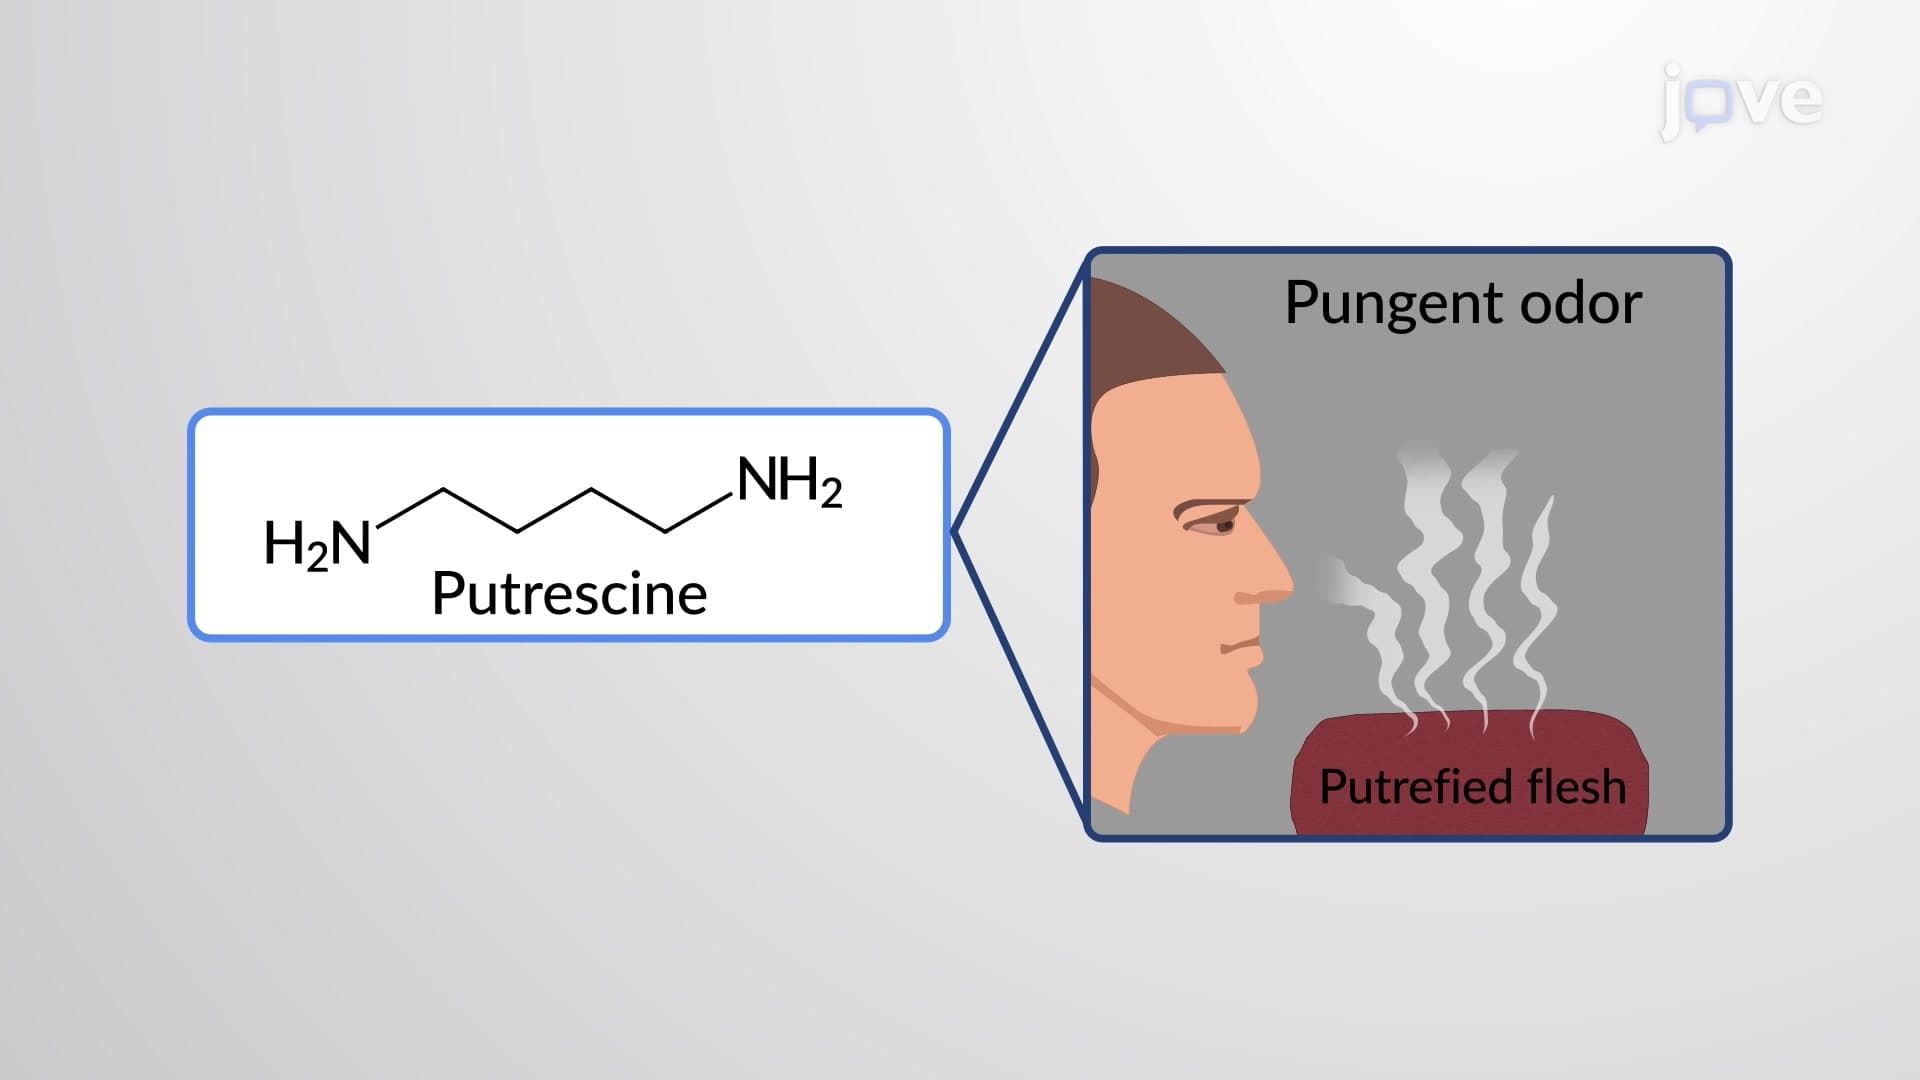
Physical Properties of Amines
 02:35
Noncovalent Attractions in Biomolecules
Four types of noncovalent interactions are hydrogen bonds, van der Waals forces, ionic bonds, and hydrophobic interactions.
Hydrogen bonding results from the electrostatic attraction of a hydrogen atom covalently bonded to a strong-electronegative atom like oxygen,...
 02:35
Peptide Bonds