Flexible protein-ligand docking by global energy optimization in internal coordinates
Related Experiment Video
Updated: Aug 17, 2026
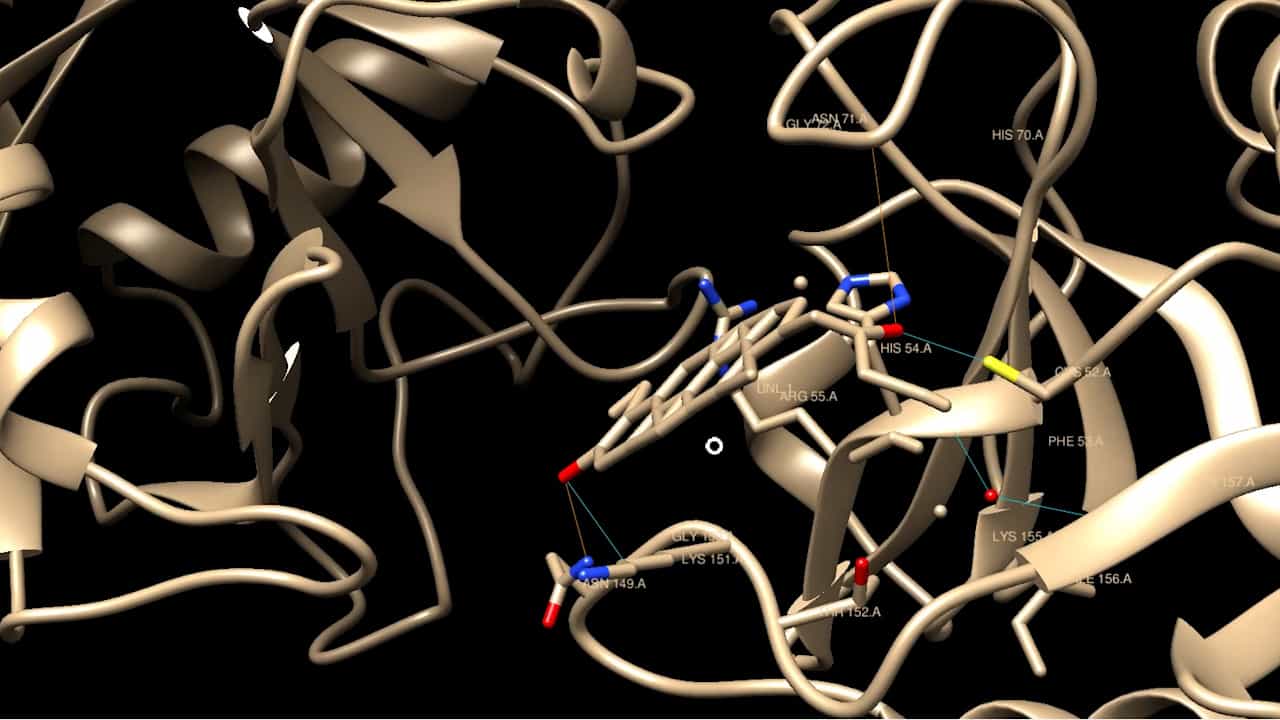
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
1Skirball Institute of Biomolecular Medicine, Biochemistry Department of New York University Medical College, New York 10016, USA.

Abstract:
Eight protein-ligand complexes were simulated by using global optimization of a complex energy function, including solvation, surface tension, and side-chain entropy in the internal coordinate space of the flexible ligand and the receptor side chains [Abagyan, R.A., Totrov, M.M. J. Mol. Biol. 235: 983-1002, 1994]. The procedure uses two types of efficient random moves, a pseudobrownian positional move [Abagyan, R.A., Totrov, M.M., Kuznetsov, D.A. J. Comp. Chem. 15:488-506, 1994] and a Biased-Probability multitorsion move [Abagyan, R.A., Totrov, M.M. J. Mol. Biol. 235: 983-1002, 1994], each accompanied by full local energy minimization. The best docking solutions were further ranked according to the interaction energy, which included intramolecular deformation energies of both receptor and ligand, the interaction energy, surface tension, side-chain entropic contribution, and an electrostatic term evaluated as a boundary element solution of the Poisson equation with the molecular surface as a dielectric boundary. The geometrical accuracy of the docking solutions ranged from 30% to 70% according to the relative displacement error measure at a 1.5 A scale. Similar results were obtained when the explicit receptor atoms were replaced with a grid potential.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:49
Ligand Binding and Linkage
00:49Ligand Binding and Linkage
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...