Optimized representations and maximal information in proteins
Related Experiment Video
Updated: Oct 17, 2026

Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
1Department of Biomathematical Sciences, Mount Sinai School of Medicine, New York, New York 10029, USA.

Abstract:
In an effort to quantify loss of information in the processing of protein bioinformatic data, we examine how representations of amino acid sequence and backbone conformation affect the quantity of accessible structural information from local sequence. We propose a method to extract the maximum amount of peptide backbone structural information available in local sequence fragments, given a finite structural data set. Using methods of information theory, we develop an unbiased measure of local structural information that gauges changes in structural distributions when different representations of secondary structure and local sequence are used. We find that the manner in which backbone structure is represented affects the amount and quality of structural information that may be extracted from local sequence. Representations based on virtual bonds capture more structural information from local sequence than a three-state assignment scheme (helix/strand/loop). Furthermore, we find that amino acids show significant kinship with respect to the backbone structural information they carry, so that a collapse of the amino acid alphabet can be accomplished without severely affecting the amount of extractable information. This strategy is critical in optimizing the utility of a limited database of experimentally solved protein structures. Finally, we discuss the similarities within and differences between groups of amino acids in their roles in the local folding code and recognize specific amino acids critical in the formation of local structure.
More Related Videos


Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:04
Protein-protein Interfaces
02:04Protein-Protein Interfaces
 01:24
Protein Organization
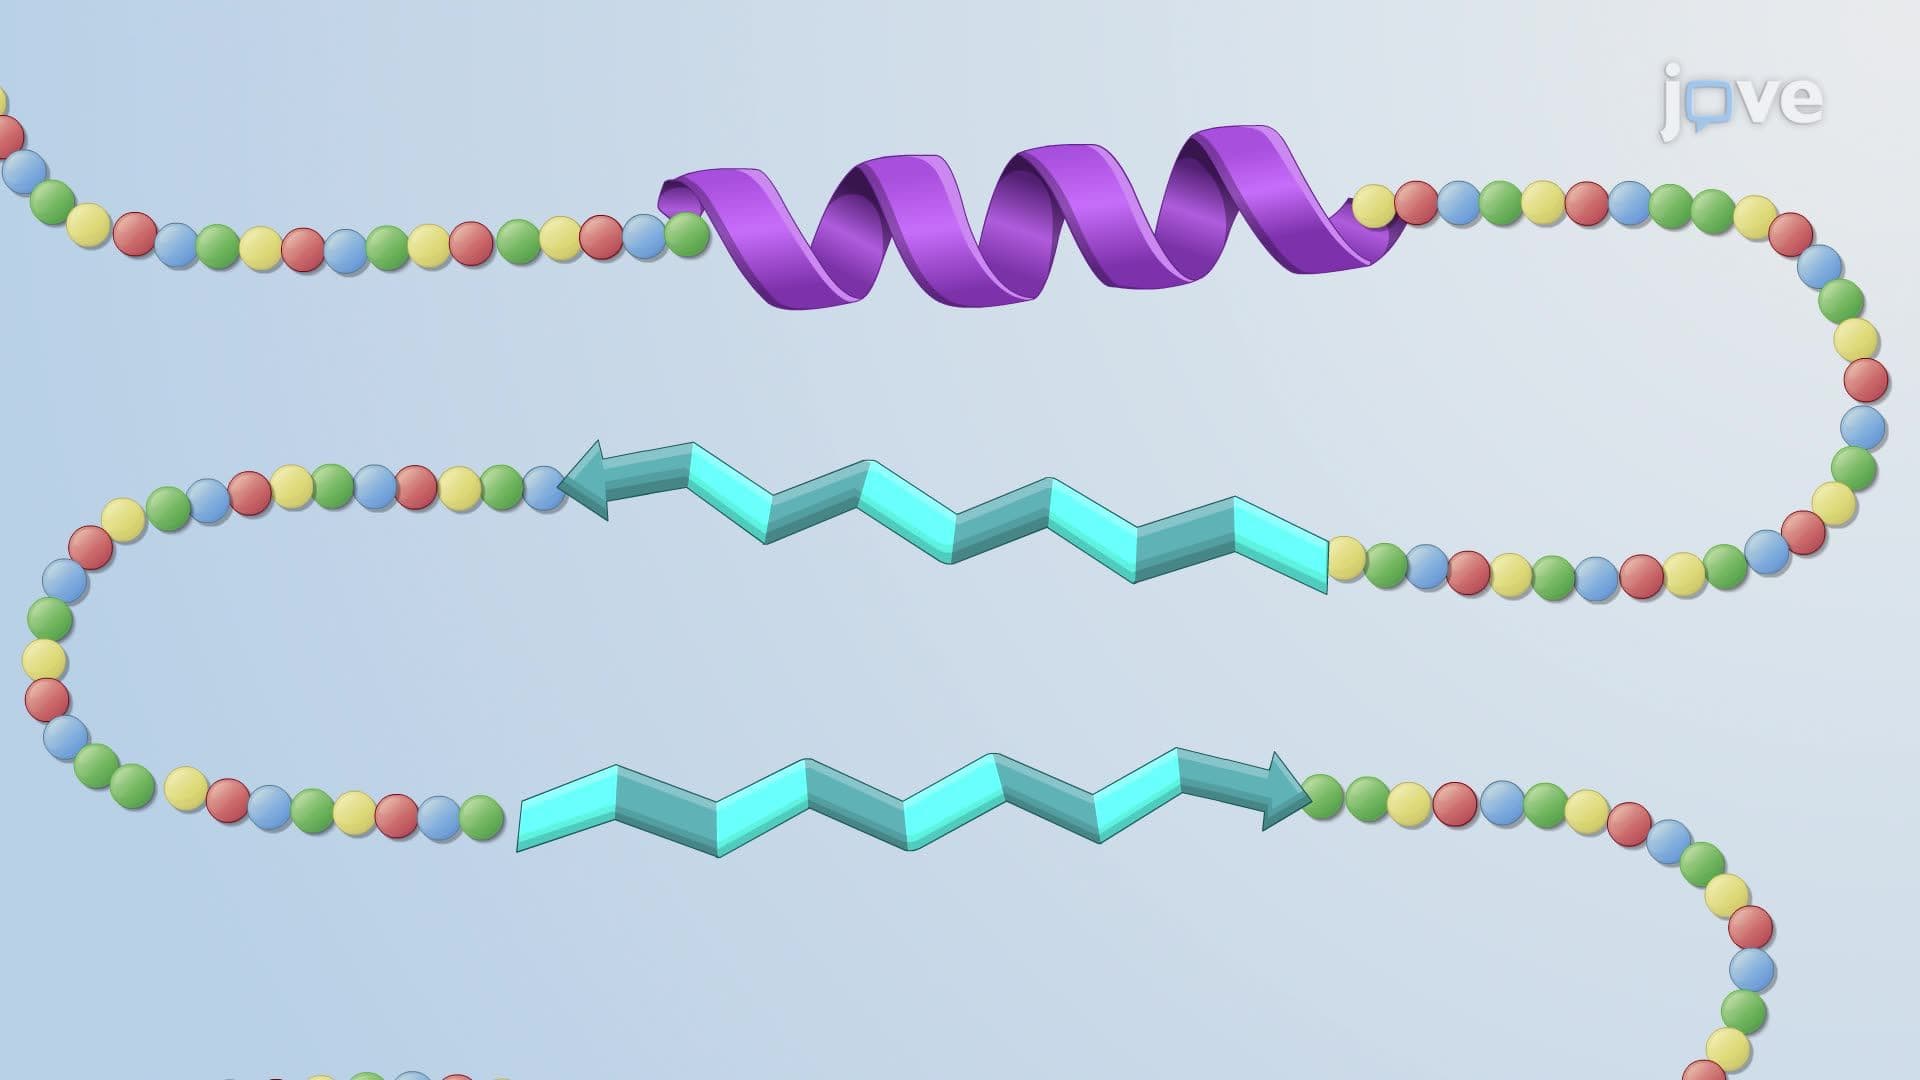
The primary structure of a protein is its amino acid sequence.
 01:28
Protein Organization