Related Experiment Video
Updated: Oct 9, 2026
Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
An algorithm for statistical alignment of sequences related by a binary tree
1Department of Ecology and Genetics, University of Aarhus, Ny Munkegade, Aarhus, Denmark. jotun.hein@biology.au.dk

Abstract:
An algorithm is presented that allows the calculation of the probability of a set of sequences related by a binary tree that has evolved according to the Thorne-Kishino-Felsenstein model (1991) for a fixed set of parameters. There are two ideas underlying this algorithm. Firstly, a markov chain is defined that generates ancestral sequences and their alignment at two neighboring nodes in a tree. Secondly, a stochastic walk on the binary tree, that defines a markov chain generating ancestral sequences and their alignment at the internal nodes in the tree is described. The running time of this algorithm is O(l2 kappa), where l is the geometric average of the sequence lengths and kappa the number of sequences--leaves at the binary tree. This could be improved to O(l kappa).
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 01:29
Modern Molecular Taxonomy
 03:21
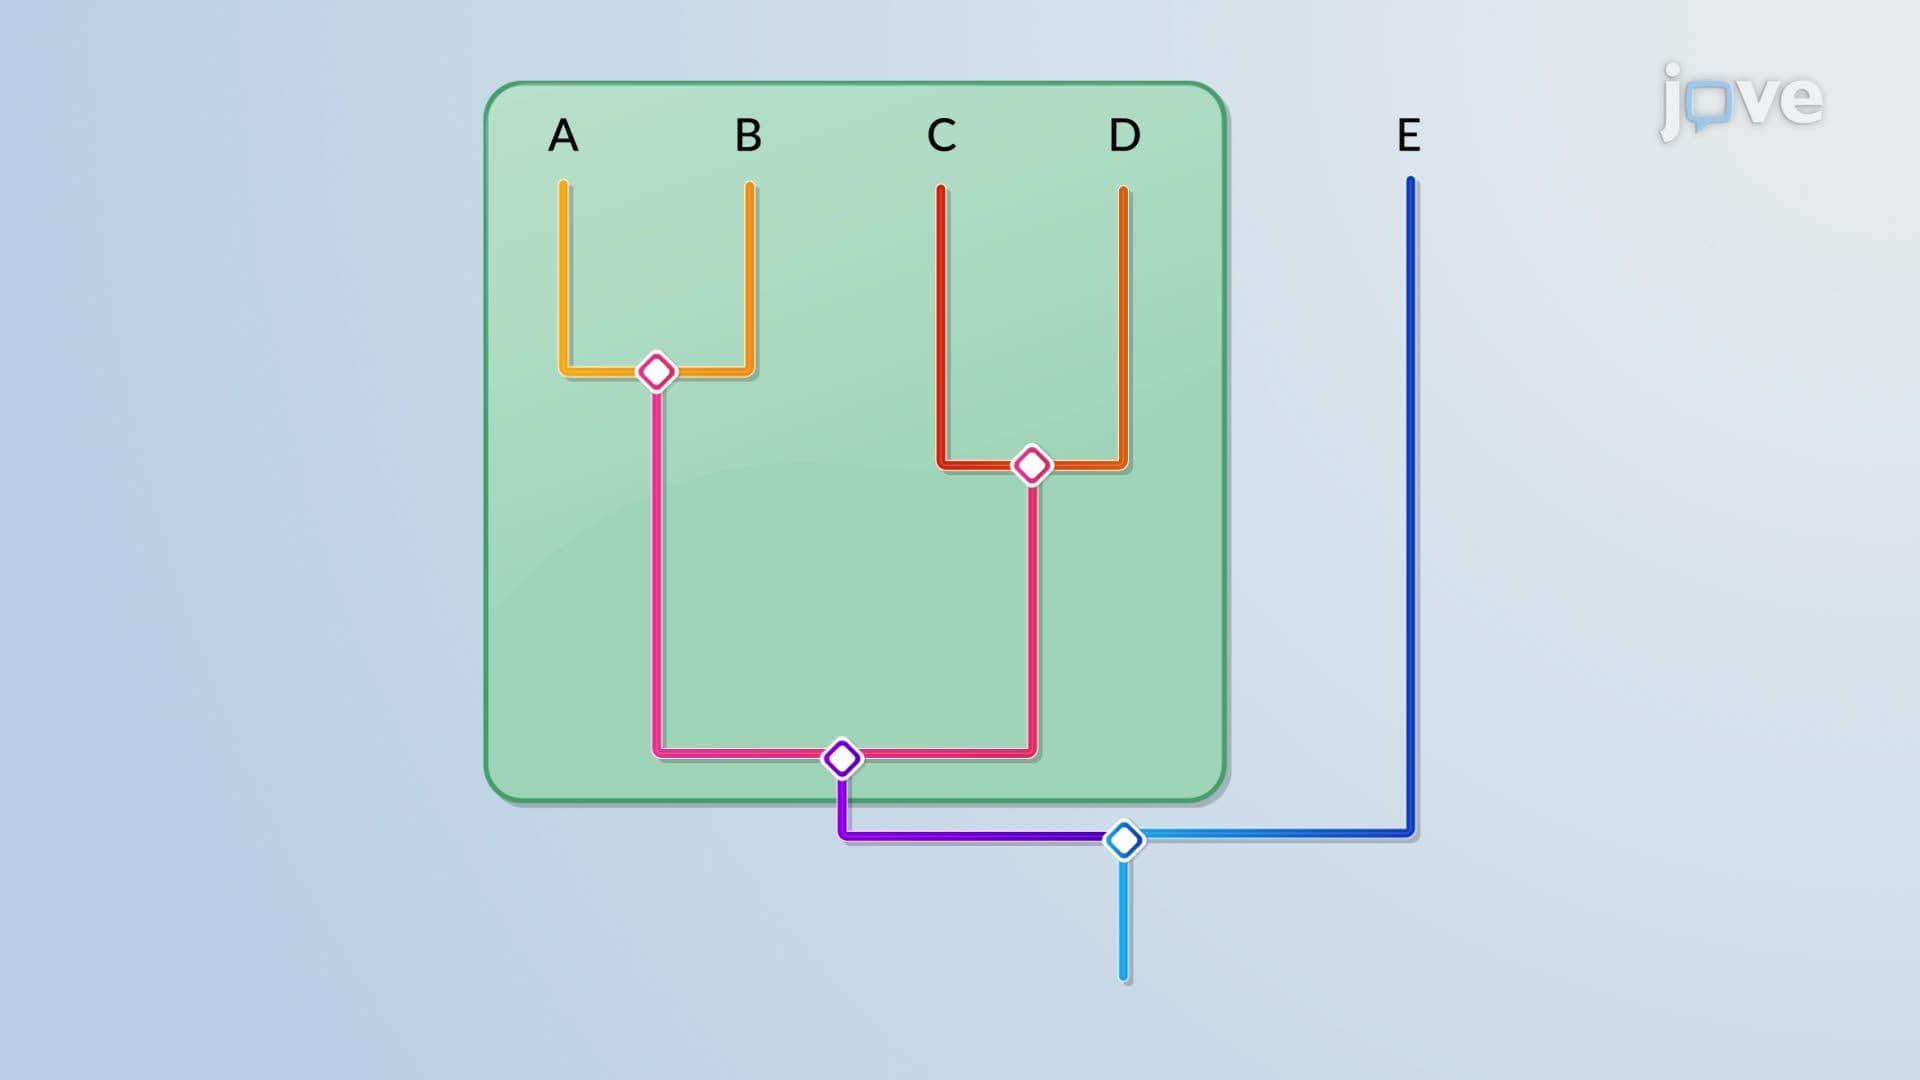
Phylogenetic Trees
03:21Phylogenetic Trees
 03:36
Genome Annotation and Assembly