Atomistic protein folding simulations on the submillisecond time scale using worldwide distributed computing
Related Experiment Video
Updated: Sep 19, 2026
Microfluidic Mixers for Studying Protein Folding
Published on: April 10, 2012
Vijay S Pande1, Ian Baker, Jarrod Chapman
1Department of Chemistry, Stanford University, Stanford, CA 94305-5080, USA. pande@stanford.edu

Abstract:
Atomistic simulations of protein folding have the potential to be a great complement to experimental studies, but have been severely limited by the time scales accessible with current computer hardware and algorithms. By employing a worldwide distributed computing network of tens of thousands of PCs and algorithms designed to efficiently utilize this new many-processor, highly heterogeneous, loosely coupled distributed computing paradigm, we have been able to simulate hundreds of microseconds of atomistic molecular dynamics. This has allowed us to directly simulate the folding mechanism and to accurately predict the folding rate of several fast-folding proteins and polymers, including a nonbiological helix, polypeptide alpha-helices, a beta-hairpin, and a three-helix bundle protein from the villin headpiece. Our results demonstrate that one can reach the time scales needed to simulate fast folding using distributed computing, and that potential sets used to describe interatomic interactions are sufficiently accurate to reach the folded state with experimentally validated rates, at least for small proteins.
More Related Videos

Related Concept Videos
 01:49
Protein Folding
01:49Protein Folding
 02:21
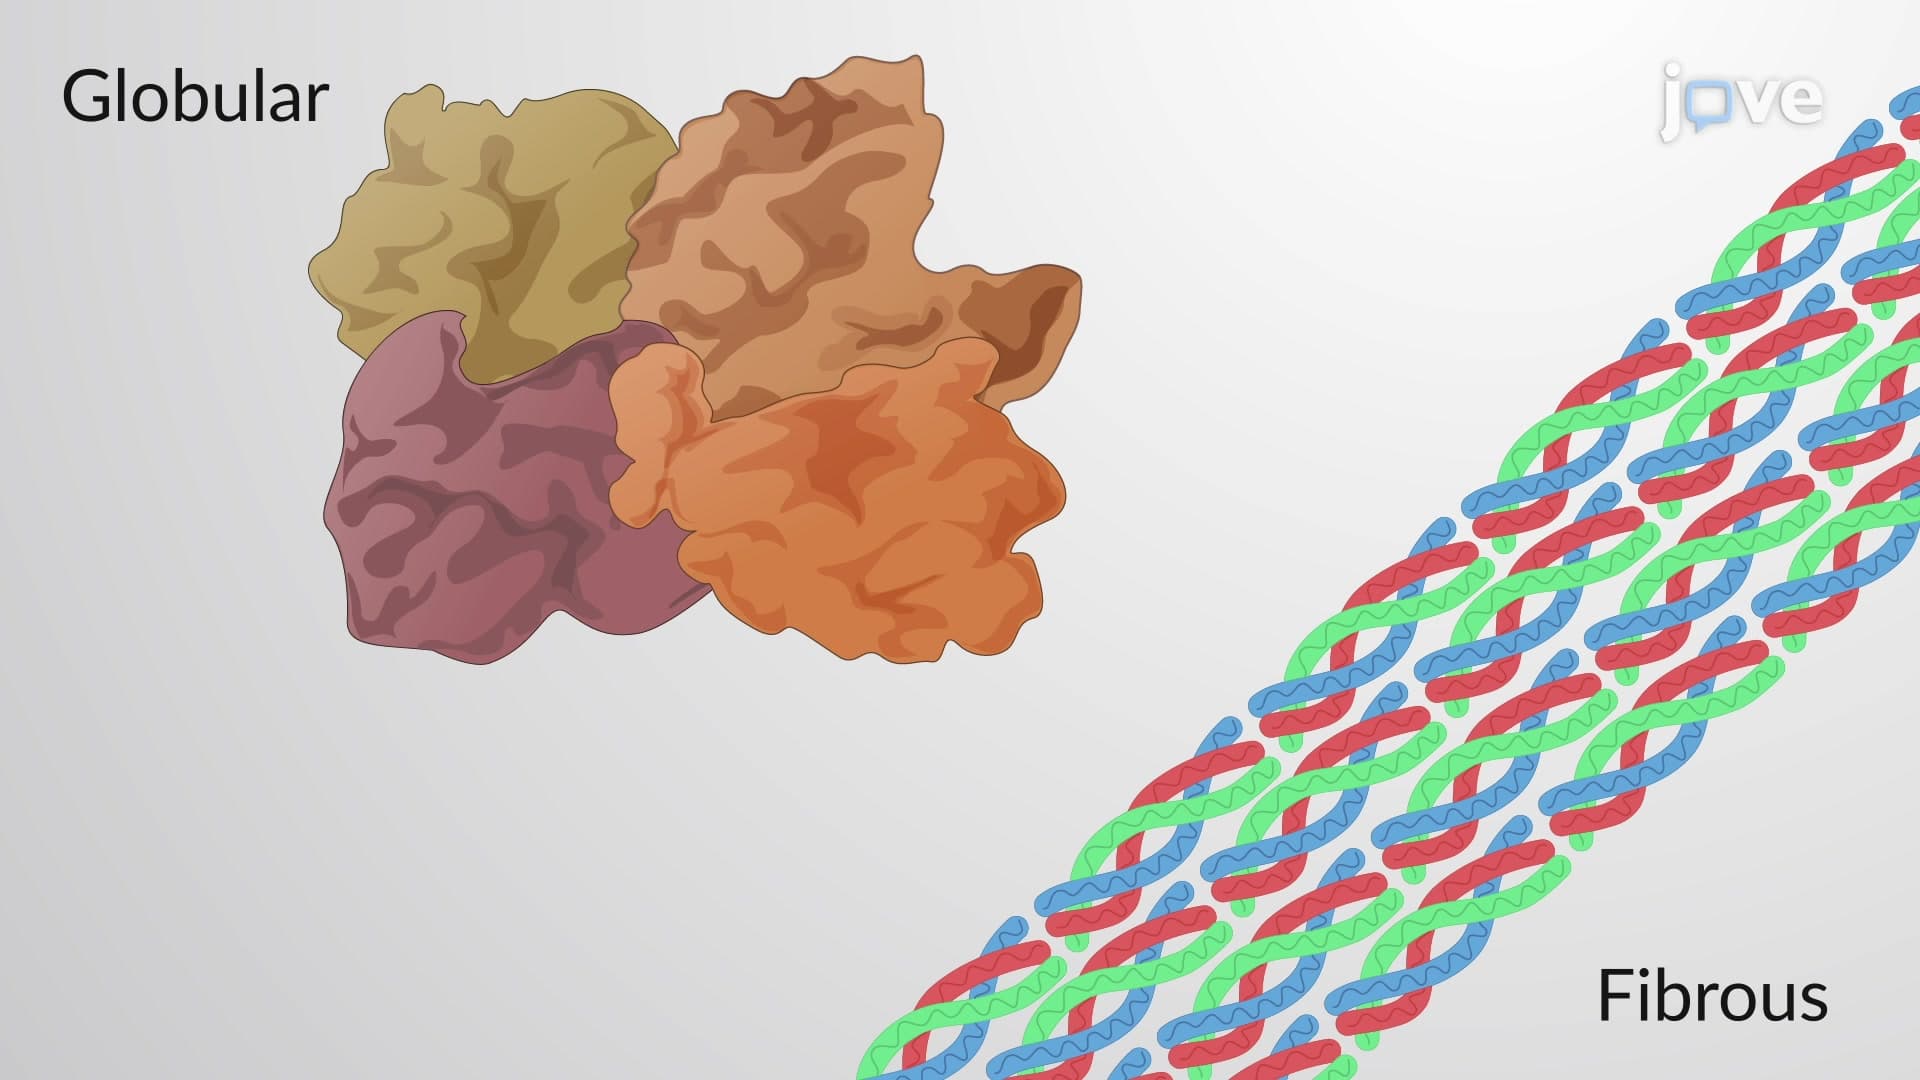
Globular and Fibrous Proteins
Globular proteins are also known as spheroproteins and typically are approximately round in shape. They contain a mix of amino acid types and contain differing sequences in their primary structures. Globular proteins have many different functions, such as enzymes, cellular messengers, and molecular transporters. These roles often require the proteins to be...
 03:00
Molecular Chaperones and Protein Folding
The...