Combining local-structure, fold-recognition, and new fold methods for protein structure prediction
Related Experiment Video
Updated: Aug 30, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
Kevin Karplus1, Rachel Karchin, Jenny Draper
1Computer Engineering Department, University of California, Santa Cruz 95064, USA. karplus@soe.ucsc.edu

Abstract:
This article presents an overview of the SAM-T02 method for protein fold recognition and the UNDERTAKER program for ab initio predictions. The SAM-T02 server is an automatic method that uses two-track hidden Markov models (HMMS) to find and align template proteins from PDB to the target protein. The two-track HMMs use an amino acid alphabet and one of several different local structure alphabets. The UNDERTAKER program is a new fragment-packing program that can use short or long fragments and alignments to create protein conformations. The HMMs and fold-recognition alignments from the SAM-T02 method were used to generate the fragment and alignment libraries used by UNDERTAKER. We present results on a few selected targets for which this combined method worked particularly well: T0129, T0181, T0135, T0130, and T0139.


Related Concept Videos
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 01:49
Protein Folding
01:49Protein Folding
 01:29
Protein Folding Quality Check in the RER
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:15
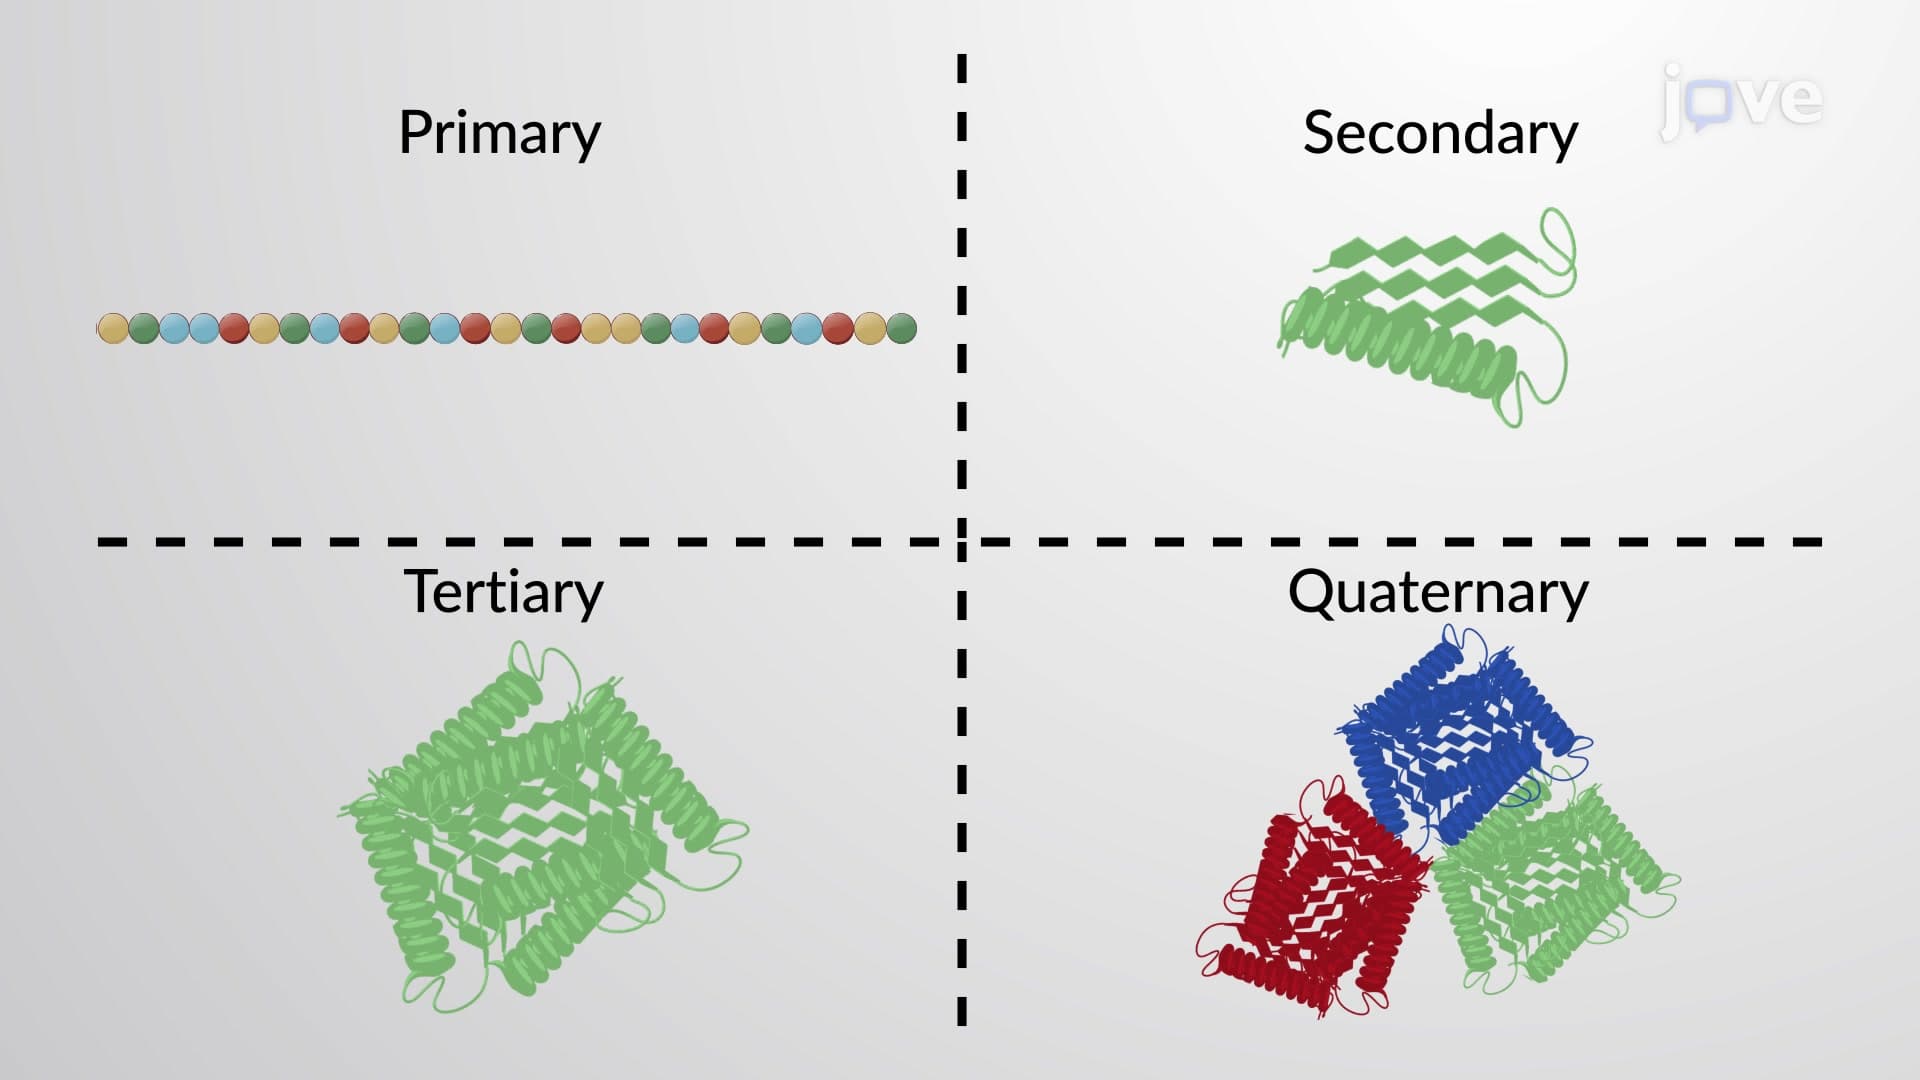
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme can...