Related Experiment Video
Updated: Aug 3, 2026
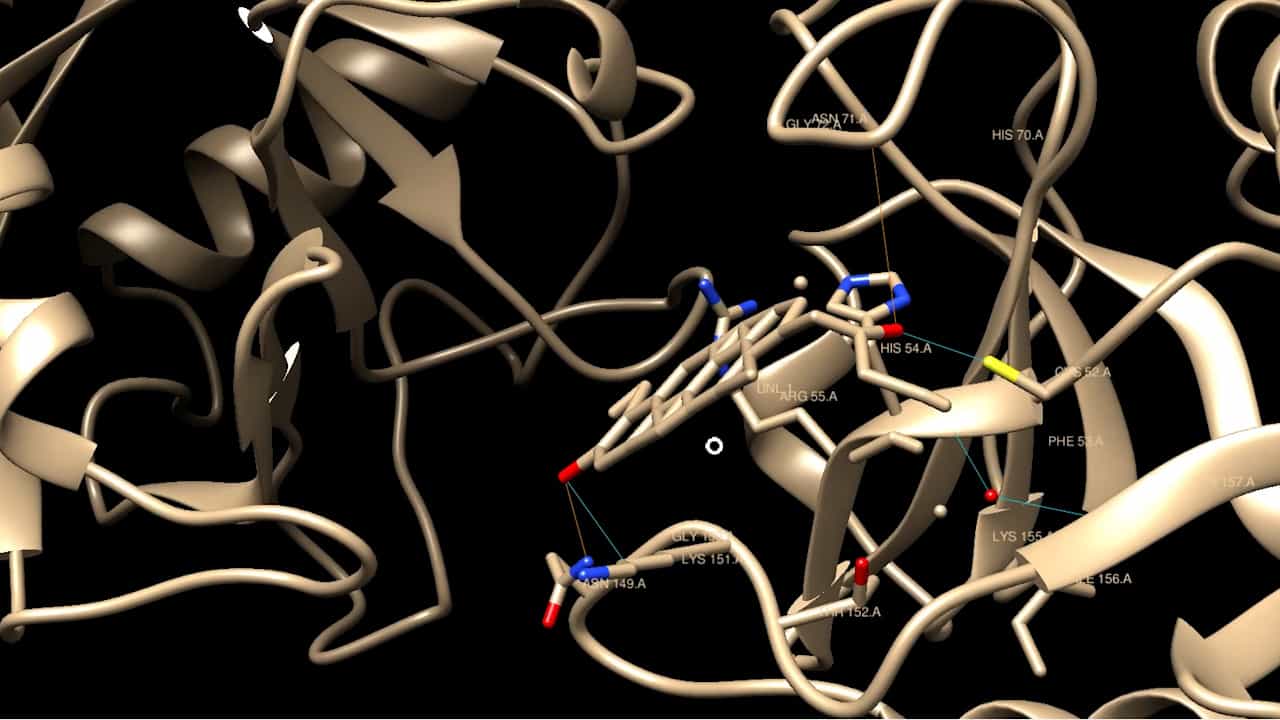

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Guided docking approaches to structure-based design and screening
Xavier Fradera1, Jordi Mestres
1Chemogenomics Laboratory, Research Group on Biomedical Informatics, Institut Municipal Investigació Medica and Universitat Pompeu Fabra, Passeig Maritim de la Barceloneta, 37-49, 08003 Barcelona (Catalonia), Spain.

Abstract:
With the number of protein-ligand complexes available in the Protein Data Bank constantly growing, structure-based approaches to drug design and screening have become increasingly important. Alongside this explosion of structural information, a number of molecular docking methods have been developed over the last years with the aim of maximally exploiting all available structural and chemical information that can be derived from proteins, from ligands, and from protein-ligand complexes. In this respect, the term 'guided docking' is introduced to refer to docking approaches that incorporate some degree of chemical information to actively guide the orientation of the ligand into the binding site. To reflect the focus on the use of chemical information, a classification scheme for guided docking approaches is proposed. In general terms, guided docking approaches can be divided into indirect and direct approaches. Indirect approaches incorporate chemical information implicitly, having an effect on scoring but not on orienting the ligand during sampling. In contrast, direct approaches incorporate chemical information explicitly, thus actively guiding the orientation of the ligand during sampling. Direct approaches can be further divided into protein-based, mapping-based, and ligand-based approaches to reflect the source used to derive the features capturing the chemical information inside the protein cavity. Within each category, a representative list of docking approaches is discussed. In view of the limitations of current scoring functions, it was generally found that making optimal use of chemical information represents an efficient knowledge-based strategy for improving binding affinity estimations, ligand binding-mode predictions, and virtual screening enrichments obtained from protein-ligand docking.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:26
Drug Discovery: Overview
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...