Related Experiment Video
Updated: Jul 15, 2026
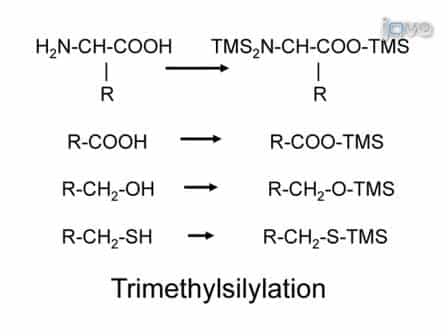

One-step Metabolomics: Carbohydrates, Organic and Amino Acids Quantified in a Single Procedure
Published on: June 26, 2010
Creatine and guanidinoacetate: diagnostic markers for inborn errors in creatine biosynthesis and transport
Lígia S Almeida1, Nanda M Verhoeven, Birthe Roos
1Department of Clinical Chemistry, Metabolic Unit, VU University Medical Center, De Boelelaan 1117, Amsterdam 1081 HV, The Netherlands.

Abstract:
In this study, measurements of guanidinoacetate (GAA) and creatine (Cr) in urine, plasma, and cerebrospinal fluid (CSF) were performed using stable isotope dilution gas chromatography-mass spectrometry. Both compounds were analyzed in a single analysis. Reference values were established for GAA and Cr. These values were age dependent. No differences with gender were observed. Eight guanidinoacetate methyltransferase (GAMT) deficient patients and eight creatine transporter SLC6A8 deficient patients were investigated. In urine, plasma, and CSF of GAMT deficient patients increased levels of GAA are present. The SLC6A8 deficient patients all show increased creatine/creatinine (Cr/Crn) ratio in urine demonstrating the importance of the Cr/Crn ratio as a pathognomonic marker of the SLC6A8 deficiency.
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:42
The ADP/ATP Carrier Protein
 01:27
Protein Import into the Peroxisomes
Peroxisomal Protein Import:
Peroxisomes lack the genetic machinery required to code for their own proteins. Hence, most peroxisomal membrane, lumenal and transmembrane proteins are synthesized in the cytoplasm or ER and transported to the peroxisome...
 01:20
Inborn Errors of Metabolism
 01:25
Drug Dosing in Renal Diseases: Measurement of Serum Creatinine Concentration and Clearance
 01:20
Blood Studies for Cardiovascular System I: Cardiac Biomarkers
The essential diagnostic tools for detecting myocardial necrosis and monitoring individuals suspected of having acute coronary syndrome (ACS) include:
Troponins
Troponins, particularly cardiac troponins I and T, are the most precise and sensitive markers of myocardial injury. They are detectable within 4-6 hours of myocardial injury and remain...