Classifying disease chromosomes arising from multiple founders, with application to fine-scale haplotype mapping
Related Experiment Video
Updated: Aug 22, 2026

Candidate Gene Testing in Clinical Cohort Studies with Multiplexed Genotyping and Mass Spectrometry
Published on: June 21, 2018
K Yu1, R B Martin, A S Whittemore
1Division of Biostatistics, School of Medicine, Washington University, St. Louis, Missouri 63110, USA. kai@wubios.wustl.edu

Abstract:
The availability of high-density haplotype data has motivated several fine-scale linkage disequilibrium mapping methods for locating disease-causing mutations. These methods identify loci around which haplotypes of case chromosomes exhibit greater similarity than do those of control chromosomes. A difficulty arising in such mapping is the possibility that case chromosomes have inherited disease-causing mutations from different ancestral chromosomes (founder heterogeneity). Such heterogeneity dilutes measures of case haplotype similarity. This dilution can be mitigated by separating case chromosomes into subsets according to their putative mutation origin, and searching for an area with excessive haplotype similarity within each subset. We propose a nonparametric method for identifying subsets of case chromosomes likely to share a common ancestral progenitor. By simulation studies and application to published data, we show that the method accurately identifies relatively large subsets of chromosomes that share a common founder. We also show that the method allows more precise estimates of the disease mutation loci than obtained by other fine-scale mapping methods.

Related Concept Videos
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
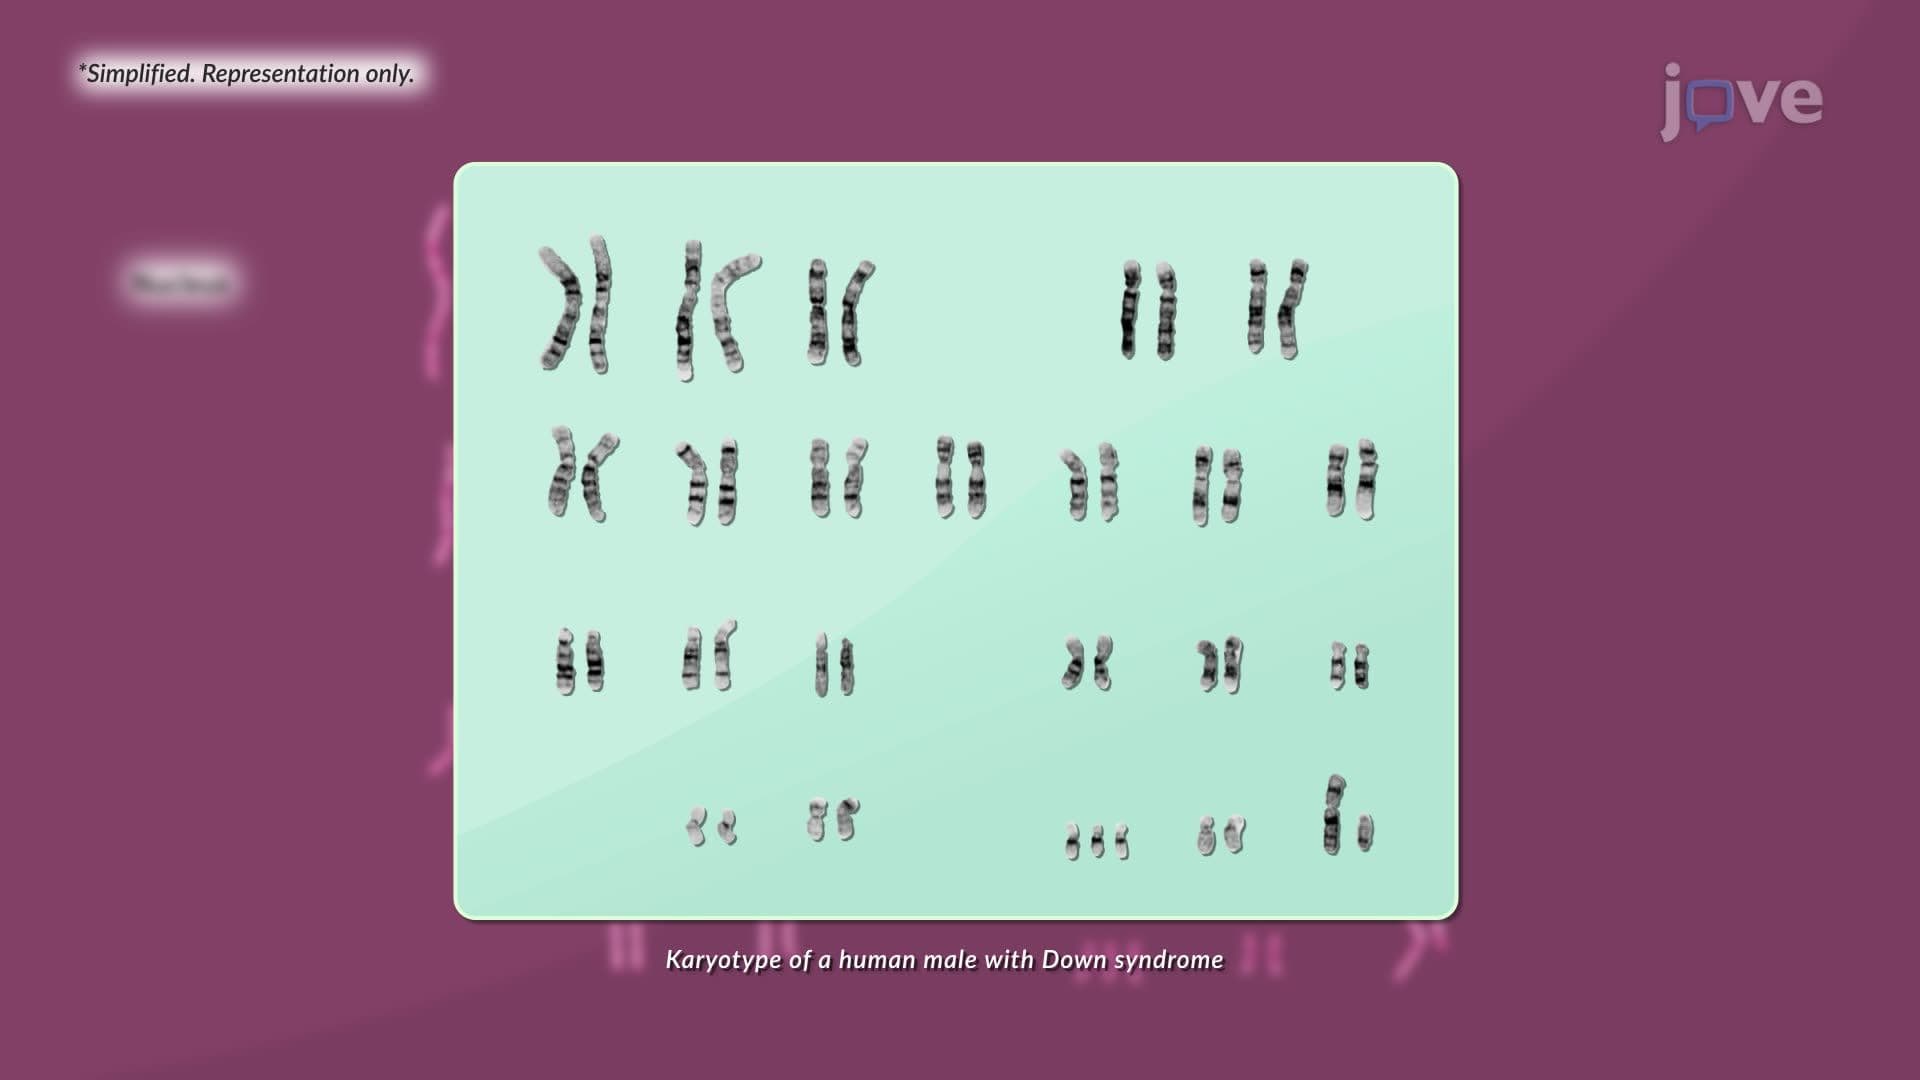 01:17
Karyotyping
 01:35
Pedigree Analysis
 01:43
Incomplete Dominance
 01:29
Modern Molecular Taxonomy
 01:39
Chromosomal Theory of Inheritance