Comparing hypothetical structures generated in the third Cambridge blind test of crystal structure prediction
Related Experiment Video
Updated: Aug 15, 2026

A Protocol for Computer-Based Protein Structure and Function Prediction
Published on: November 3, 2011
1Utrecht University, Department of Crystal and Structural Chemistry, Padualaan 8, 3584 CH Utrecht, The Netherlands. b.p.vaneijck@chem.uu.nl

Abstract:
In the third Cambridge blind test of crystal structure prediction, participants submitted extended lists of up to 100 hypothetical structures. In this paper these lists are analyzed for the two small semi-rigid molecules, hydantoin and azetidine, by performing a new energy minimization using an accurate force field, and grouping these newly minimized structures into clusters of equivalent structures. Many participants found the same low-energy structures, but no list appeared to be complete even for the structures with one independent molecule in the asymmetric unit. This may well be due to the fact that a cutoff at even 100 structures cannot ensure the presence of a structure that has a relatively high ranking in another force field. Moreover, some structures should have possibly been discarded because they correspond to transition states rather than true energy minima. The r.m.s. deviation between energies in corresponding clusters was calculated to compare the reported relative crystal energies for each pair of participants. Some groups of force fields show a reasonably good correspondence, yet the order of magnitude of their discrepancies is comparable to the energy differences between, say, the first ten structures of lowest energy. Therefore, even if we assume that energy is a sufficient criterion, it is not surprising that crystal structure predictions are still inconsistent and unreliable.
Related Concept Videos
 01:29
Determination of Crystal Structures
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:22
Structures of Solids
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:00
Molecular Models
 02:18
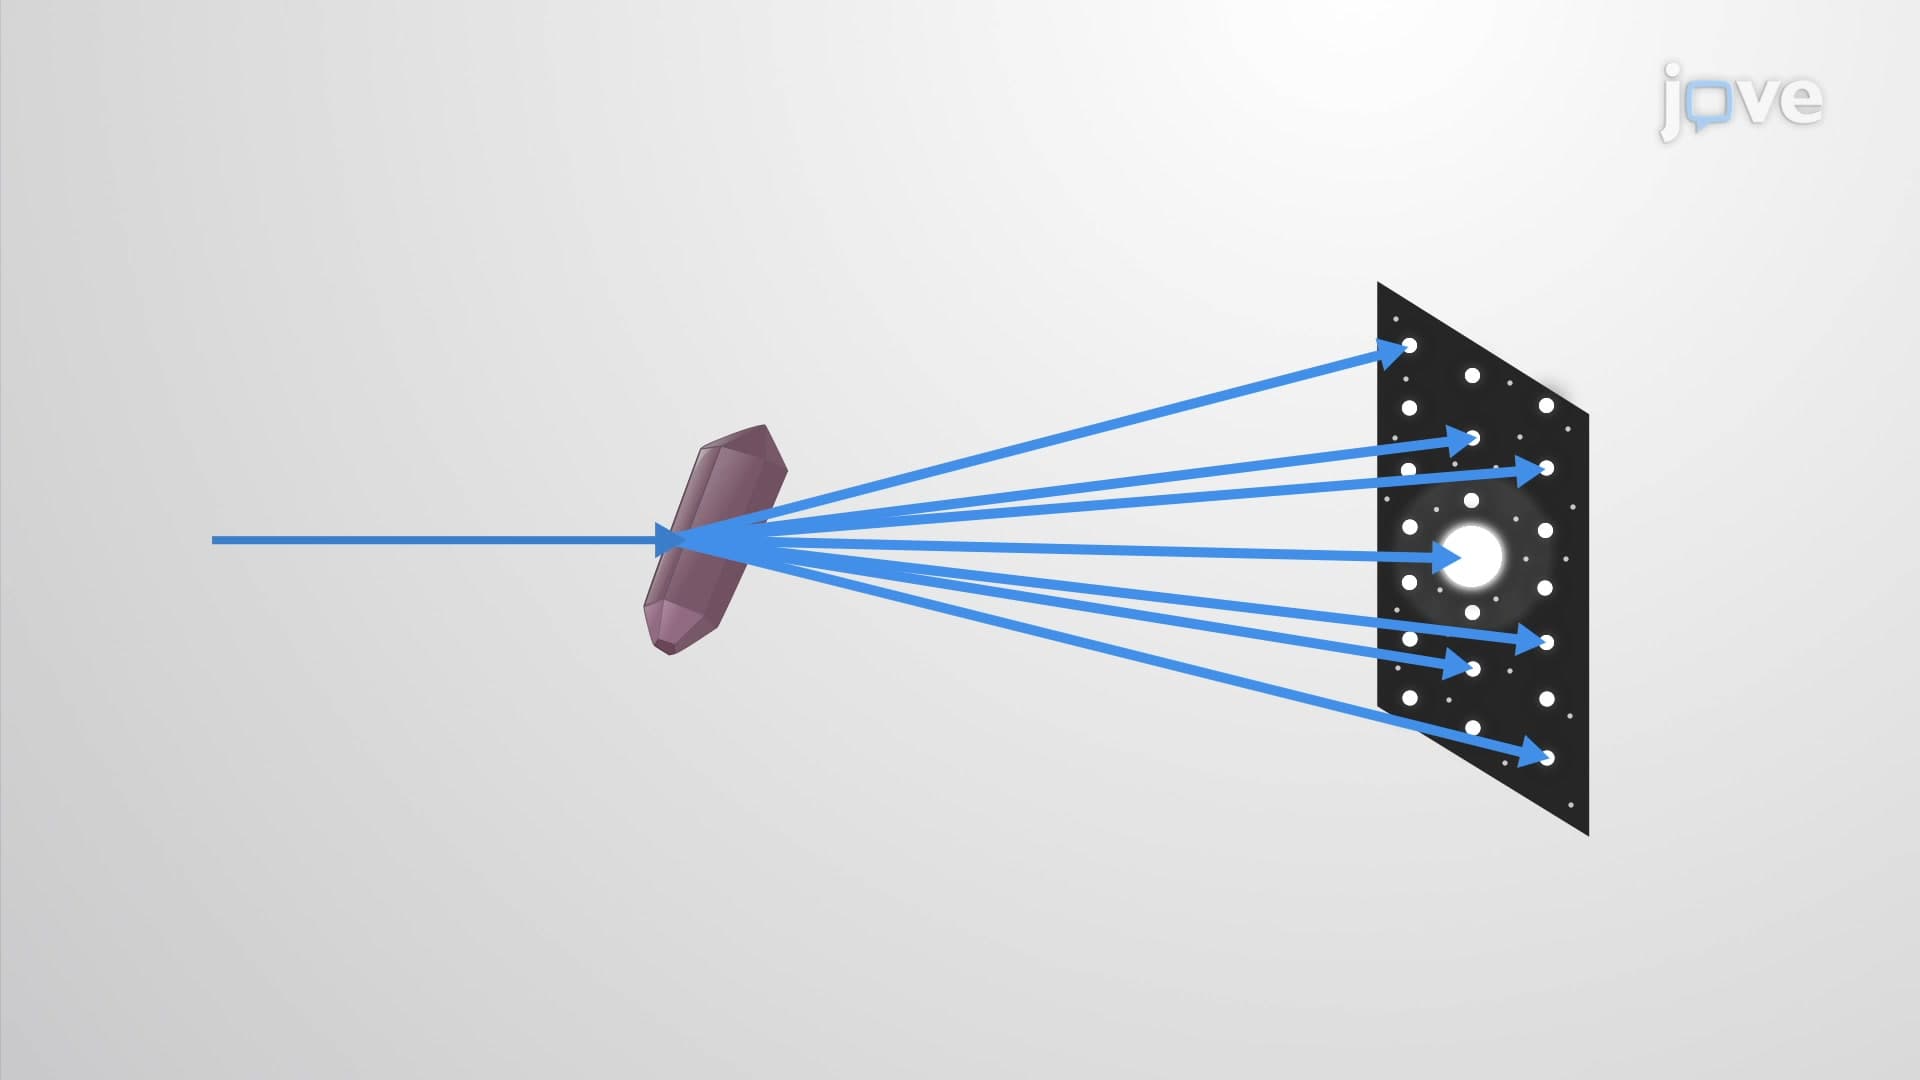
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...