Structure prediction of ordered and disordered multiple octahedral cation perovskites using SPuDS
Related Experiment Video
Updated: Aug 8, 2026
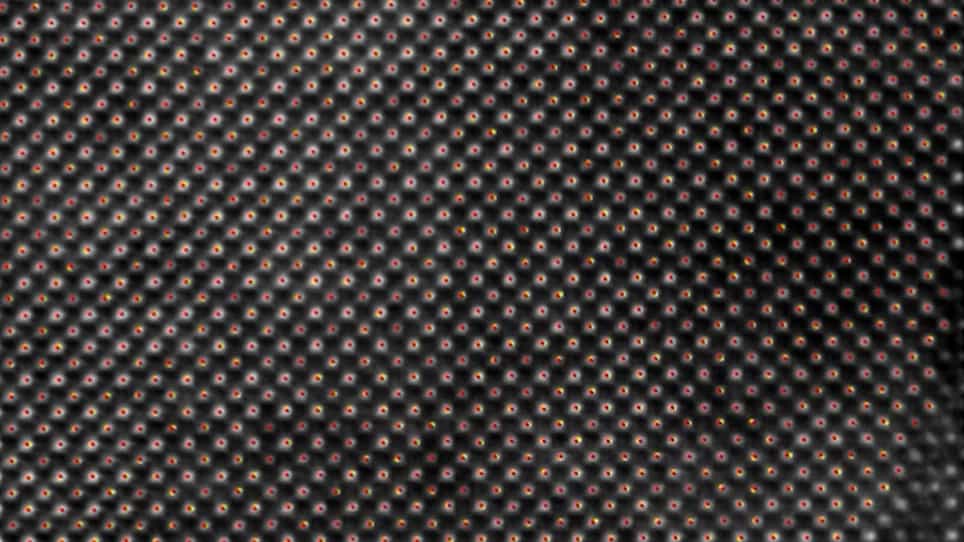
Picometer-Precision Atomic Position Tracking through Electron Microscopy
Published on: July 3, 2021
Michael W Lufaso1, Paris W Barnes, Patrick M Woodward
1Department of Chemistry, The Ohio State University, Columbus, OH 43210, USA.

Abstract:
The software package SPuDS has previously been shown to accurately predict crystal structures of AMX(3) and A(1 - x)A'(x)MX(3) perovskites that have undergone octahedral tilting distortions. This paper describes the extension of this technique and its accuracy for A(2)MM'X(6) ordered double perovskites with the aristotype Fm\overline 3m cubic structure, as well as those that have undergone octahedral tilting distortions. A survey of the literature shows that roughly 70% of all ordered double perovskites undergo octahedral tilting distortions. Of the 11 distinct types of octahedral tilting that can occur in ordered perovskites, five tilt systems account for approximately 97% of the reported structures. SPuDS can calculate structures for the five dominant tilt systems, Fm\overline 3m (a(0)a(0)a(0)), I4/m (a(0)a(0)c(-)), R\overline 3 (a(-)a(-)a(-)), I2/m (a(0)b(-)b(-)) and P2(1)/n (a(-)a(-)b(+)), as well as two additional tilt systems, Pn\overline 3 (a(+)a(+)a(+)) and P4/mnc (a(0)a(0)c(+)). Comparison with reported crystal structures shows that SPuDS is quite accurate at predicting distortions driven by octahedral tilting. The favored modes of octahedral tilting in ordered double perovskites are compared and contrasted with those in AMX(3) perovskites. Unit-cell pseudosymmetry in Sr- and Ca-containing double perovskites is also examined. Experimentally, Sr(2)MM'O(6) compounds show a much stronger tendency toward pseudosymmetry than do Ca(2)MM'O(6) compounds with similar tolerance factors.
Related Concept Videos
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:27
Predicting Molecular Geometry
 02:42
Valence Bond Theory
 02:52
VSEPR Theory and the Basic Shapes