Related Experiment Video
Updated: Jul 17, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Ab initio based potential energy surfaces and Franck-Condon analysis of ionization thresholds of cyclic-C3H and
Yimin Wang1, Bastiaan J Braams, Joel M Bowman
1Department of Chemistry and Cherry L. Emerson Center for Scientific Computation and Department of Mathematics and Computer Science, Emory University, Atlanta, Georgia 30322, USA.

Abstract:
We report a Franck-Condon analysis in reduced dimensionality of the ionization thresholds of linear(l)-C3H and cyclic(c)-C3H using MP2-based potential energy surfaces and CCSD(T)/aug-cc-pVTZ calculations of electronic energies at selected geometries. The potential energy surfaces are fits to tens of thousands of MP2/aug-cc-pVTZ energies for the neutral and cation systems. These fits properly describe the invariance of the potential with respect to all permutations of the three C atoms. The realism of the potential surfaces is assessed by comparing stationary-point structures, energies, and normal-mode frequencies with previous high-level ab initio calculations. Several key vibrational modes in this ionization process are located at saddle points and so a numerical approach to obtain the Franck-Condon factors for those modes is done. On the basis of this analysis combined with a simple harmonic treatment of the energies of the remaining modes and key electronic energy differences obtained with CCSD(T)/aug-cc-pVTZ calculations, we find the threshold ionization energy of l-C3H to be 9.06 eV and for c-C3H we estimate the threshold to be in the range 9.70-9.76 eV. We estimate these values are accurate to within +/-0.05 eV.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 07:53
Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
Related Concept Videos
 01:21
Chemical Ionization (CI) Mass Spectrometry
 01:18
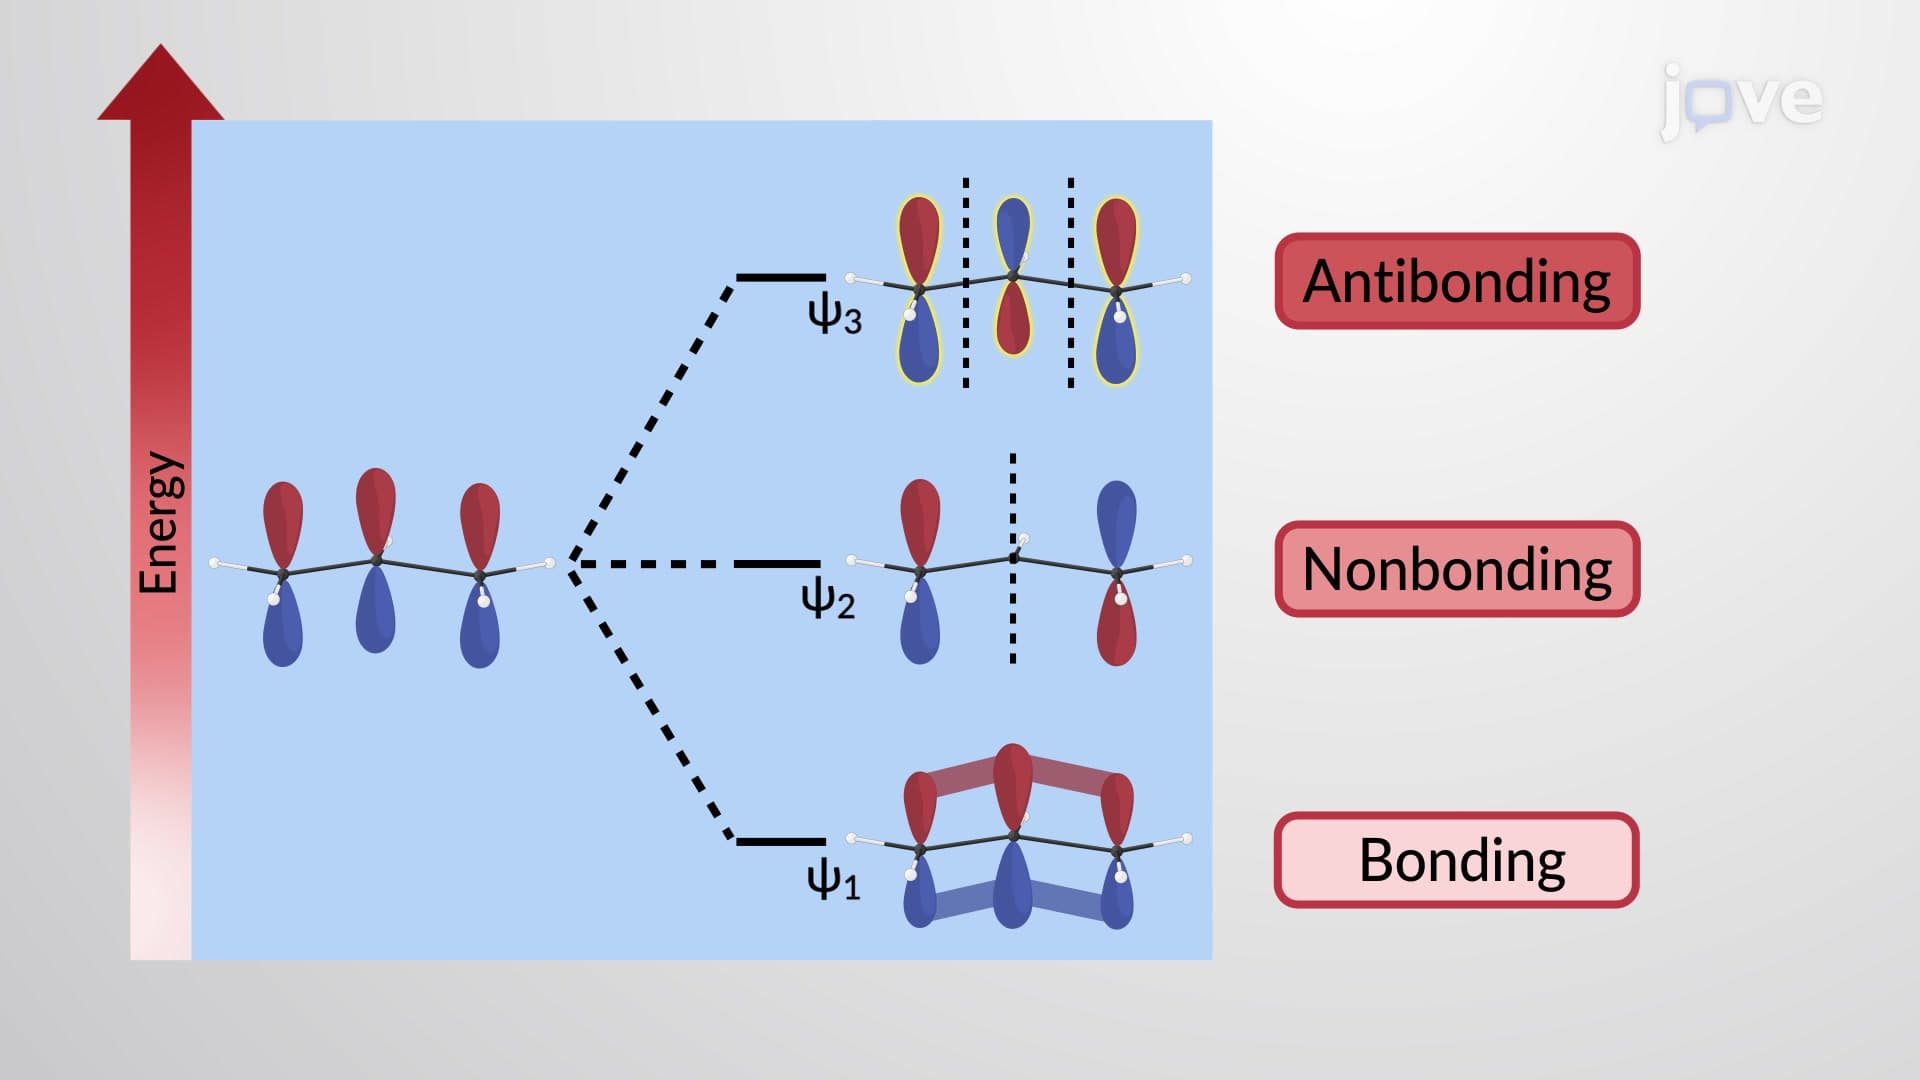
π Molecular Orbitals of the Allyl Cation and Anion
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:05
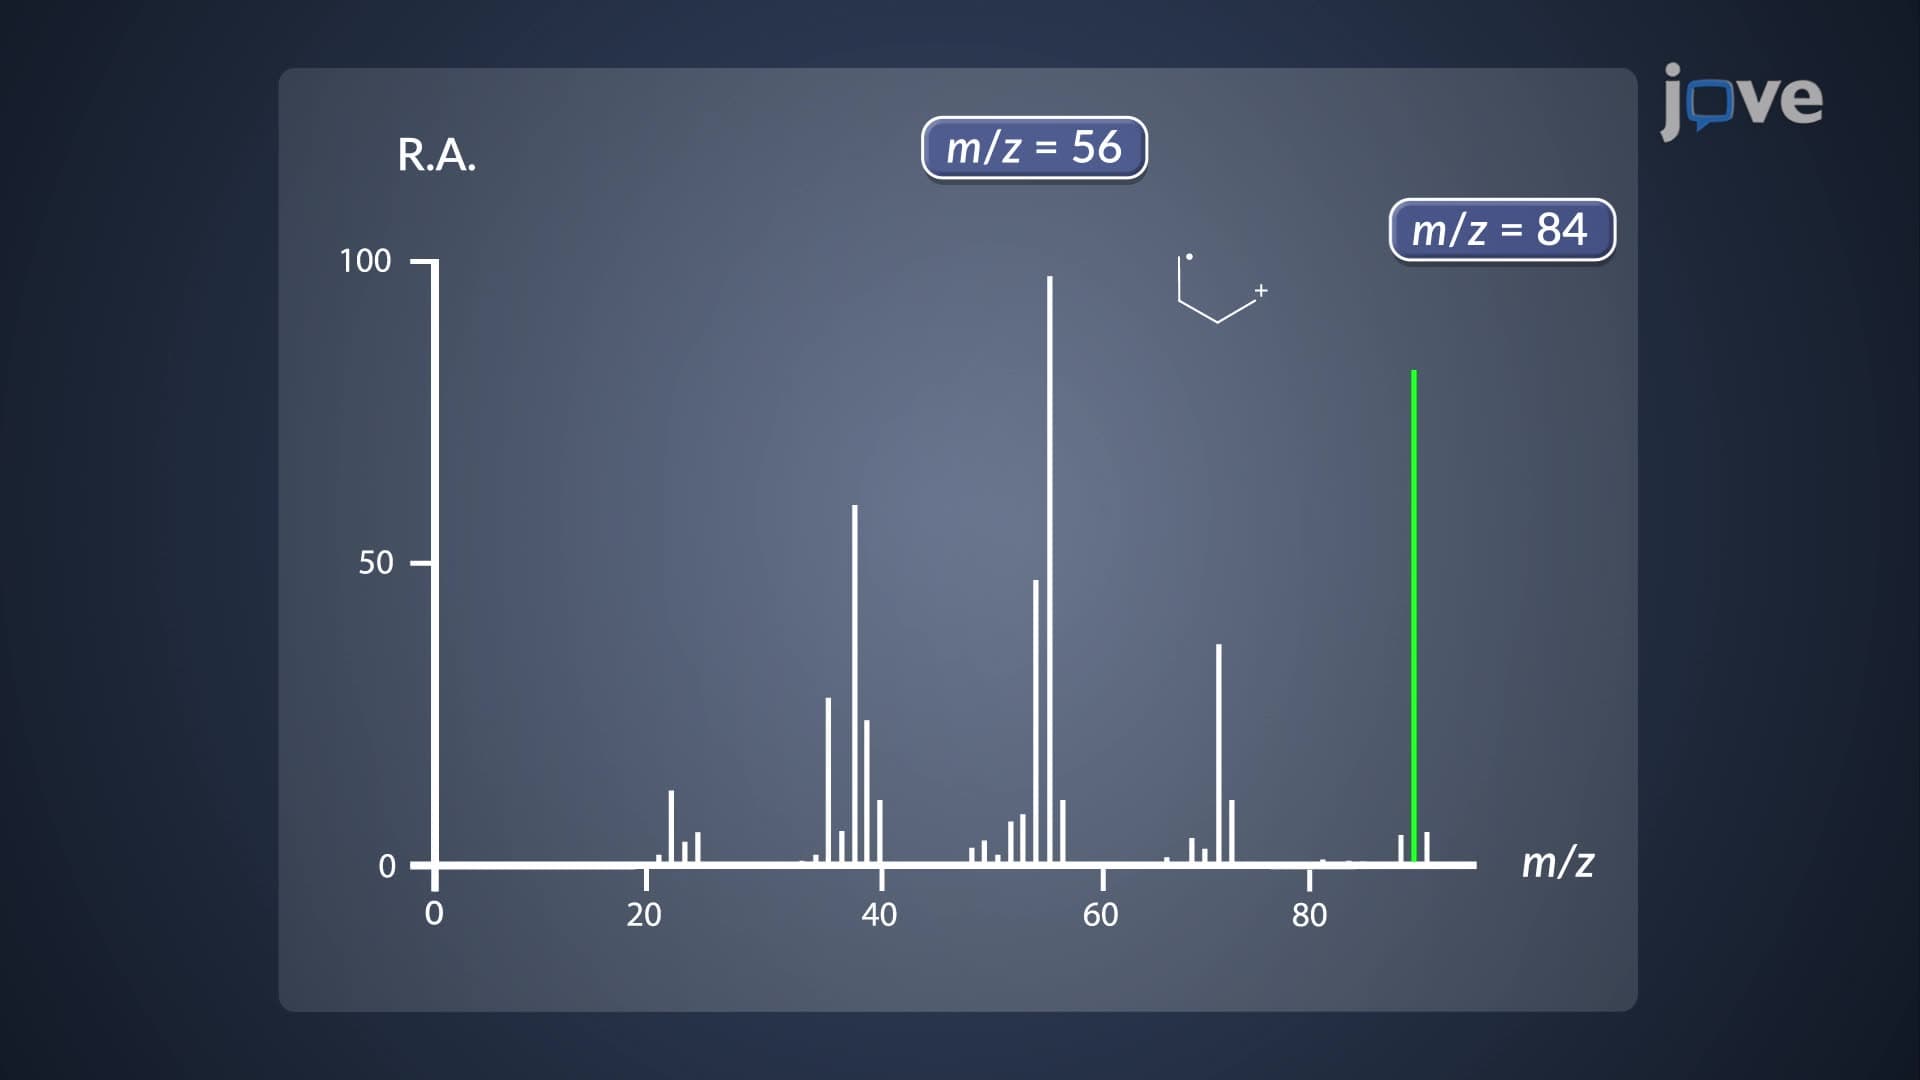
Mass Spectrometry: Cycloalkane Fragmentation
For example, cyclohexane molecular ions have a mass-to-charge ratio (m/z) of 84, which tends to produce a stronger signal than linear alkanes like hexane. This stability comes from...
 01:18
Aromatic Hydrocarbon Cations: Structural Overview
Removing one hydrogen from the intervening CH2 group with both...
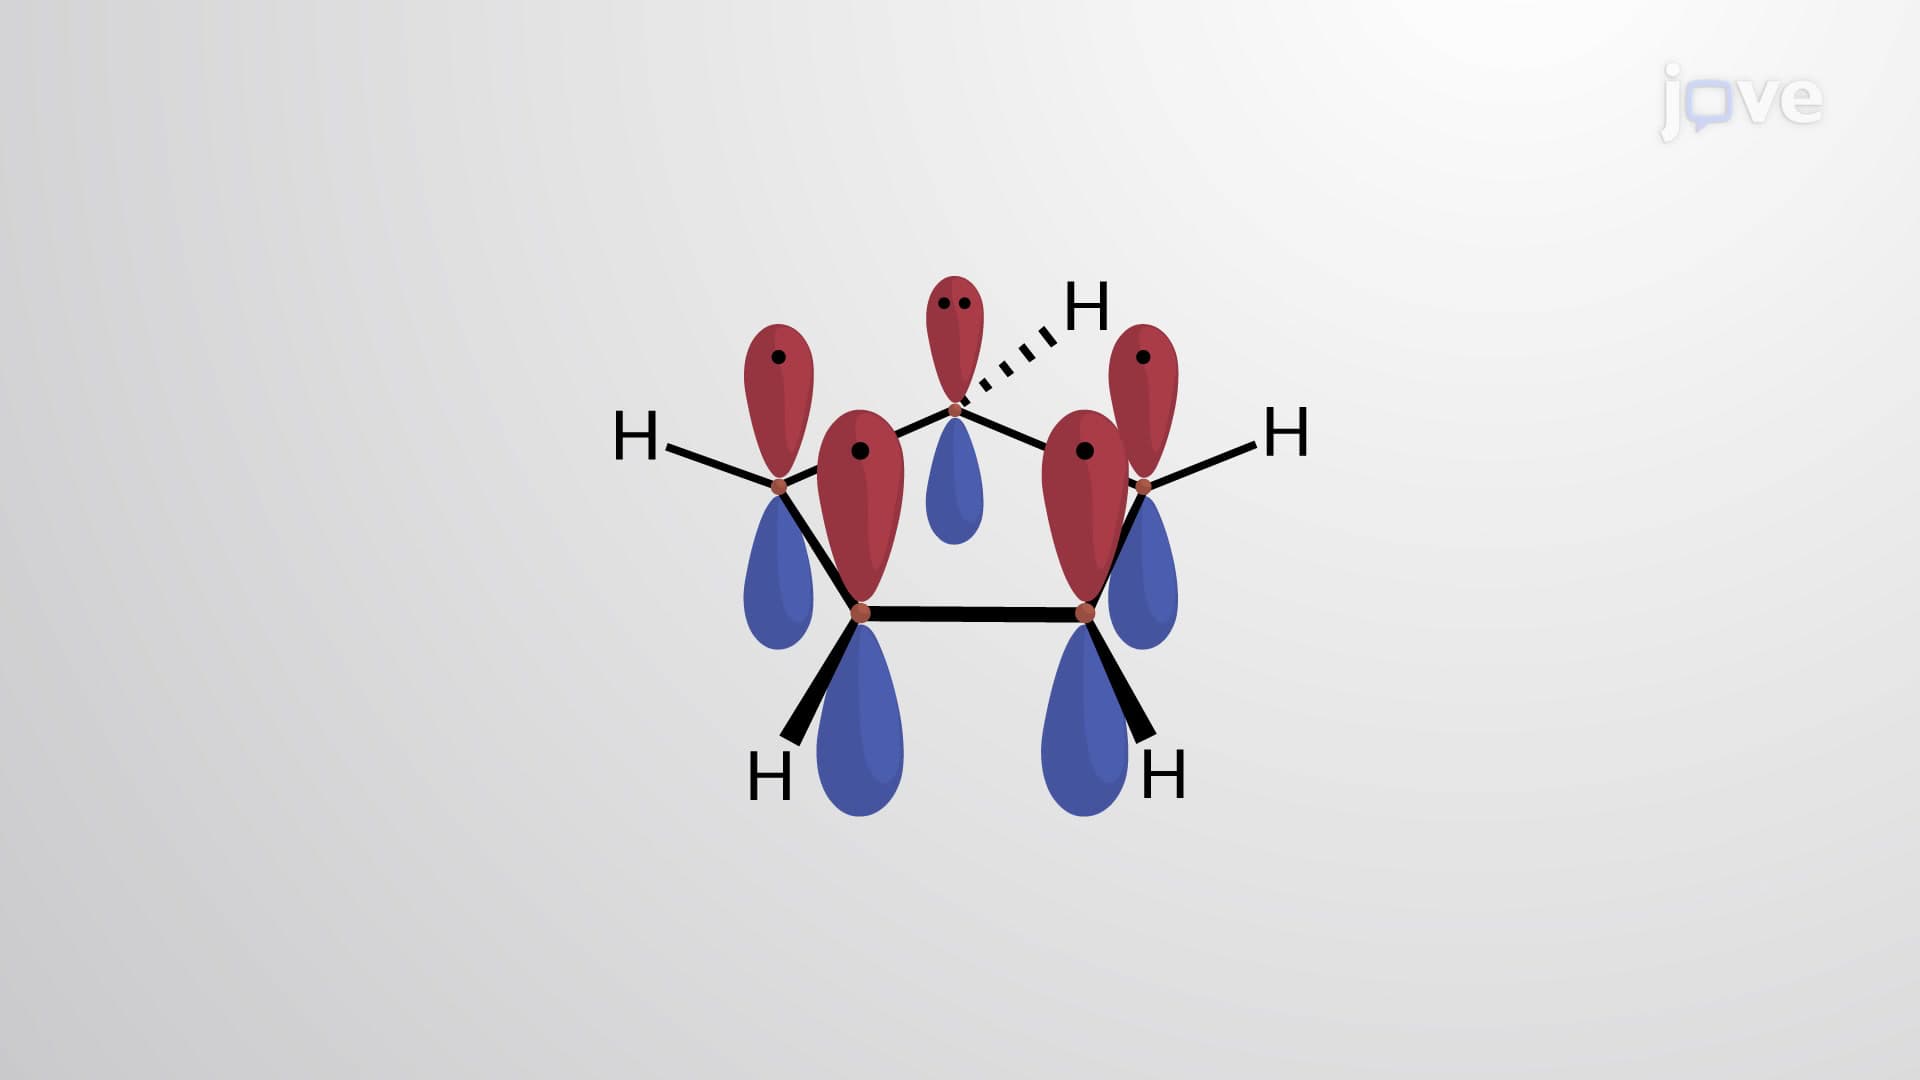 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...