Related Experiment Video
Updated: Jul 5, 2026

Thermochemical Studies of Ni(II) and Zn(II) Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
The role of exchange-correlation functionals in the potential energy surface and dynamics of N(2) dissociation on W
G A Bocan1, R Díez Muiño, M Alducin
1Donostia International Physics Center (DIPC), P Manuel de Lardizabal 4, San Sebastián, Spain.

Abstract:
We study the dissociative adsorption of N(2) on W(100) and W(110) by means of density functional theory and classical dynamics. Working with a full six-dimensional adiabatic potential energy surface (PES), we find that the theoretical results of the dynamical problem strongly depend on the choice of approximate exchange-correlation functional for the determination of the PES. We consider the Perdew-Wang-91 [Perdew et al., Phys. Rev. B 46, 6671 (1992)] and Perdew-Burke-Ernzerhof (RPBE) [Hammer et al., Phys. Rev. B 59, 7413 (1999)] functionals and carry out a systematic comparison between the dynamics determined by the respective PESs. Even though it has been shown in earlier works that the RPBE may provide better values for the chemisorption energies, our study brings evidence that it gives rise to a PES with excessive repulsion far from the surface.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 07:53
Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:13
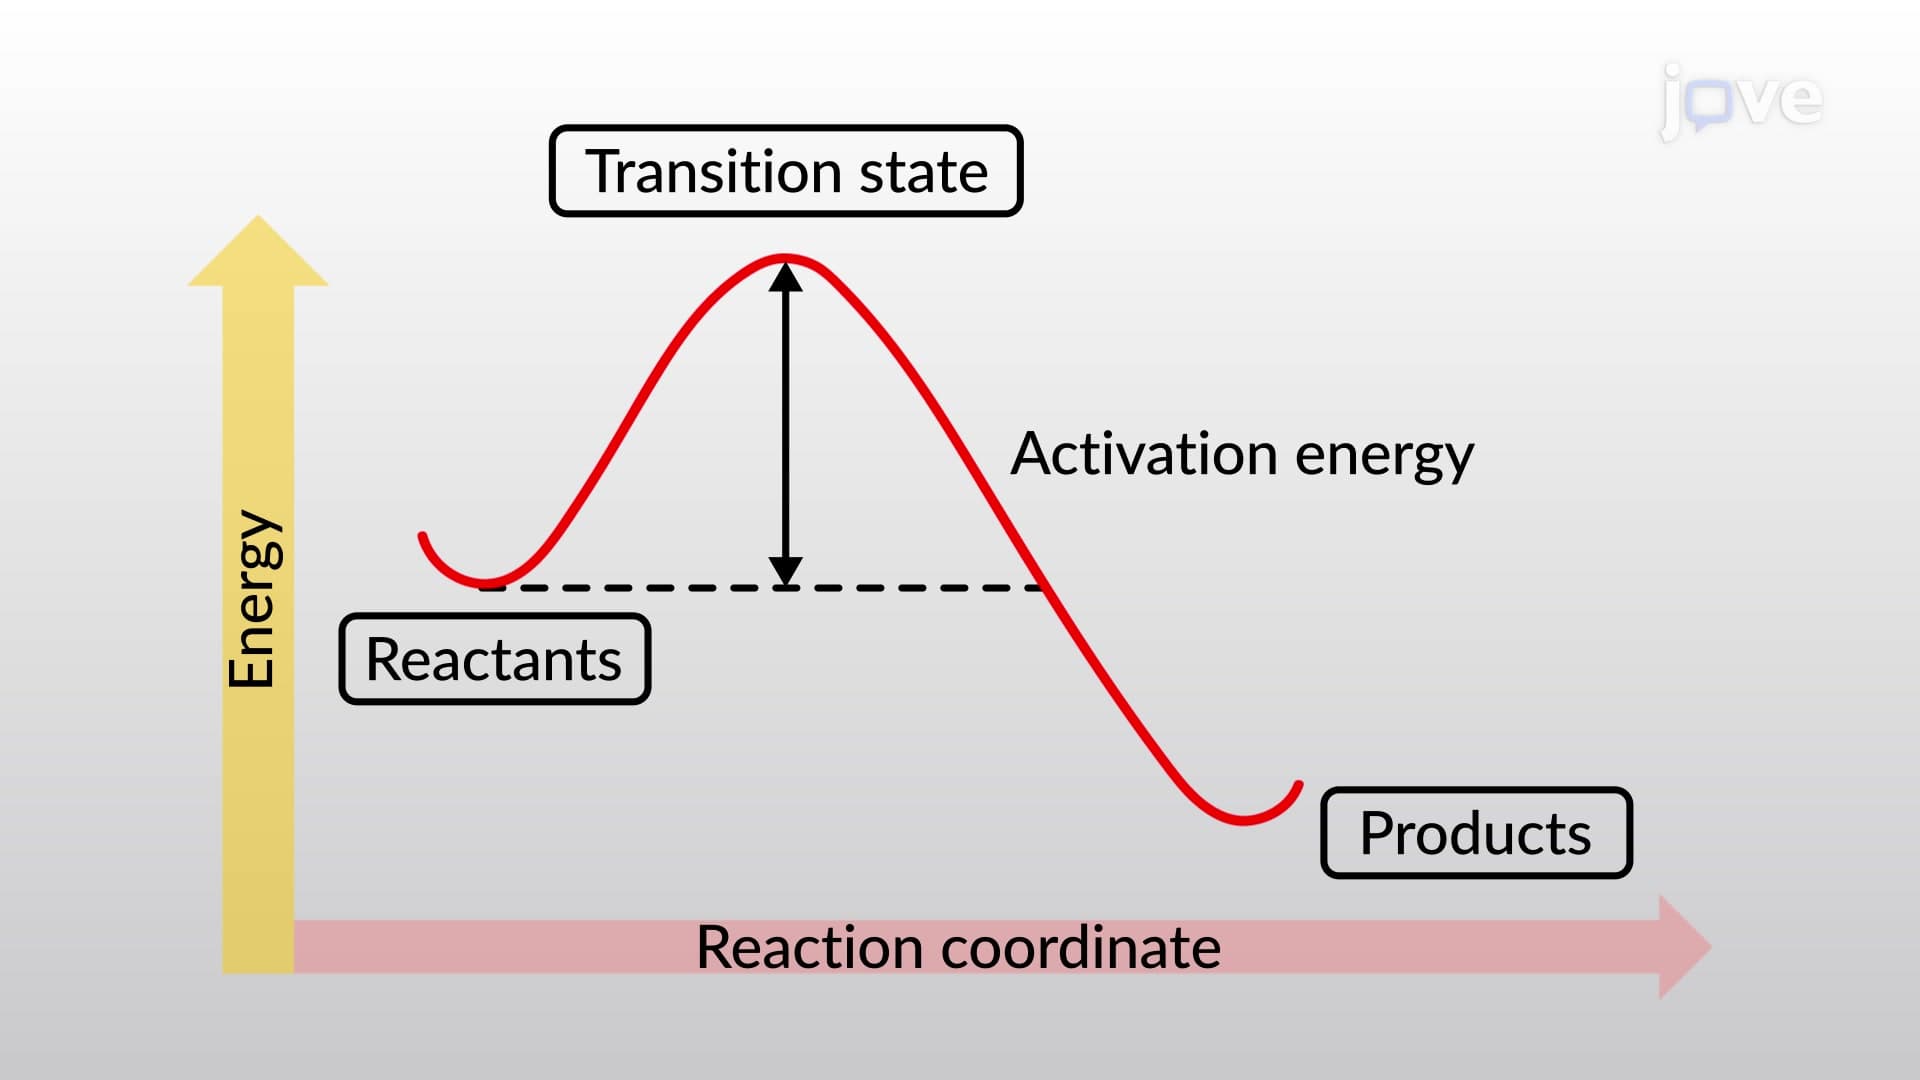
Bond Dissociation Energy and Activation Energy
 02:45
Valence Bond Theory
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:40
MO Theory and Covalent Bonding