Related Experiment Video
Updated: Jun 25, 2026

Interactive Molecular Model Assembly with 3D Printing
Published on: August 13, 2020
Conformational stability from rare gas solutions, r0 structural parameters, barriers to internal rotation, and ab
Yasser E Nashed1, Mohammad A Qtaitat, Chao Zheng
1Department of Chemistry, University of Missouri-Kansas City, Kansas City, Missouri 64110, USA.

Abstract:
The Raman (3300-10 cm(-1)) and infrared (3300-40 cm(-1)) spectra of gaseous and solid vinyl silyl fluoride, CH(2)=CHSiH(2)F, have been recorded. Raman spectrum of the liquid has also been recorded and depolarization values obtained. Variable-temperature studies of the infrared spectra of the sample dissolved in liquid krypton (-110 to -150 degrees C) and liquid xenon (-60 to -100 degrees C) have been carried out. From these studies, the enthalpy difference has been determined to be 76 +/- 7 cm(-1) (0.91 +/- 0.08 kJ/mol) from the krypton solutions and 69 +/- 7 cm(-1) (0.82 +/- 0.08 kJ/mol) from the xenon solutions, with the gauche conformer the more stable form. From the far-infrared spectrum of the gas, the asymmetric torsional fundamentals for the cis and gauche conformers have been observed at 102.34 and 86.56 cm(-1), respectively, with each having several "hot bands" falling to lower frequencies. From these frequencies along with the experimentally determined conformational enthalpy difference, as well as the gauche skeletal dihedral angle, the potential function governing the conformational interchange has been determined with the following Fourier cosine potential coefficients: V(1) = -80 +/- 11, V(2) = -42 +/- 15, V(3) = 622 +/- 5, V(4) = 34 +/- 5, and V(6) = -31 +/- 2 cm(-1). The gauche-to-cis and gauche-to-gauche barriers are 664 cm(-1) (7.94 kJ/mol) and 608 cm(-1) (7.27 kJ/mol), respectively. Complete vibrational assignments are provided for both conformers. In addition, equilibrium geometries and electronic energies have been determined for both rotamers from ab initio calculations using restricted Hartree-Fock and Møller-Plesset perturbation method to the second order (MP2), as well as density functional theory by the B3LYP methods, employing a number of basis sets up to 6-311+G(2df,2pd). All levels of calculation predict the gauche conformer to be the more stable form. By systematically adjusting the ab initio predicted structural values to fit the previously reported microwave rotational constants, adjusted r(0) parameters have been obtained for both conformers. These values are compared to those for the corresponding chloride and methyl compounds. The spectroscopic and theoretical results are discussed and compared to the corresponding quantities for some similar molecules.

Related Concept Videos
 02:18
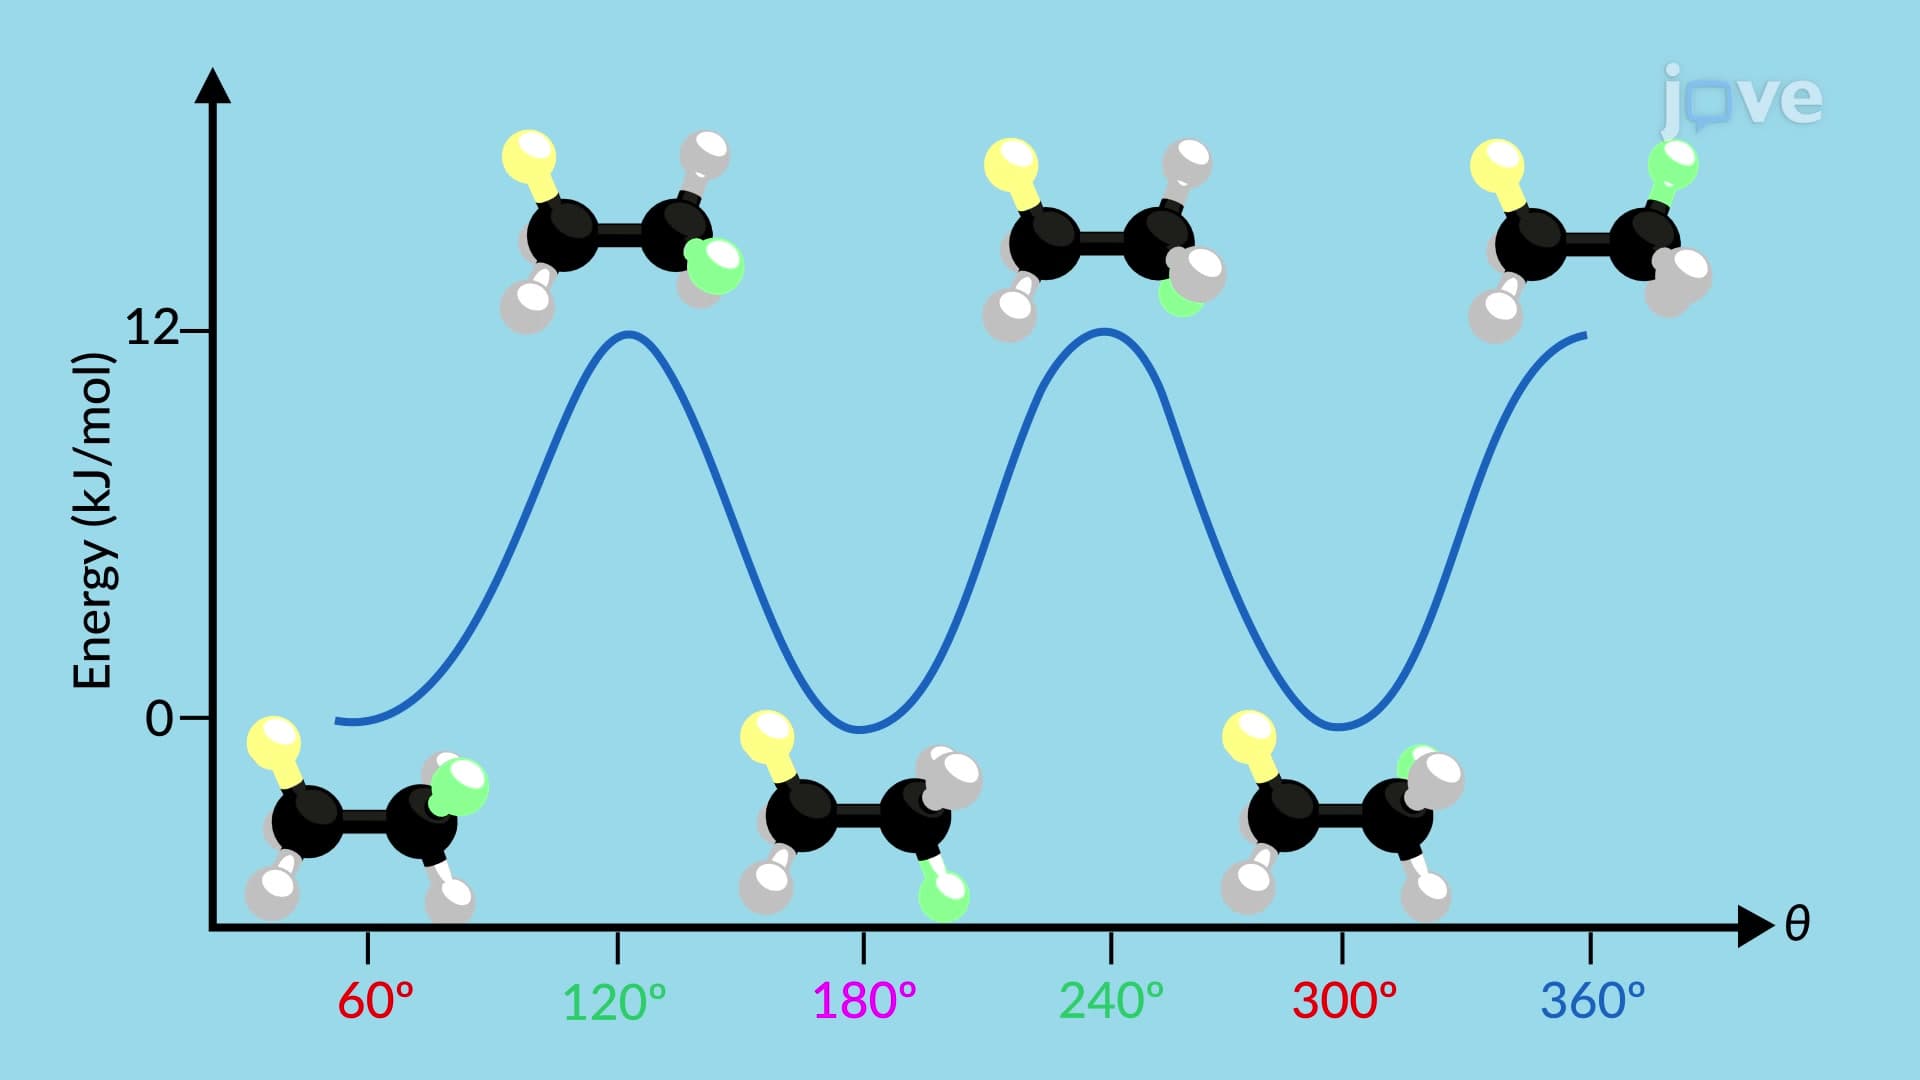
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...
 02:29
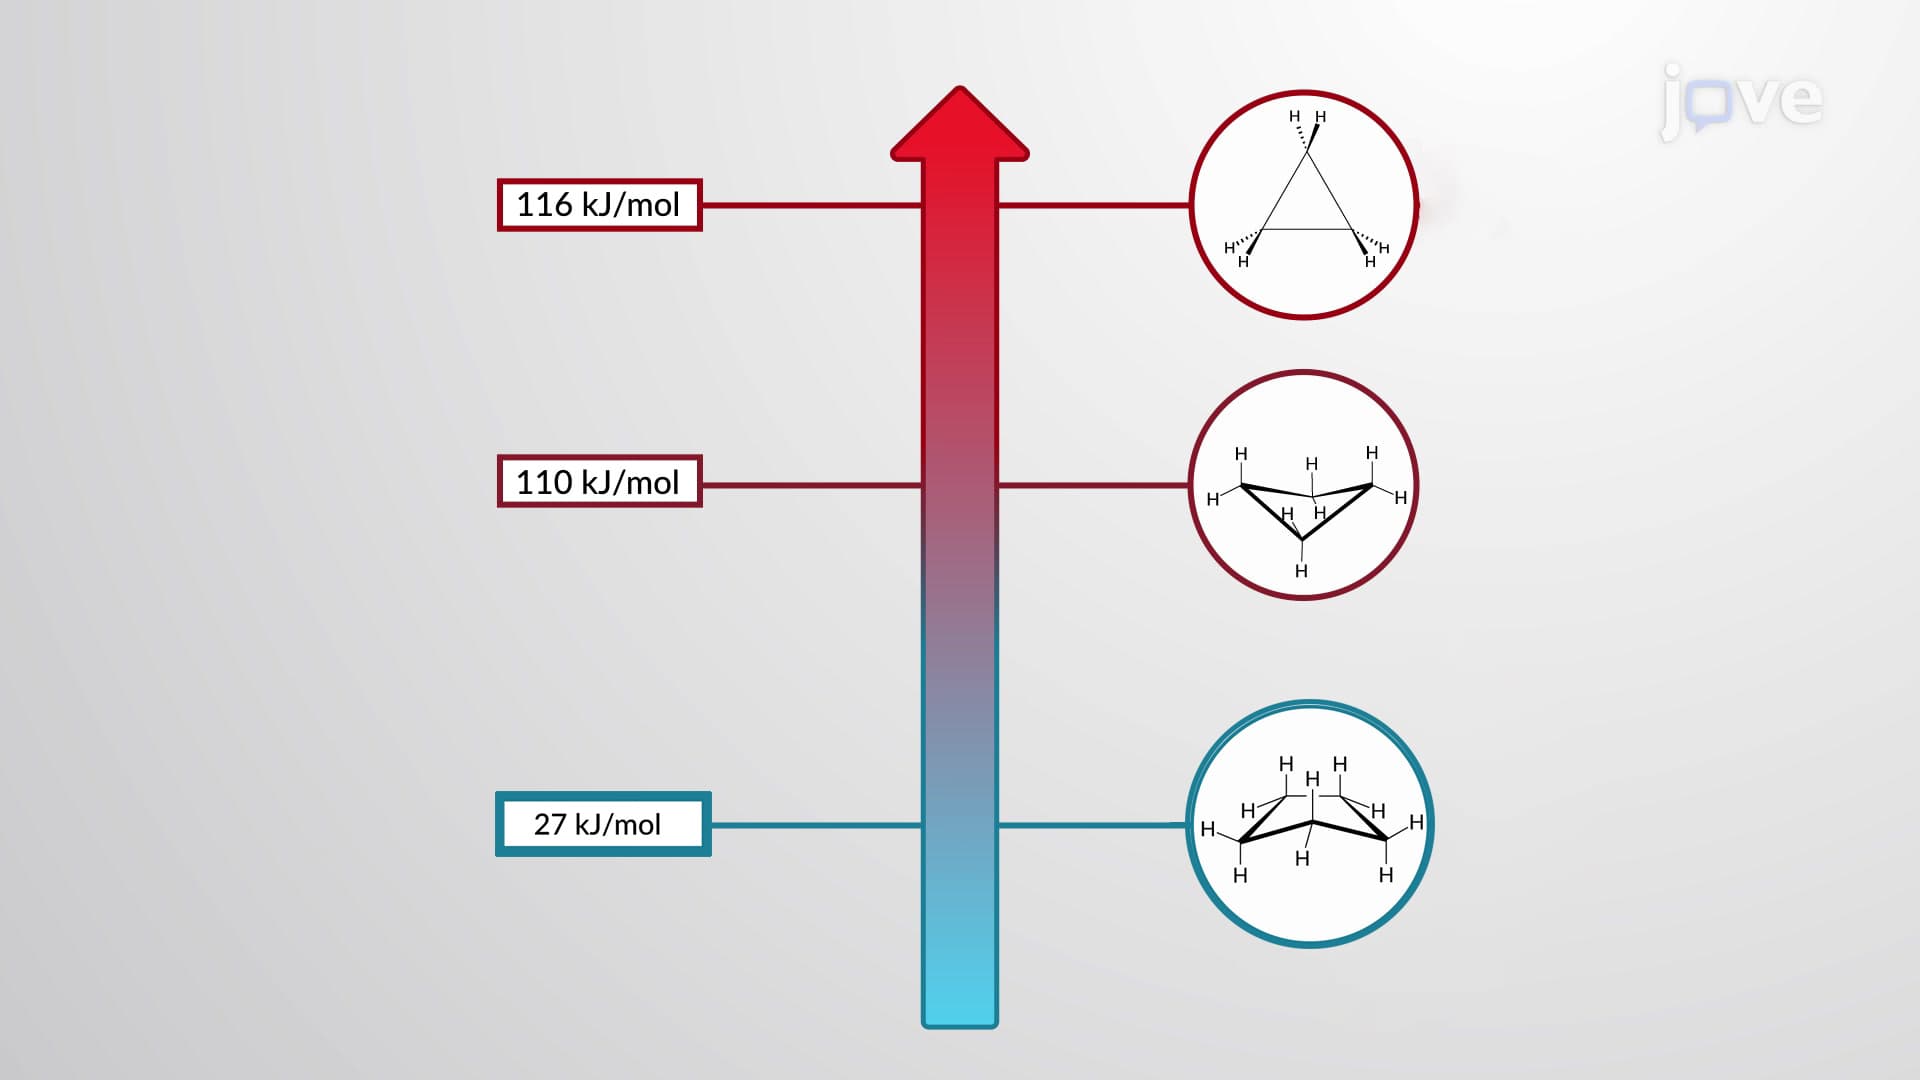
Conformations of Cycloalkanes