Related Experiment Video
Updated: Jun 21, 2026

Visualizing the Conformational Dynamics of Membrane Receptors Using Single-Molecule FRET
Published on: August 17, 2022
Mutational analyses of HAMP helices suggest a dynamic bundle model of input-output signalling in chemoreceptors
Qin Zhou1, Peter Ames, John S Parkinson
1Biology Department, University of Utah, Salt Lake City, UT 84112, USA.

Abstract:
To test the gearbox model of HAMP signalling in the Escherichia coli serine receptor, Tsr, we generated a series of amino acid replacements at each residue of the AS1 and AS2 helices. The residues most critical for Tsr function defined hydrophobic packing faces consistent with a four-helix bundle. Suppression patterns of helix lesions conformed to the predicted packing layers in the bundle. Although the properties and patterns of most AS1 and AS2 lesions were consistent with both proposed gearbox structures, some mutational features specifically indicate the functional importance of an x-da bundle over an alternative a-d bundle. These genetic data suggest that HAMP signalling could simply involve changes in the stability of its x-da bundle. We propose that Tsr HAMP controls output signals by modulating destabilizing phase clashes between the AS2 helices and the adjoining kinase control helices. Our model further proposes that chemoeffectors regulate HAMP bundle stability through a control cable connection between the transmembrane segments and AS1 helices. Attractant stimuli, which cause inward piston displacements in chemoreceptors, should reduce cable tension, thereby stabilizing the HAMP bundle. This study shows how transmembrane signalling and HAMP input-output control could occur without the helix rotations central to the gearbox model.
Related Concept Videos
 01:09
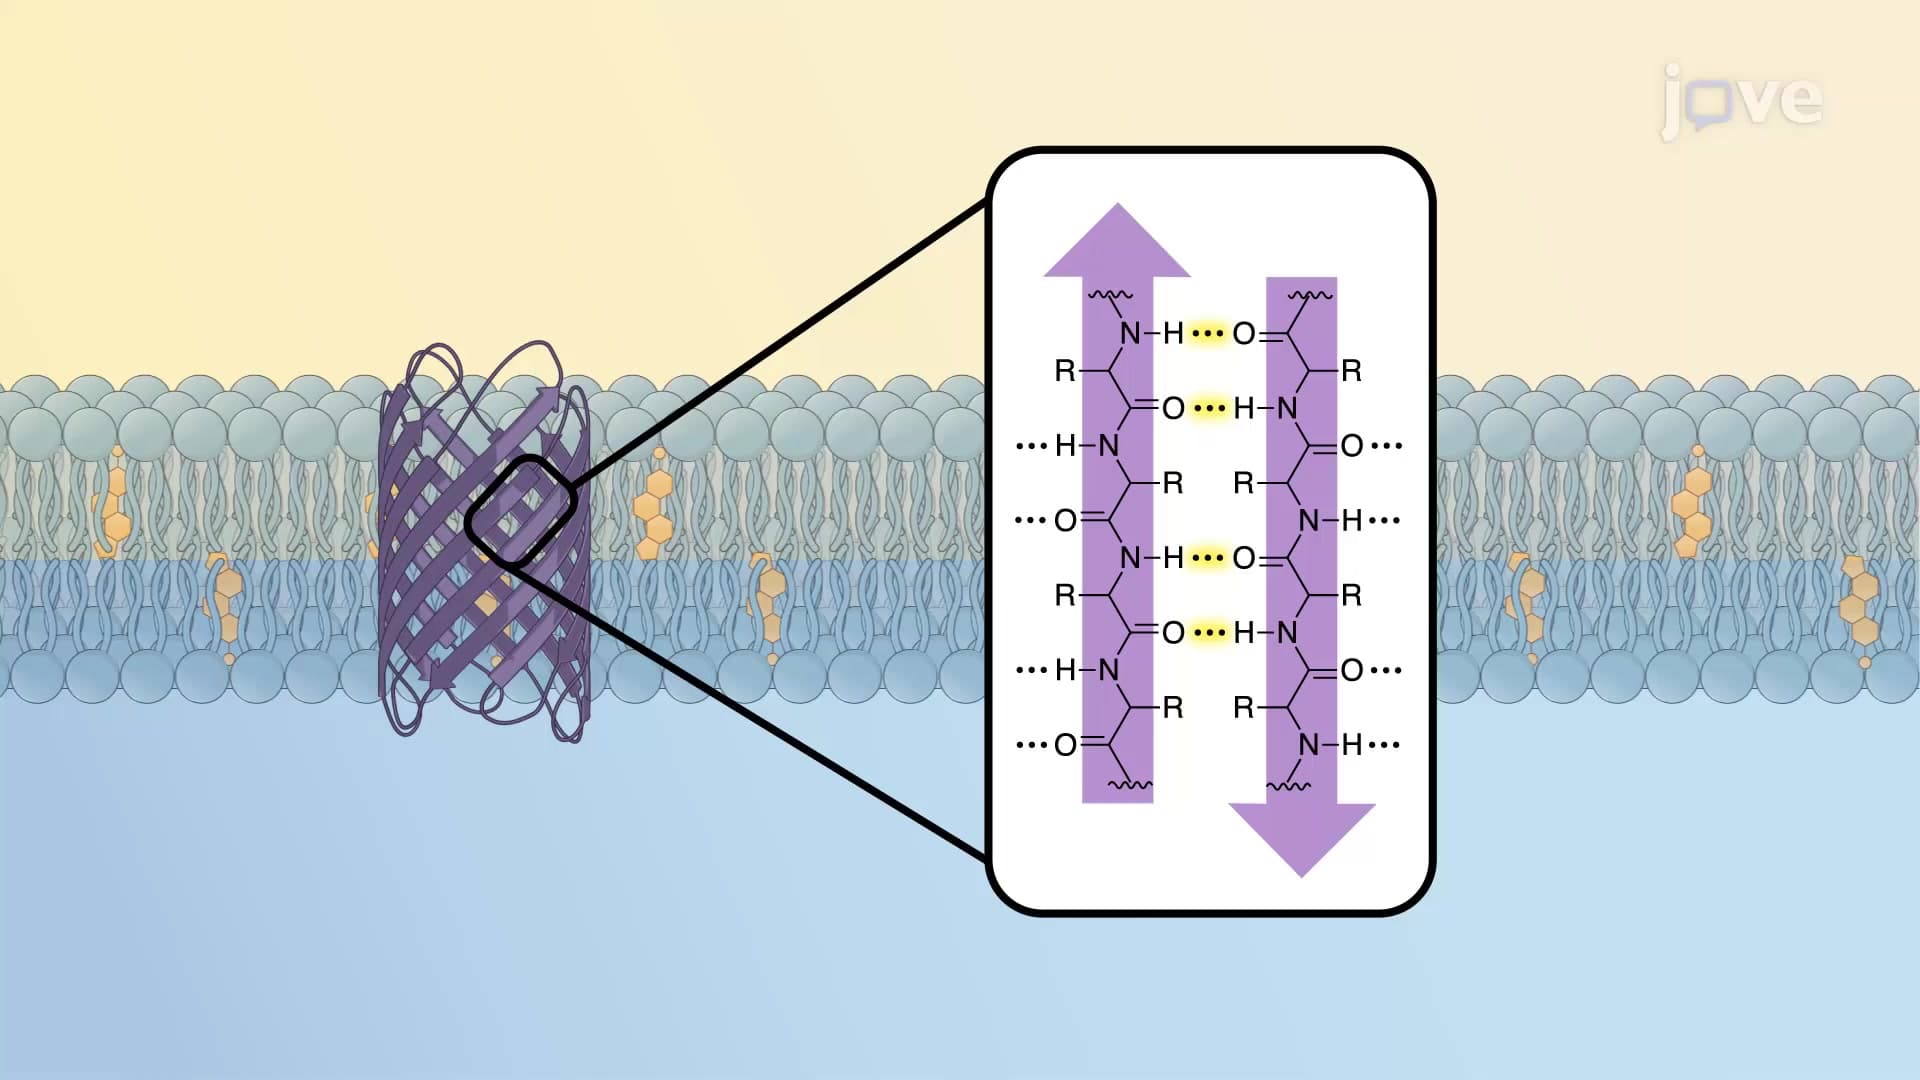
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as G-protein-linked receptors (GPCRs) and...
 01:30
Assembly of Signaling Complexes
Interaction domains in cell signaling
Interaction domains recognize exposed features of their binding partners containing post-translationally modified sequences,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:58
Cooperative Allosteric Transitions
 03:00
Molecular Chaperones and Protein Folding
The...
 01:22
Amplifying Signals via Enzymatic Cascade