Related Experiment Video
Updated: Jun 4, 2026
A Practical Guide to Phylogenetics for Nonexperts
Published on: February 5, 2014

Abstract:
INTRODUCTIONThree methods--maximum parsimony, distance, and maximum likelihood--are generally used to find the evolutionary tree or trees that best account for the observed variation in a group of sequences. Each of these methods uses a different type of analysis. Programs based on distance methods are commonly used in the molecular biology laboratory because they are straightforward and can be used with a large number of sequences. Maximum likelihood methods are more challenging and require a greater understanding of the evolutionary models on which they are based. Because they involve so many computational steps and because the number of steps increases dramatically with the number of sequences, maximum likelihood programs are limited to a smaller number of sequences. They can be implemented on a supercomputer in order to analyze a greater number of sequences. This article presents an overview for the researcher who has a set of related sequences and wants to analyze them to predict the best trees that depict the phylogenetic relationships among the sequences.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 01:28
Microbial Phylogeny
 02:05
Gene Evolution - Fast or Slow?
In contrast, regions which code...
02:05Gene Evolution - Fast or Slow?
In contrast, regions which code...
 03:21
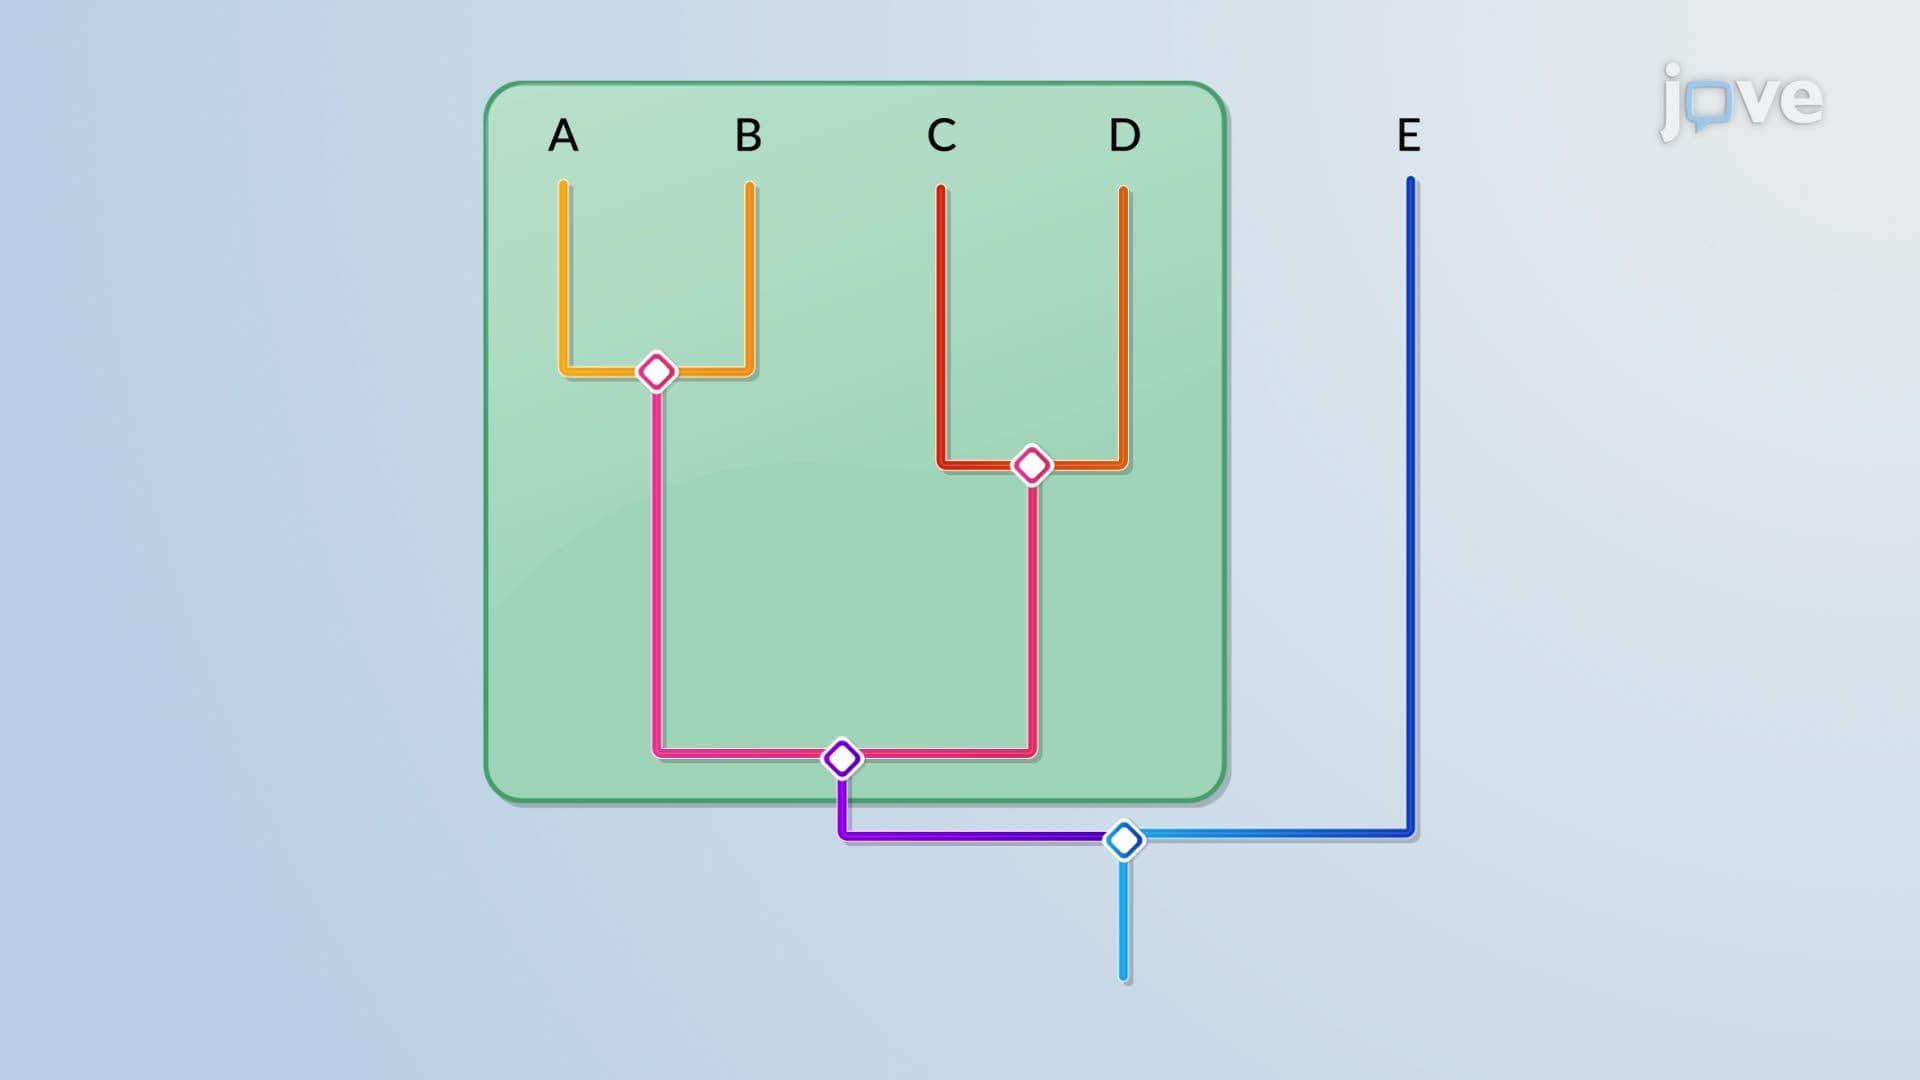
Phylogenetic Trees
03:21Phylogenetic Trees