MDAnalysis: a toolkit for the analysis of molecular dynamics simulations
Related Experiment Video
Updated: Jun 2, 2026
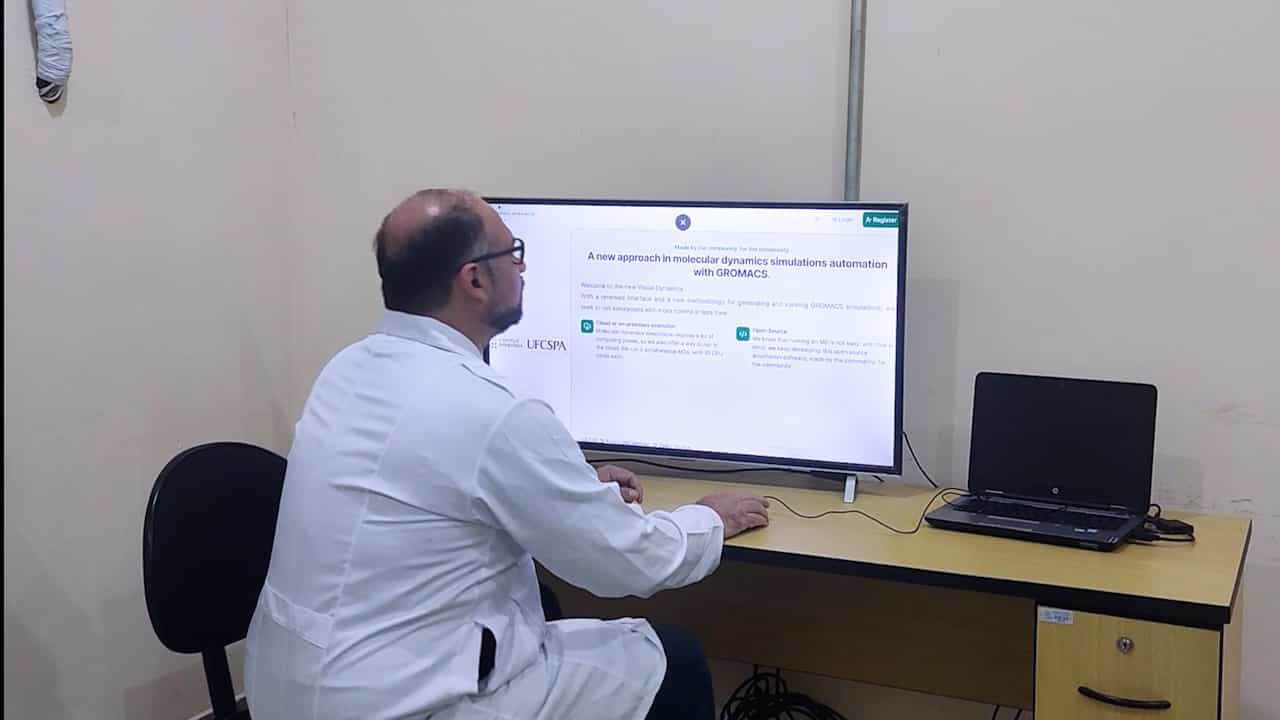
New Features in Visual Dynamics 3.0
Published on: August 9, 2024
Naveen Michaud-Agrawal1, Elizabeth J Denning, Thomas B Woolf
1Department of Biophysics, Johns Hopkins School of Medicine, Baltimore, Maryland 21205.

Abstract:
MDAnalysis is an object-oriented library for structural and temporal analysis of molecular dynamics (MD) simulation trajectories and individual protein structures. It is written in the Python language with some performance-critical code in C. It uses the powerful NumPy package to expose trajectory data as fast and efficient NumPy arrays. It has been tested on systems of millions of particles. Many common file formats of simulation packages including CHARMM, Gromacs, Amber, and NAMD and the Protein Data Bank format can be read and written. Atoms can be selected with a syntax similar to CHARMM's powerful selection commands. MDAnalysis enables both novice and experienced programmers to rapidly write their own analytical tools and access data stored in trajectories in an easily accessible manner that facilitates interactive explorative analysis. MDAnalysis has been tested on and works for most Unix-based platforms such as Linux and Mac OS X. It is freely available under the GNU General Public License from http://mdanalysis.googlecode.com.