Related Experiment Video
Updated: May 26, 2026
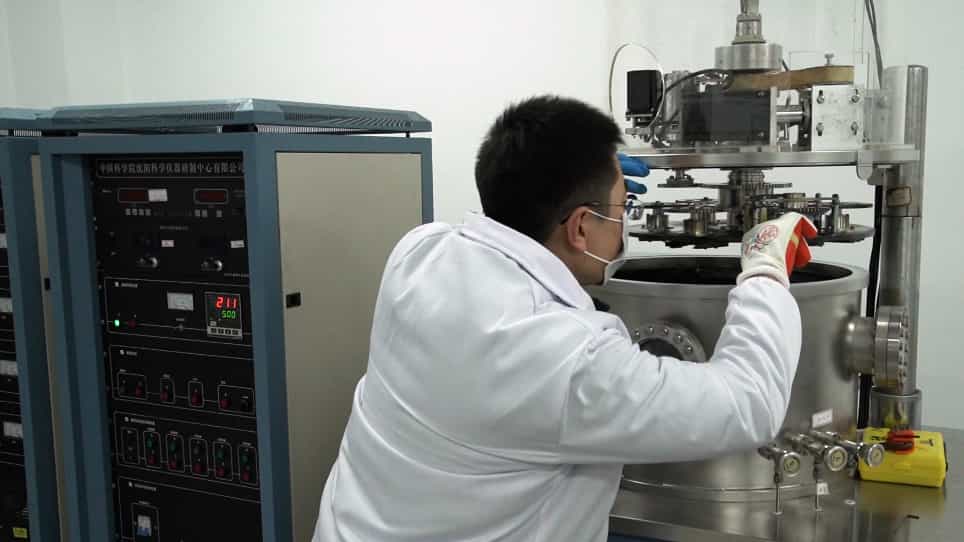

Radio Frequency Magnetron Sputtering of GdBa2Cu3O7−δ/ La0.67Sr0.33MnO3 Quasi-bilayer Films on SrTiO3 (STO) Single-crystal Substrates
Published on: April 12, 2019
Defect structure and Fermi-level pinning of BaTiO3 co-doped with a variable-valence acceptor (Mn) and a fixed-valence
Yun-Young Yeoh1, Hyejin Jang, Han-Ill Yoo
1WCU Hybrid Materials Program, Department of Materials Science and Engineering, Seoul National University, Seoul 151-744, Korea.

Abstract:
Defect structures of BaTiO(3) and the like co-doped with variable-valence acceptors and donors are not clear particularly in transition from acceptor domination to donor domination with increasing oxygen activity. We have, thus, examined the electrical conductivity and thermoelectric power of BaTiO(3) co-doped with a variable-valence acceptor Mn(Mn(Ti)'', Mn(Ti)') and a fixed-valence donor Y(Y(ba)·) in different co-doping ratios (m(d)/m(a)) as functions of oxygen activity in the range of -20 < log a(O(2))≤ 0 at elevated temperatures of 900-1100 °C. Their systematic variations with m(d)/m(a) and log a(O(2)) are reported, and thereby defect structures of the co-doped BaTiO(3) depending on m(d)/m(a) are determined. It is found that for the co-doping ratio 1 < m(d)/m(a) < 2, the Fermi level is pinned at a few kT's around the deep level of Mn(Ti)'' across the otherwise p-type semiconducting log a(O(2))-region of Mn-singly doped BaTiO(3), and attributed to deep acceptor-shallow donor mutual compensation 2[Mn(Ti)''] + [Mn(Ti)'] ≈ [Y(ba)·], thus turning otherwise p-type semiconducting BaTiO(3) semi-insulating.
More Related Videos
 07:24
Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
 08:00
Chemical Synthesis of Porous Barium Titanate Thin Film and Thermal Stabilization of Ferroelectric Phase by Porosity-Induced Strain
Published on: March 27, 2018
Related Concept Videos
 02:42
Valence Bond Theory
 01:29
Imperfections in Crystal Structure: Non-Stoichiometric Defects
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human eye.
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...