Markov Chain Ontology Analysis (MCOA)
H Robert Frost1, Alexa T McCray
1Center for Biomedical Informatics, Harvard Medical School, Boston, MA 02115, USA. rob_frost@hms.harvard.edu

Background:
Biomedical ontologies have become an increasingly critical lens through which researchers analyze the genomic, clinical and bibliographic data that fuels scientific research. Of particular relevance are methods, such as enrichment analysis, that quantify the importance of ontology classes relative to a collection of domain data. Current analytical techniques, however, remain limited in their ability to handle many important types of structural complexity encountered in real biological systems including class overlaps, continuously valued data, inter-instance relationships, non-hierarchical relationships between classes, semantic distance and sparse data.
Results:
In this paper, we describe a methodology called Markov Chain Ontology Analysis (MCOA) and illustrate its use through a MCOA-based enrichment analysis application based on a generative model of gene activation. MCOA models the classes in an ontology, the instances from an associated dataset and all directional inter-class, class-to-instance and inter-instance relationships as a single finite ergodic Markov chain. The adjusted transition probability matrix for this Markov chain enables the calculation of eigenvector values that quantify the importance of each ontology class relative to other classes and the associated data set members. On both controlled Gene Ontology (GO) data sets created with Escherichia coli, Drosophila melanogaster and Homo sapiens annotations and real gene expression data extracted from the Gene Expression Omnibus (GEO), the MCOA enrichment analysis approach provides the best performance of comparable state-of-the-art methods.
Conclusion:
A methodology based on Markov chain models and network analytic metrics can help detect the relevant signal within large, highly interdependent and noisy data sets and, for applications such as enrichment analysis, has been shown to generate superior performance on both real and simulated data relative to existing state-of-the-art approaches.
More Related Videos
 09:37
Navigating MARRVEL, a Web-Based Tool that Integrates Human Genomics and Model Organism Genetics Information
Published on: August 15, 2019
 09:47
DeepOmicsAE: Representing Signaling Modules in Alzheimer's Disease with Deep Learning Analysis of Proteomics, Metabolomics, and Clinical Data
Published on: December 15, 2023
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 01:20
Applications of Molecular Taxonomy
 02:00
Molecular Models
 01:29
Modern Molecular Taxonomy
 01:13
Mass Analyzers: Overview
 01:08
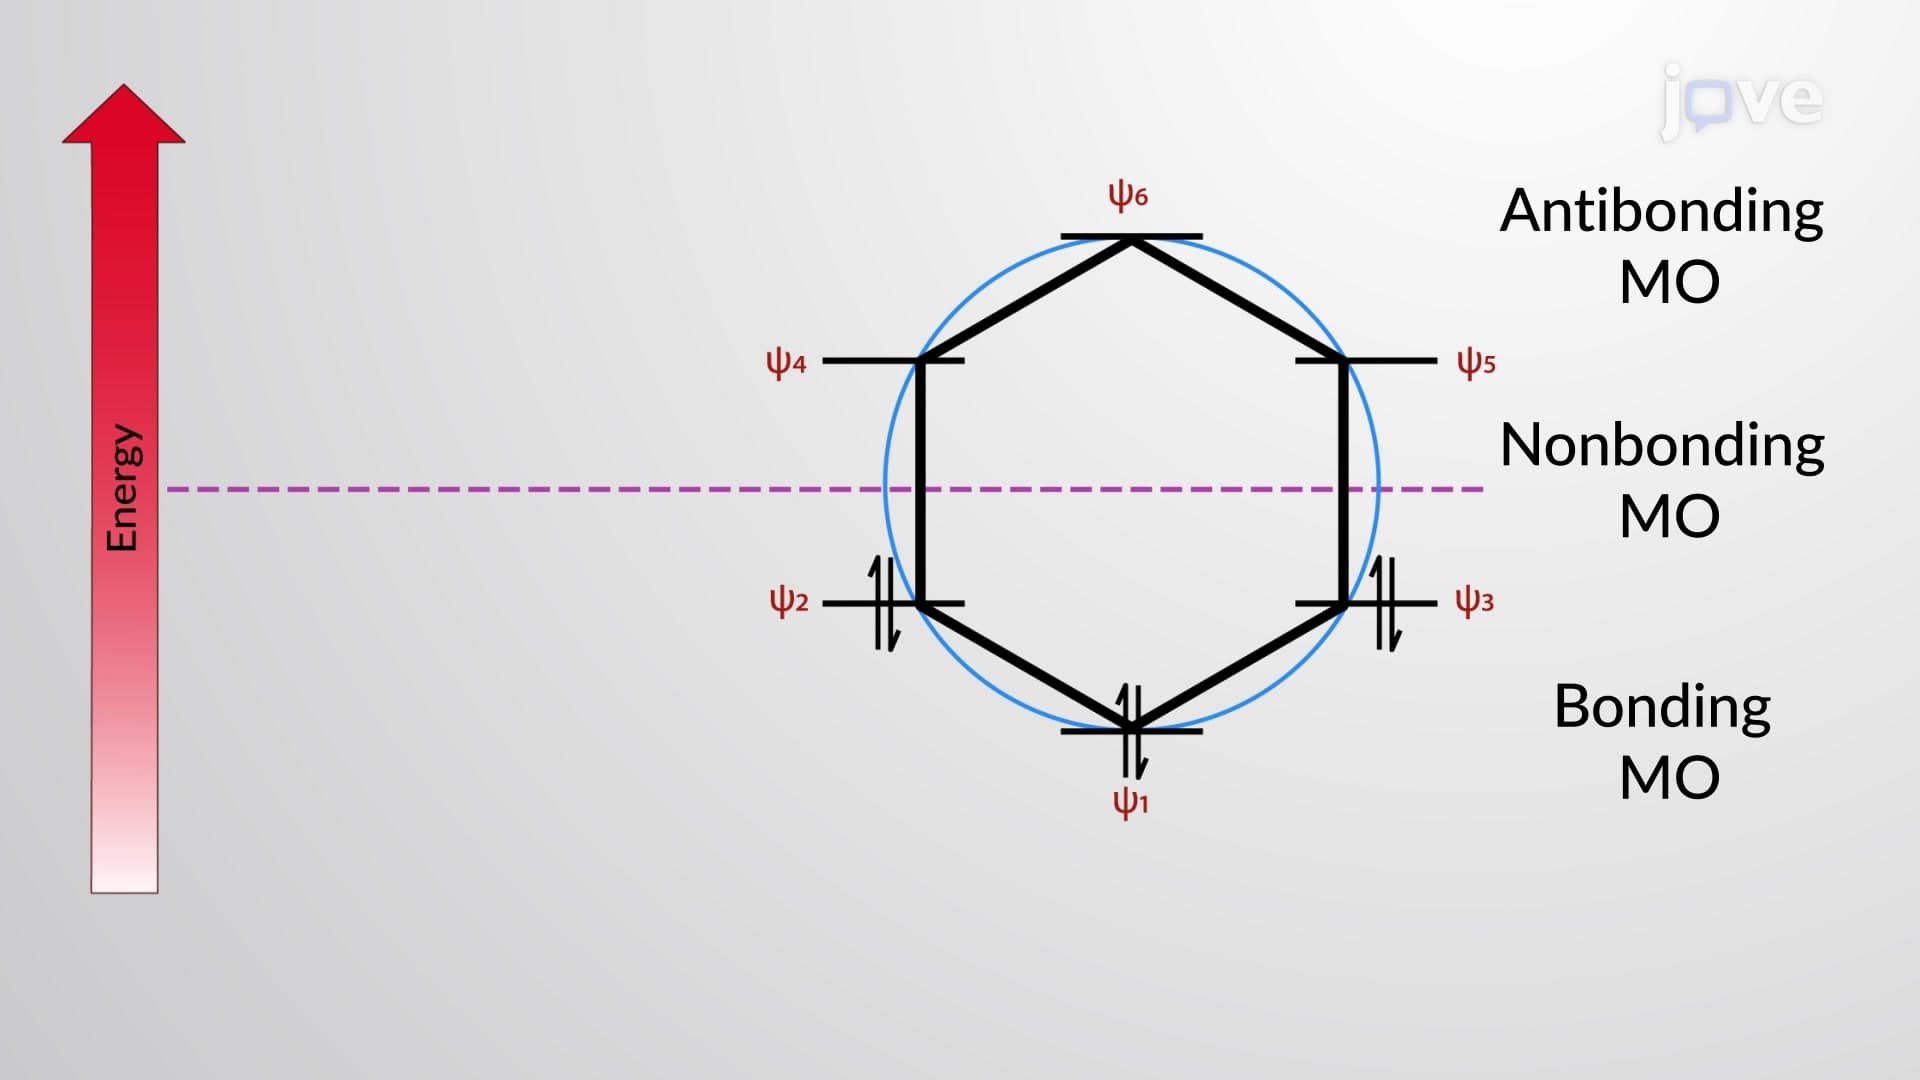
Hückel's Rule Diagram of π MOs: Frost Circle
A Frost circle is constructed by drawing a polygon whose number of edges is equal to the number of carbons of the given cyclic system, with one of the vertices pointing down. Then, a circle is drawn enclosing the polygon so that...