Related Experiment Video
Updated: May 20, 2026

Novel Techniques for Observing Structural Dynamics of Photoresponsive Liquid Crystals
Published on: May 29, 2018
A pass too far: dissociation of internal energy selected paracyclophane cations, theory and experiment
Patrick Hemberger1, Andras Bodi, Christof Schon
1Molecular Dynamics Group, Paul Scherrer Institute, 5234 Villigen PSI, Switzerland. patrick.hemberger@psi.ch

Abstract:
The vacuum ultraviolet (VUV) photoionization and dissociative photoionization of three hydroxy-substituted [2.2]paracyclophane derivatives were studied yielding adiabatic ionization energies and dissociative photoionization energies (appearance energies). The simplest dissociation pathway is the breaking of both CH(2)-CH(2) bridge units and fragmenting the molecular ion in half to yield xylylene neutral and cationic fragments. The experimental data show that this process is outcompeted by a faster, higher energy channel, possibly yielding cyclooctatetraene derivatives. The role of the reaction coordinate, the effect of large amplitude motions on the density of states function at low and high energies and the temperature dependent 'population gap' in the internal energy distribution in large molecules are discussed in the context of applying statistical models to the dissociation. Computational approaches to the binding energy of paracyclophanes are marred with pitfalls. Noncovalent interactions play a major role in keeping paracyclophanes bound by some 200 kJ mol(-1) with respect to the two xylylene motifs, and the covalent CH(2)-CH(2) bonds are mostly counteracted by the geometric strain. The stabilizing effects are twofold: first, paracyclophanes are aromatic compounds, whereas xylylenes are not. Thus, the aromaticity of the molecule is induced by dimerization. Second, dispersive π-π interactions also stabilize the molecule. We evaluated 23 different computational chemistry approaches, and found that very few of the favorably scaling ones give an adequate description of this system. Among the DFT functionals tested, only PBE-D3 and perhaps M06-2X yielded consistently accurate results, comparable with MP3 and CCSD, or the G4 and CBS-QB3 composite methods. MP2 results in general suffer from significant overbonding.
More Related Videos

Related Concept Videos
 01:13
Force and Potential Energy in One Dimension
 01:25
Deactivation Processes: Jablonski Diagram
 01:23
Theory of Strong Electrolytes
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
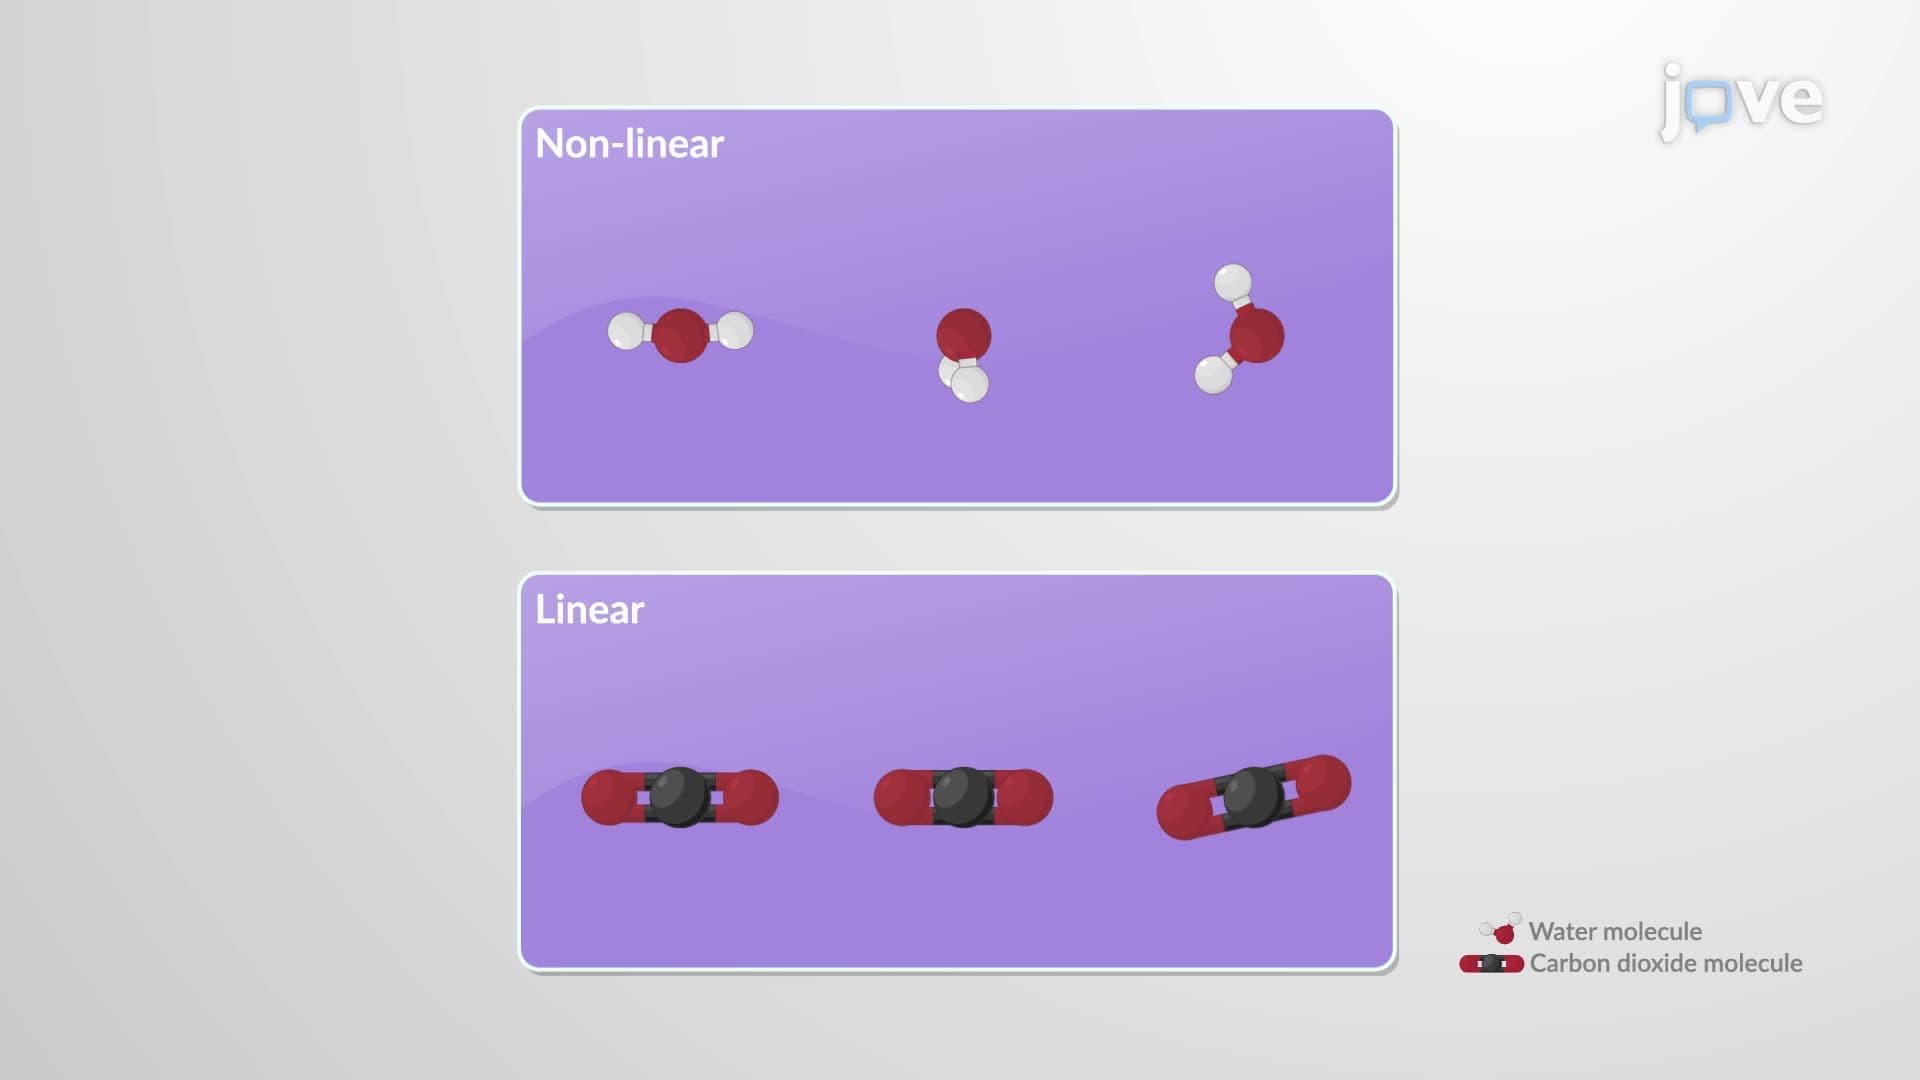 01:27
The Molecular Nature of Internal Energy