Related Experiment Video
Updated: May 18, 2026

Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
A parallel systematic-Monte Carlo algorithm for exploring conformational space
Yasset Perez-Riverol1, Roberto Vera, Yuliet Mazola
1Department of Proteomics and Bioinformatics, Center for Genetic Engineering and Biotechnology, Havana, Cuba. yasset.perez@biocomp.cigb.edu.cu

Abstract:
Computational algorithms to explore the conformational space of small molecules are complex and computer demand field in chemoinformatics. In this paper a hybrid algorithm to explore the conformational space of organic molecules is presented. This hybrid algorithm is based in a systematic search approach combined with a Monte Carlo based method in order to obtain an ensemble of low-energy conformations simulating the flexibility of small chemical compounds. The Monte Carlo method uses the Metropolis criterion to accept or reject a conformation through an in-house implementation of the MMFF94s force field to calculate the conformational energy. The parallel design of this algorithm, based on the message passing interface (MPI) paradigm, was implemented. The results showed a performance increase in the terms of speed and efficiency.
Related Concept Videos
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...
 02:29
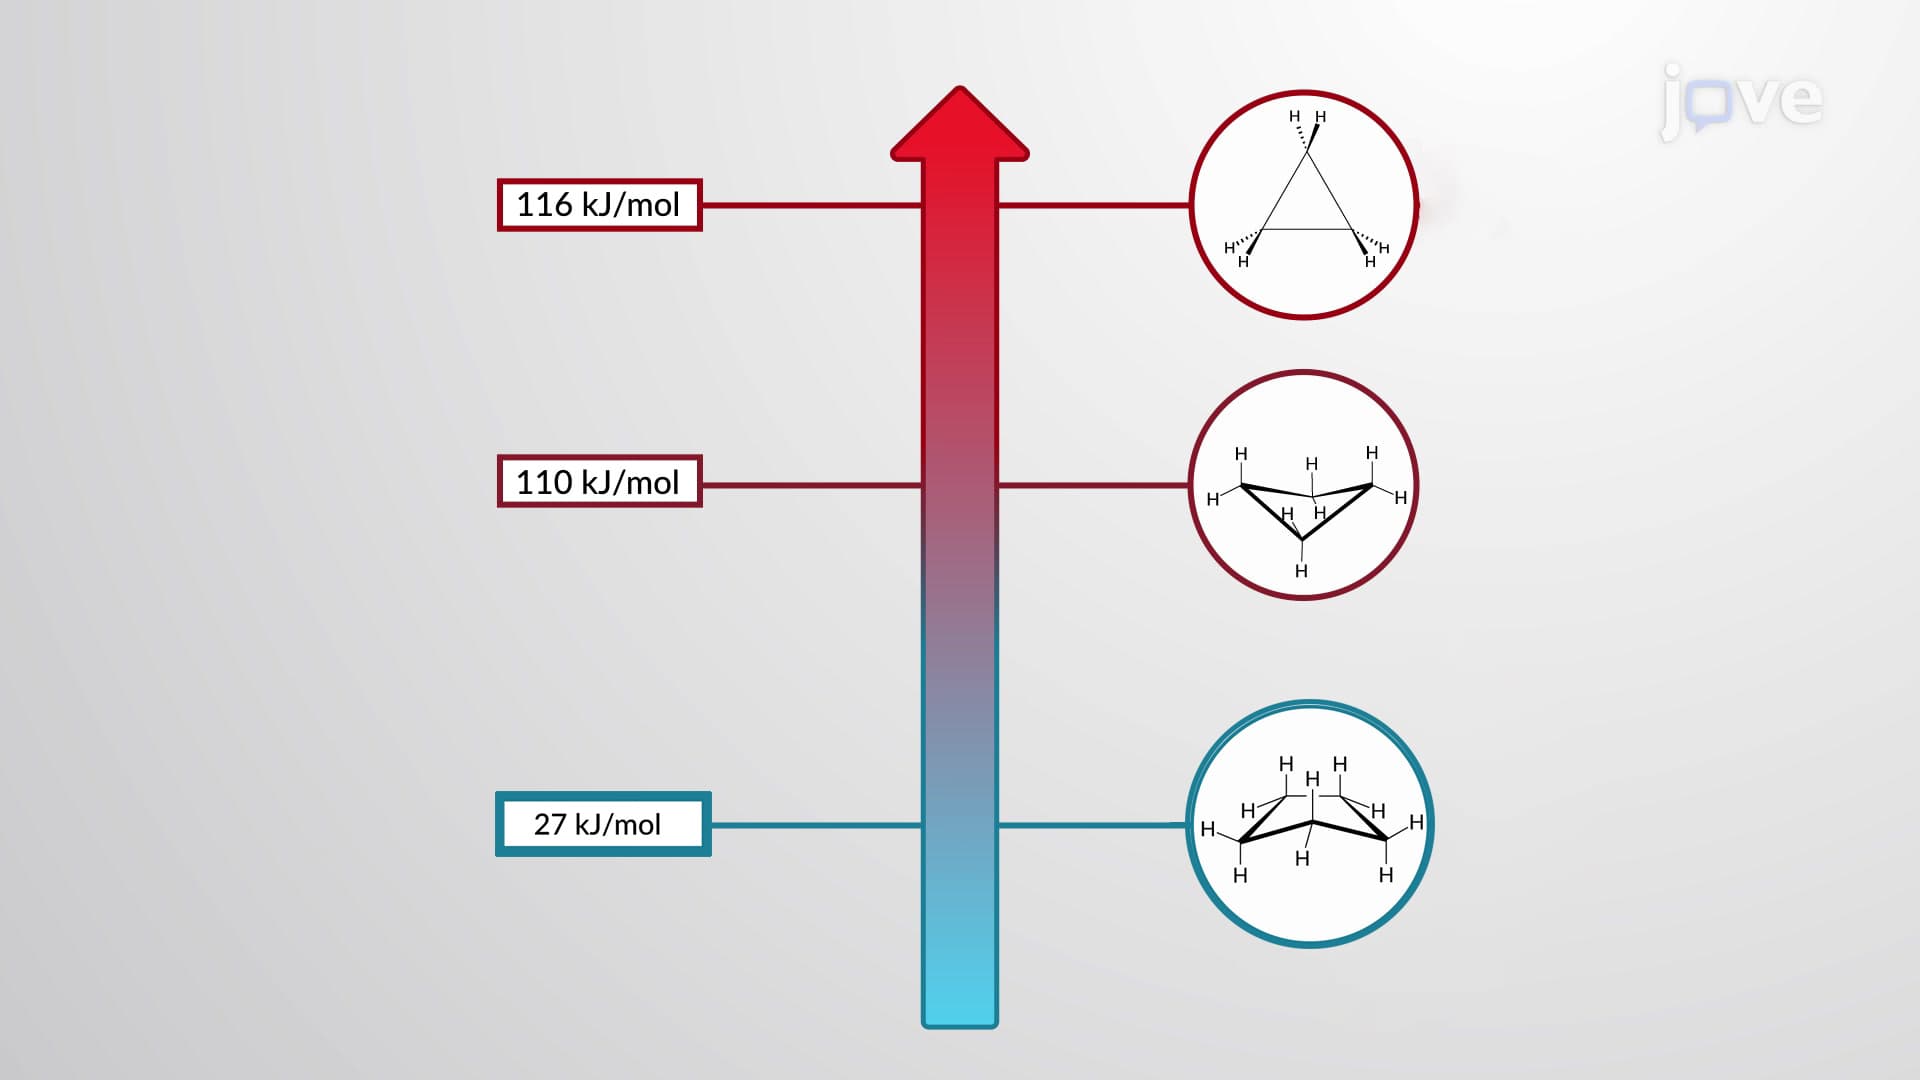
Conformations of Cycloalkanes
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:02
Chair Conformation of Cyclohexane
The hydrogen atoms linked to carbons are arranged in two different axial and equatorial orientations to achieve this staggered...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR