Related Experiment Video
Updated: May 16, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Ab initio calculations of the Ar-ethane intermolecular potential energy surface using bond function basis sets
Jian-Dong Zhang1, Shu-Jin Li, Fu-Ming Tao
1Department of Chemistry, School of Chemistry, Chemical Engineering and Materials Science, Soochow University, Suzhou, Jiangsu 215123, People's Republic of China.

Abstract:
The intermolecular potential energy surface (PES) of argon with ethane has been studied by ab initio calculations at the levels of second-order Møller-Plesset perturbation (MP2) theory and coupled-cluster theory with single, double, and noniterative triple configurations (CCSD(T)) using a series of augmented correlation-consistent basis sets. Two sets of bond functions, bf1 (3s3p2d) and bf2 (6s6p4d2f), have been added to the basis sets to show a dramatic and systematic improvement in the convergence of the entire PES. The PES of Ar-ethane is characterized by a global minimum at a near T-shaped configuration with a well depth of 0.611 kcal mol(-1), a second minimum at a collinear configuration with a well depth of 0.456 kcal mol(-1), and a saddle point connecting the two minima. It is shown that an augmented correlation-consistent basis set with a set of bond functions, either bf1 or bf2, can effectively produce results equivalent to the next larger augmented correlation-consistent basis set, that is, aug-cc-pVDZ-bf1 ≈ aug-cc-pVTZ, aug-cc-pVTZ-bf1 ≈ aug-cc-pVQZ. Very importantly, the use of bond functions improves the PES globally, resulting accurate potential anisotropy. Finally, MP2 method is inadequate for accurate calculations, because it gives a potentially overestimated well depth and, more seriously, a poor potential anisotropy.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 13:56
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 03:51
Molecular Orbital Theory II
 02:49
Bond Energies and Bond Lengths
 03:35
Hybridization of Atomic Orbitals II
 02:37
Standard Enthalpy of Formation
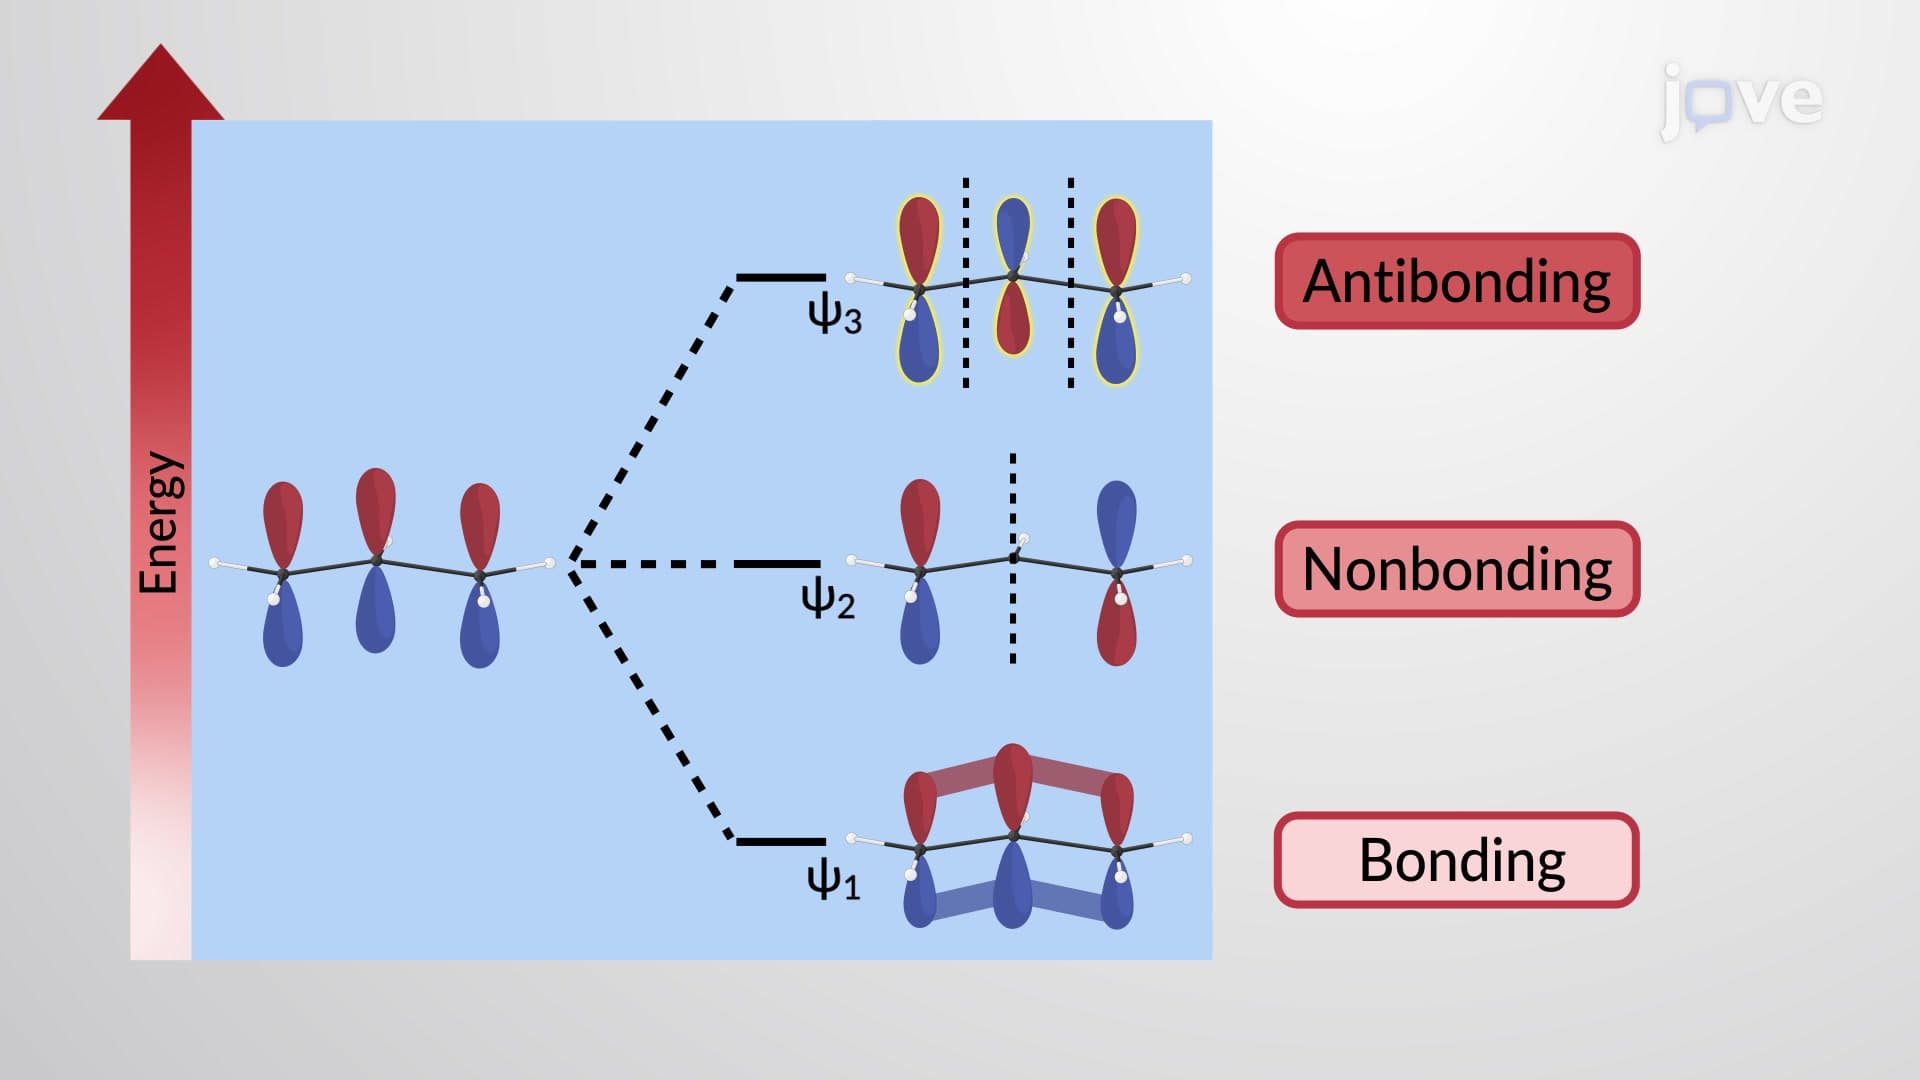 01:18
π Molecular Orbitals of the Allyl Cation and Anion