Related Experiment Video
Updated: Apr 26, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Computationally Efficient Multiconfigurational Reactive Molecular Dynamics
Takefumi Yamashita1, Yuxing Peng2, Chris Knight3
1Laboratory for Systems Biology and Medicine, Research Center for Advanced Science and Technology, the University of Tokyo, 4-6-1 Komaba, Tokyo, Japan.

Abstract:
It is a computationally demanding task to explicitly simulate the electronic degrees of freedom in a system to observe the chemical transformations of interest, while at the same time sampling the time and length scales required to converge statistical properties and thus reduce artifacts due to initial conditions, finite-size effects, and limited sampling. One solution that significantly reduces the computational expense consists of molecular models in which effective interactions between particles govern the dynamics of the system. If the interaction potentials in these models are developed to reproduce calculated properties from electronic structure calculations and/or ab initio molecular dynamics simulations, then one can calculate accurate properties at a fraction of the computational cost. Multiconfigurational algorithms model the system as a linear combination of several chemical bonding topologies to simulate chemical reactions, also sometimes referred to as "multistate". These algorithms typically utilize energy and force calculations already found in popular molecular dynamics software packages, thus facilitating their implementation without significant changes to the structure of the code. However, the evaluation of energies and forces for several bonding topologies per simulation step can lead to poor computational efficiency if redundancy is not efficiently removed, particularly with respect to the calculation of long-ranged Coulombic interactions. This paper presents accurate approximations (effective long-range interaction and resulting hybrid methods) and multiple-program parallelization strategies for the efficient calculation of electrostatic interactions in reactive molecular simulations.
More Related Videos
 05:00
Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
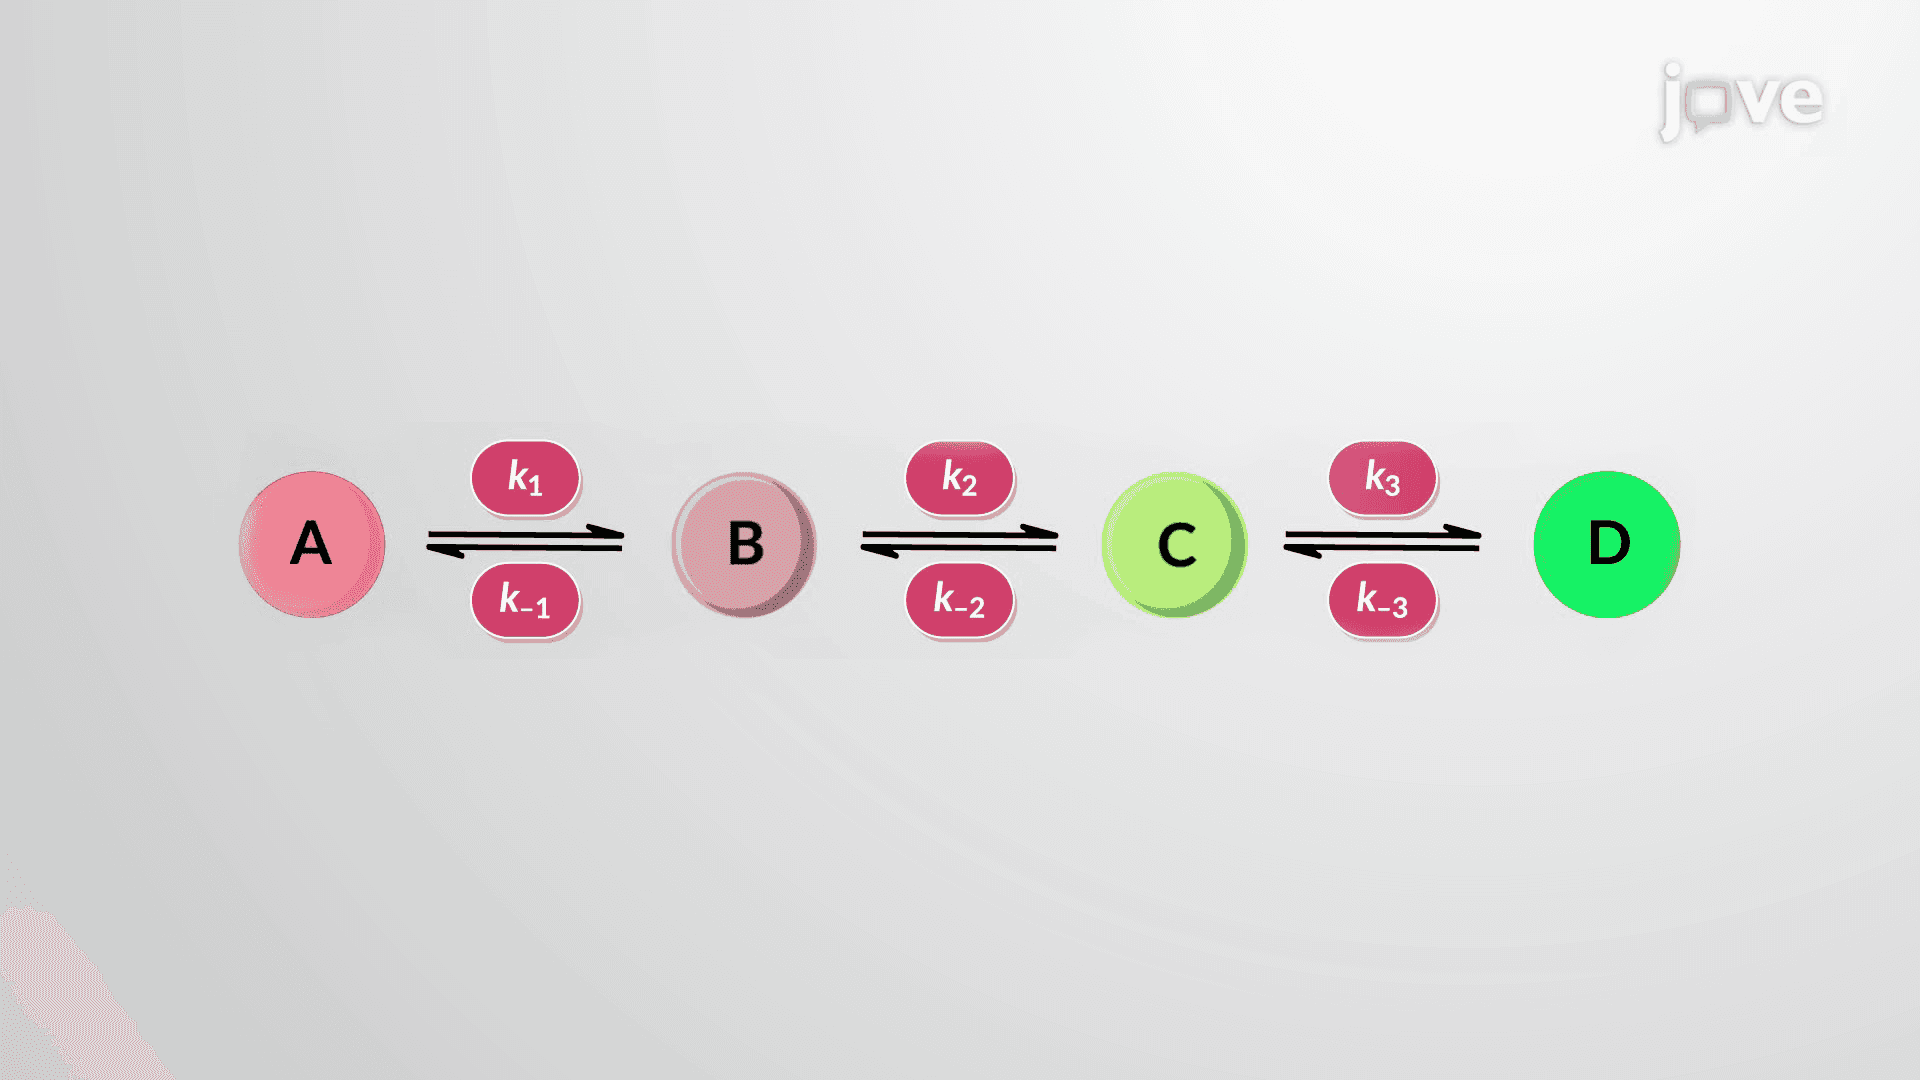 01:29
Reaction Mechanisms: Rate-limiting Step Approximation
 01:17
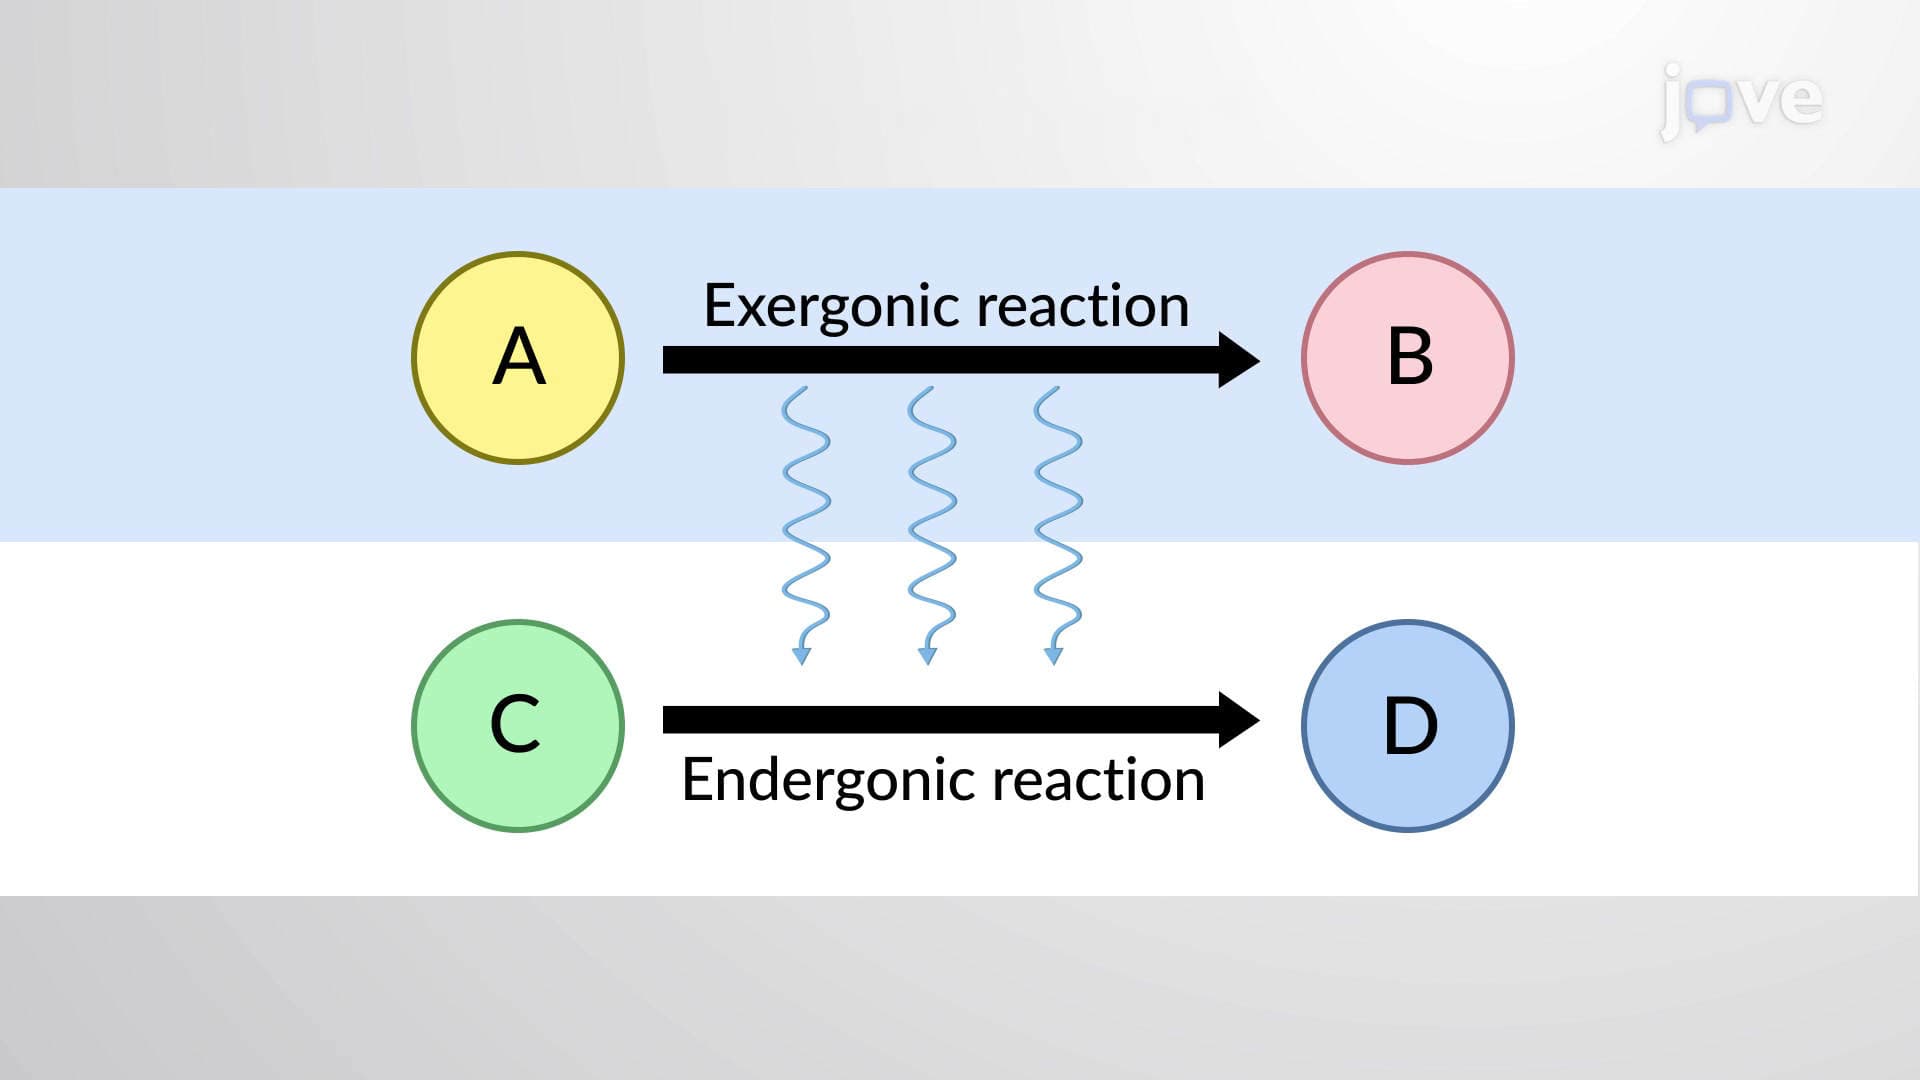
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 02:31
Multi-Step Reactions
 01:34
Woodward–Hoffmann Selection Rules and Microscopic Reversibility
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:16
Cycloaddition Reactions: MO Requirements for Thermal Activation