
Rare loss-of-function mutation in complement component C3 provides insight into molecular and pathophysiological
Georgia Sfyroera1, Daniel Ricklin1, Edimara S Reis1
1Department of Pathology and Laboratory Medicine, University of Pennsylvania, Philadelphia, PA 19104;

Insights
A novel C3 gene mutation (M373T) was identified, causing a complete loss of alternative pathway complement activation. This discovery clarifies the molecular basis of C3 dysfunction and its varied clinical presentations.
Area of Science:
- Immunology
- Molecular Biology
- Genetics
Background:
- The complement system's central protein, C3, is crucial for immune responses.
- A hereditary C3 dysfunction affecting the alternative pathway (AP) was previously noted but lacked genetic and molecular explanation.
Purpose of the Study:
- To identify the genetic cause and molecular consequences of a hereditary C3 dysfunction.
- To elucidate the role of C3 in alternative pathway activation and its clinical relevance.
Main Methods:
- Genetic sequencing to pinpoint a point mutation in the C3 β-chain (c.1180T > C; p.Met(373)Thr).
- Functional assays to assess AP activity in patient plasma.
- Structural analysis and binding studies involving compstatin and C3b.
Main Results:
- The identified M373T mutation completely abolished AP activity, requiring normal C3 for reconstitution.
- The mutation impacts C3's macroglobulin domain 4, altering compstatin and C3b binding.
- C3 activation and opsonization were impaired by the M373T mutation.
Conclusions:
- This study reveals the molecular mechanism of a C3 loss-of-function mutation.
- Findings provide insights into C3 convertase function, C3's role in diverse clinical conditions, and implications for complement inhibition therapies.
Abstract:
The plasma protein C3 is a central element in the activation and effector functions of the complement system. A hereditary dysfunction of C3 that prevents complement activation via the alternative pathway (AP) was described previously in a Swedish family, but its genetic cause and molecular consequences have remained elusive. In this study, we provide these missing links by pinpointing the dysfunction to a point mutation in the β-chain of C3 (c.1180T > C; p.Met(373)Thr). In the patient's plasma, AP activity was completely abolished and could only be reconstituted with the addition of normal C3. The M373T mutation was localized to the macroglobulin domain 4 of C3, which contains a binding site for the complement inhibitor compstatin and is considered critical for the interaction of C3 with the AP C3 convertase. Structural analyses suggested that the mutation disturbs the integrity of macroglobulin domain 4 and induces conformational changes that propagate into adjacent regions. Indeed, C3 M373T showed an altered binding pattern for compstatin and surface-bound C3b, and the presence of Thr(373) in either the C3 substrate or convertase-affiliated C3b impaired C3 activation and opsonization. In contrast to known gain-of-function mutations in C3, patients affected by this loss-of-function mutation did not develop familial disease, but rather showed diverse and mostly episodic symptoms. Our study therefore reveals the molecular mechanism of a relevant loss-of-function mutation in C3 and provides insight into the function of the C3 convertase, the differential involvement of C3 activity in clinical conditions, and some potential implications of therapeutic complement inhibition.
Related Concept Videos
 01:27
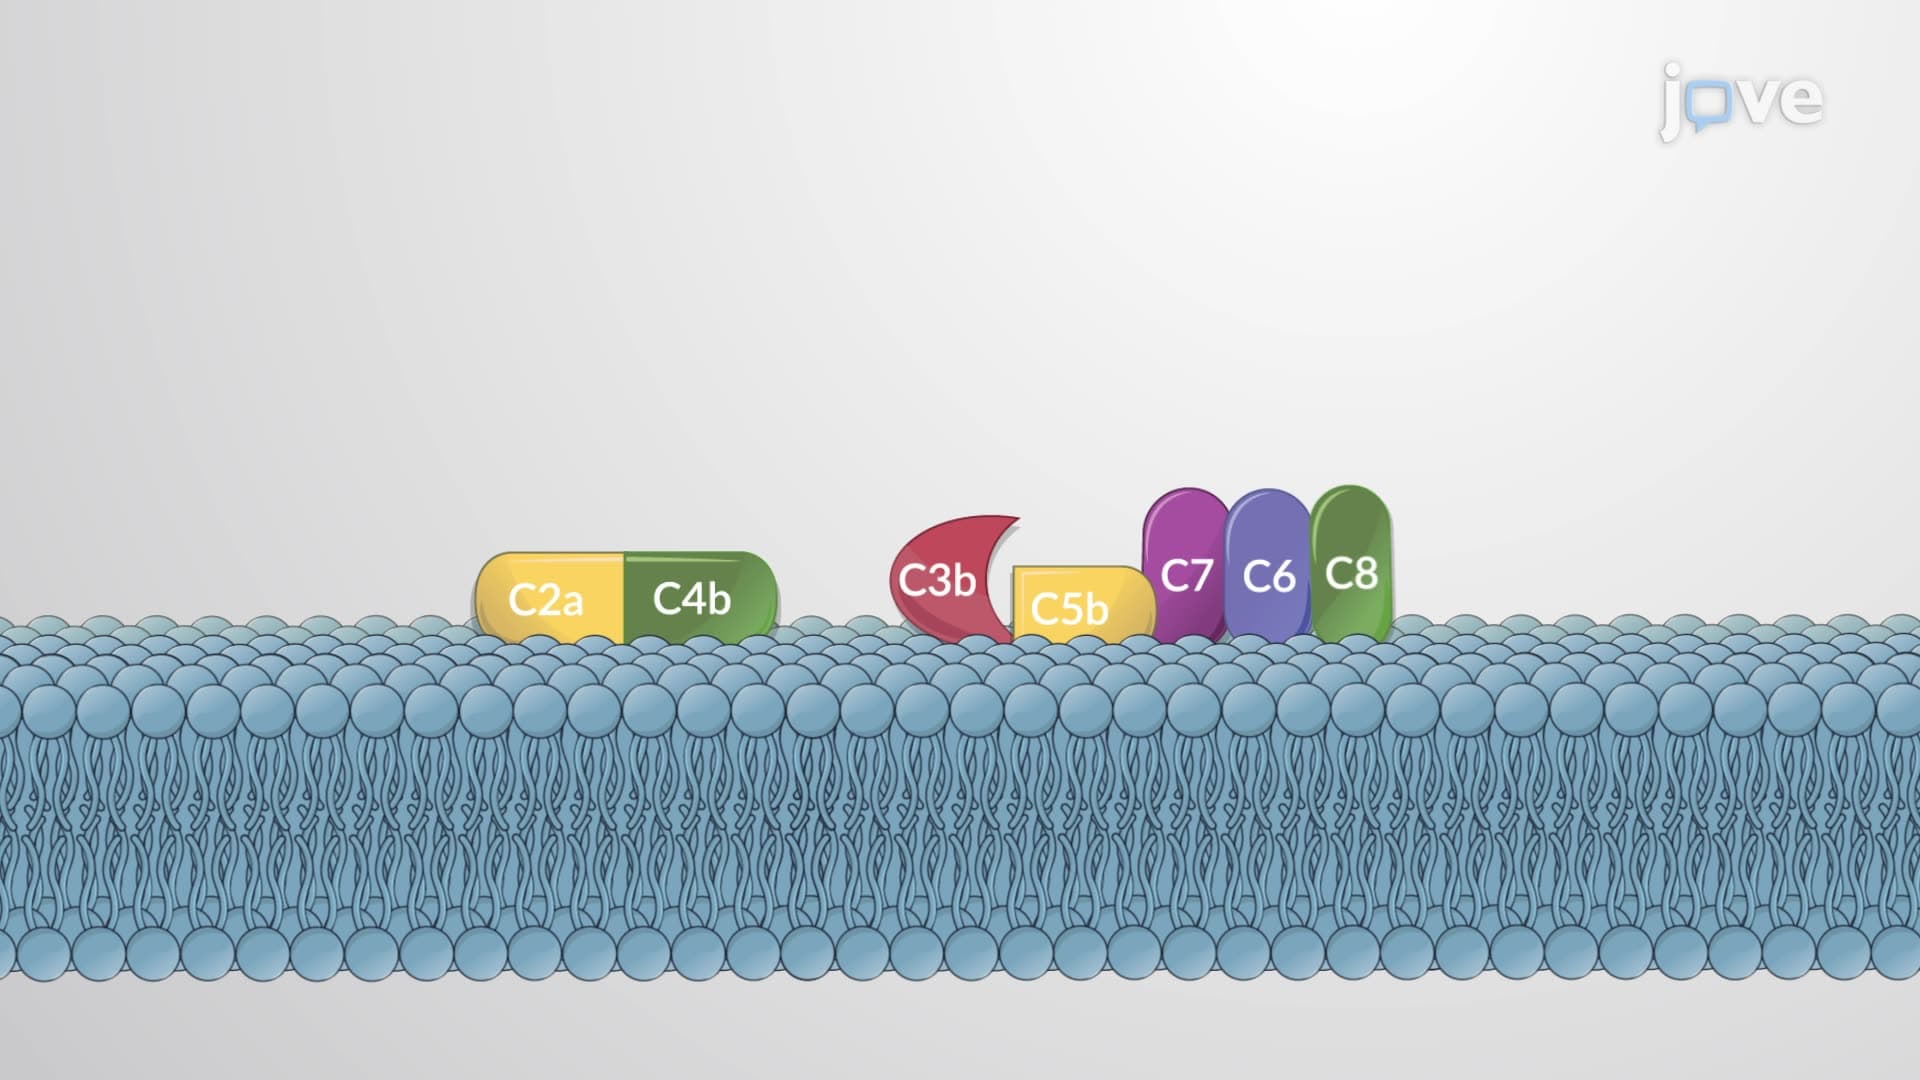
Complement System
 00:49
Complementation Tests
Organisms heterozygous for different mutations are crossed pairwise in all combinations. If present on different genes, the mutations can complement each other by providing the missing...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:29
Pharmacogenomics: Identification of New Drug Targets