Related Experiment Video
Updated: Apr 15, 2026

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Spatial dispersion of lone electron pairs?--experimental charge density of cubic arsenic(III) oxide
Piotr A Guńka1, Zygmunt Gontarz, Janusz Zachara
1Faculty of Chemistry, Warsaw University of Technology, Noakowskiego 3, 00-664 Warsaw, Poland. piogun@ch.pw.edu.pl.

Abstract:
The first experimental charge density study of arsenolite, a cubic polymorph of arsenic(III) oxide, extended by periodic DFT calculations is reported. The presence of weak AsO interactions is confirmed and their topological characterization based on experimental electron density is provided. Spatial dispersion of arsenic lone electron pairs into three domains is observed in the Laplacian of electron density as well as in the electron localization function. It results from the clustering of As atomic cores in the crystal structure and/or from the presence of strong As-O bonds. A similar phenomenon is recognized in the crystal structures of antimonates(III) and bismuthates(III) of alkaline metals indicating that this could be a more general feature worthy of further investigations.
Related Concept Videos
 04:01
VSEPR Theory and the Effect of Lone Pairs
 02:42
Ionic Crystal Structures
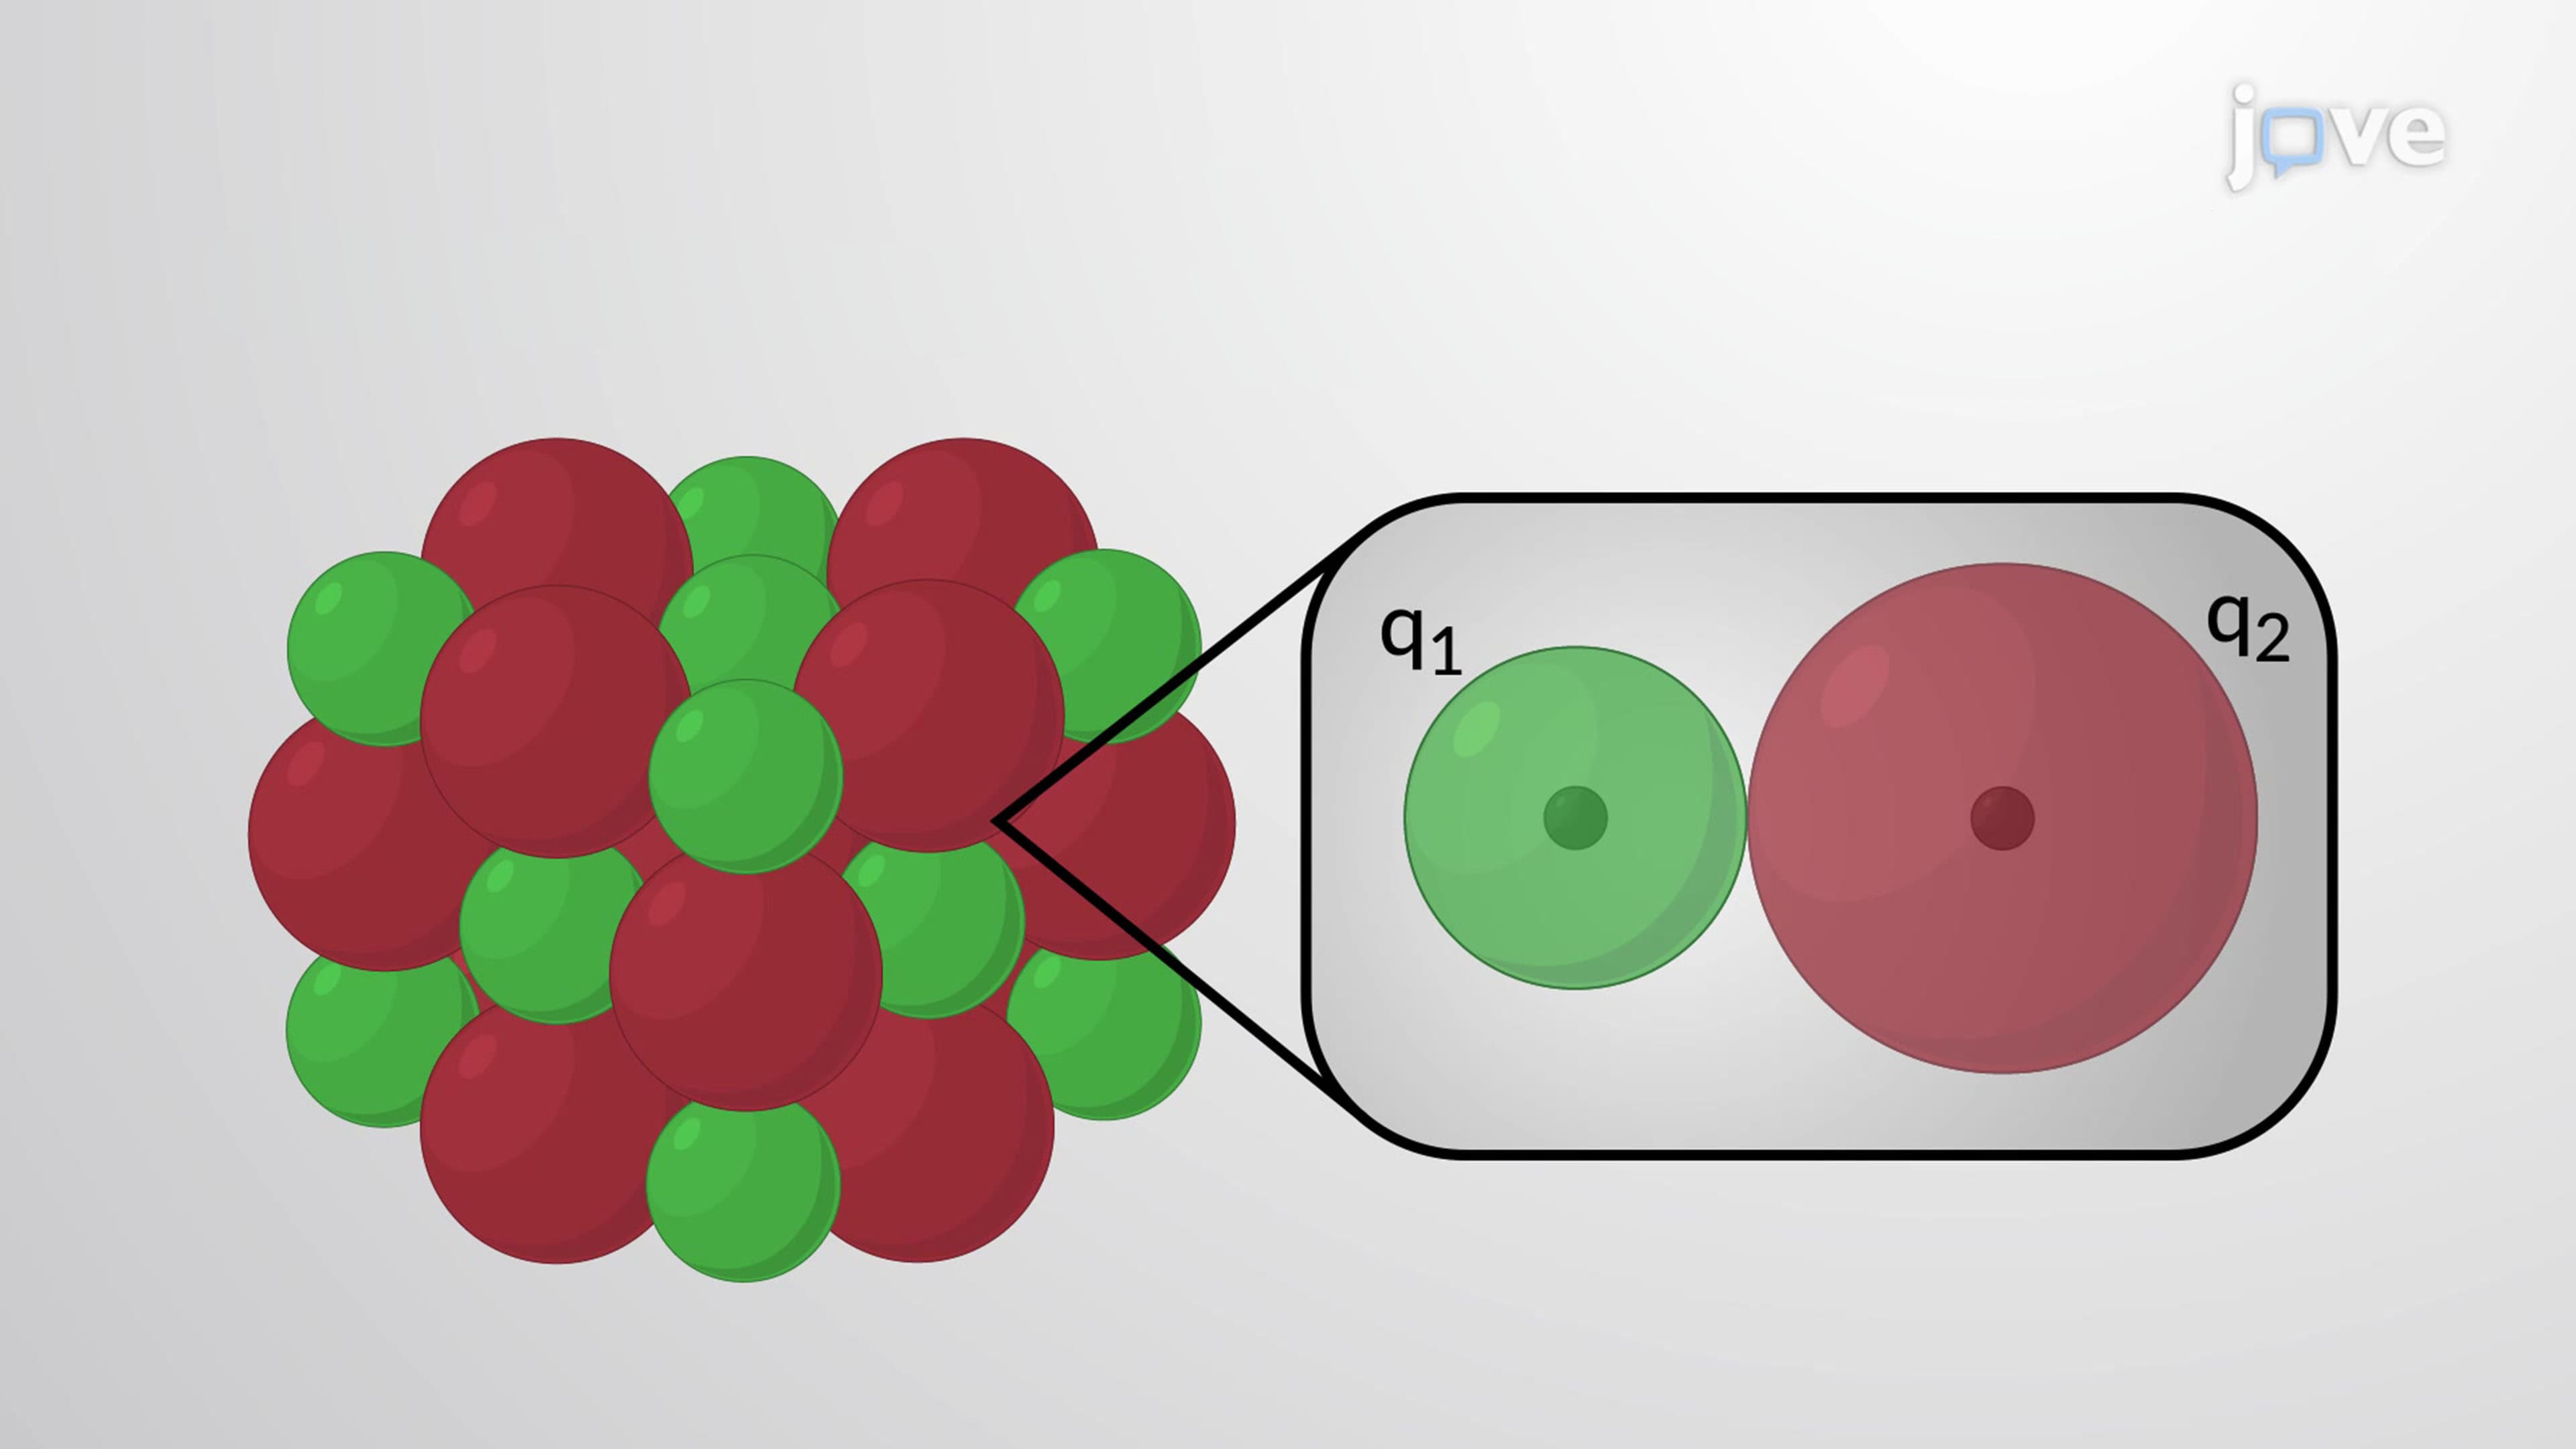
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than...
 02:54
Trends in Lattice Energy: Ion Size and Charge
 02:27
Predicting Molecular Geometry
 02:42
Valence Bond Theory