Computational Thermochemistry: Scale Factor Databases and Scale Factors for Vibrational Frequencies Obtained from Electronic Model Chemistries
Related Experiment Video
Updated: Mar 29, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
I M Alecu1, Jingjing Zheng1, Yan Zhao1
1Department of Chemistry and Supercomputing Institute, University of Minnesota, Minneapolis, Minnesota 55455-0431 and Commercial Print Engine Lab, HP Laboratories, Hewlett-Packard Company, Palo Alto, California 94304.

Abstract:
Optimized scale factors for calculating vibrational harmonic and fundamental frequencies and zero-point energies have been determined for 145 electronic model chemistries, including 119 based on approximate functionals depending on occupied orbitals, 19 based on single-level wave function theory, three based on the neglect-of-diatomic-differential-overlap, two based on doubly hybrid density functional theory, and two based on multicoefficient correlation methods. Forty of the scale factors are obtained from large databases, which are also used to derive two universal scale factor ratios that can be used to interconvert between scale factors optimized for various properties, enabling the derivation of three key scale factors at the effort of optimizing only one of them. A reduced scale factor optimization model is formulated in order to further reduce the cost of optimizing scale factors, and the reduced model is illustrated by using it to obtain 105 additional scale factors. Using root-mean-square errors from the values in the large databases, we find that scaling reduces errors in zero-point energies by a factor of 2.3 and errors in fundamental vibrational frequencies by a factor of 3.0, but it reduces errors in harmonic vibrational frequencies by only a factor of 1.3. It is shown that, upon scaling, the balanced multicoefficient correlation method based on coupled cluster theory with single and double excitations (BMC-CCSD) can lead to very accurate predictions of vibrational frequencies. With a polarized, minimally augmented basis set, the density functionals with zero-point energy scale factors closest to unity are MPWLYP1M (1.009), τHCTHhyb (0.989), BB95 (1.012), BLYP (1.013), BP86 (1.014), B3LYP (0.986), MPW3LYP (0.986), and VSXC (0.986).
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...
 01:26
Thermodynamic Potentials
 01:26
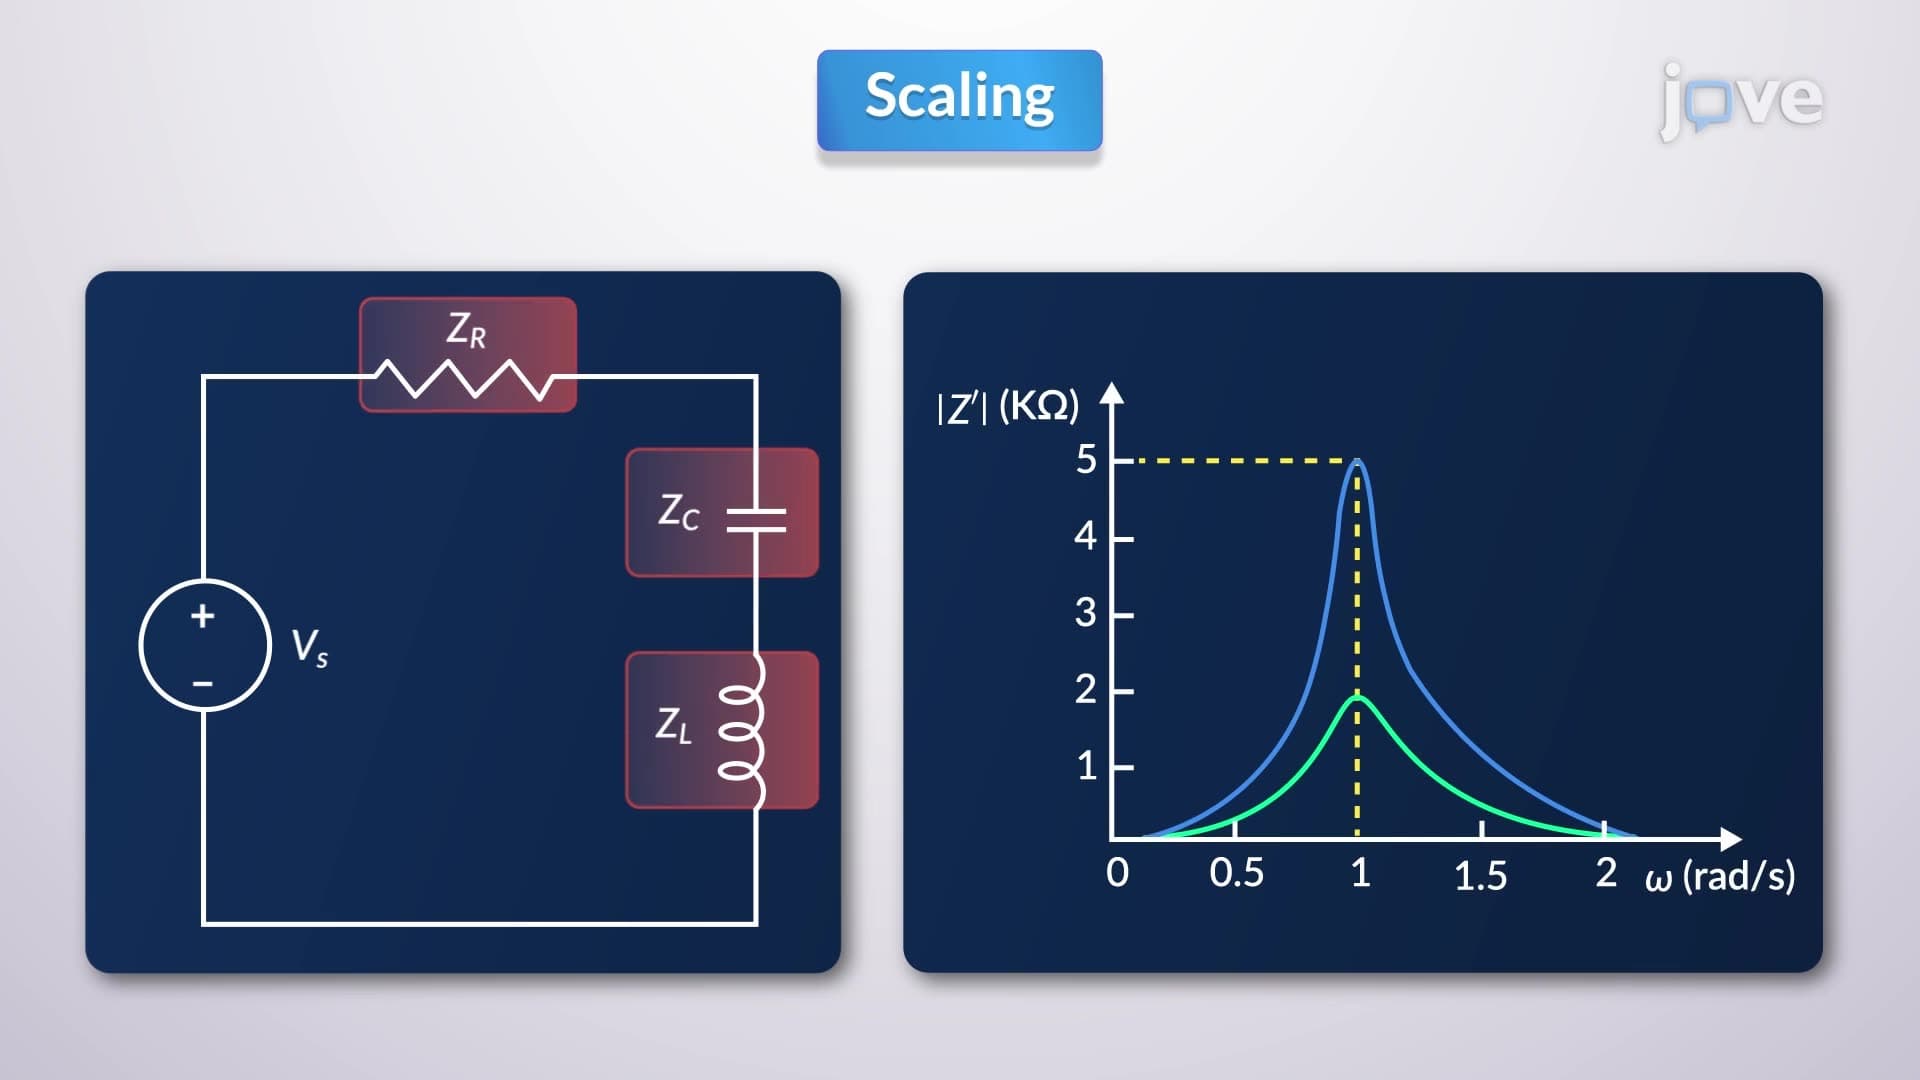
Scaling
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
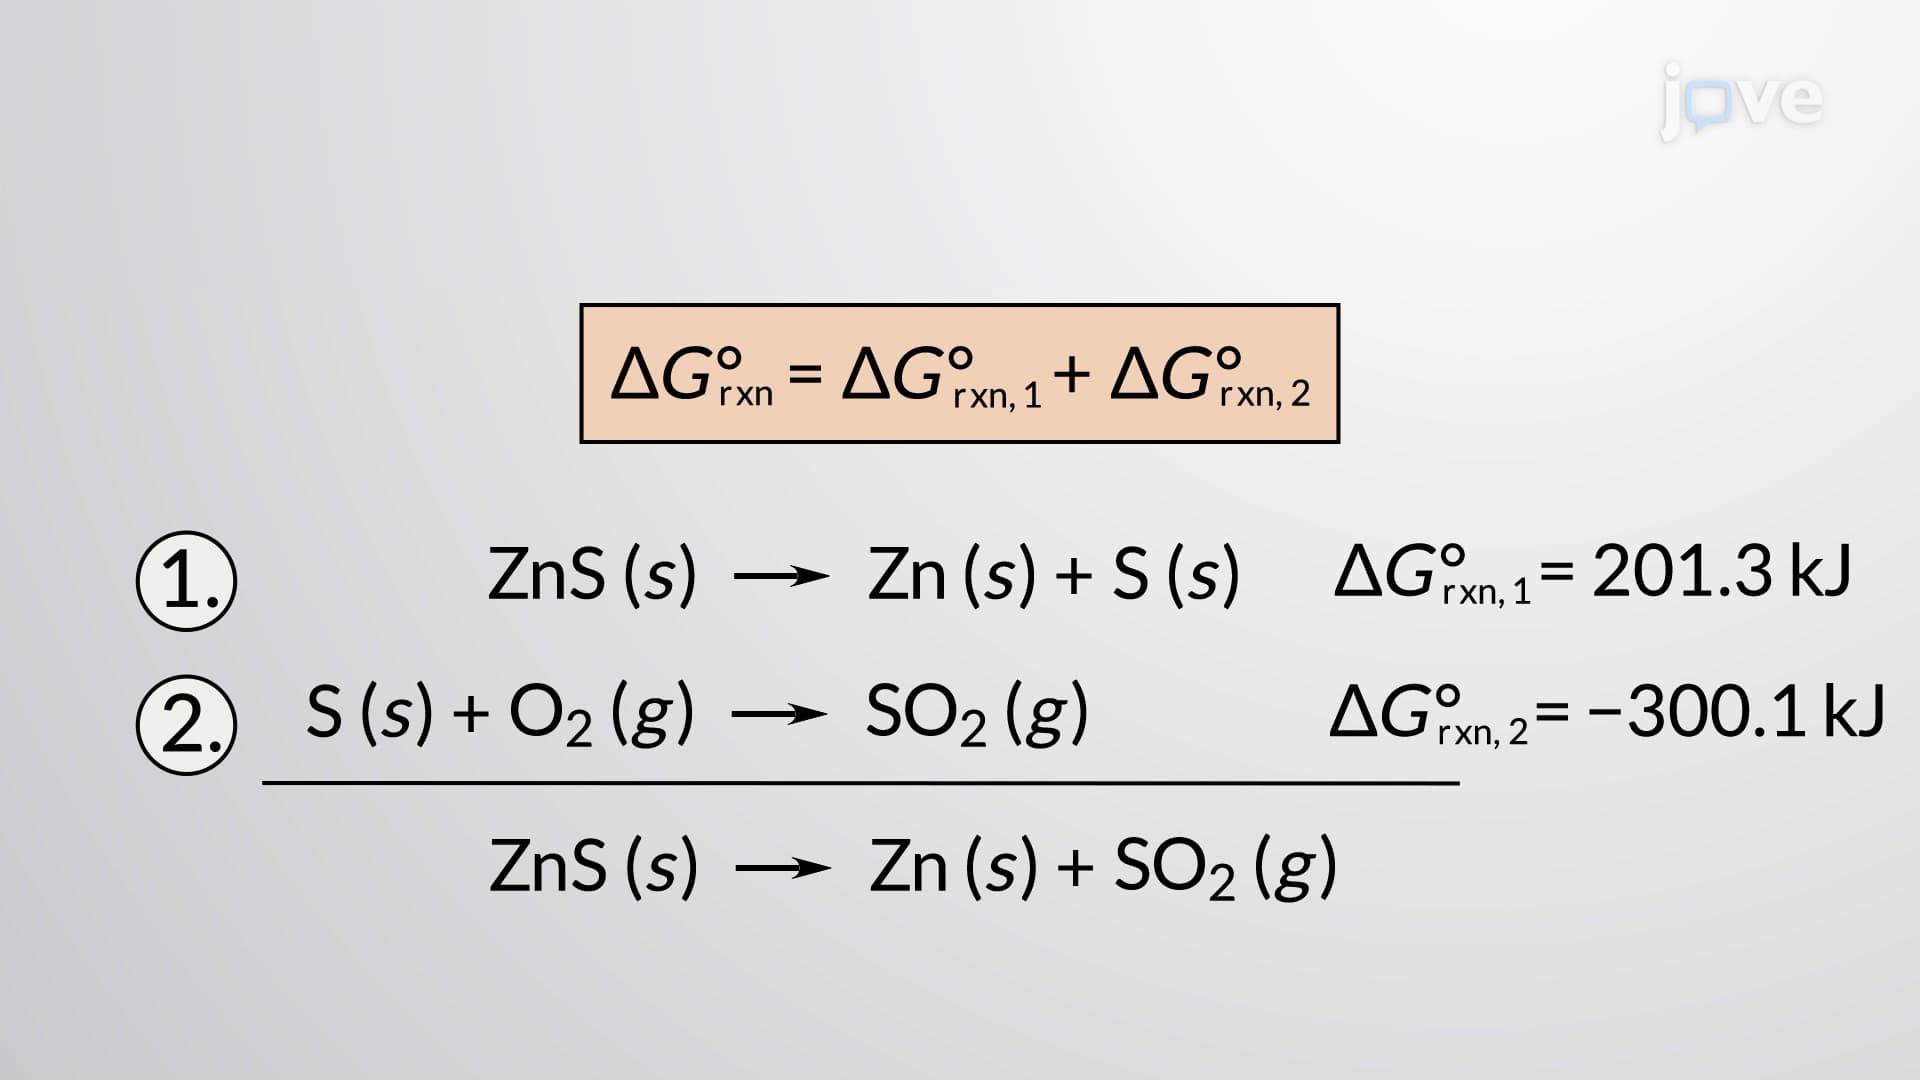 02:49
Calculating Standard Free Energy Changes
 01:25
Heat Capacities of an Ideal Gas III