Related Experiment Video
Updated: Mar 7, 2026

2 in 1: One-step Affinity Purification for the Parallel Analysis of Protein-Protein and Protein-Metabolite Complexes
Published on: August 6, 2018
msPurity: Automated Evaluation of Precursor Ion Purity for Mass Spectrometry-Based Fragmentation in Metabolomics
Thomas N Lawson1, Ralf J M Weber1, Martin R Jones1
1School of Biosciences and ‡Phenome Centre Birmingham, College of Life and Environmental Sciences, University of Birmingham , Edgbaston, Birmingham, B15 2TT, United Kingdom.

Abstract:
Tandem mass spectrometry (MS/MS or MS2) is a widely used approach for structural annotation and identification of metabolites in complex biological samples. The importance of assessing the contribution of the precursor ion within an isolation window for MS2 experiments has been previously detailed in proteomics, where precursor ion purity influences the quality and accuracy of matching to mass spectral libraries, but to date, there has been little attention to this data-processing technique in metabolomics. Here, we present msPurity, a vendor-independent R package for liquid chromatography (LC) and direct infusion (DI) MS2 that calculates a simple metric to describe the contribution of the selected precursor. The precursor purity metric is calculated as "intensity of a selected precursor divided by the summed intensity of the isolation window". The metric is interpolated at the recorded point of MS2 acquisition using bordering full-scan spectra. Isotopic peaks of the selected precursor can be removed, and low abundance peaks that are believed to have limited contribution to the resulting MS2 spectra are removed. Additionally, the isolation efficiency of the mass spectrometer can be taken into account. The package was applied to Data Dependent Acquisition (DDA)-based MS2 metabolomics data sets derived from three metabolomics data repositories. For the 10 LC-MS2 DDA data sets with > ±1 Da isolation windows, the median precursor purity score ranged from 0.67 to 0.96 (scale = 0 to +1). The R package was also used to assess precursor purity of theoretical isolation windows from LC-MS data sets of differing sample types. The theoretical isolation windows being the same width used for an anticipated DDA experiment (±0.5 Da). The most complex sample had a median precursor purity score of 0.46 for the 64,498 XCMS determined features, in comparison to the less spectrally complex sample that had a purity score of 0.66 for 5071 XCMS features. It has been previously reported in proteomics that a purity score of <0.5 can produce unreliable spectra matching results. With this assumption, we show that for complex samples there will be a large number of metabolites where traditional DDA approaches will struggle to provide reliable annotations or accurate matches to mass spectral libraries.
More Related Videos
 11:17
A Simple Fractionated Extraction Method for the Comprehensive Analysis of Metabolites, Lipids, and Proteins from a Single Sample
Published on: June 1, 2017
 08:07
Sample Preparation for Single Cell Mass Spectrometry Metabolomics Studies: Combined Cell Washing, Quenching, Drying, and Storage
Published on: September 16, 2025
Related Concept Videos
 01:21
Tandem Mass Spectrometry
 01:33
Peptide Identification Using Tandem Mass Spectrometry
This technique helps gather information regarding the protein from which the peptide was obtained and to study the peptides’ amino acid sequence. Identifying peptides from a complex mixture is an important component of the growing field of...
 01:01
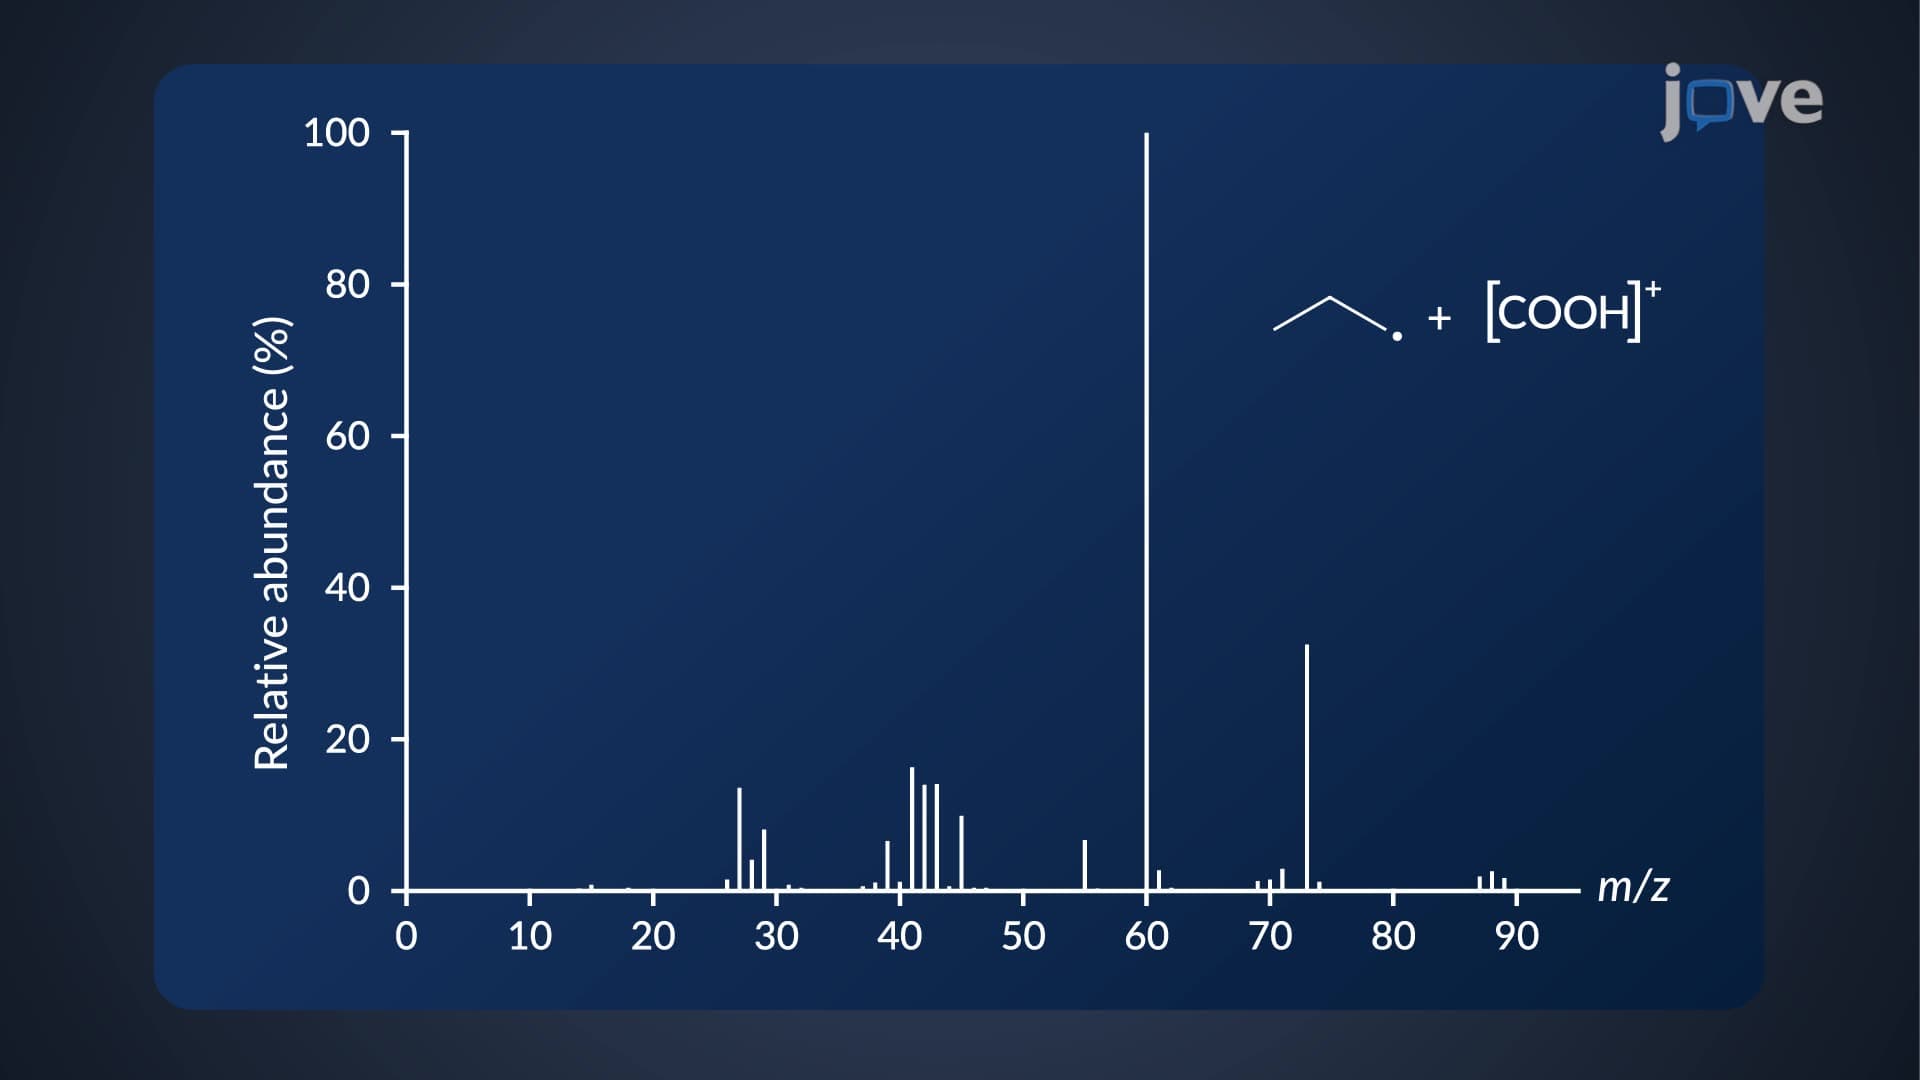
Mass Spectrometry: Carboxylic Acid, Ester, and Amide Fragmentation
For example, the...
 01:20
Mass Spectrometry: Molecular Fragmentation Overview
One type of fragmentation pattern is the cleavage of a single bond in the molecular ion. The cleavage leads to a radical and a cation. The cleavage can occur at...
 01:19
Mass Spectrometry: Overview
 01:16
Mass Spectrometers