Related Experiment Video
Updated: Feb 28, 2026

Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Extension of the SAFT-VR Mie EoS To Model Homonuclear Rings and Its Parametrization Based on the Principle of
Erich A Müller1, Andrés Mejía2
1Department of Chemical Engineering, Imperial College London , South Kensington Campus, London SW7 2AZ, United Kingdom.

Abstract:
The statistical associating fluid theory of variable range employing a Mie potential (SAFT-VR-Mie) proposed by Lafitte et al. (J. Chem Phys. 2013, 139, 154504) is one of the latest versions of the SAFT family. This particular version has been shown to have a remarkable capability to connect experimental determinations, theoretical calculations, and molecular simulations results. However, the theoretical development restricts the model to chains of beads connected in a linear fashion. In this work, the capabilities of the SAFT-VR Mie equation of state for modeling phase equilibria are extended for the case of planar ring compounds. This modification proposed replaces the Helmholtz energy of chain formation by an empirical contribution based on a parallelism to the second-order thermodynamic perturbation theory for hard sphere trimers. The proposed expression is given in terms of an extra parameter, χ, that depends on the number of beads, ms, and the geometry of the ring. The model is used to describe the phase equilibrium for planar ring compounds formed of Mie isotropic segments for the cases of ms equals to 3, 4, 5 (two configurations), and 7 (two configurations). The resulting molecular model is further parametrized, invoking a corresponding states principle resulting in sets of parameters that can be used indistinctively in theoretical calculations or in molecular simulations without any further refinements. The extent and performance of the methodology has been exemplified by predicting the phase equilibria and vapor pressure curves for aromatic hydrocarbons (benzene, hexafluorobenzene, toluene), heterocyclic molecules (2,5-dimethylfuran, sulfolane, tetrahydro-2H-pyran, tetrahydrofuran), and polycyclic aromatic hydrocarbons (naphthalene, pyrene, anthracene, pentacene, and coronene). An important aspect of the theory is that the parameters of the model can be used directly in molecular dynamics (MD) simulations to calculate equilibrium phase properties and interfacial tensions with an accuracy that rivals other coarse grained and united atom models, for example, liquid densities, are predicted, with a maximum absolute average deviation of 3% from both the theory and the MD simulations, while the interfacial tension is predicted, with a maximum absolute average of 8%. The extension to mixtures is exemplified by considering a binary system of hexane (chain fluid) and tetrahydro-2H-pyran (ring fluid).
More Related Videos
 11:28
A Coupled Experiment-finite Element Modeling Methodology for Assessing High Strain Rate Mechanical Response of Soft Biomaterials
Published on: May 18, 2015
 08:55
Methods of Ex Situ and In Situ Investigations of Structural Transformations: The Case of Crystallization of Metallic Glasses
Published on: June 7, 2018
Related Concept Videos
 01:07
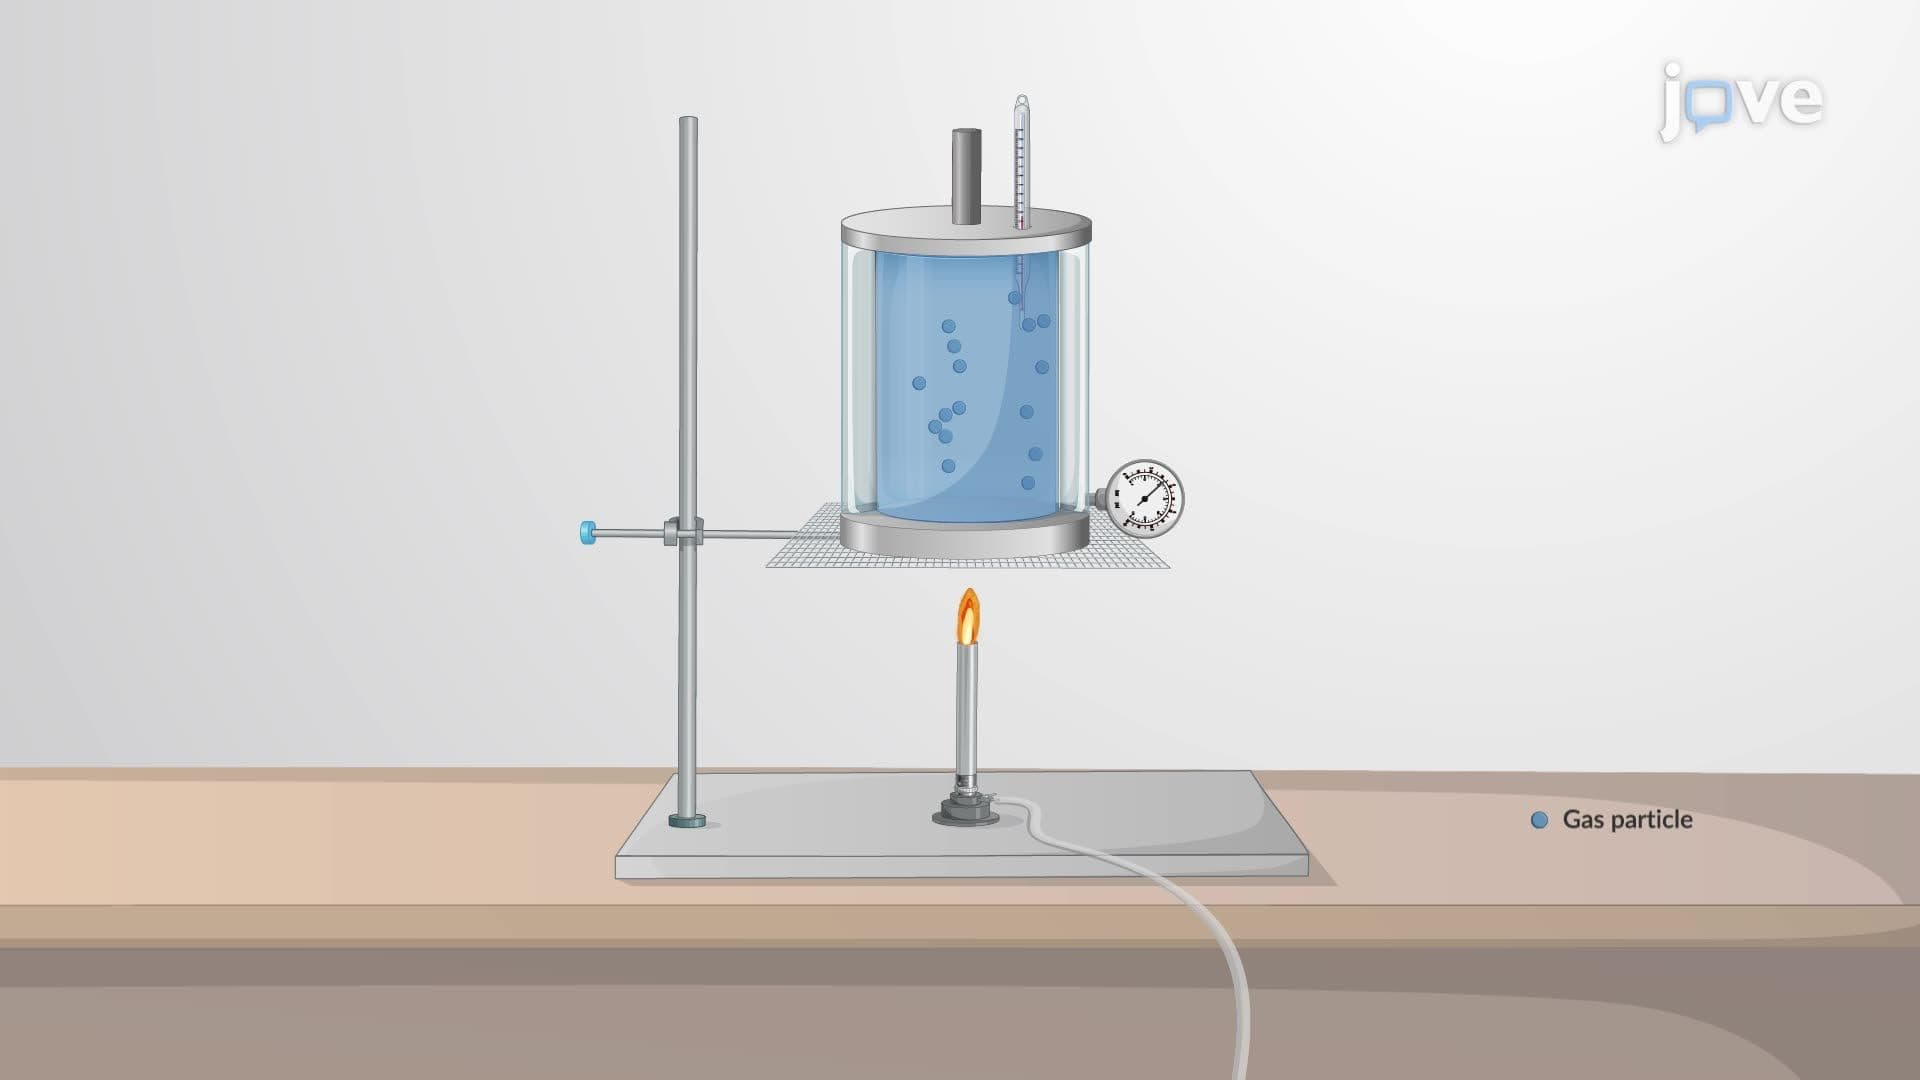
Equation of State
 01:27
Variables and Equations of State
 01:22
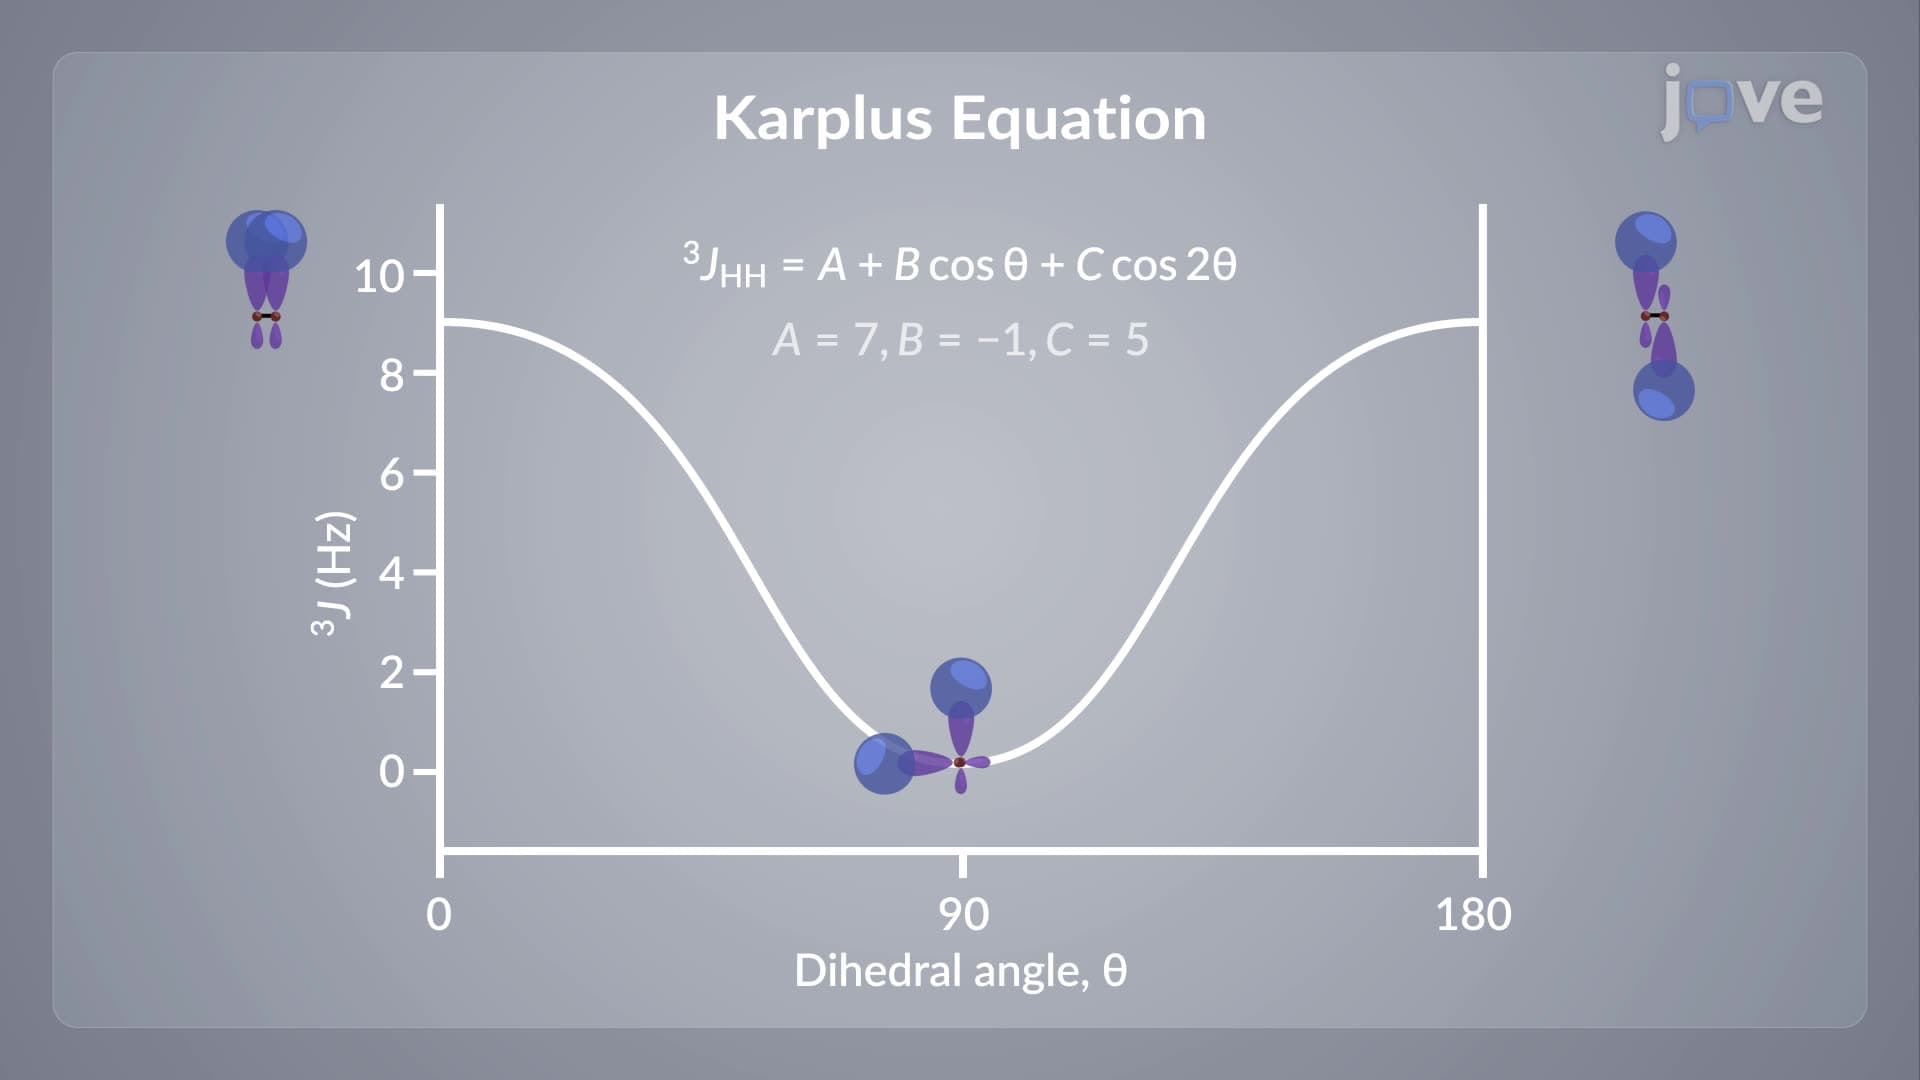
Spin–Spin Coupling: Three-Bond Coupling (Vicinal Coupling)
The extent of coupling depends on the C‑C bond length, the two H‑C‑C angles, any electron-withdrawing substituents, and the dihedral angle between the involved orbitals. The...
 01:10
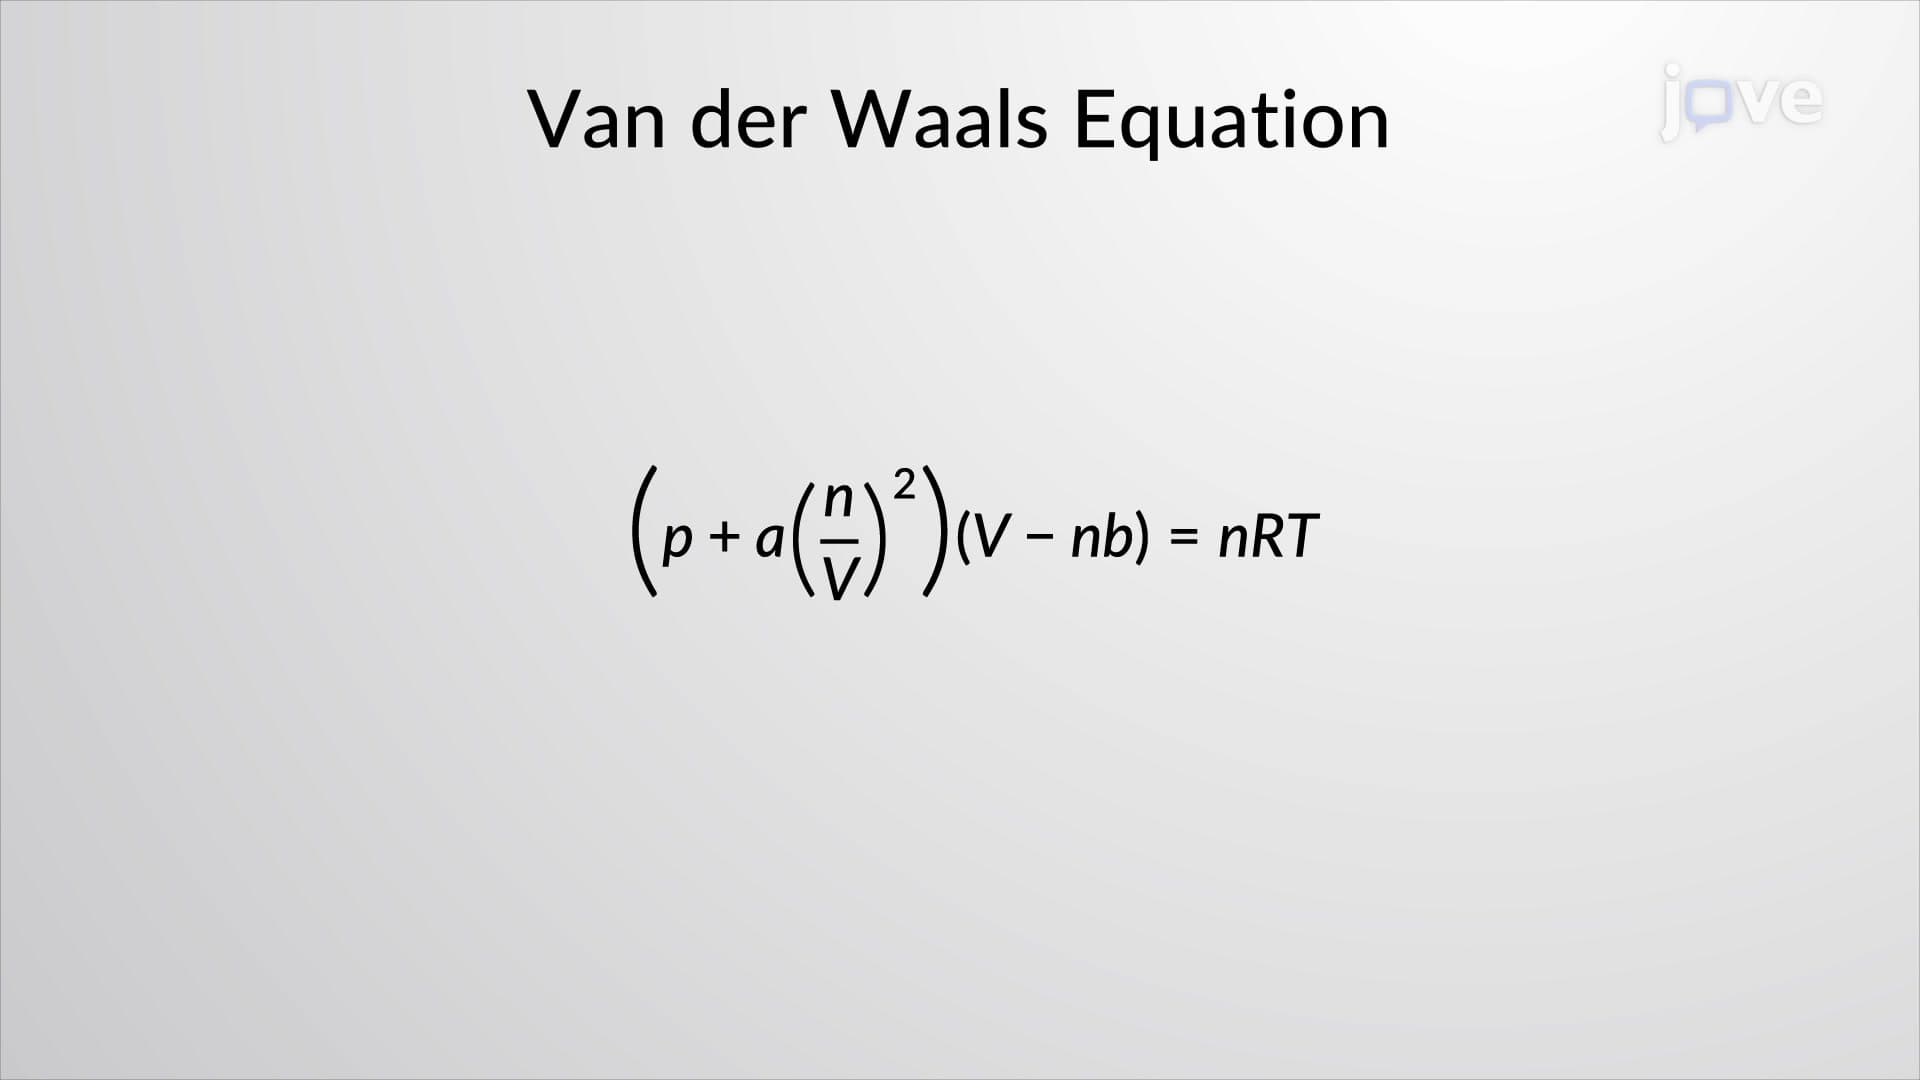
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...