Related Experiment Video
Updated: Feb 25, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A machine learning approach to graph-theoretical cluster expansions of the energy of adsorbate layers
Emanuele Vignola1, Stephan N Steinmann1, Bart D Vandegehuchte2
1Université Lyon, ENS de Lyon, CNRS, Université Lyon 1, Laboratoire de Chimie UMR 5182, F-69342 Lyon, France.

Abstract:
The accurate description of the energy of adsorbate layers is crucial for the understanding of chemistry at interfaces. For heterogeneous catalysis, not only the interaction of the adsorbate with the surface but also the adsorbate-adsorbate lateral interactions significantly affect the activation energies of reactions. Modeling the interactions of the adsorbates with the catalyst surface and with each other can be efficiently achieved in the cluster expansion Hamiltonian formalism, which has recently been implemented in a graph-theoretical kinetic Monte Carlo (kMC) scheme to describe multi-dentate species. Automating the development of the cluster expansion Hamiltonians for catalytic systems is challenging and requires the mapping of adsorbate configurations for extended adsorbates onto a graphical lattice. The current work adopts machine learning methods to reach this goal. Clusters are automatically detected based on formalized, but intuitive chemical concepts. The corresponding energy coefficients for the cluster expansion are calculated by an inversion scheme. The potential of this method is demonstrated for the example of ethylene adsorption on Pd(111), for which we propose several expansions, depending on the graphical lattice. It turns out that for this system, the best description is obtained as a combination of single molecule patterns and a few coupling terms accounting for lateral interactions.
More Related Videos
Related Concept Videos
 02:34
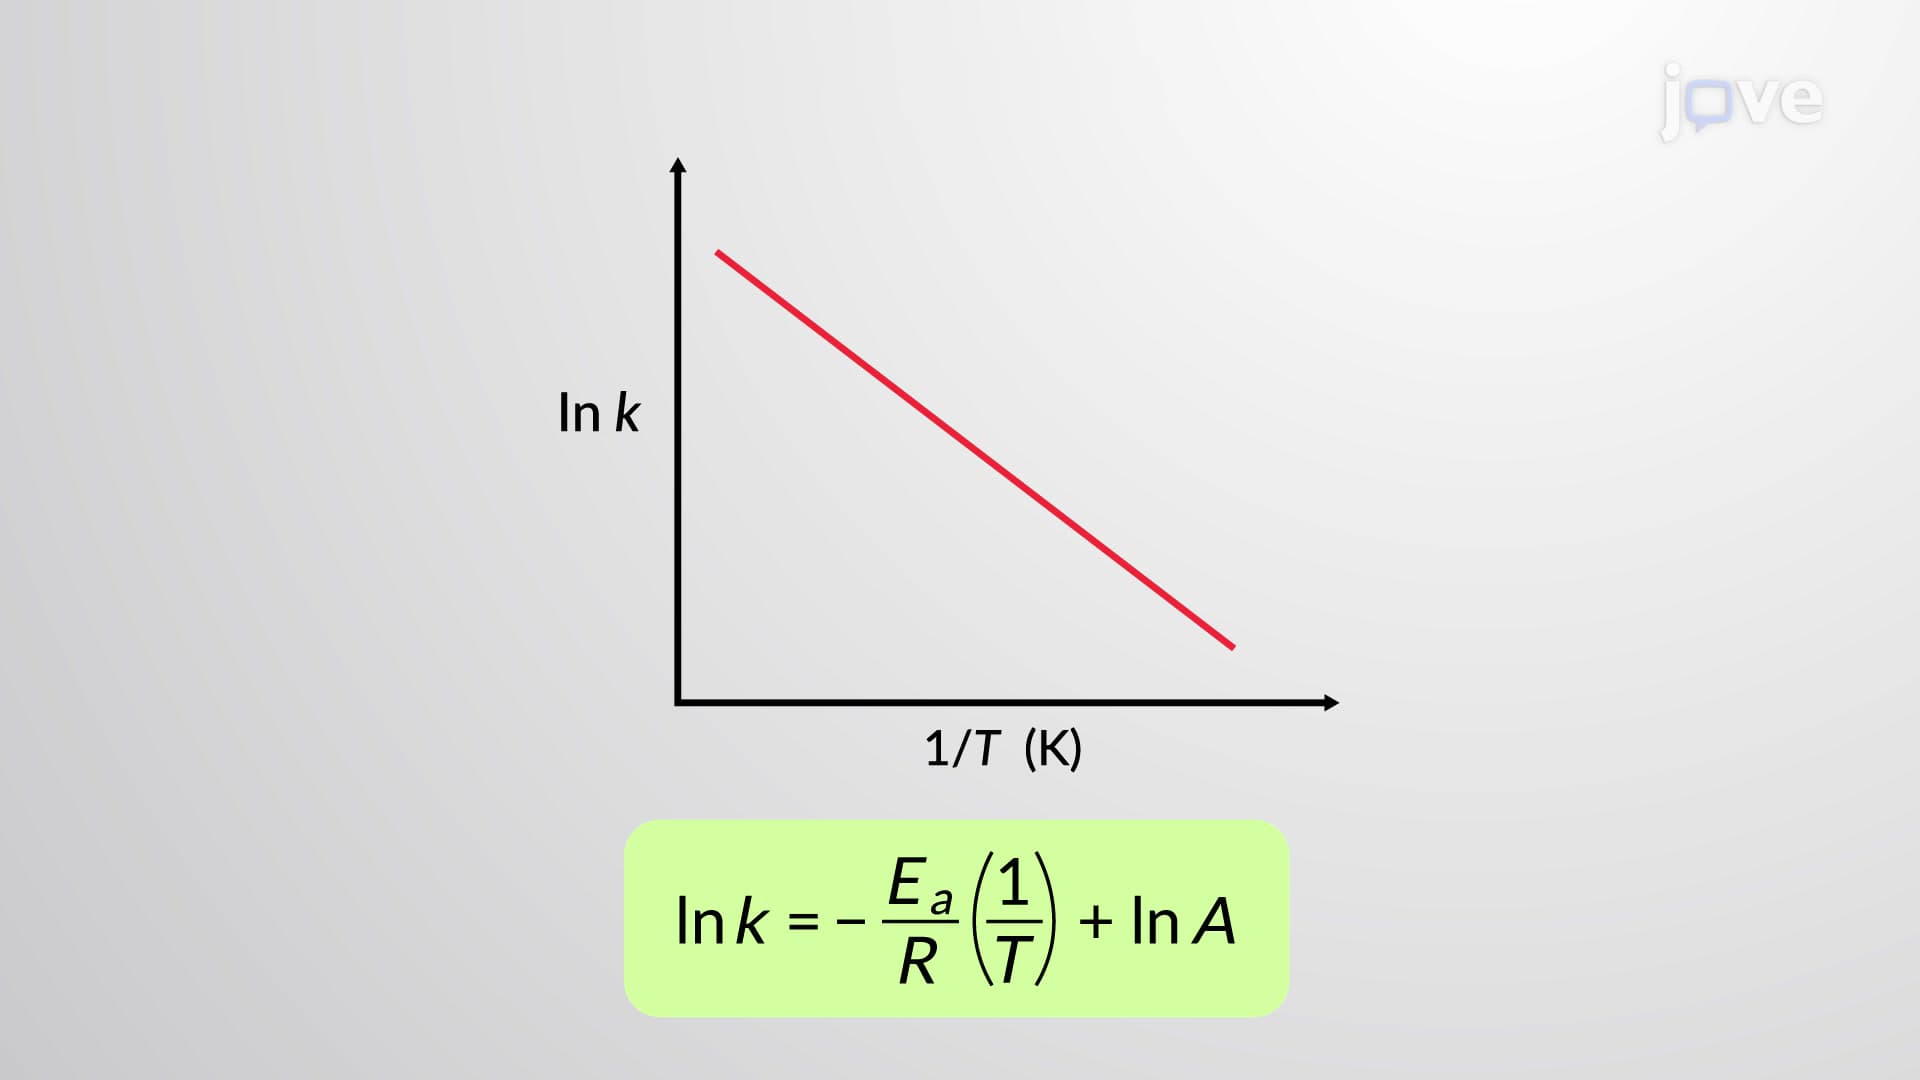
Arrhenius Plots
The Arrhenius equation can be used...
 01:09
Analyte Adsorption and Distribution
 03:51
Molecular Orbital Theory II
 02:13
Energy Diagrams, Transition States, and Intermediates
 02:27
Predicting Molecular Geometry
 02:54
Trends in Lattice Energy: Ion Size and Charge