Differential methylation analysis of reduced representation bisulfite sequencing experiments using edgeR
Related Experiment Video
Updated: Feb 15, 2026
Enhanced Reduced Representation Bisulfite Sequencing for Assessment of DNA Methylation at Base Pair Resolution
Published on: February 24, 2015
Yunshun Chen1,2, Bhupinder Pal1,2, Jane E Visvader1,2
1The Walter and Eliza Hall Institute of Medical Research, Parkville, VIC, 3052, Australia.

Abstract:
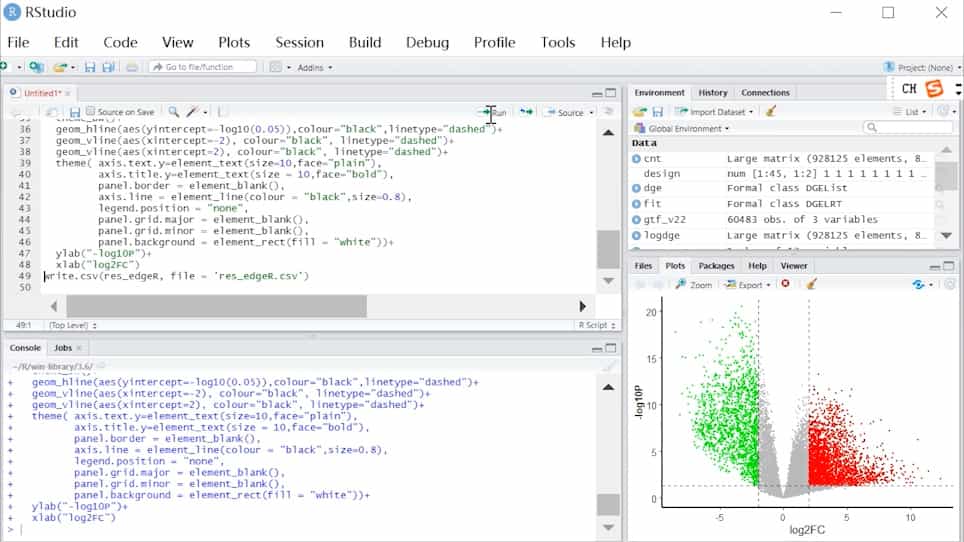
Cytosine methylation is an important DNA epigenetic modification. In vertebrates, methylation occurs at CpG sites, which are dinucleotides where a cytosine is immediately followed by a guanine in the DNA sequence from 5' to 3'. When located in the promoter region of a gene, DNA methylation is often associated with transcriptional silencing of the gene. Aberrant DNA methylation is associated with the development of various diseases such as cancer. Bisulfite sequencing (BS-seq) is the current "gold-standard" technology for high-resolution profiling of DNA methylation. Reduced representation bisulfite sequencing (RRBS) is an efficient form of BS-seq that targets CpG-rich DNA regions in order to save sequencing costs. A typical bioinformatics aim is to identify CpGs that are differentially methylated (DM) between experimental conditions. This workflow demonstrates that differential methylation analysis of RRBS data can be conducted using software and methodology originally developed for RNA-seq data. The RNA-seq pipeline is adapted to methylation by adding extra columns to the design matrix to account for read coverage at each CpG, after which the RRBS and RNA-seq pipelines are almost identical. This approach is statistically natural and gives analysts access to a rich collection of analysis tools including generalized linear models, gene set testing and pathway analysis. The article presents a complete start to finish case study analysis of RRBS profiles of different cell populations from the mouse mammary gland using the Bioconductor package edgeR. We show that lineage-committed cells are typically hyper-methylated compared to progenitor cells and this is true on all the autosomes but not the sex chromosomes. We demonstrate a strong negative correlation between methylation of promoter regions and gene expression as measured by RNA-seq for the same cell types, showing that methylation is a regulatory mechanism involved in epithelial linear commitment.
Related Concept Videos
 01:27
State Space Representation
Consider an RLC circuit, a...
 01:28
Graphical Representation of Inequalities
 01:16
Control Volume and System Representations
The control volume approach considers a stationary region in space through which fluid flows. This region is bounded by a control surface. For instance, in the case of water...
 01:16
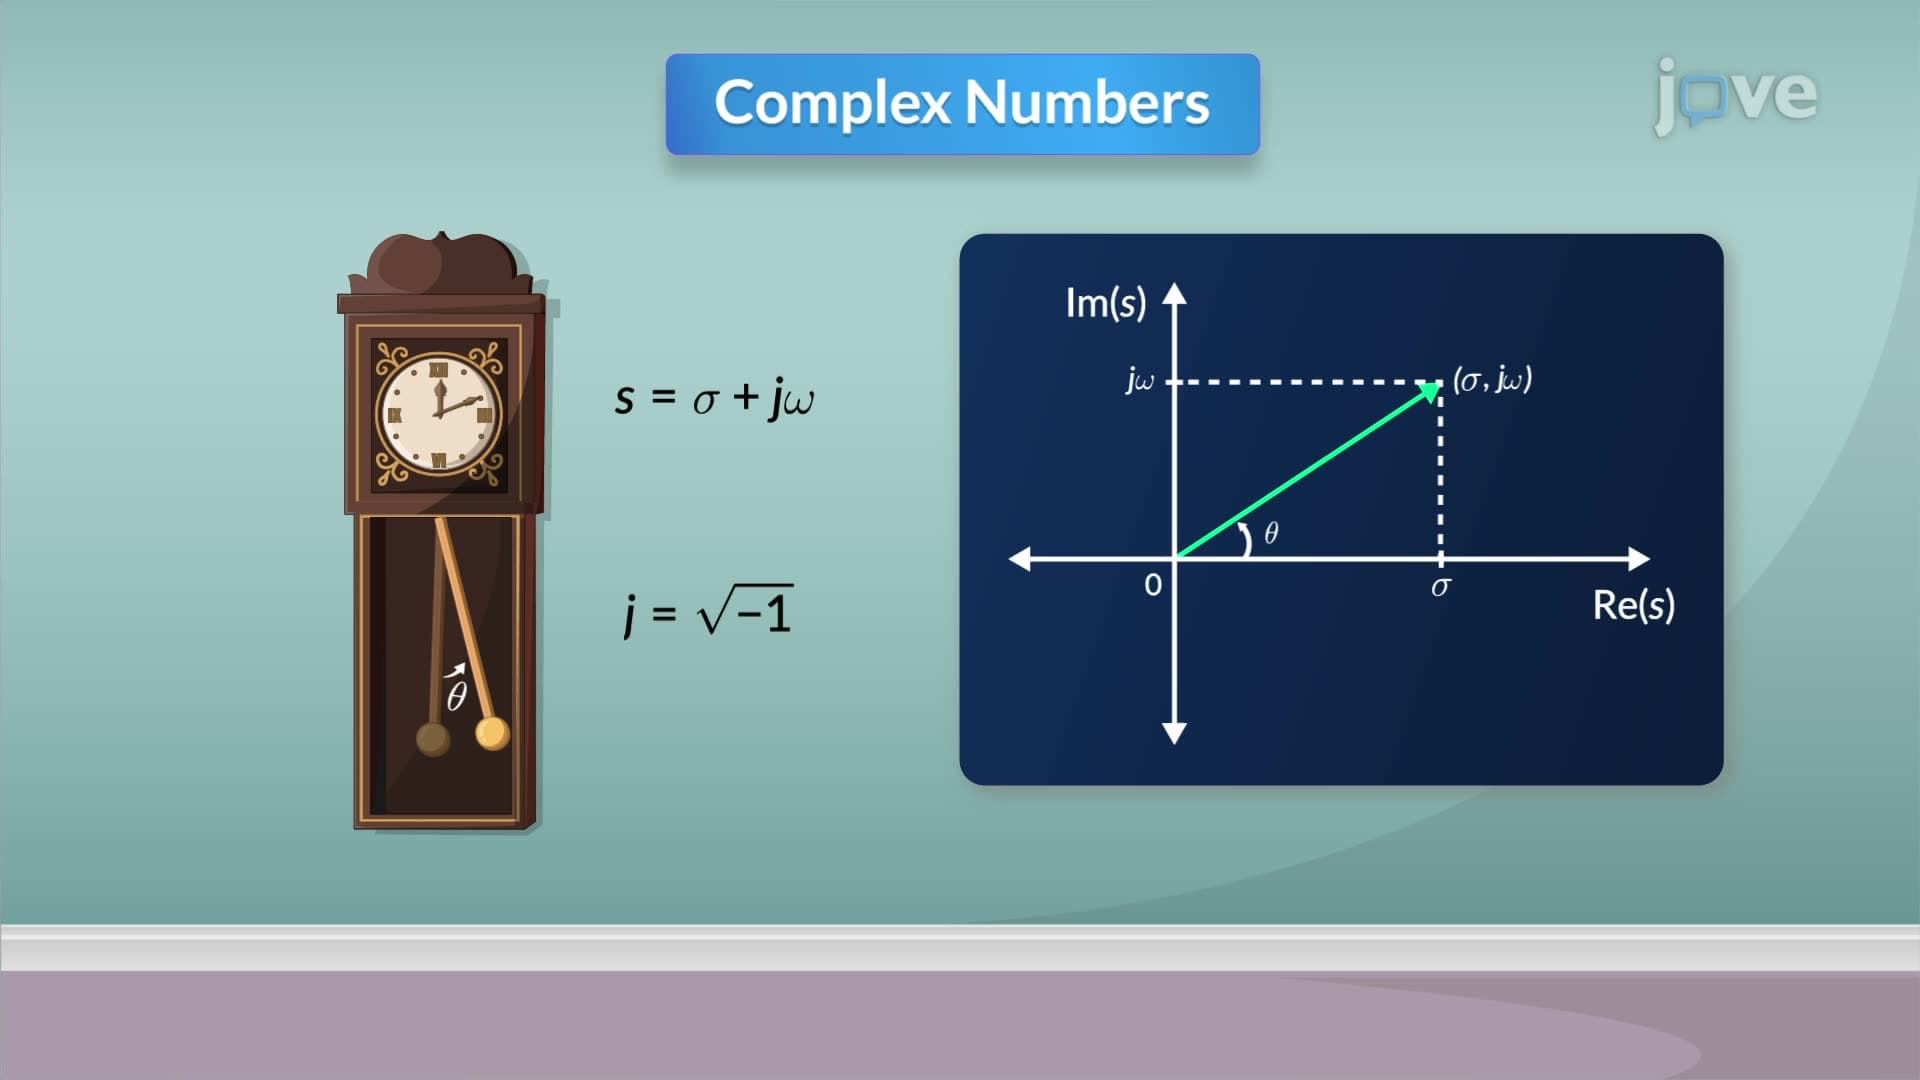
Vector Representation of Complex Numbers
Consider a function defined as the product of the complex factors in the numerator divided by the product of the complex factors in the...
 01:20
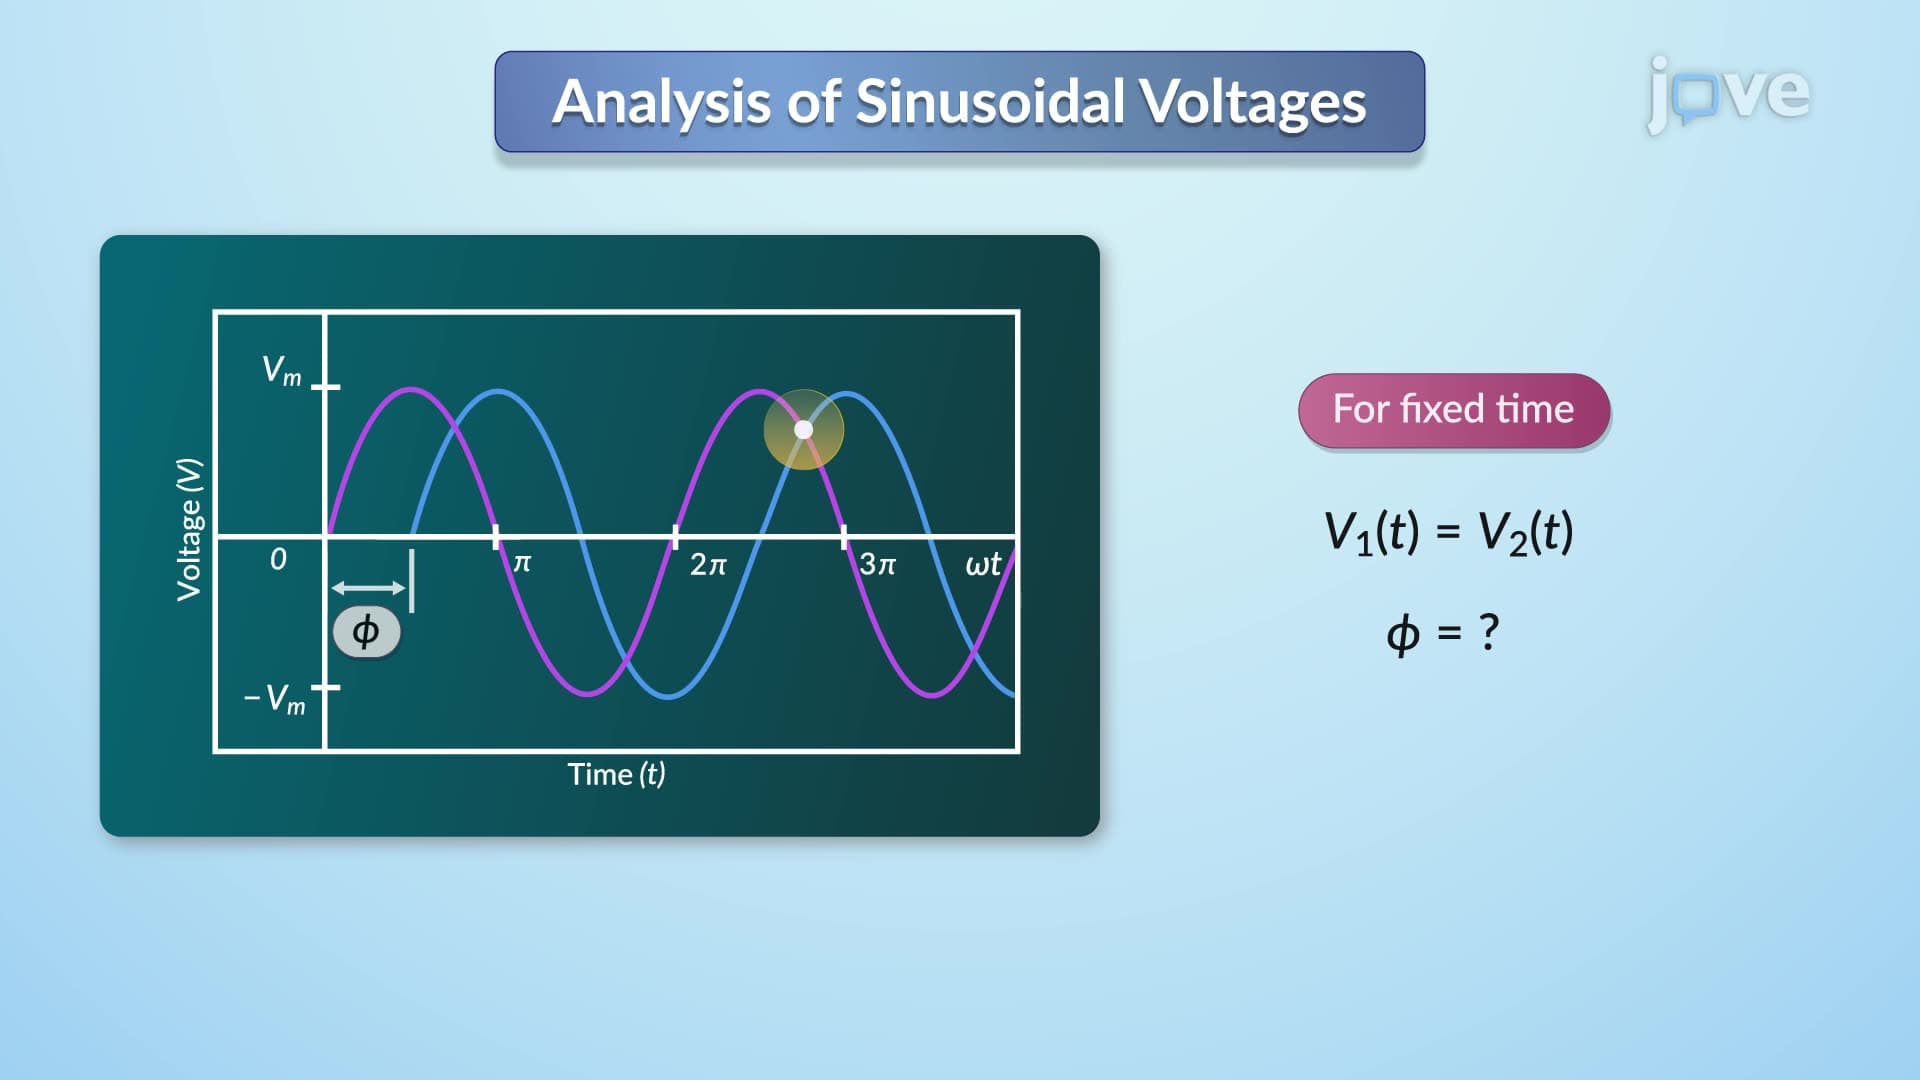
Graphical and Analytic Representation of Sinusoids
The first step is measuring the peak-to-peak value, which is twice the amplitude of the sinusoid. This provides information about the maximum voltage swing of the waveform.
Secondly, the period and angular frequency are determined. The period is the time taken for one complete cycle of the waveform, while...
 01:18
Reducing Line Loss
With a step-up transformer at the source, the voltage is increased, thereby reducing the current in the transmission lines since power loss in...