Related Experiment Video
Updated: Feb 12, 2026
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Chemical Reactivity and Spectroscopy Explored From QM/MM Molecular Dynamics Simulations Using the LIO Code
Juan P Marcolongo1, Ari Zeida1,2, Jonathan A Semelak1
1DQIAyQF, INQUIMAE-CONICET, Facultad de Ciencias Exactas y Naturales, Universidad de Buenos Aires, Buenos Aires, Argentina.

Abstract:
In this work we present the current advances in the development and the applications of LIO, a lab-made code designed for density functional theory calculations in graphical processing units (GPU), that can be coupled with different classical molecular dynamics engines. This code has been thoroughly optimized to perform efficient molecular dynamics simulations at the QM/MM DFT level, allowing for an exhaustive sampling of the configurational space. Selected examples are presented for the description of chemical reactivity in terms of free energy profiles, and also for the computation of optical properties, such as vibrational and electronic spectra in solvent and protein environments.
More Related Videos
Related Concept Videos
 02:00
Molecular Models
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 01:14
Molecular Spectroscopy: Absorption and Emission
 01:15
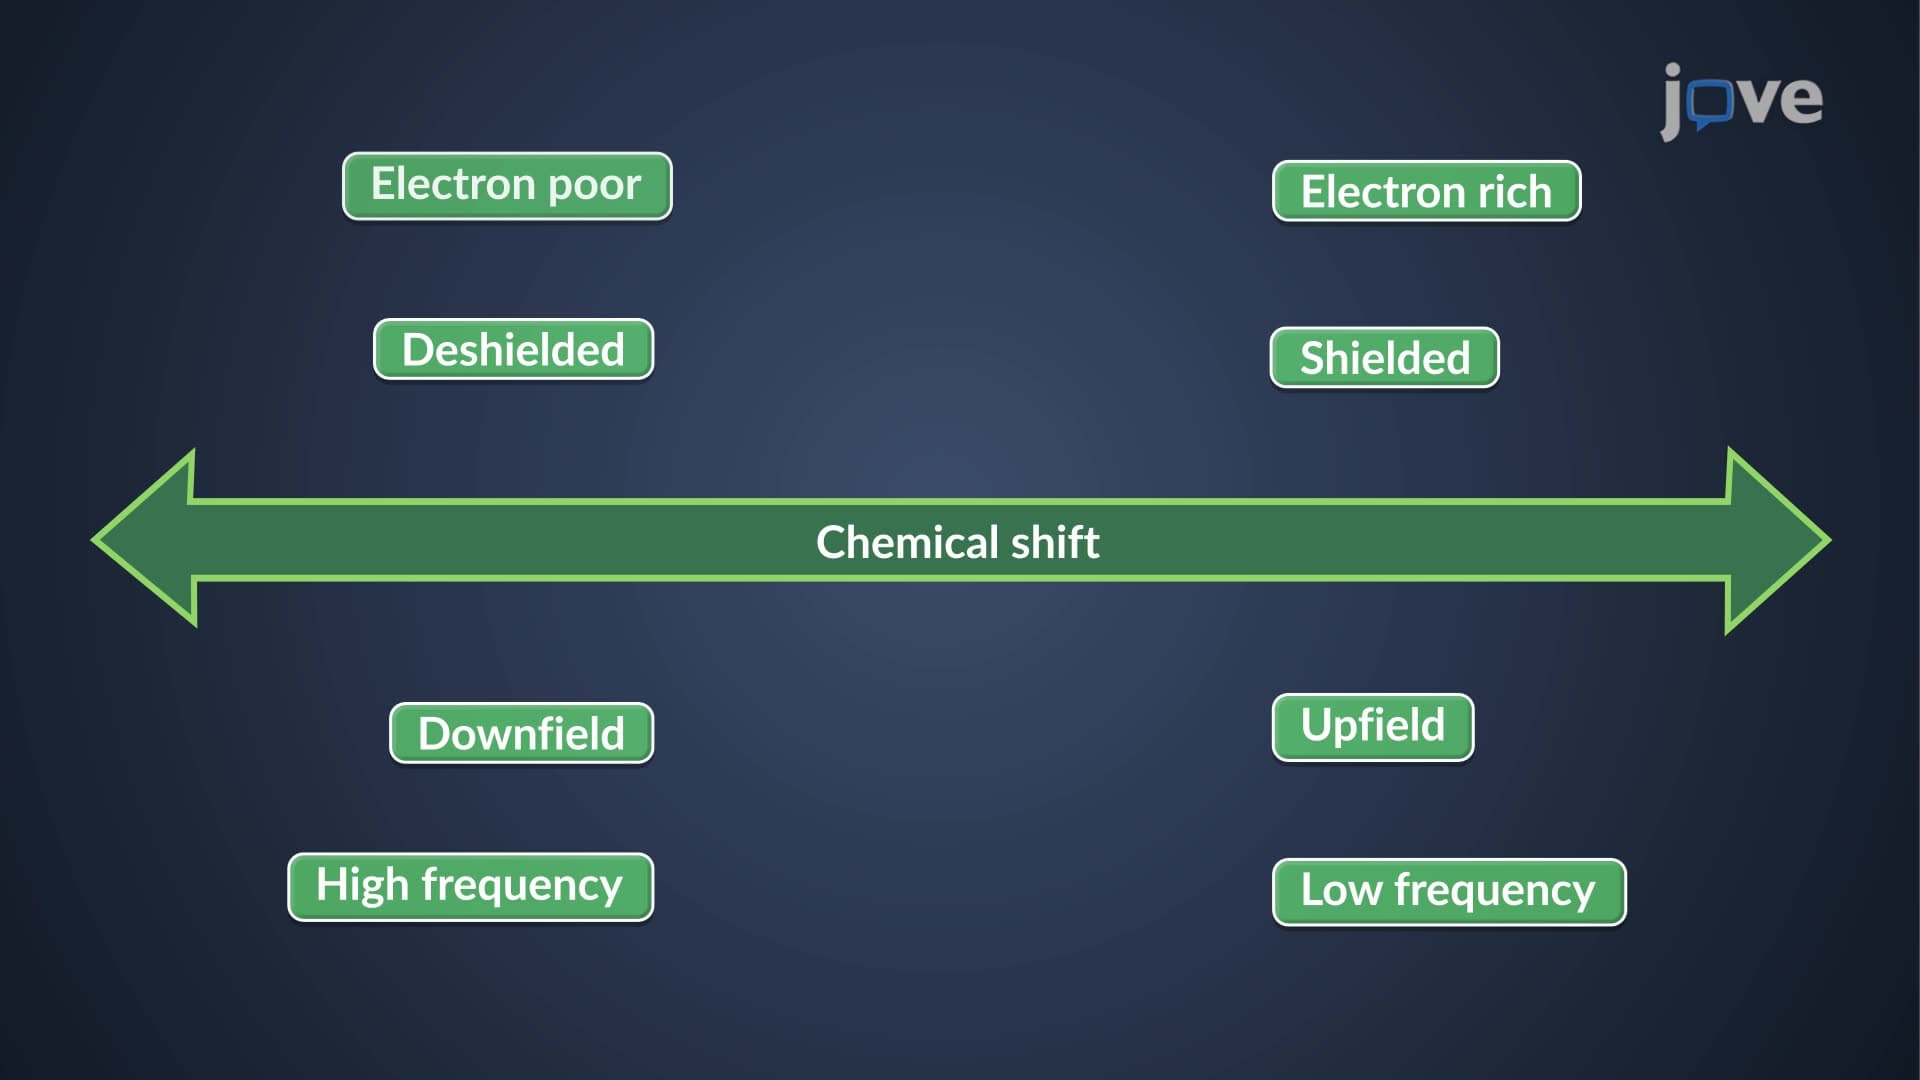
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 02:20
Dynamic Equilibrium
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions