Related Experiment Video
Updated: Feb 5, 2026

Author Spotlight: Isolating Biomolecules from Mouse Tears — A Methodology for Molecular Analysis and Biomarker Research
Published on: July 19, 2024
Assessment and optimization of collective variables for protein conformational landscape: GB1 β-hairpin as a case
Navjeet Ahalawat1, Jagannath Mondal1
1Tata Institute of Fundamental Research, Center for Interdisciplinary Sciences, Hyderabad 500107, India.

Abstract:
Collective variables (CVs), when chosen judiciously, can play an important role in recognizing rate-limiting processes and rare events in any biomolecular systems. However, high dimensionality and inherent complexities associated with such biochemical systems render the identification of an optimal CV a challenging task, which in turn precludes the elucidation of an underlying conformational landscape in sufficient details. In this context, a relevant model system is presented by a 16-residue β-hairpin of GB1 protein. Despite being the target of numerous theoretical and computational studies for understanding the protein folding, the set of CVs optimally characterizing the conformational landscape of the β-hairpin of GB1 protein has remained elusive, resulting in a lack of consensus on its folding mechanism. Here we address this by proposing a pair of optimal CVs which can resolve the underlying free energy landscape of the GB1 hairpin quite efficiently. Expressed as a linear combination of a number of traditional CVs, the optimal CV for this system is derived by employing the recently introduced time-structured independent component analysis approach on a large number of independent unbiased simulations. By projecting the replica-exchange simulated trajectories along these pair of optimized CVs, the resulting free energy landscape of this system is able to resolve four distinct well-separated metastable states encompassing the extensive ensembles of folded, unfolded, and molten globule states. Importantly, the optimized CVs were found to be capable of automatically recovering a novel partial helical state of this protein, without needing to explicitly invoke helicity as a constituent CV. Furthermore, a quantitative sensitivity analysis of each constituent in the optimized CV provided key insights on the relative contributions of the constituent CVs in the overall free energy landscapes. Finally, the kinetic pathways connecting these metastable states, constructed using a Markov state model, provide an optimum description of the underlying folding mechanism of the peptide. Taken together, this work offers a quantitatively robust approach toward comprehensive mapping of the underlying folding landscape of a quintessential model system along its optimized CV.
More Related Videos
Related Concept Videos
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:20
Conformity
 02:15
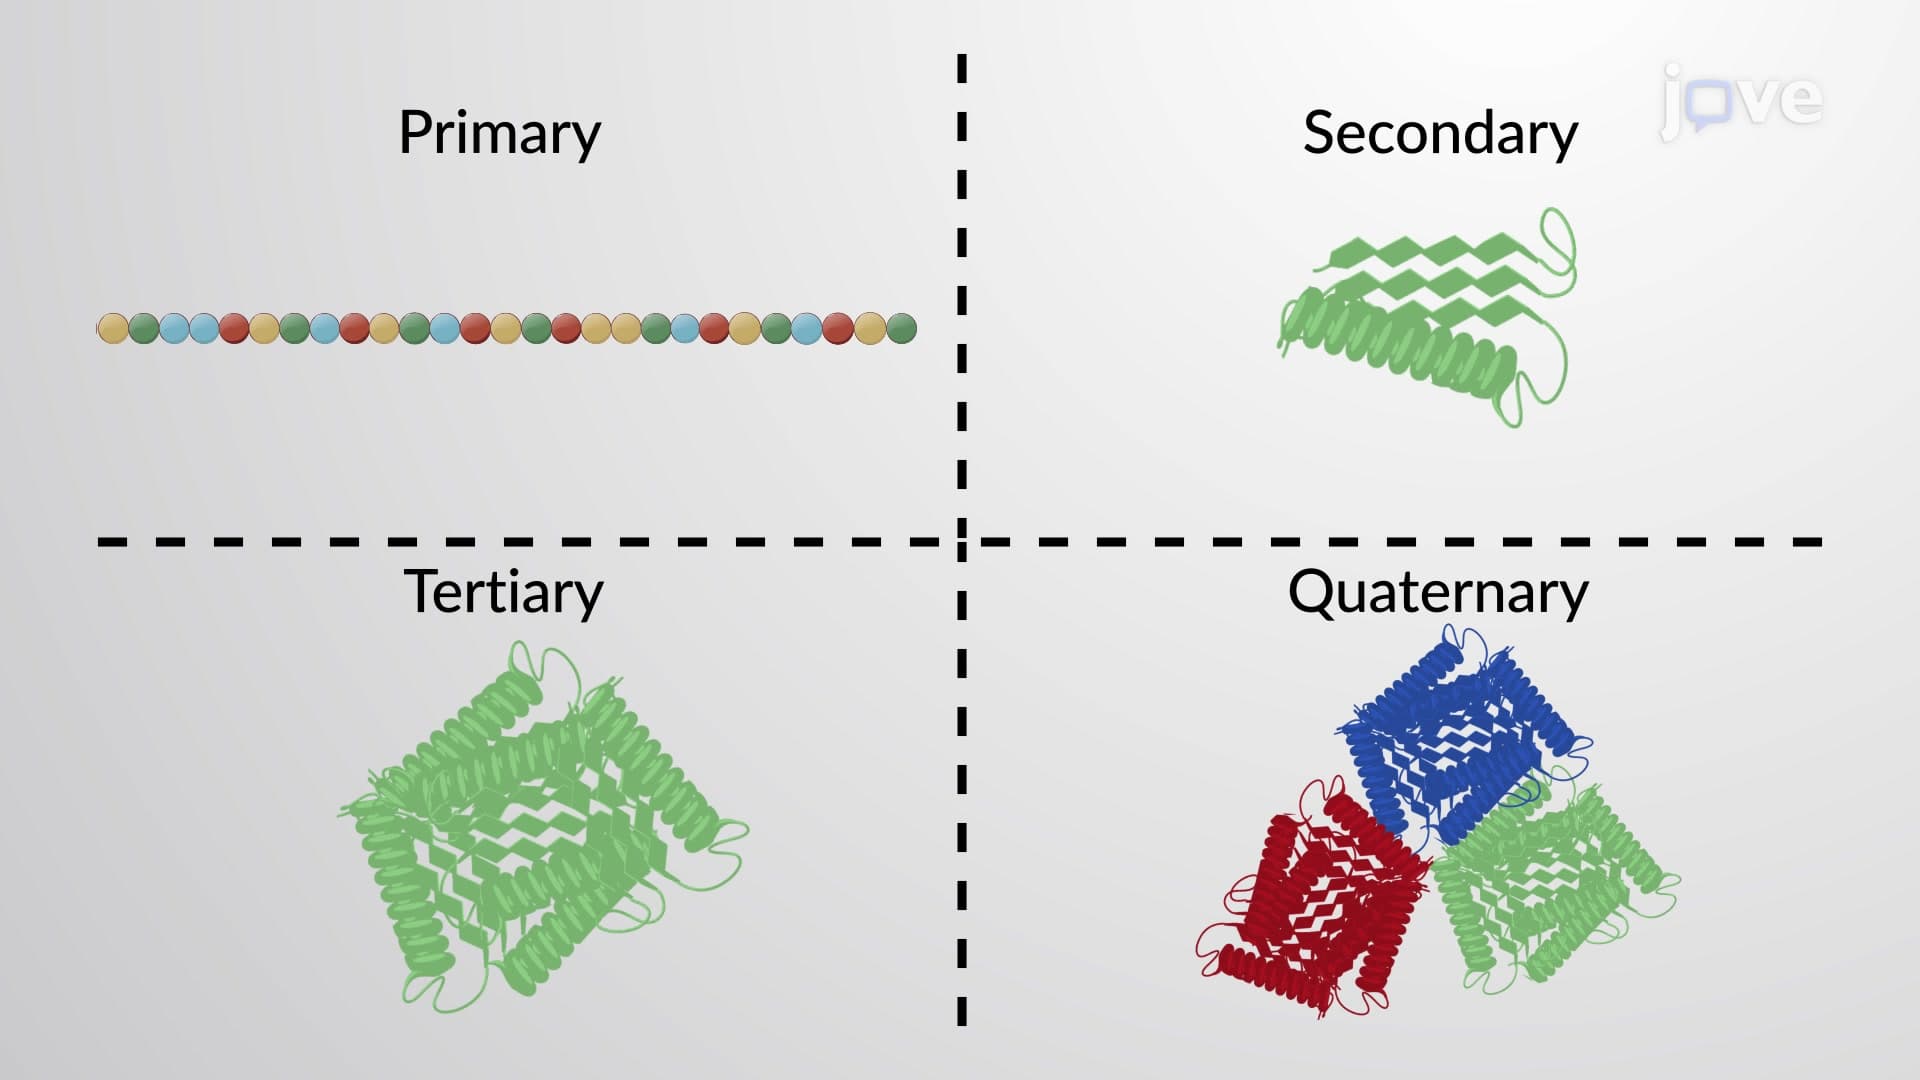
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 01:09
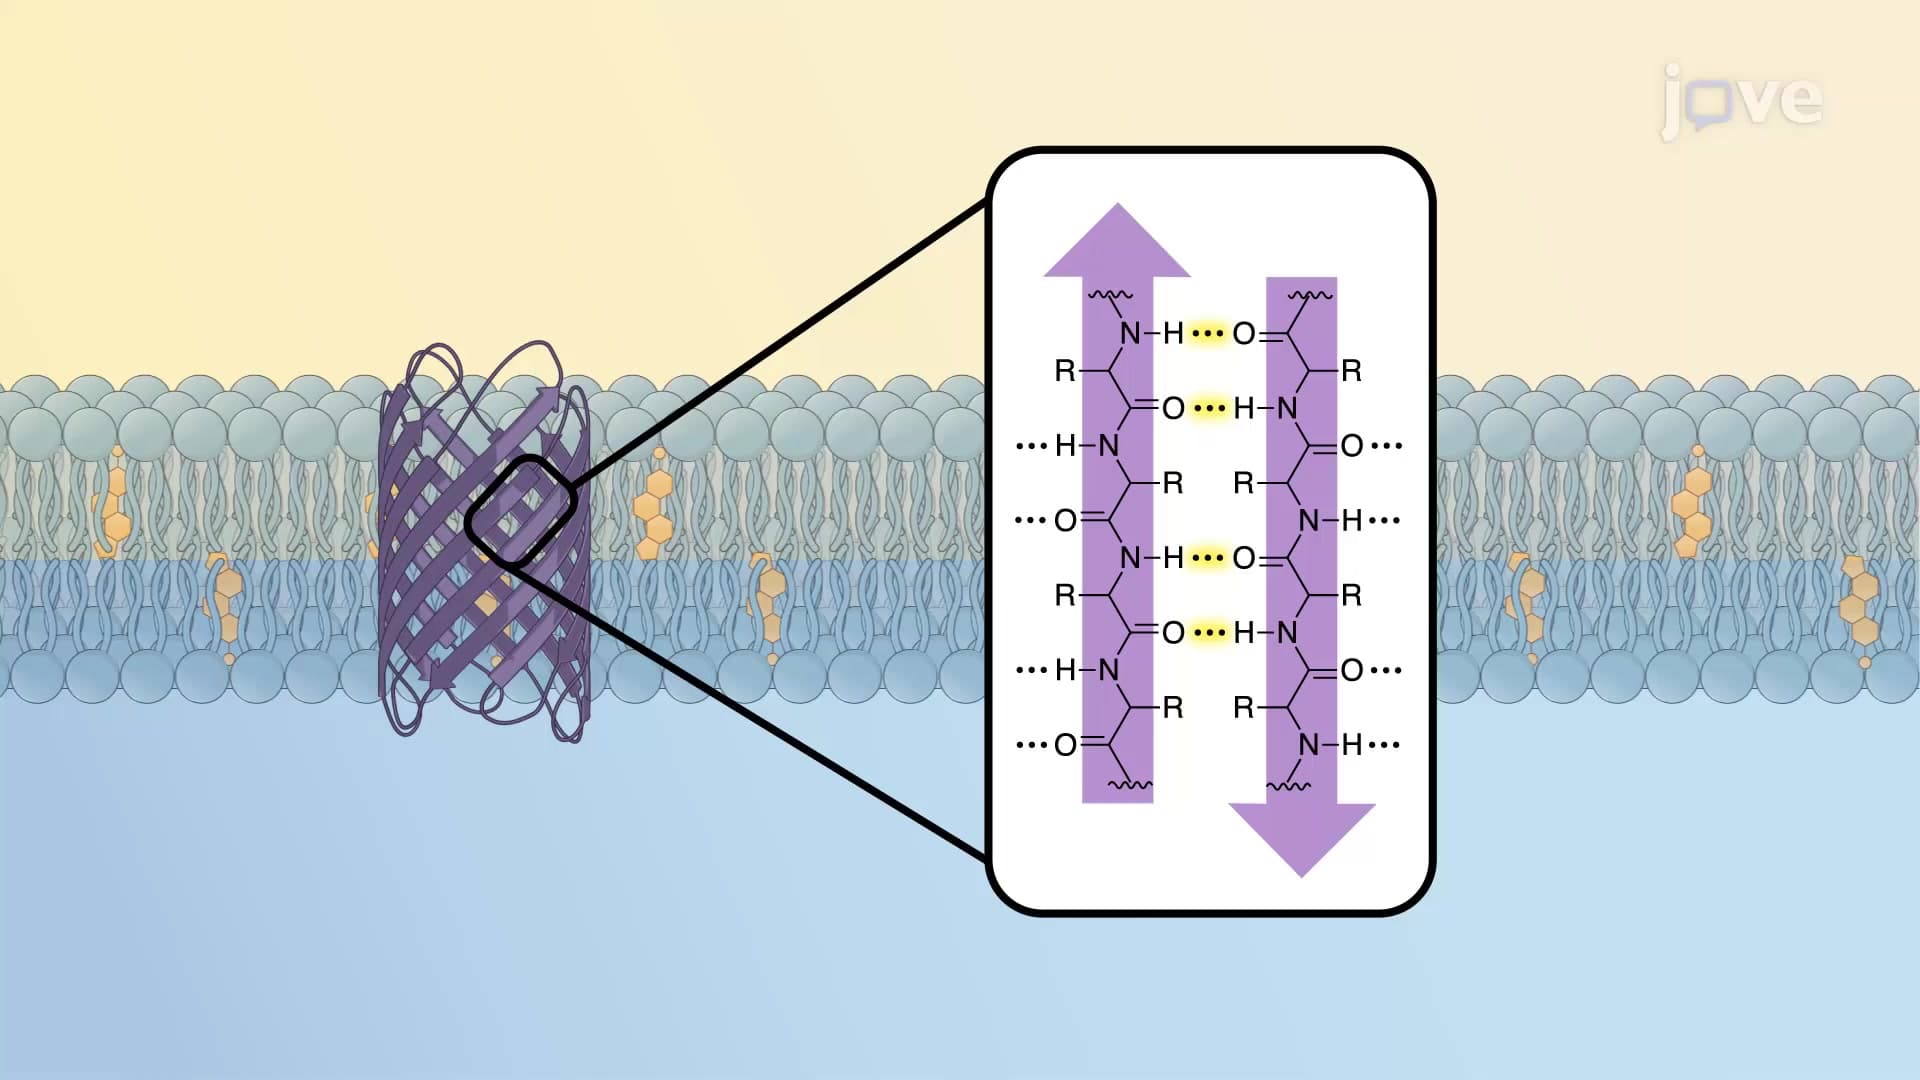
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as...
 02:20
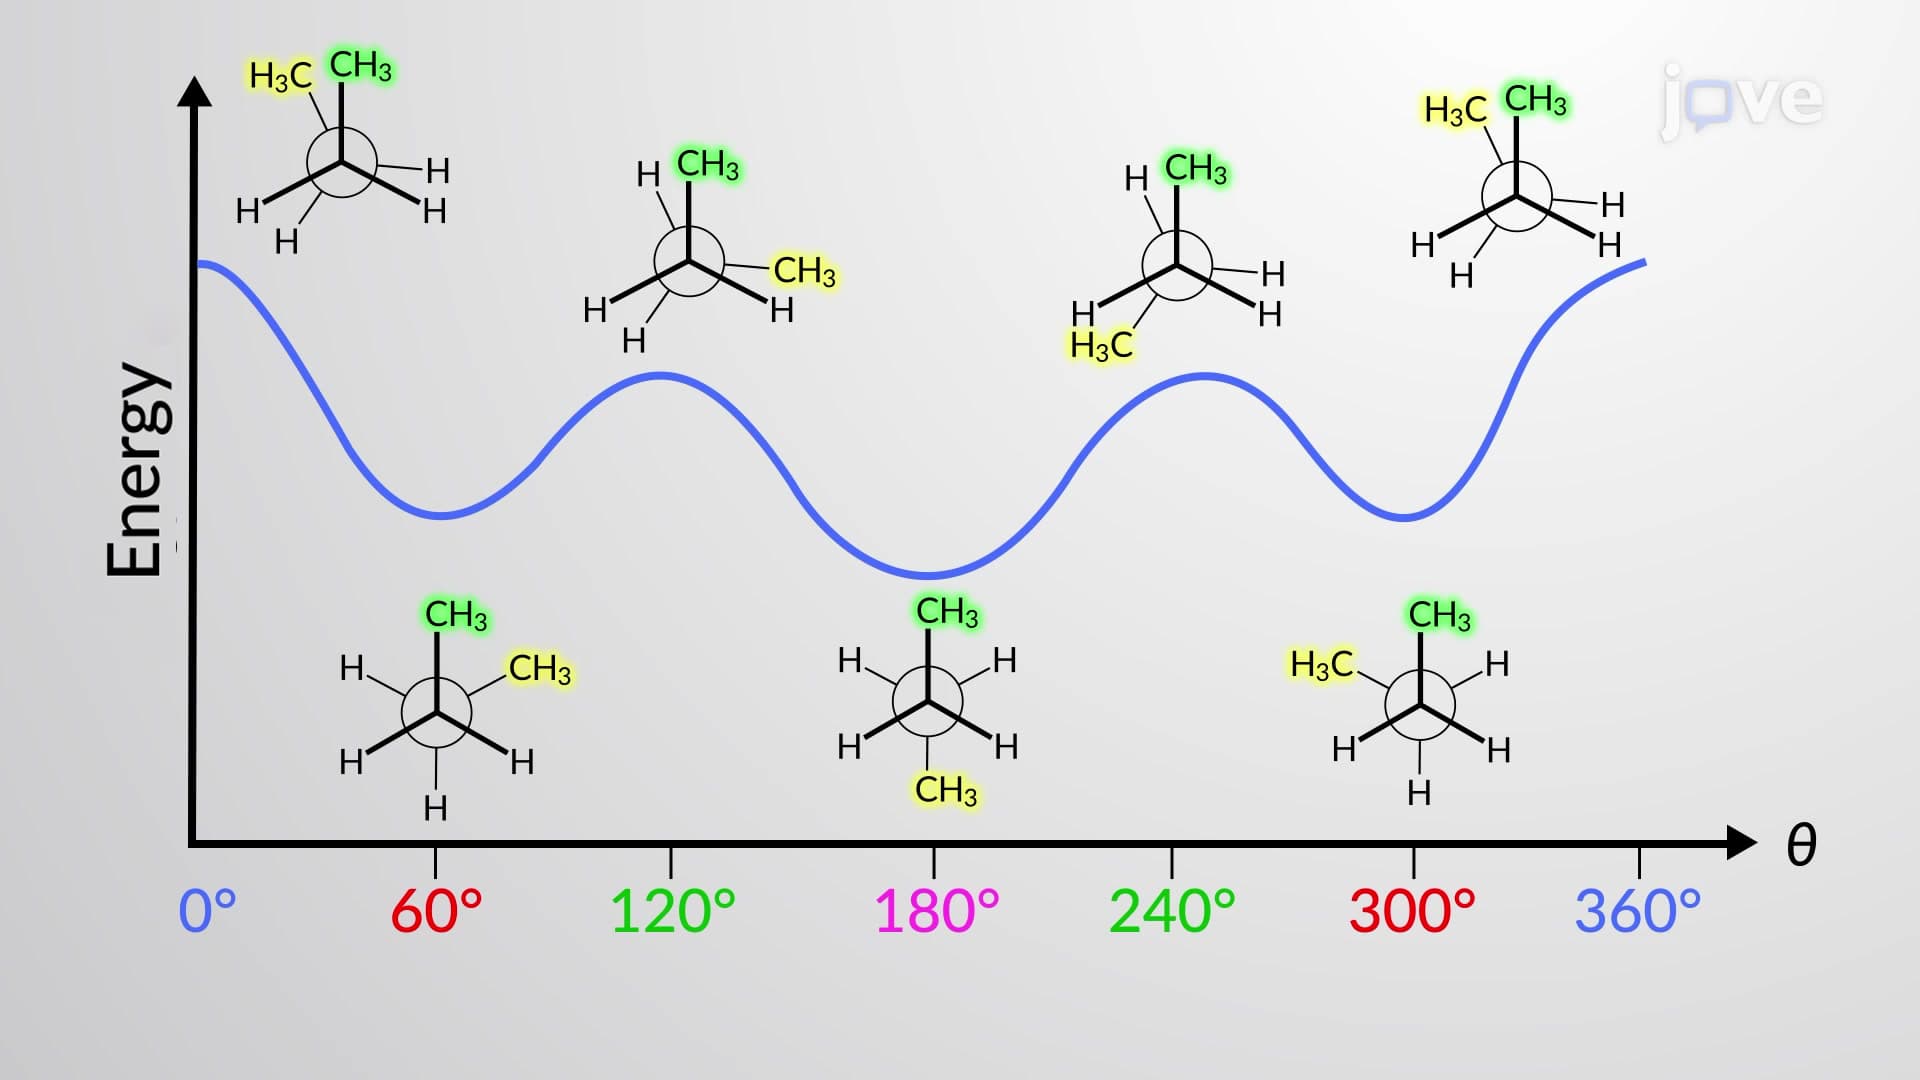
Conformations of Butane
 01:11
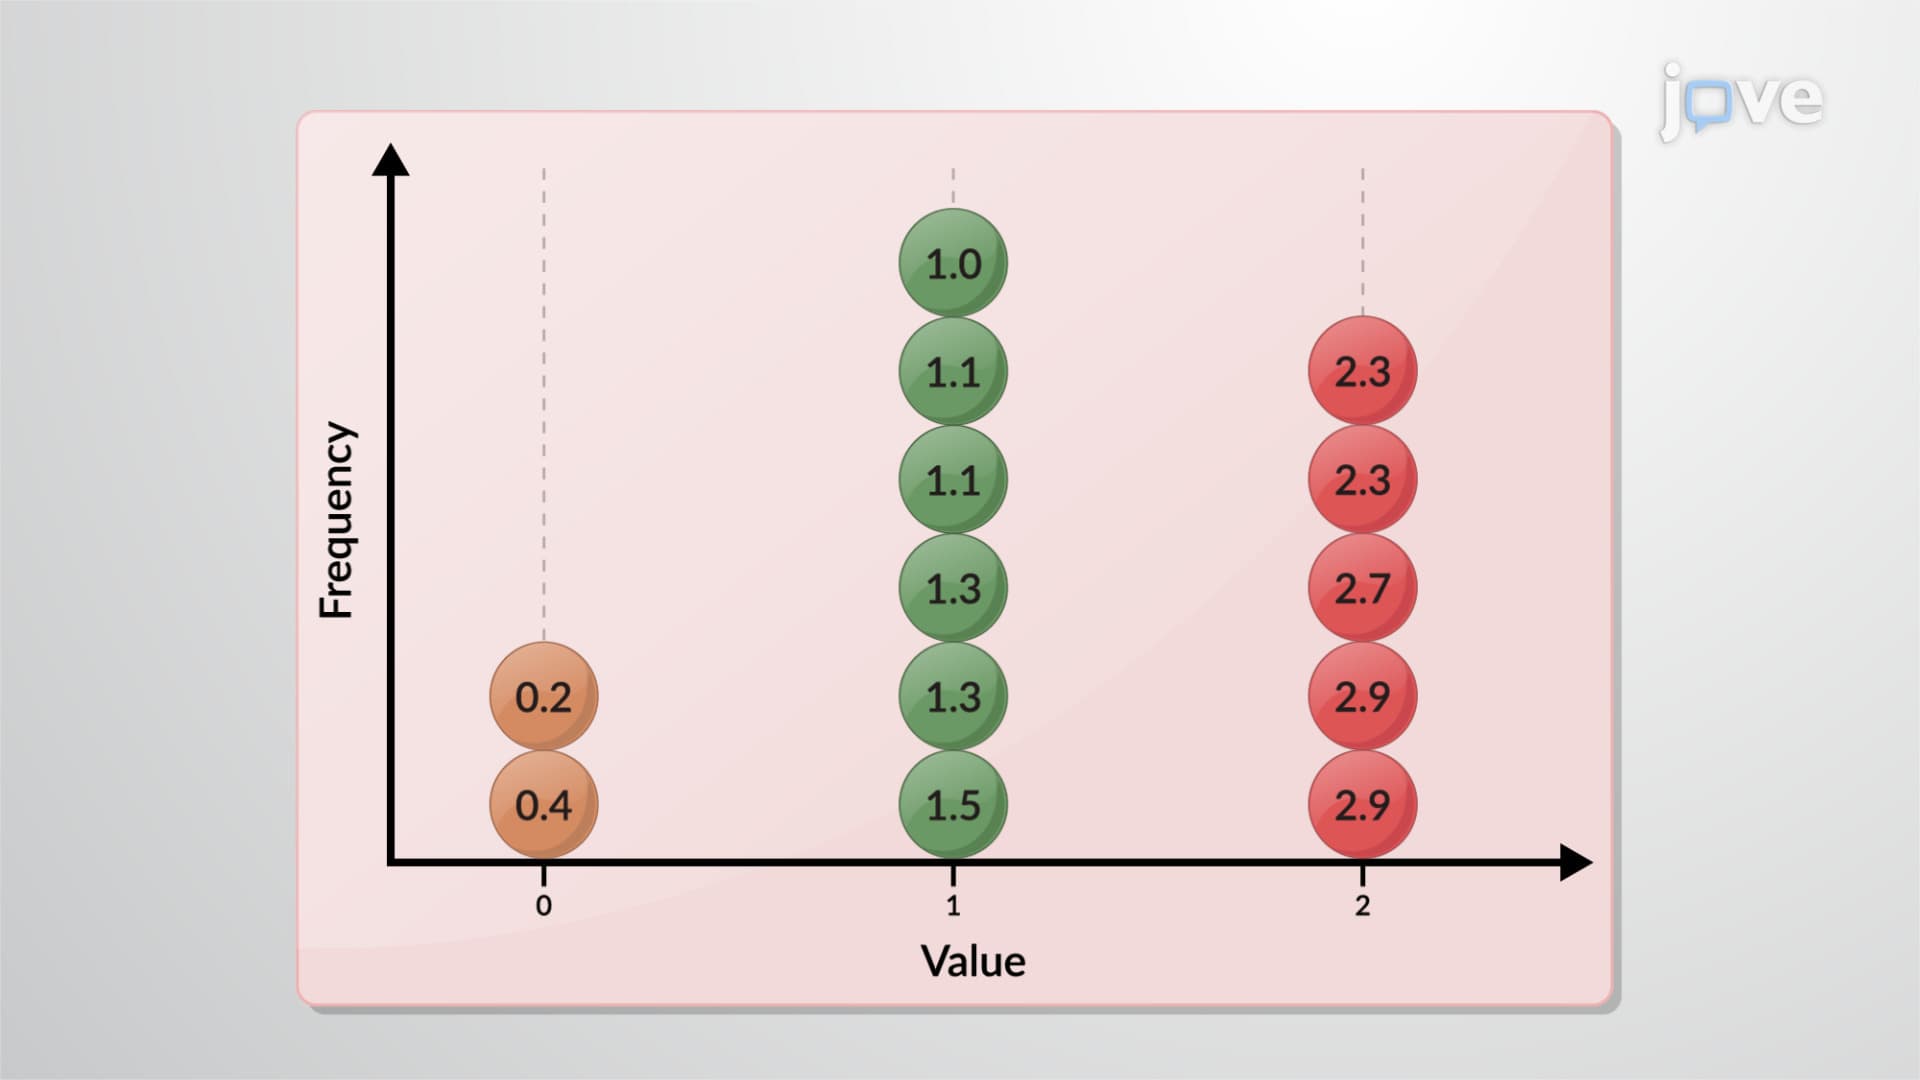
Variability: Analysis
The range is a simple measure of variability, indicating the difference between the highest and...