Related Experiment Video
Updated: Jan 6, 2026

Microscopic Visualization of Porous Nanographenes Synthesized through a Combination of Solution and On-Surface Chemistry
Published on: March 4, 2021
Externally predictive quantum-mechanical models for the adsorption of aromatic organic compounds by graphene-oxide
1Quantum Chemistry Group, Department of Chemistry and Centre of Advanced Studies in Chemistry, Panjab University, Chandigarh, India.

Abstract:
Graphene oxide is most often chosen as an alternative to graphene in the applications of carbon-based nanomaterials where adsorption is the primary process. However, its adsorption properties are poorly understood. The existing reports on the adsorption mechanism of graphene oxide rely on the linear free-energy/solvation-energy relationship (LFER/LSER) models. This computational work explores the role of quantum mechanical descriptors in the adsorption of aromatic organic compounds by graphene-oxide. For this, externally predictive quantitative models based on quantum-mechanical descriptors are developed and compared with the existing LSERs for the prediction of adsorption coefficients of organic compounds at three different adsorbate concentrations. The predictivity of the models is assessed using an external prediction set of compounds not used for developing the models. Notably, the mean polarizability, but originating from the quantum mechanical exchange interactions (between electrons of parallel spin), is found to be the most significant factor in driving the adsorption on graphene oxide. The present work also proposes quantum-mechanical-LSER models based on a combination of quantum-mechanical and LSER descriptors, which are in fact found to be equally predictive as the existing LSERs. The quantum-mechanical models proposed in this work are further utilized for the prediction of adsorption coefficients of aliphatic compounds.
More Related Videos
Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 03:51
Molecular Orbital Theory II
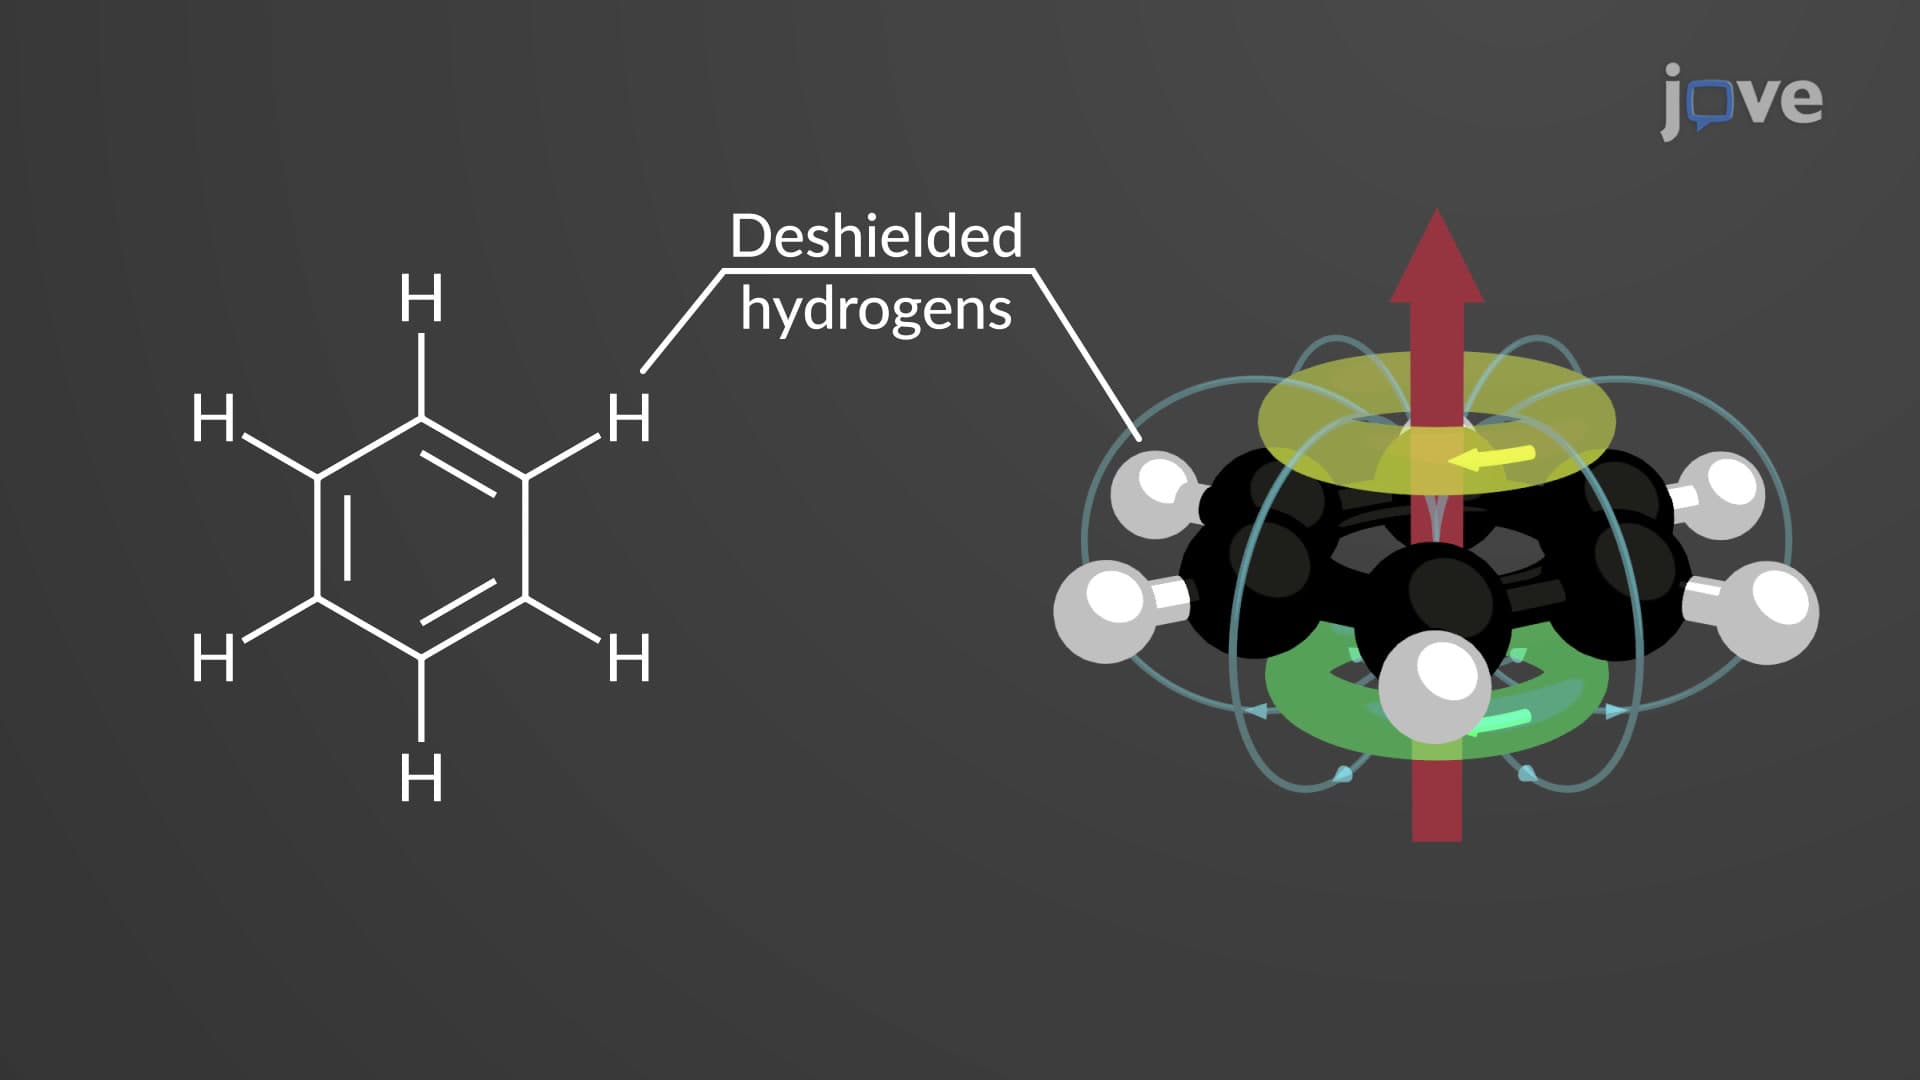 01:14
NMR Spectroscopy of Aromatic Compounds
 01:29
UV–Vis Spectroscopy: Woodward–Fieser Rules
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to...