Related Experiment Video
Updated: Dec 6, 2025

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Topological Phase and Strong Correlation in Rare-Earth Hexaborides XB6 (X = La, Ce, Pr, Nd, Pm, Sm, Eu)
Sheng-Hsiung Hung1, Horng-Tay Jeng1,2,3
1Department of Physics, National Tsing Hua University, Hsinchu 30013, Taiwan.

Abstract:
The rare-earth hexaboride SmB6, known as the topological Kondo insulator, has attracted tremendous attention in recent years. It was revealed that the topological phase of SmB6 is insensitive to the value of on-site Coulomb interactions (Hubbard U), indicating that the topological phase in SmB6 is robust against strong correlations. On the contrary, the isostructural YbB6 displays a sensitivity to the Hubbard U value. As U increases, YbB6 transforms from topological Kondo insulator to trivial insulator, showing the weak robustness of the topological phase of YbB6 against U. Consequently, the dependence of the topological phase on Hubbard U is a crucial issue in the rare-earth hexaboride family. In this work, we investigate the structural and electronic properties of rare-earth hexaboride compounds through first-principles calculations based on density functional theory. By taking the strong correlations into consideration using a wide range of on-site U values, we study the evolution of the topological phases in rare-earth hexaboride (XB6, X = La, Ce, Pr, Nd, Pm, Sm, Eu). Unlike YbB6, the topological trends in all the examples of XB6 studied in this work are insensitive to the U values. We conclude that in addition to the well-known SmB6, PmB6, NdB6 and EuB6 are also topologically nontrivial compounds, whereas LaB6, CeB6 and PrB6 are topologically trivial metal.
Related Concept Videos
 02:42
Valence Bond Theory
 04:01
VSEPR Theory and the Effect of Lone Pairs
 03:02
Colors and Magnetism
When atoms or molecules absorb light at the proper frequency, their electrons are excited to higher-energy orbitals. For many main group atoms and molecules, the absorbed photons are in the ultraviolet range of the electromagnetic spectrum, which cannot be detected by the human eye. For coordination compounds, the energy difference between the d orbitals often allows photons in the visible range to be absorbed and emitted, which is seen as colors by the human...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 03:26
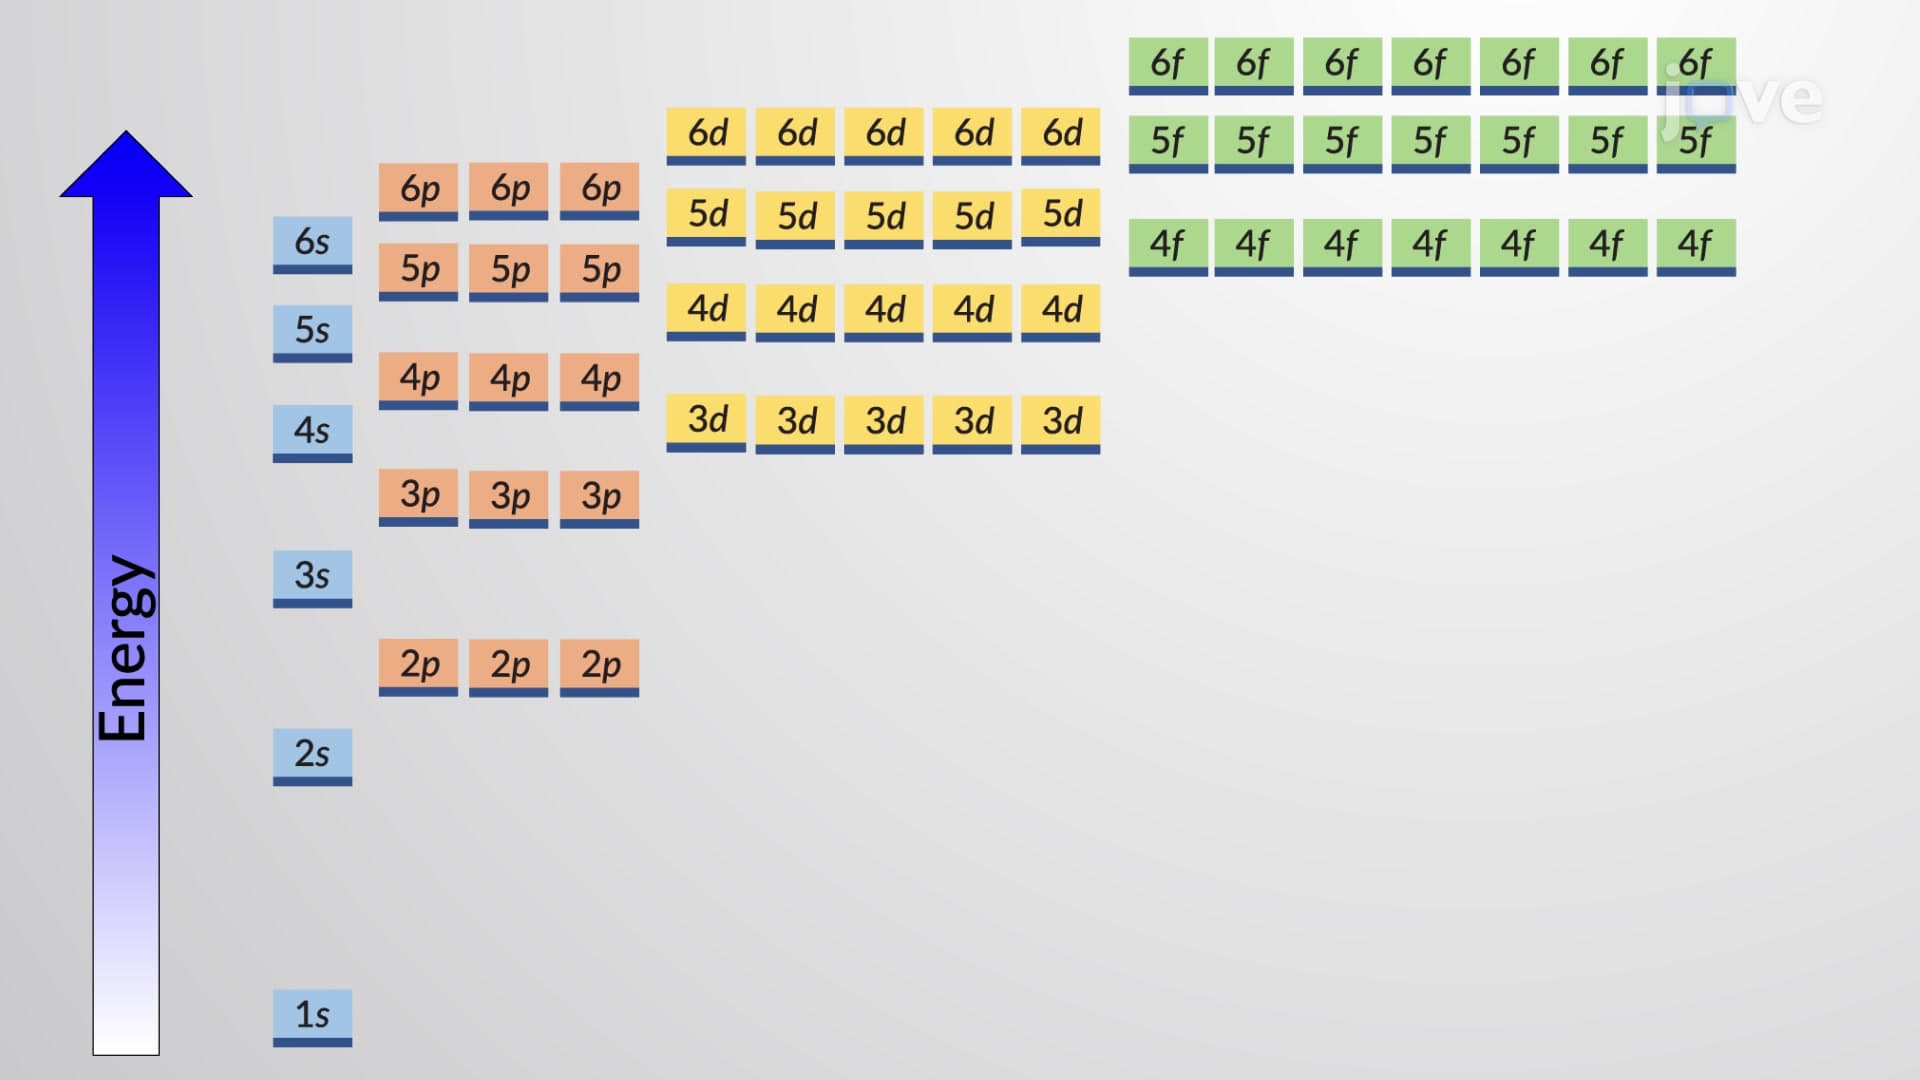
Electron Configuration of Multielectron Atoms
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...