Related Experiment Video
Updated: Nov 24, 2025
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
The effective isotropy of the hydrogen local potential in biphenyl and other hydrocarbons
Pierfrancesco Ulpiani1, Giovanni Romanelli2, Dalila Onorati3
1Università degli Studi di Roma "Tor Vergata," Dipartimento di Scienze e Tecnologie Chimiche, Via della Ricerca Scientifica 1, Rome 00133, Italy.

Abstract:
We present an experimental investigation of the hydrogen nuclear momentum distribution in biphenyl using deep inelastic neutron scattering. Our experimental results suggest that the local potential affecting hydrogen is both harmonic and isotropic within experimental uncertainties. This feature is interpreted as a consequence of the central limit theorem, whereby the three-dimensional momentum distribution is expected to become a purely Gaussian function as the number of independent vibrational modes in a system increases. We also performed ab initio phonon calculations on biphenyl and other saturated hydrocarbons, from methane to decane. From the results of the simulations, one can observe that the nuclear momentum distribution becomes more isotropic as the number of atoms and normal modes in the molecule increases. Moreover, the predicted theoretical anisotropy in biphenyl is clearly larger than in the experiment. The reason is that the total number of normal modes necessary to reproduce the experimental results is much larger than the number of normal modes encompassed by a single unit cell due to the presence of structural disorder and intermolecular interactions in the real crystal, as well as coupling of different normal modes. Finally, experimental data were collected, over a subset of detectors on the VESUVIO spectrometer at ISIS, with a novel setup to increase the count rate and signal-to-background ratio. We envision that such an optimized experimental setup can provide faster measurements and more stringent constraints for phonon calculations.
Related Concept Videos
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:03
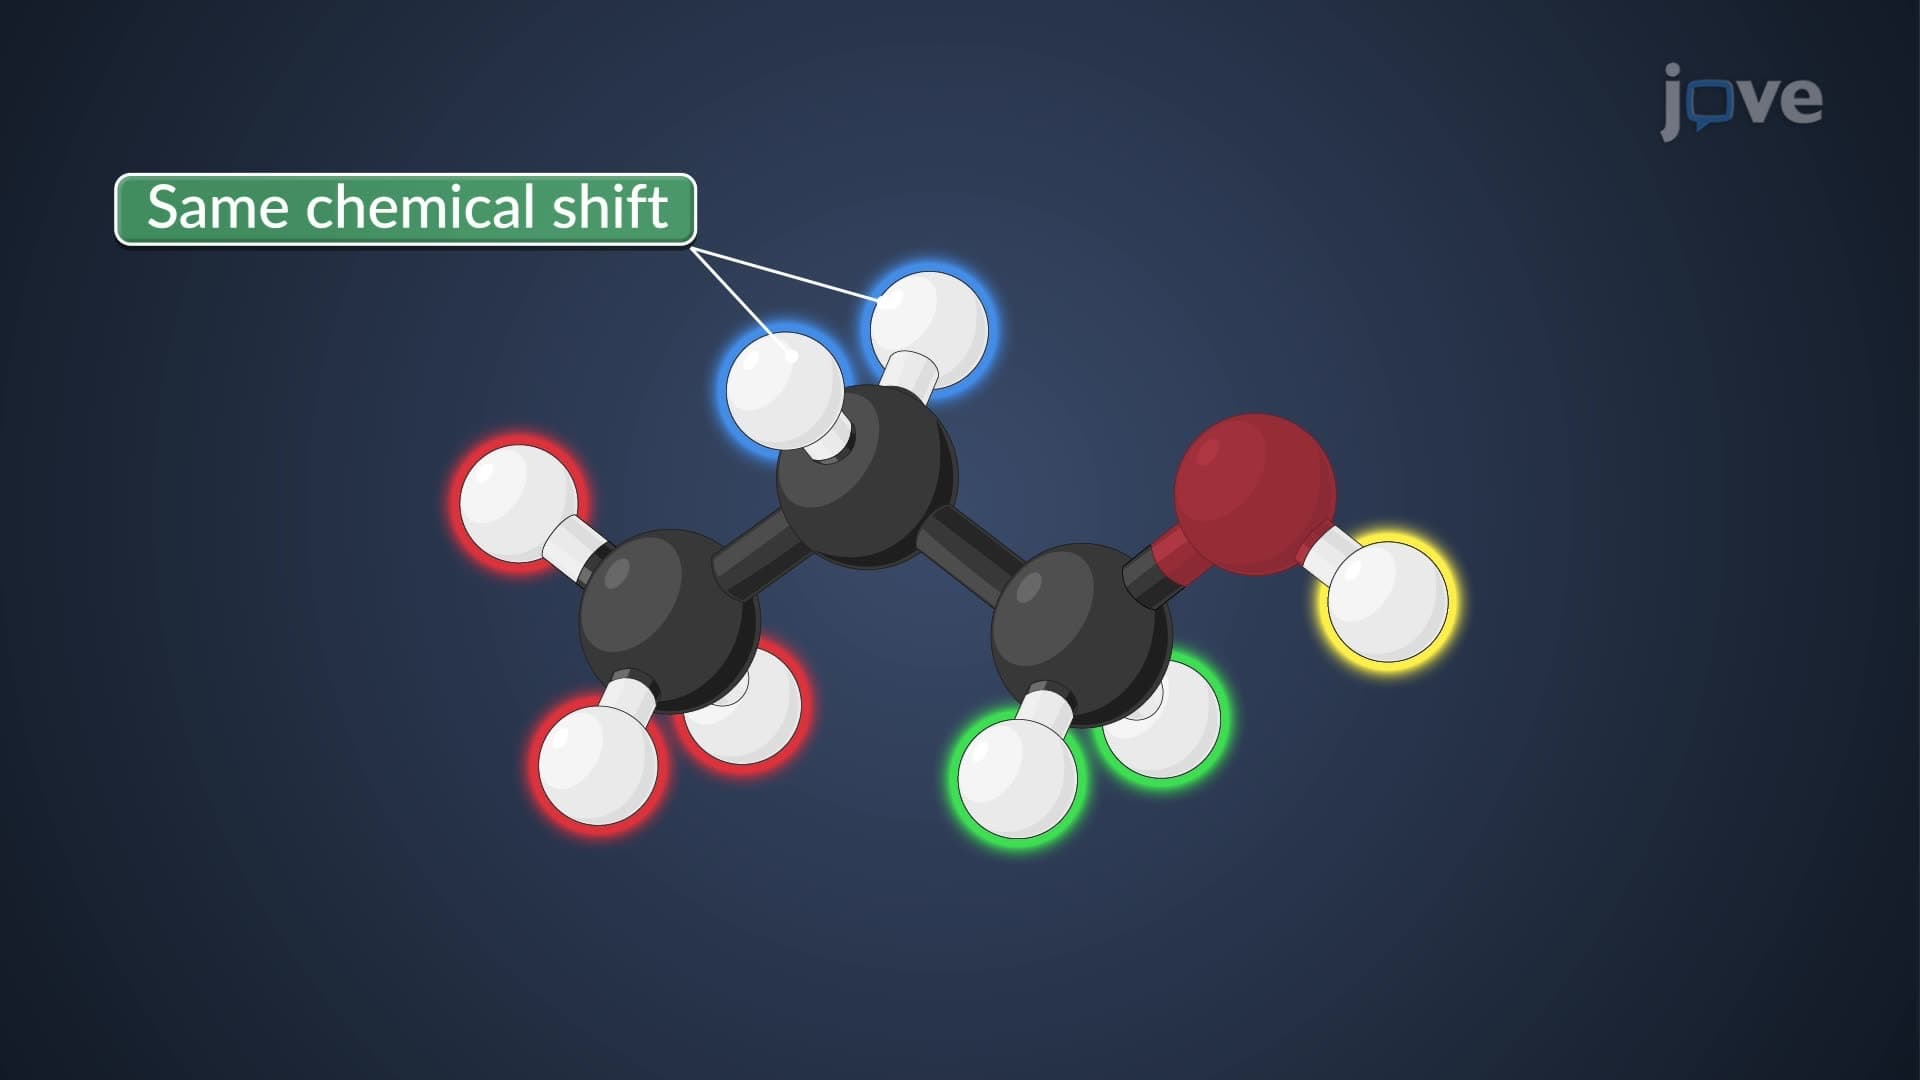
¹H NMR Chemical Shift Equivalence: Homotopic and Heterotopic Protons
 03:35
Hybridization of Atomic Orbitals II
 01:27
π Electron Effects on Chemical Shift: Overview
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 01:18
Structure of Benzene: Molecular Orbital Model