Related Experiment Video
Updated: Sep 22, 2025
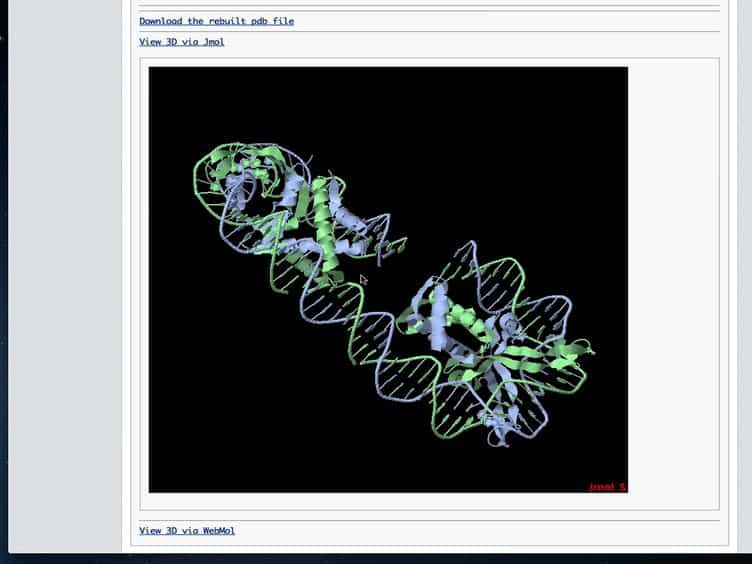

Analyzing and Building Nucleic Acid Structures with 3DNA
Published on: April 26, 2013
Understanding the role of non-Watson-Crick base pairs in DNA-protein recognition: Structural and energetic aspects
Soumi Das1, Siddhartha Roy1, Dhananjay Bhattacharyya2
1Department of Biophysics, Bose Institute, Kolkata, India.

Abstract:
Specific recognition of DNA base sequences by proteins is vital for life-cycles of all organisms. In a large number of crystal structures of protein-DNA complexes, DNA conformation significantly deviates from the canonical B-DNA structure. A key question is whether such alternate conformations exist prior to protein binding and one is selected for complexation or the structure observed is induced by protein binding. Non-canonical base pairs, such as Hoogsteen base pairs, are often observed in crystal structures of protein-DNA complexes. We decided to explore whether the occurrence of such non-canonical base pairs in protein-DNA complexes is induced by the protein or is selected from pre-existing conformations. Detailed quantum chemical calculations with dispersion-corrected density functional theory (DFT-D) indicated that most of the non-canonical base pairs with DNA bases are stable even in the absence of the interacting amino acids. However, the G:G Hoogsteen base pair, which also appears in the telomere structure, appears to be unstable in the absence of other stabilizing agents, such as positively charged amino acids. Thus, the stability of many of the non-canonical base pair containing duplexes may be close to the canonical B-DNA structure and hence energetically accessible in the ground state; suggesting that the selection from pre-existing conformations may be an important mechanism for observed non-canonical base pairs in protein-DNA complexes.

Related Concept Videos
 02:05
DNA as a Genetic Template
 02:27
DNA Base Pairing
 01:07
The DNA Helix
 01:24
Protein Organization
The primary structure of a protein is its amino acid sequence....
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:04
Protein-protein Interfaces