Related Experiment Video
Updated: Aug 16, 2025


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
A truncated Davidson method for the efficient "chemically accurate" calculation of full configuration interaction
1Quantum Artificial Intelligence Laboratory (QuAIL), NASA Ames Research Center, Moffett Field, California 94035, USA and Employed By KBR, 601 Jefferson St., Houston, Texas 77002, USA.

Abstract:
This work develops and illustrates a new method of calculating "chemically accurate" electronic wavefunctions (and energies) via a truncated full configuration interaction (CI) procedure, which arguably circumvents the large matrix diagonalization that is the core problem of full CI and is also central to modern selective CI approaches. This is accomplished simply by following the standard/ubiquitous Davidson method in its "direct" form-wherein, in each iteration, the electronic Hamiltonian operator is applied directly in second quantization to the Ritz vector/wavefunction from the prior iteration-except that (in this work) only a small portion of the resultant expansion vector is actually even computed (through the application of only a similarly small portion of the Hamiltonian). Specifically, at each iteration of this truncated Davidson approach, the new expansion vector is taken to be twice as large as that from the prior iteration. In this manner, a small set of highly truncated expansion vectors (say 10-30) of increasing precision is incrementally constructed, forming a small subspace within which diagonalization of the Hamiltonian yields clear, consistent, and monotonically variational convergence to the approximate full CI limit. The good efficiency in which convergence to the level of chemical accuracy (1.6 mhartree) is achieved suggests, at least for the demonstrated problem sizes-Hilbert spaces of 1018 and wavefunctions of 108 determinants-that this truncated Davidson methodology can serve as a replacement of standard CI and complete-active space approaches in circumstances where only a few chemically significant digits of accuracy are required and/or meaningful in view of ever-present basis set limitations.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 05:00
Author Spotlight: Streamlining Visual Dynamics to Simplify Molecular Dynamics Simulations Using Gromacs
Published on: August 9, 2024
Related Concept Videos
 01:13
Graphing the Wave Function
 01:24
Fast Decoupled and DC Powerflow
 01:24
Electromagnetic Wave Equation
However, although electric and magnetic fields were first introduced as mathematical constructs to simplify the description of mutual forces between charges, a natural question emerges from Maxwell's equations:...
 03:35
Hybridization of Atomic Orbitals II
 01:10
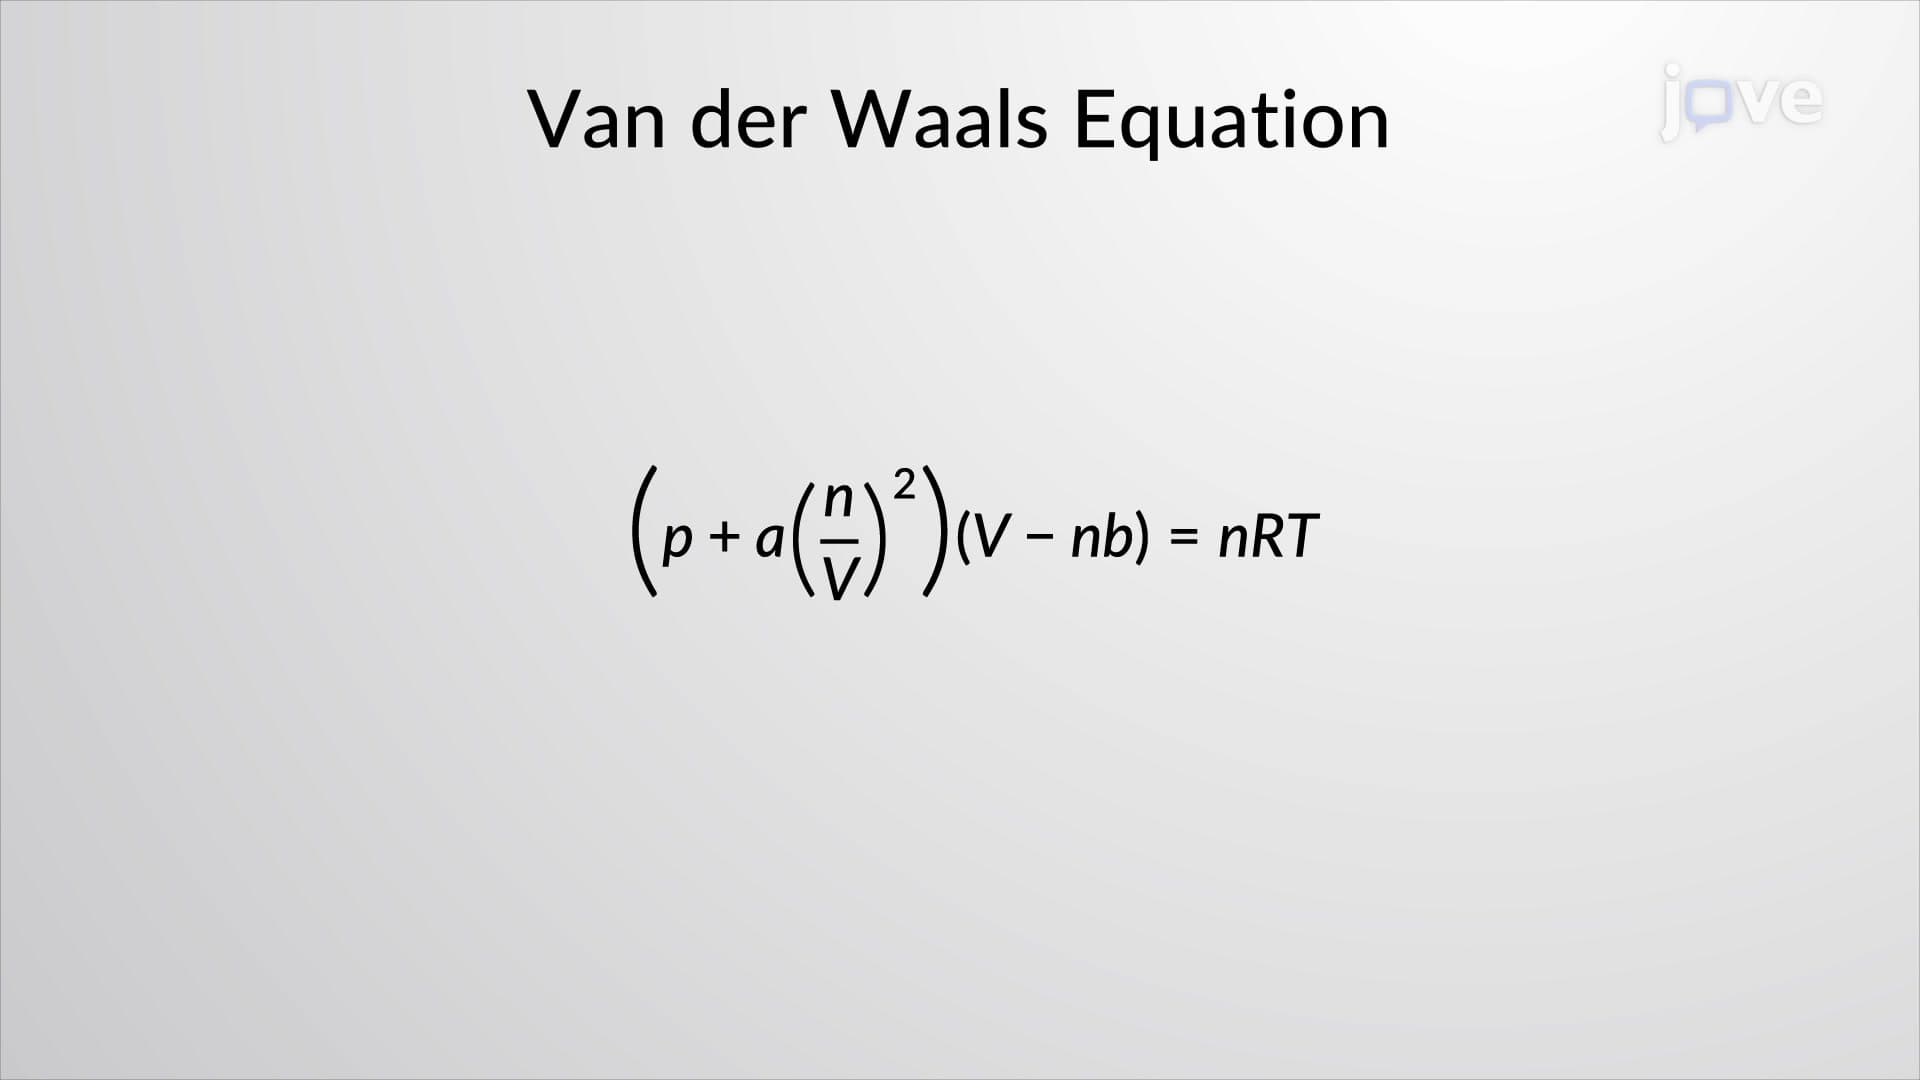
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 01:24
One-Compartment Open Model: Wagner-Nelson and Loo Riegelman Method for ka Estimation
On...