Anisotropic Interlayer Force Field for Group-VI Transition Metal Dichalcogenides
Related Experiment Video
Updated: Jul 11, 2025
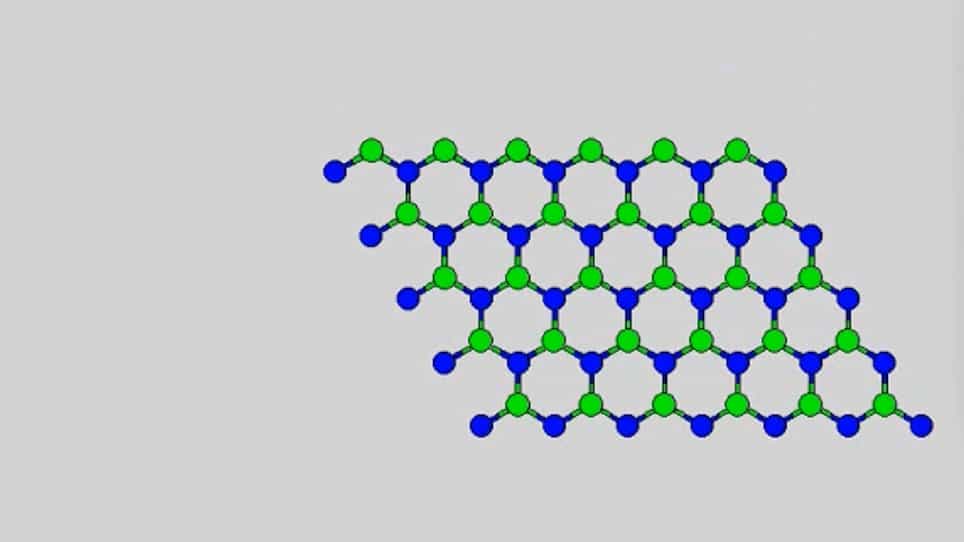
Probe Type II Band Alignment in One-Dimensional Van Der Waals Heterostructures Using First-Principles Calculations
Published on: October 12, 2019
Wenwu Jiang1, Reut Sofer2, Xiang Gao2
1Department of Engineering Mechanics, School of Civil Engineering, Wuhan University, Wuhan, Hubei 430072, China.

Abstract:
An anisotropic interlayer force field that describes the interlayer interactions in homogeneous and heterogeneous interfaces of group-VI transition metal dichalcogenides (MX2, where M = Mo, W, and X = S, Se) is presented. The force field is benchmarked against density functional theory calculations for bilayer systems within the Heyd-Scuseria-Ernzerhof hybrid density functional approximation, augmented by a nonlocal many-body dispersion treatment of long-range correlation. The parametrization yields good agreement with the reference calculations of binding energy curves and sliding potential energy surfaces. It is found to be transferable to transition metal dichalcogenide (TMD) junctions outside of the training set that contain the same atom types. Calculated bulk moduli agree with most previous dispersion-corrected density functional theory predictions, which underestimate the available experimental values. Calculated phonon spectra of the various junctions under consideration demonstrate the importance of appropriately treating the anisotropic nature of the layered interfaces. Considering our previous parametrization for MoS2, the anisotropic interlayer potential enables accurate and efficient large-scale simulations of the dynamical, tribological, and thermal transport properties of a large set of homogeneous and heterogeneous TMD interfaces.
More Related Videos
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability....
 01:27
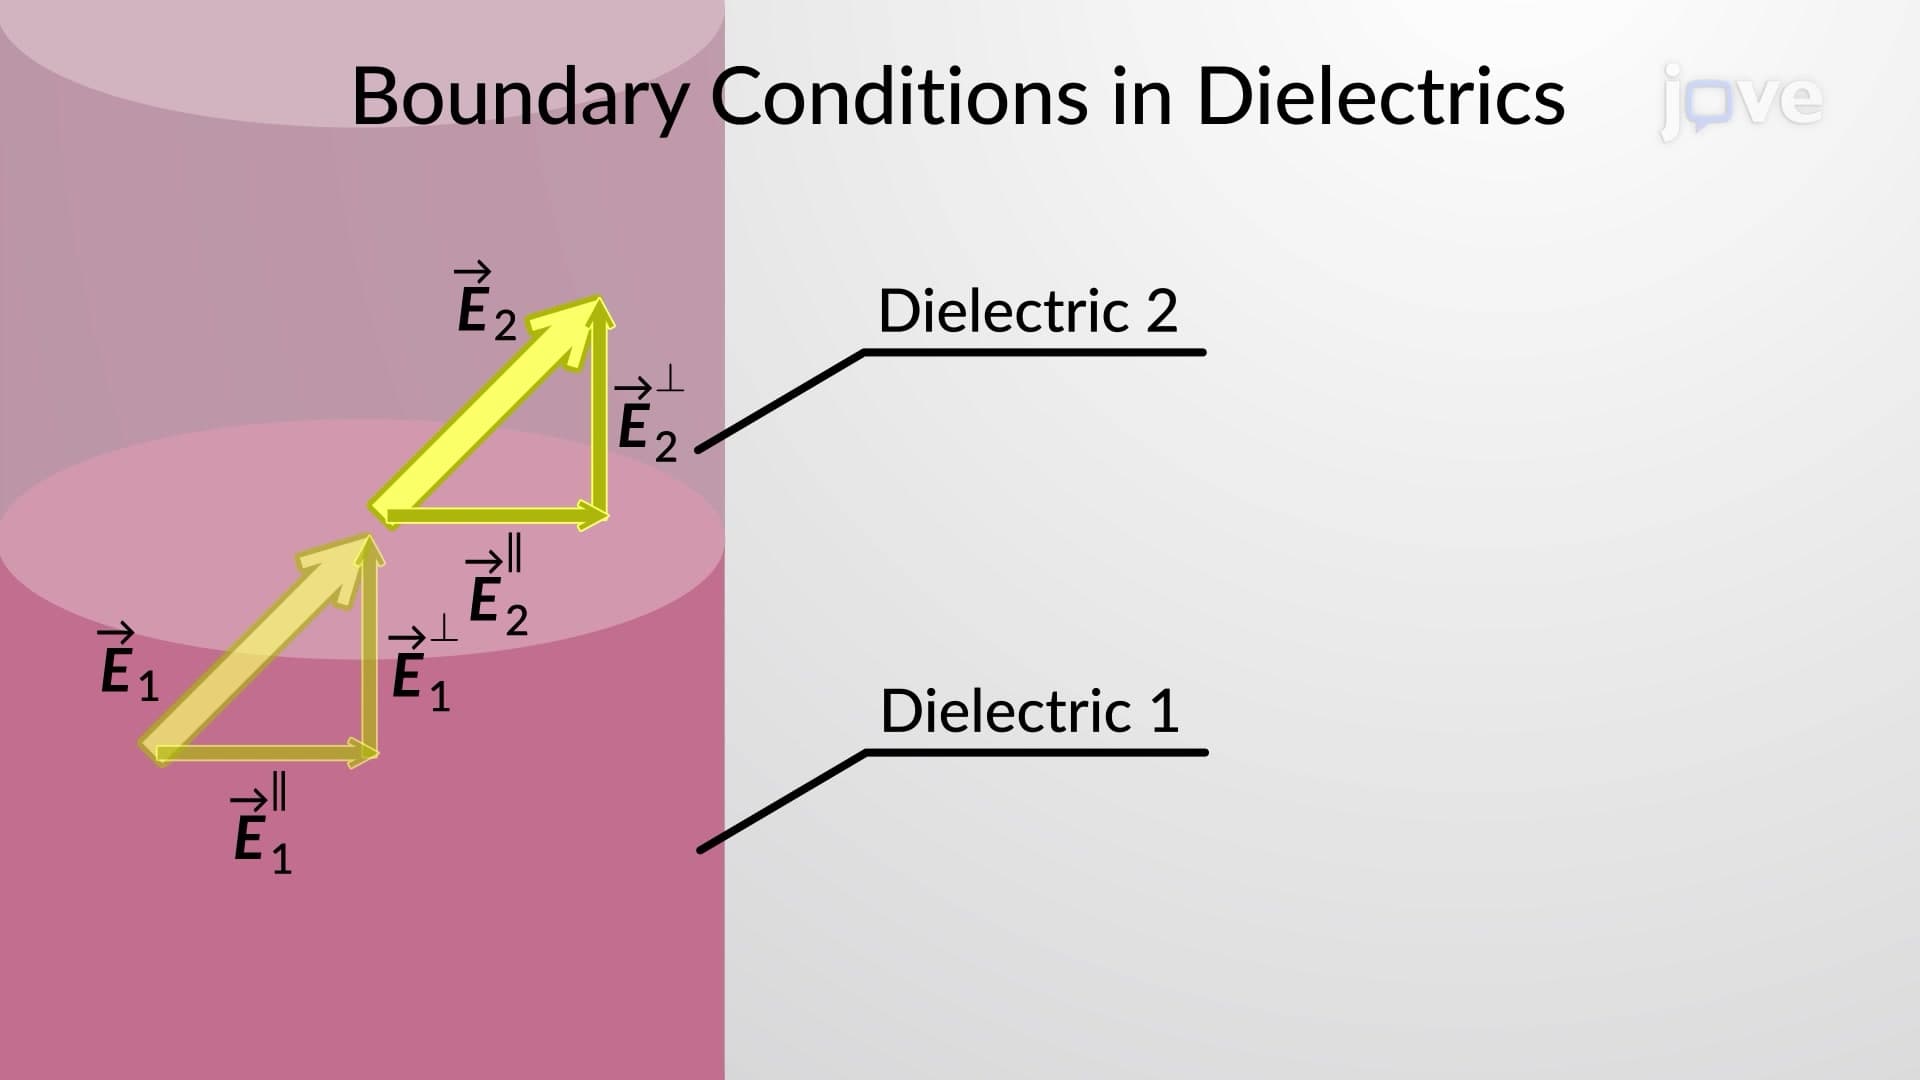
Electrostatic Boundary Conditions in Dielectrics
Consider a case where both the mediums across a boundary are two different dielectric materials. Recall that the electric field and electric displacement are proportional and related through the material's...
 01:24
Van der Waals Interactions
 02:42
Valence Bond Theory