Related Experiment Video
Updated: Jul 10, 2025
Combining Solid-state and Solution-based Techniques: Synthesis and Reactivity of ChalcogenidoplumbatesII or IV
Published on: December 29, 2016
Computational Insight into the Nature and Strength of the π-Hole Type Chalcogen∙∙∙Chalcogen Interactions in the
Fengying Lei1, Qingyu Liu1, Yeshuang Zhong1
1School of Basic Medical Sciences/School of Biology and Engineering, Guizhou Medical University, Guiyang 550025, China.

Abstract:
In recent years, the non-covalent interactions between chalcogen centers have aroused substantial research interest because of their potential applications in organocatalysis, materials science, drug design, biological systems, crystal engineering, and molecular recognition. However, studies on π-hole-type chalcogen∙∙∙chalcogen interactions are scarcely reported in the literature. Herein, the π-hole-type intermolecular chalcogen∙∙∙chalcogen interactions in the model complexes formed between XO2 (X = S, Se, Te) and CH3YCH3 (Y = O, S, Se, Te) were systematically studied by using quantum chemical computations. The model complexes are stabilized via one primary X∙∙∙Y chalcogen bond (ChB) and the secondary C-H∙∙∙O hydrogen bonds. The binding energies of the studied complexes are in the range of -21.6~-60.4 kJ/mol. The X∙∙∙Y distances are significantly smaller than the sum of the van der Waals radii of the corresponding two atoms. The X∙∙∙Y ChBs in all the studied complexes except for the SO2∙∙∙CH3OCH3 complex are strong in strength and display a partial covalent character revealed by conducting the quantum theory of atoms in molecules (QTAIM), a non-covalent interaction plot (NCIplot), and natural bond orbital (NBO) analyses. The symmetry-adapted perturbation theory (SAPT) analysis discloses that the X∙∙∙Y ChBs are primarily dominated by the electrostatic component.
More Related Videos
 06:44
From Molecules to Materials: Engineering New Ionic Liquid Crystals Through Halogen Bonding
Published on: March 24, 2018
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Related Concept Videos
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:42
Valence Bond Theory
 02:27
Predicting Molecular Geometry
 04:01
VSEPR Theory and the Effect of Lone Pairs
 01:18
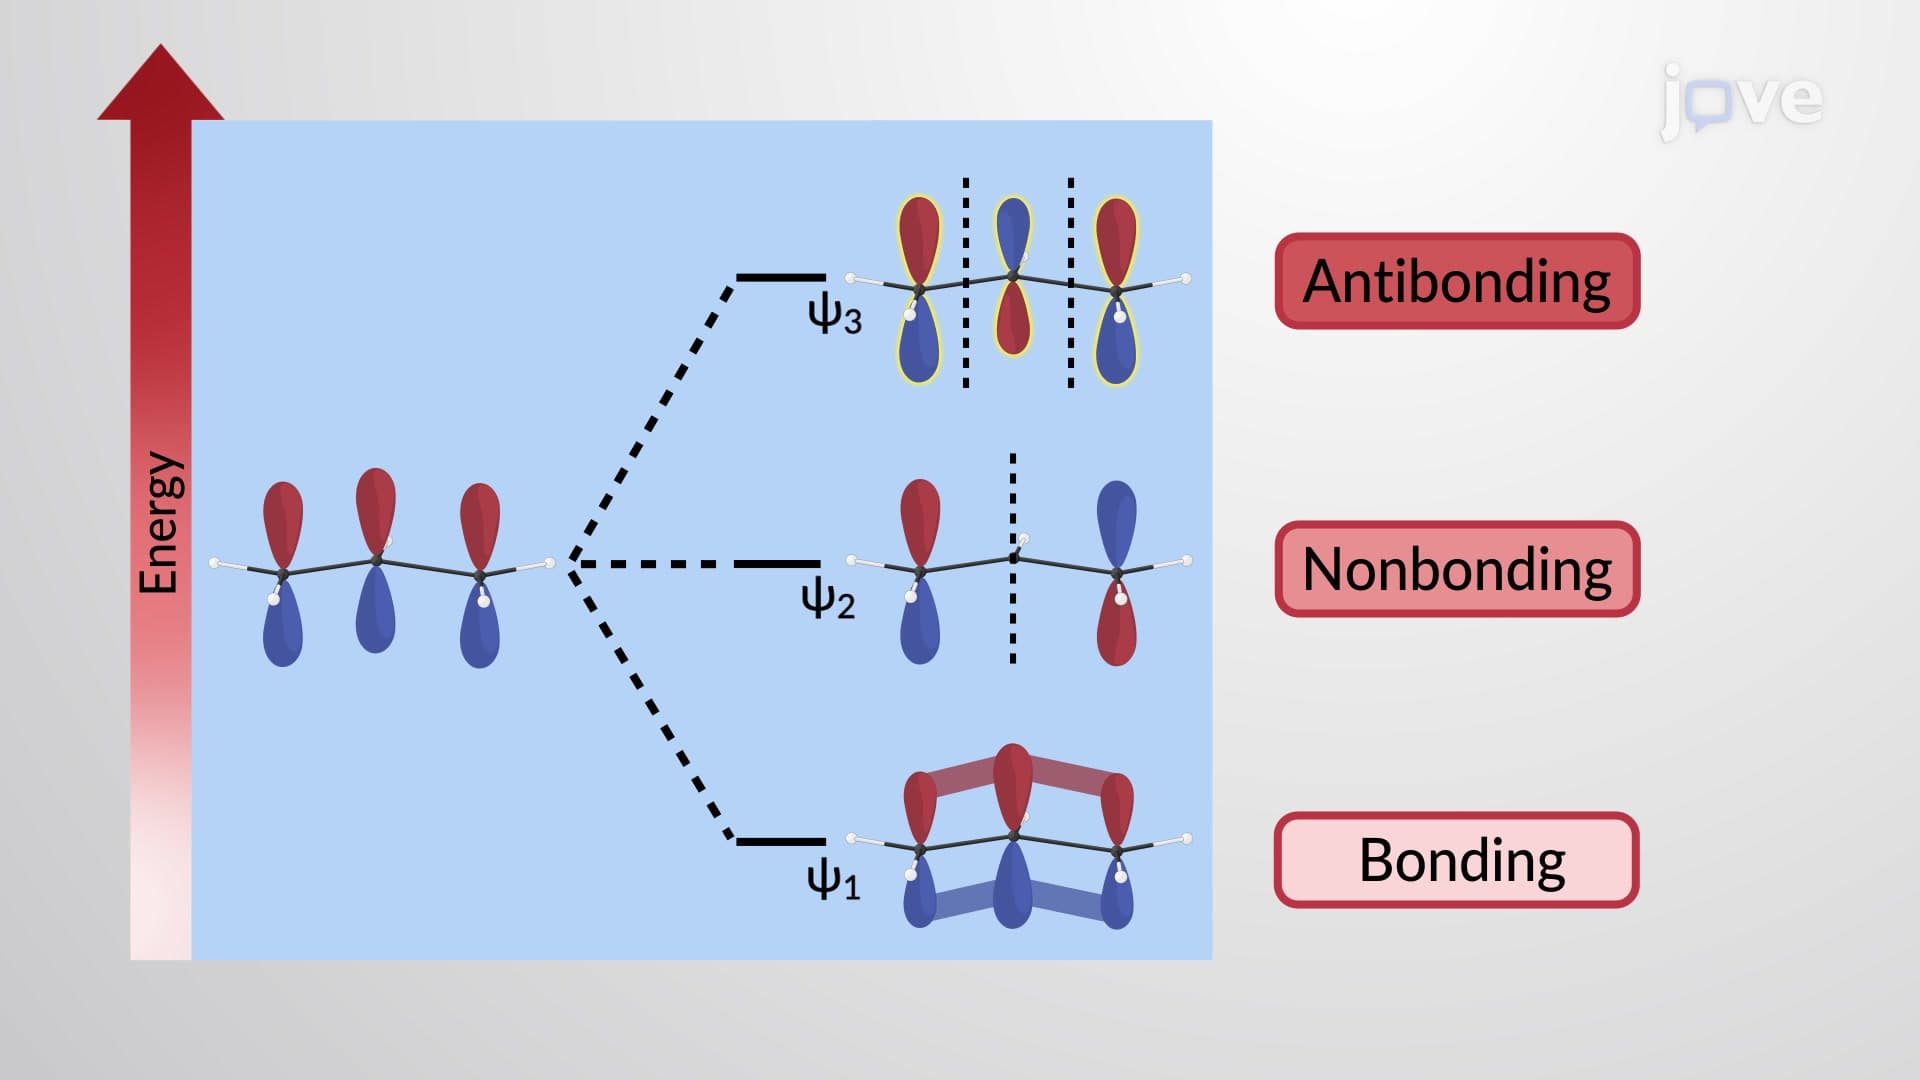
π Molecular Orbitals of the Allyl Cation and Anion