Related Experiment Video
Updated: Jul 7, 2025

Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
The CN(X 2Σ+) + C2H6 reaction: Dynamics study based on an analytical full-dimensional potential energy surface
Joaquin Espinosa-Garcia1, Cipriano Rangel1
1Área de Química Física and Instituto de Computación Científica Avanzada, Universidad de Extremadura, 06071 Badajoz, Spain.

Abstract:
The hydrogen abstraction reaction of the cyano radical with molecules of ethane presents some interesting points in the chemistry from ultra-cold to combustion environments especially with regard to HCN(v) product vibrational distribution. In order to understand its dynamics, a new analytical full-dimensional potential energy surface was developed, named PES-2023. It uses a combination of valence bond and mechanic molecular terms as the functional form, fitted to high-level ab initio calculations at the explicitly correlated CCSD(T)-F12/aug-cc-pVTZ level on a reduced and selected number of points describing the reactive process. The new surface showed a continuous and smooth behavior, describing reasonably the topology of the reaction: high exothermicity, low barrier, and presence of intermediate complexes in the entrance and exit channels. Using quasi-classical trajectory calculations (QCT) on the new PES-2023, a dynamics study was performed at room temperature with special emphasis on the HCN(v1,v2,v3) product stretching and bending vibrational excitations, and the results were compared with the experimental evidence, which presented discrepancies in the bending excitation. The available energy was mostly deposited as HCN(v) vibrational energy with the vibrational population inverted in the CH stretching mode and not inverted in the CN stretching and bending modes, thus simulating the experimental evidence. Other dynamics properties at room temperature were also analyzed; cold rotational energy distribution was found, associated with a linear and soft transition state, and backward scattering distribution was found, associated with a rebound mechanism.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:13
Force and Potential Energy in One Dimension
 01:23
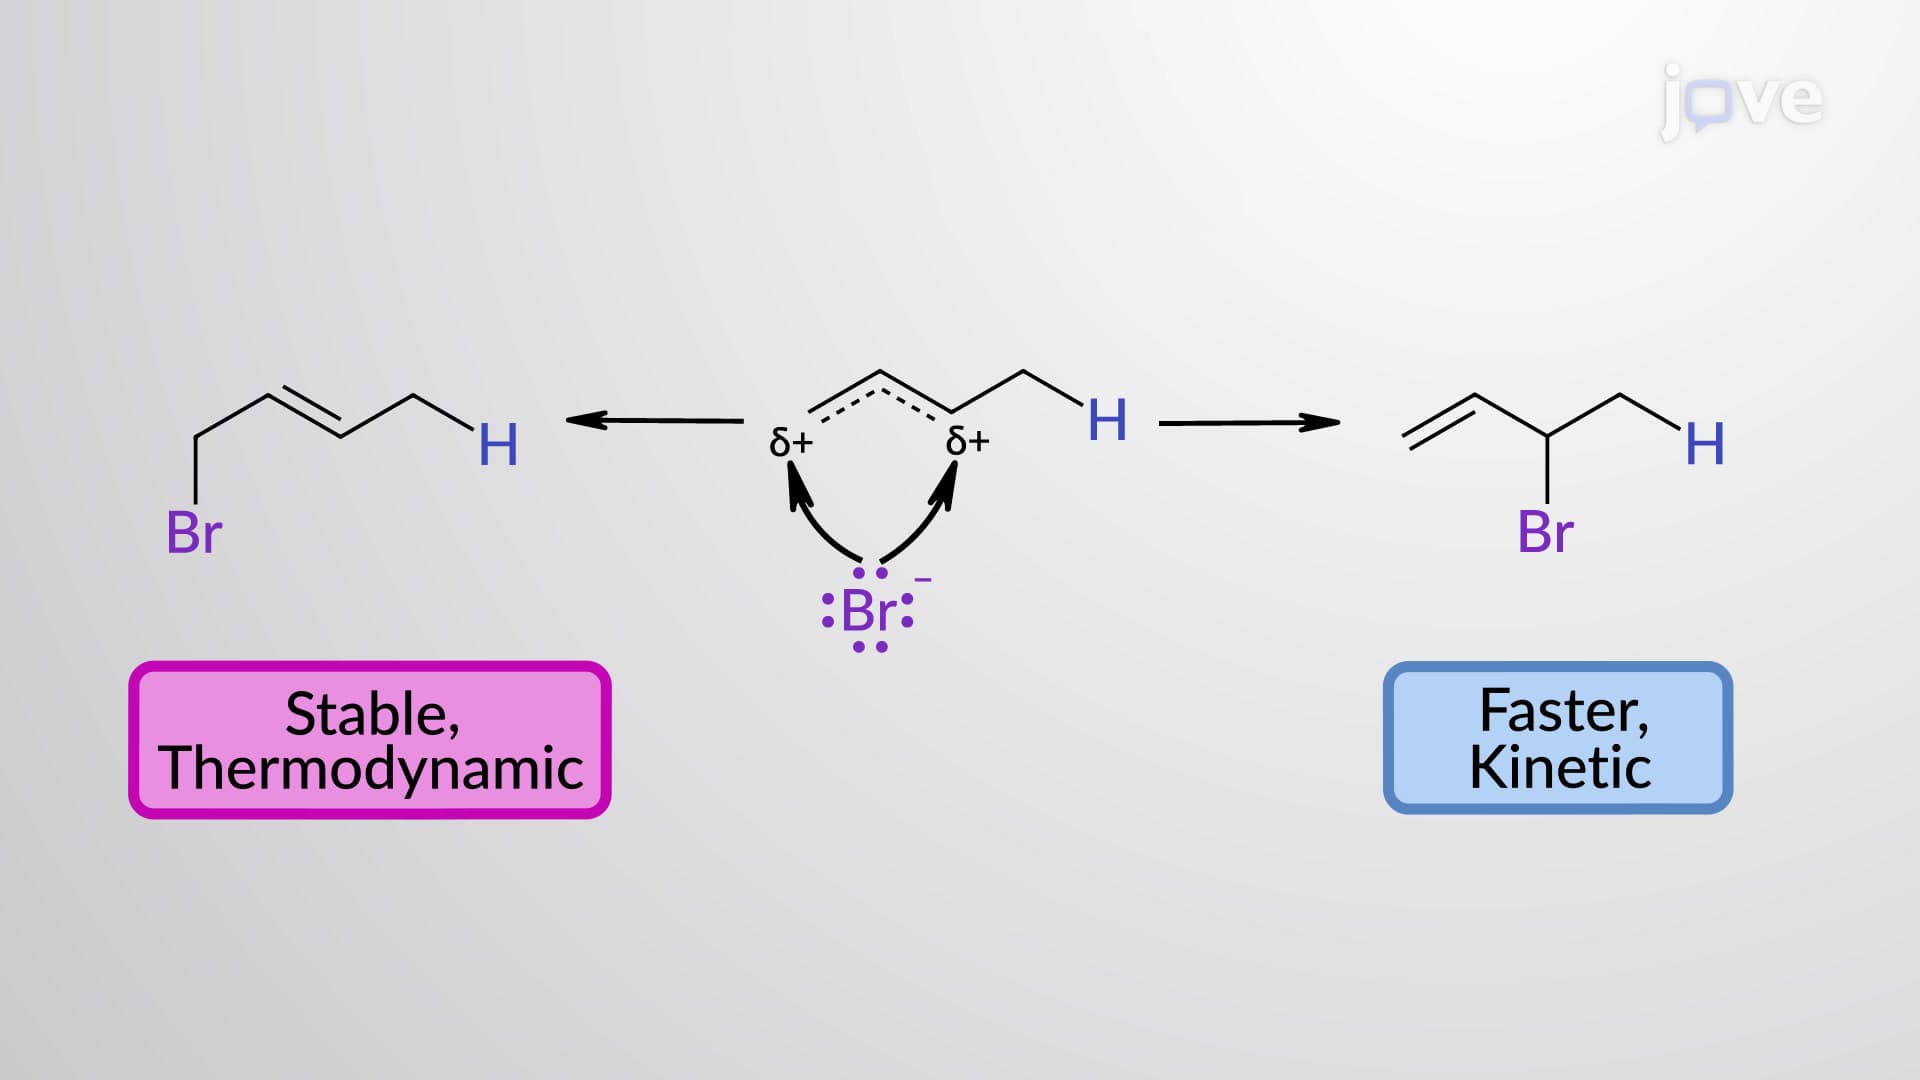
Electrophilic Addition of HX to 1,3-Butadiene: Thermodynamic vs Kinetic Control
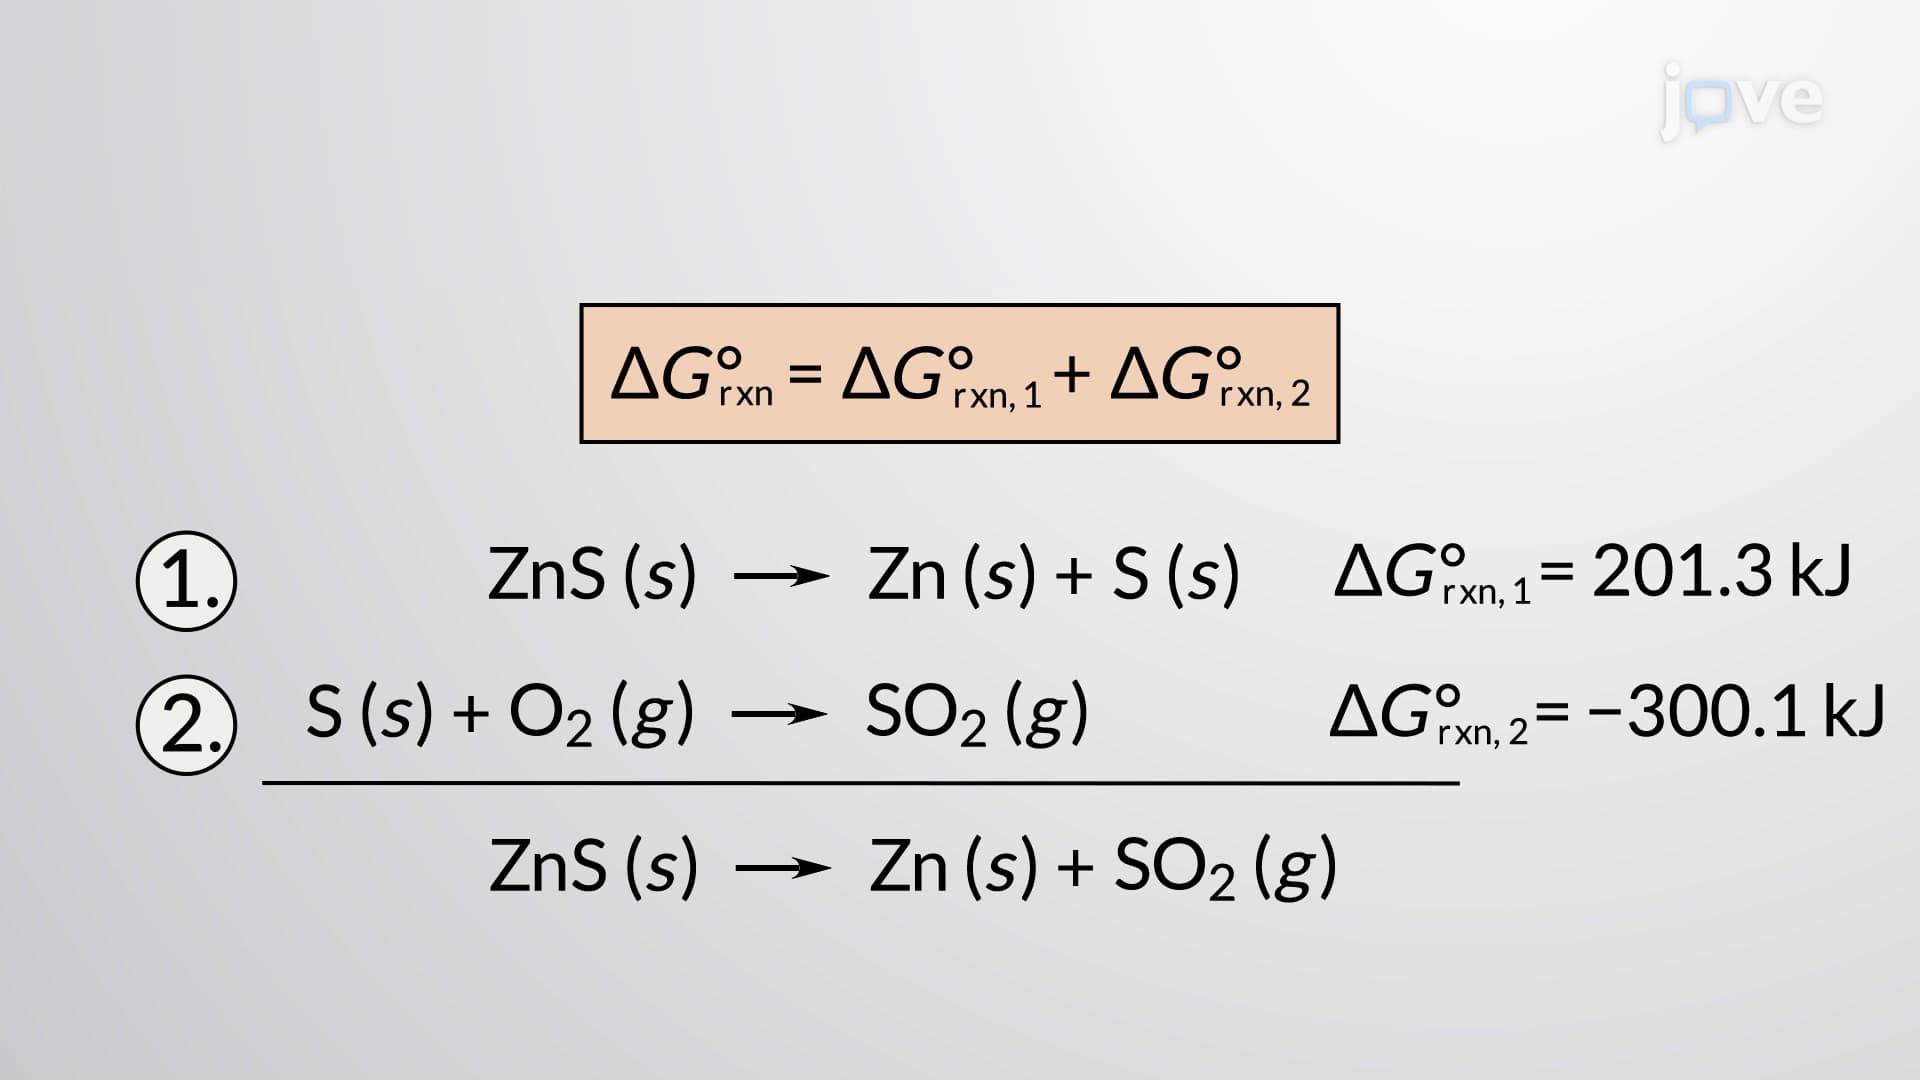 02:49
Calculating Standard Free Energy Changes
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...
 01:21
[3,3] Sigmatropic Rearrangement of 1,5-Dienes: Cope Rearrangement