Related Experiment Video
Updated: Sep 16, 2025

Author Spotlight: A Computational Approach to Decipher Amino Acid Preferences in Multispecific Protein-Protein Interactions
Published on: January 26, 2024
Human protein interactome structure prediction at scale with Boltz-2
Alexander M Ille1,2, Christopher Markosian1,2, Stephen K Burley3,4,5,6,7
1Rutgers Cancer Institute, Newark, NJ, USA.

Abstract:
In humans, protein-protein interactions mediate numerous biological processes and are central to both normal physiology and disease. Extensive research efforts have aimed to elucidate the human protein interactome, and comprehensive databases now catalog these interactions at scale. However, structural coverage of the human protein interactome is limited and remains challenging to resolve through experimental methodology alone. Recent advances in artificial intelligence/machine learning (AI/ML)-based approaches for protein interaction structure prediction present opportunities for large-scale structural characterization of the human interactome. One such model, Boltz-2, which is capable of predicting the structures of protein complexes, may serve this objective. Here, we present de novo computed models of 1,394 binary human protein interaction structures predicted using Boltz-2 based on biochemically determined interaction data sourced from the IntAct database. We assessed the predicted interaction structures through different confidence metrics, which consider both overall structure and the interaction interface. These analyses indicated that prediction confidence tended to be greater for smaller complexes, while increased multiple sequence alignment (MSA) depth tended to improve prediction confidence. Additionally, we examined annotated protein domains and found that 679 of the predicted structural complexes contained a variety of domains with putative interaction involvement on the basis of interaction interface proximity. Furthermore, our analyses revealed intricate interaction networks within the context of biological function and cancer. This work demonstrates the utility of Boltz-2 for in silico structural modeling of the human protein interactome, highlighting both strengths and limitations, while also providing a novel view of broad functional contextualization. Ultimately, such modeling is expected to yield broad structural insights with relevance across multiple domains of biomedical research.


Related Concept Videos
 02:04
Protein-protein Interfaces
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 01:57
Protein Complexes with Interchangeable Parts
The SCF ubiquitin ligase is a protein complex of five individual proteins. This complex attaches ubiquitin to other target proteins to mark them for degradation. In order...
 01:24
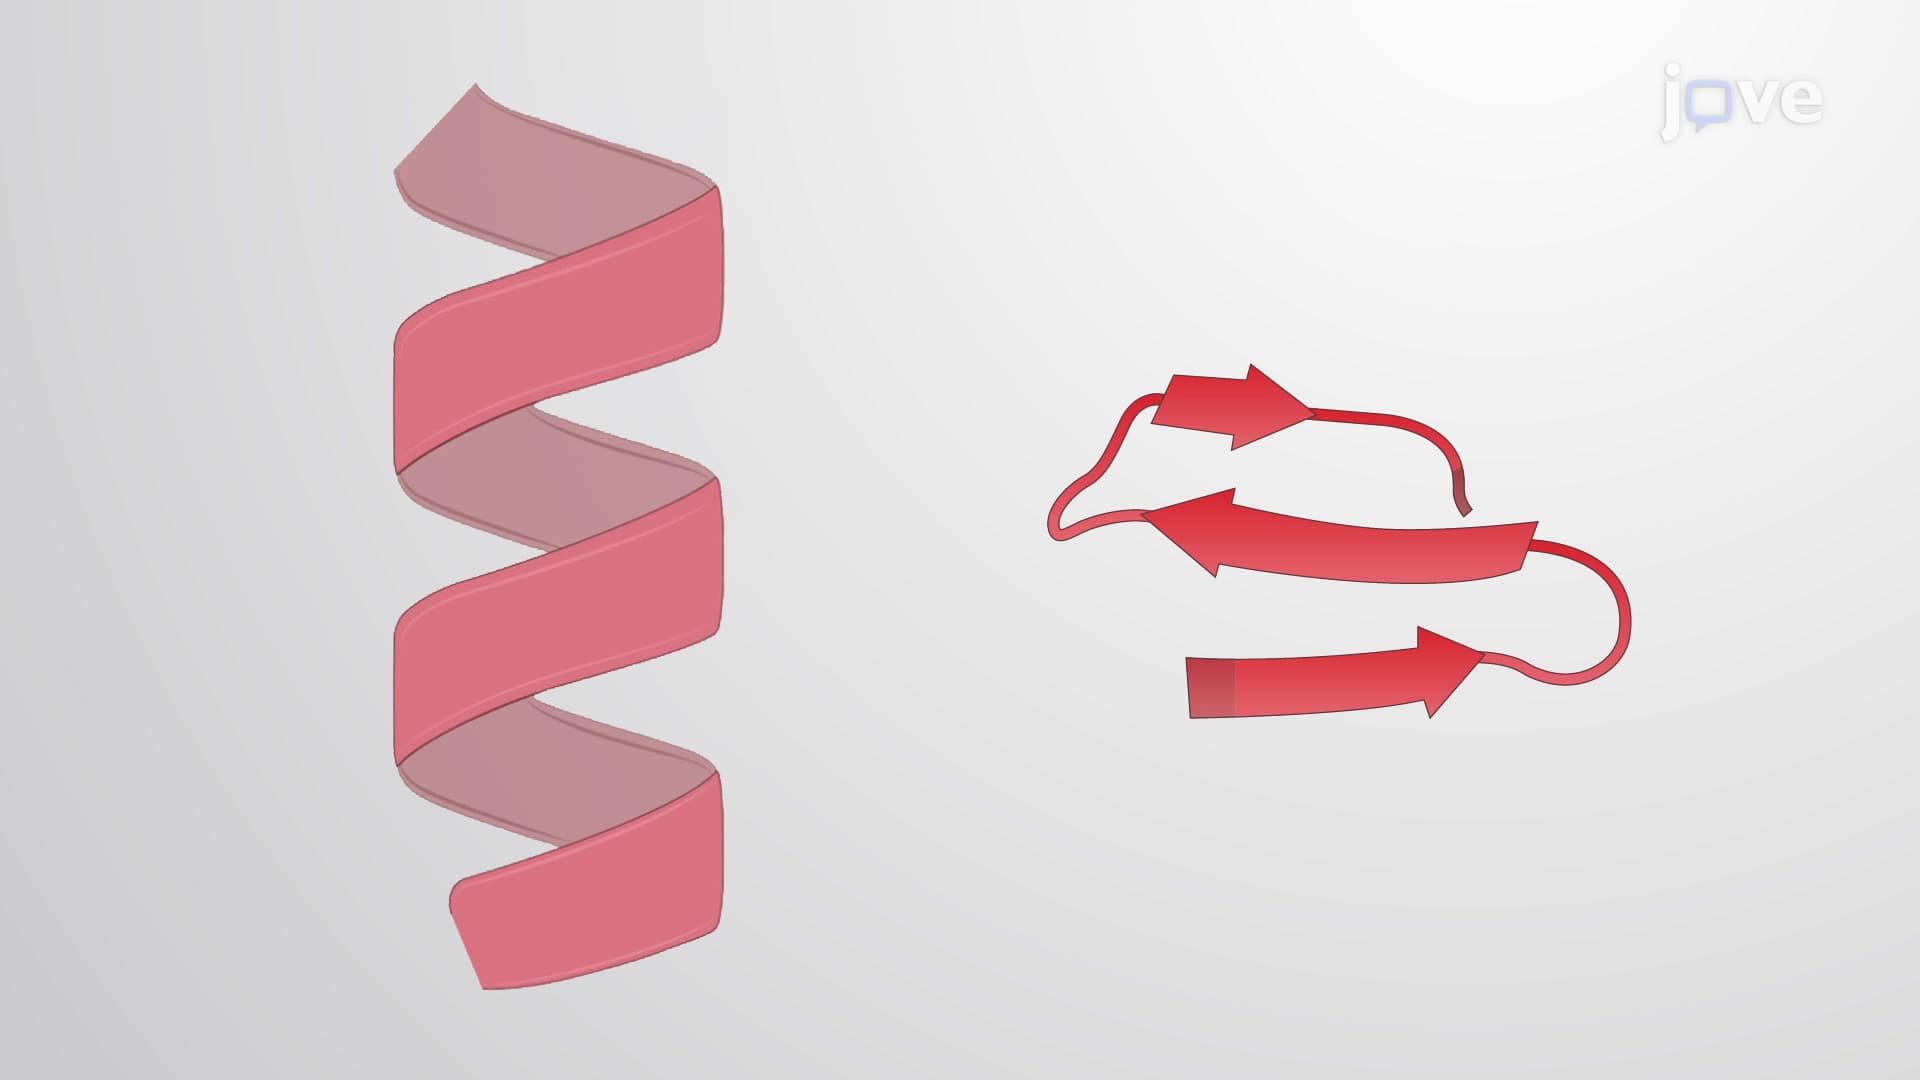
Protein Organization
The primary structure of a protein is its amino acid sequence....
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...